Q.Would you expect benzaldehyde to be more reactive or less reactive in nucleophilic addition reactions than propanal? Explain your answer.
Concept understanding — Resonance Stabilization Effect
Resonance Stabilization Effect
Imagine you're holding a rubber band stretched between two fingers. The moment you let go, it snaps back to its relaxed shape. That relaxed shape is the lowest-energy state — the most stable one. Now think about a molecule that can't decide which single structure it "wants" to be in. It's like the rubber band being pulled in two different directions at once, but instead of snapping, it finds a middle ground that is more stable than either extreme.
That middle ground is resonance stabilization.
The Intuition: Why "Delocalization" Lowers Energy
In chemistry, electrons (especially π electrons and lone pairs) like to be spread out. When an electron is confined to a small space between two atoms, it has high energy — like a child bouncing off the walls of a tiny room. But if you give that electron more space to move — delocalize it over several atoms — its energy drops. The system becomes more stable.
Resonance is the formal way we describe this delocalization. We draw multiple Lewis structures (called resonance contributors or canonical forms) that differ only in the arrangement of π electrons and lone pairs. The real molecule is not any one of these structures — it is a hybrid of all of them, with electron density spread out.
The key point: resonance structures are not real. They are imaginary snapshots. The real molecule is the resonance hybrid, which has lower energy than any single contributor would predict.
The Precise Statement
Resonance stabilization is the extra stability a molecule gains because its electrons are delocalized over multiple atoms via conjugation (alternating single and multiple bonds) or through the involvement of lone pairs or empty orbitals. This stabilization energy is the difference between the actual energy of the molecule and the energy of the most stable resonance contributor (if it existed alone).
ΔEresonance=Emost stable contributor−Eactual molecule
This ΔE is always positive — the actual molecule is always more stable (lower in energy) than any single contributor.
A Concrete Example: The Carbonate Ion (CO32−)
Draw the carbonate ion. You'll find three equivalent Lewis structures, each with one C=O double bond and two C–O⁻ single bonds. The double bond can be placed on any of the three oxygen atoms.
- If the molecule were truly one of these structures, the C–O bond lengths would be different (one short double, two long singles).
- But experiment shows all three C–O bonds are identical — exactly 1.28 Å, intermediate between a single and double bond.
- The negative charge is not on any one oxygen; it is delocalized equally over all three oxygens.
The resonance hybrid looks like this: each C–O bond has a bond order of 131, and each oxygen carries a partial negative charge of −32. The molecule is about 150 kJ/mol more stable than any single contributor.
When resonance contributors are equivalent (same energy), the stabilization is largest. When they are unequal (one is much more stable than others), the hybrid resembles the most stable contributor, and the stabilization is smaller.
How to Recognize Resonance Stabilization
Look for these features in a molecule:
- Conjugated π systems — alternating single and double bonds (e.g., 1,3-butadiene)
- Lone pairs adjacent to π bonds (e.g., the oxygen in an ester, or the nitrogen in an amide)
- Empty p orbitals adjacent to π bonds (e.g., carbocations, carbonyl groups)
- Atoms with π bonds and adjacent charges (e.g., allyl anion, allyl cation)
Resonance does not involve the movement of σ bonds or atoms. Only π electrons and lone pairs (in p orbitals) are delocalized. The positions of all atoms remain fixed.
Why It Matters for Exams
Resonance stabilization explains:
- Why carboxylic acids are more acidic than alcohols — the conjugate base (carboxylate) has resonance, spreading the negative charge over two oxygens, making it more stable.
- Why amides are planar and have restricted rotation — the C–N bond has partial double-bond character due to resonance.
- Why benzene is unusually stable — its six π electrons are delocalized over the ring, giving it a resonance stabilization energy of about 150 kJ/mol.
- Why allylic carbocations are more stable than primary carbocations — the positive charge is delocalized over two carbons.
The more resonance contributors you can draw (especially equivalent ones), the greater the stabilization. But the quality of contributors matters more than quantity — contributors with complete octets and minimal charge separation are more important.
A Final Mental Model
Think of resonance stabilization like a group of people holding a heavy rope. If one person pulls alone, the rope is taut and high-energy. But if everyone holds the rope together, the tension is spread out — each person feels less pull, and the whole system is more relaxed. The electrons are the people; the rope is the molecule. Spreading the "pull" (electron density) over more atoms lowers the energy of the whole system.
That's resonance stabilization.
Resonance and resonance stabilization energy are staple topics in the NCERT Class 11 Chemistry chapter on organic chemistry basic principles, and questions comparing the acidity of carboxylic acids, phenols and alcohols using this concept appear frequently in CBSE board exams and JEE Main. If you are looking for "resonance effect in organic chemistry definition and examples" or practising important questions on the carbonate ion and benzene stability, this is precisely the reasoning examiners expect.
Why this formula?
Resonance Stabilization Effect: Why It Works
The Resonance Stabilization Effect explains why certain molecules or ions are more stable than a single Lewis structure would suggest. Let's build the reasoning from the ground up.
1. The Core Problem: Localized vs. Delocalized Electrons
In a simple Lewis structure, we draw localized bonds — electrons are assigned to specific atoms or bonds. But in reality, for molecules like benzene (C6H6) or the carboxylate ion (RCOO−), the electrons are delocalized over multiple atoms.
- Localized picture: One double bond, one single bond — but this doesn't match experimental bond lengths or stability.
- Delocalized reality: All bonds are identical (e.g., benzene's C–C bonds are all 1.39 Å, between single and double).
Key insight: Delocalization lowers the energy of the system. This energy lowering is the resonance stabilization energy.
2. The Mathematical Foundation: Linear Combination of Atomic Orbitals (LCAO)
Resonance is best understood through Molecular Orbital Theory. For a system with n atomic orbitals (AOs) that can overlap, we form n molecular orbitals (MOs) as linear combinations:
ψj=∑i=1ncjiϕi
where:
- ψj = j-th molecular orbital
- ϕi = i-th atomic orbital
- cji = coefficient (contribution of ϕi to ψj)
The energy of each MO is found by solving the secular determinant:
det∣Hij−ESij∣=0
where Hij=⟨ϕi∣H^∣ϕj⟩ (resonance integral) and Sij=⟨ϕi∣ϕj⟩ (overlap integral).
3. The Simplest Case: The Allyl System (3 Carbon Atoms)
Consider the allyl radical (CH2=CH−CH2∙) or allyl cation/anion. Three p orbitals (one per carbon) combine.
Step 1: Set up the Hückel approximation
- Assume all Sij=0 for i=j (zero overlap approximation)
- Hii=α (Coulomb integral, same for all carbons)
- Hij=β for adjacent carbons, 0 otherwise
Step 2: The secular determinant
For three atoms in a line (1–2–3):
α−Eβ0βα−Eβ0βα−E=0
Step 3: Solve for energies
Let x=βα−E. Then:
x101x101x=0
Expanding: x(x2−1)−1(x)=0⟹x3−2x=0⟹x(x2−2)=0
So x=0 or x=±2.
Thus the three MO energies are:
E1=α+2β,E2=α,E3=α−2β
(Since β<0, E1 is lowest, E3 highest.)
4. Why Stabilization Occurs: The Energy Lowering
For the allyl cation (2 π electrons):
- Electrons fill the lowest MO: E1=α+2β
- Total energy = 2(α+2β)=2α+22β
Compare to localized picture (one isolated double bond):
- One double bond = 2 electrons in a bonding MO of energy α+β
- Total energy = 2(α+β)=2α+2β
Resonance stabilization energy:
ΔE=(2α+22β)−(2α+2β)=2(2−1)β≈0.828β
Since β is negative, ΔE is negative → stabilization.
General formula for a linear conjugated system with n atoms:
The Hückel energy levels are:
Ek=α+2βcos(n+1kπ),k=1,2,…,n
The total π-electron energy for N electrons (filling from lowest up) is:
Eπ=∑occupied2Ek
The resonance stabilization energy is the difference between Eπ and the energy of the best localized structure.
5. The Key Formula: Resonance Energy
For a cyclic conjugated system (like benzene, n=6):
Ek=α+2βcos(n2πk),k=0,±1,±2,…
For benzene (n=6):
- k=0: E=α+2β
- k=±1: E=α+β
- k=±2: E=α−β
- k=3: E=α−2β
Filling 6 electrons: Eπ=2(α+2β)+4(α+β)=6α+8β
Localized (3 double bonds): Eloc=3×2(α+β)=6α+6β
Resonance energy = (6α+8β)−(6α+6β)=2β≈−36 kcal/mol (experimentally ~36 kcal/mol for benzene).
6. Why This Matters for Exam Problems
The resonance stabilization effect explains:
- Bond length equalization: Delocalization spreads bond order evenly.
- Increased stability: Lower energy than any single resonance structure.
- Reactivity patterns: More stable intermediates (e.g., allyl carbocation) form faster.
- Acidity: Carboxylate ion (RCOO−) is stabilized by resonance, making carboxylic acids more acidic than alcohols.
Remember: The actual molecule is a hybrid of all resonance structures, not a rapidly interconverting mixture. The stabilization energy is the energy difference between this hybrid and the most stable single Lewis structure.
7. Quick Summary for Revision
| Concept | Formula/Result |
|---|---|
| Hückel MO energy (linear, n atoms) | Ek=α+2βcos(n+1kπ) |
| Hückel MO energy (cyclic, n atoms) | Ek=α+2βcos(n2πk) |
| Resonance energy | ΔE=Eπ(delocalized)−Eπ(localized) |
| Allyl cation stabilization | ΔE=2(2−1)β≈0.828β |
| Benzene stabilization | ΔE=2β (per ring) |
Bottom line: Resonance stabilization arises because delocalization of π electrons into a larger system of overlapping orbitals lowers the total energy compared to confining them to isolated bonds. The mathematics of LCAO-MO theory quantifies this as a negative resonance energy.
The key idea is resonance stabilization of the carbonyl group in benzaldehyde.
-
In propanal (CH3CH2CHO), the carbonyl carbon is electron-deficient and highly susceptible to nucleophilic attack. The alkyl group has a weak +I effect, which slightly reduces the positive charge.
-
In benzaldehyde (C6H5CHO), the lone pair on the carbonyl oxygen can delocalise into the aromatic ring via resonance. This creates partial double-bond character between the ring and the carbonyl carbon, making the carbon less electrophilic.
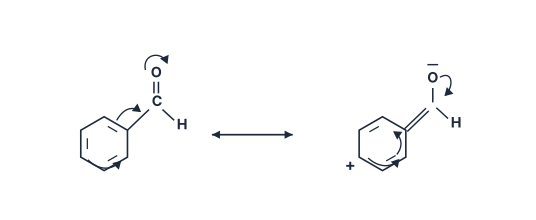
- Additionally, the resonance structures place a positive charge on the ring, which stabilises the carbonyl group and reduces its reactivity toward nucleophiles.
Benzaldehyde is less reactive than propanal in nucleophilic addition reactions due to resonance stabilisation of the carbonyl group.
Benzaldehyde is less reactive than propanal toward nucleophilic addition because the carbonyl carbon in benzaldehyde is less electrophilic — the phenyl ring stabilises the carbonyl group through resonance, reducing its partial positive charge, while propanal has no such stabilisation.
Why resonance matters here
Nucleophilic addition reactions depend on how strongly the carbonyl carbon attracts a nucleophile. That attraction comes from the partial positive charge (δ+) on the carbon. Anything that reduces this δ+ makes the carbon less electrophilic — and therefore less reactive.
In benzaldehyde, the carbonyl group is directly attached to a benzene ring. That ring can delocalise the π electrons of the C=O bond through resonance. This is the key effect.
Propanal has an alkyl group (ethyl) attached to the carbonyl. Alkyl groups are electron-donating by hyperconjugation and induction, but they cannot participate in resonance with the C=O bond. So the carbonyl carbon in propanal retains a stronger δ+ charge.
Step-by-step reasoning
-
Draw the resonance structures of benzaldehyde.
The lone pair on the carbonyl oxygen can be pushed into the C=O π bond, and that electron density can be further delocalised into the benzene ring. This gives a resonance structure where the carbonyl carbon gains electron density (it becomes a single bond to oxygen, and the ring carries a positive charge elsewhere).
The net effect: the carbonyl carbon in benzaldehyde has less partial positive charge than in a simple aldehyde.
Resonance structures of benzaldehyde: the ring's pi electrons delocalise toward the carbonyl group, giving a charge-separated structure with O-minus and a positive ring, which reduces the carbonyl carbon's electrophilicity -
Compare with propanal.
Propanal (CH3CH2CHO) has no such resonance delocalisation. The ethyl group donates electrons weakly through induction, but this is much smaller than the resonance effect in benzaldehyde. The carbonyl carbon in propanal remains significantly δ+.
-
Relate charge to reactivity.
A nucleophile attacks the electrophilic carbonyl carbon. The more positive this carbon is, the faster the attack. Since benzaldehyde’s carbonyl carbon is less positive, it is less reactive toward nucleophilic addition.
-
Consider steric hindrance (a minor factor).
The phenyl ring is bulkier than an ethyl group, but steric hindrance is not the dominant reason here — the electronic effect is far more important.
A common mistake is to think that the benzene ring “withdraws” electrons by induction (it does, weakly), and therefore benzaldehyde should be more reactive. But the resonance donation from the ring into the carbonyl dominates, making the carbon less electrophilic. Induction is secondary here.
You can remember this as: resonance stabilises the reactant (the carbonyl group) in benzaldehyde, so it is less eager to react. Propanal has no such stabilisation, so it reacts more readily.
Final answer
Benzaldehyde is less reactive than propanal in nucleophilic addition reactions because resonance delocalisation from the phenyl ring reduces the electrophilicity of the carbonyl carbon.
Method: Resonance Effect Analysis in Carbonyl Reactivity
Method name: Resonance Stabilization & Electrophilicity Comparison
Step 1 – Identify the electrophilic centre
Both benzaldehyde (CX6HX5CHO) and propanal (CHX3CHX2CHO) have a carbonyl group (C=O). In nucleophilic addition, the nucleophile attacks the electrophilic carbonyl carbon.
Step 2 – Analyse resonance in benzaldehyde
In benzaldehyde, the carbonyl group is directly attached to a benzene ring. The lone pair on the oxygen and the π electrons of the ring can participate in resonance:
- The carbonyl carbon can accept electron density from the ring via resonance.
- This delocalizes the positive charge that develops on the carbonyl carbon during attack.
Draw the resonance forms:
The C=O group conjugates with the aromatic ring, giving structures where the positive charge moves to the ortho and para positions of the ring.
Step 3 – Compare electrophilicity
- Propanal: No resonance delocalization beyond the carbonyl. The carbonyl carbon is strongly electrophilic.
- Benzaldehyde: Resonance reduces the partial positive charge on the carbonyl carbon, making it less electrophilic.
Step 4 – Conclusion on reactivity
Benzaldehyde is less reactive toward nucleophilic addition than propanal.
Step 5 – Key exam point
The resonance stabilization effect decreases the electrophilicity of the carbonyl carbon in benzaldehyde. This is why benzaldehyde does not undergo typical aldehyde reactions like the Fehling’s test or Schiff’s test as readily as propanal.
Final answer:
Benzaldehyde is less reactive than propanal in nucleophilic addition reactions due to resonance stabilization of the carbonyl group with the aromatic ring.
Here are the common mistakes students make when comparing the reactivity of benzaldehyde and propanal in nucleophilic addition reactions, along with how to avoid each.
Mistake 1: Forgetting the "Resonance Stabilization" of the Carbonyl Carbon
The Mistake: Students often remember that the benzene ring is electron-withdrawing (due to the inductive effect) and conclude that benzaldehyde must be more reactive. They forget that the key factor here is resonance.
Why it's wrong: In benzaldehyde, the lone pair on the carbonyl oxygen can delocalize into the benzene ring. This creates a resonance structure where the carbonyl carbon gains partial negative charge (making it less electrophilic). Propanal has no such resonance.
How to Avoid: Always draw the resonance structures.
- For benzaldehyde, draw the structure showing the C=O double bond moving to give the oxygen a negative charge and the adjacent carbon (in the ring) a positive charge.
- This shows that the carbonyl carbon in benzaldehyde is less electron-deficient (less positive) than in propanal.
Mistake 2: Confusing "Electrophilicity" with "Stability of the Product"
The Mistake: Students think that because the product (the tetrahedral intermediate) is stabilized by resonance with the ring, the reaction is faster. They confuse thermodynamic stability (product stability) with kinetic reactivity (rate of the first step).
Why it's wrong: The rate-determining step for nucleophilic addition to a carbonyl is the attack on the carbonyl carbon. This step depends on how electrophilic (positive) that carbon is. The resonance stabilization of the product does not speed up the initial attack; it only affects the equilibrium position.
How to Avoid: Focus on the transition state of the first step.
- The transition state resembles the carbonyl carbon. If the carbonyl carbon is already less positive (due to resonance), the transition state is higher in energy.
- Therefore, benzaldehyde reacts slower (is less reactive) than propanal.
Mistake 3: Ignoring the "Steric Hindrance" Factor
The Mistake: Students only consider electronic effects and forget that the bulky benzene ring physically blocks the approach of the nucleophile.
Why it's wrong: Even if the electronic effect were neutral, the large, flat benzene ring creates more steric hindrance around the carbonyl carbon than the small methyl group in propanal.
How to Avoid: Visualize the molecule.
- In propanal, the carbonyl carbon is attached to a small −CHX2CHX3 group.
- In benzaldehyde, it's attached to a large, planar −CX6HX5 ring.
- The nucleophile has to squeeze past this ring to attack. This makes the reaction slower.
Mistake 4: Giving a "One-Word Answer" Without Explanation
The Mistake: Students simply write "Benzaldehyde is less reactive" or "Benzaldehyde is more reactive" without explaining why.
Why it's wrong: The question explicitly asks you to "Explain your answer." A one-word answer gets zero marks.
How to Avoid: Structure your answer like this:
- State the conclusion: "Benzaldehyde is less reactive than propanal in nucleophilic addition reactions."
- Give the primary reason: "This is because the carbonyl carbon in benzaldehyde is stabilized by resonance with the benzene ring, making it less electrophilic."
- Give the secondary reason: "Additionally, the bulky benzene ring creates steric hindrance, making it harder for the nucleophile to attack."
- Contrast with propanal: "In propanal, there is no such resonance stabilization, and the alkyl group is small, so the carbonyl carbon is more electrophilic and more accessible."
Summary Table: How to Avoid Each Mistake
| Common Mistake | How to Avoid It |
|---|---|
| Forgetting resonance stabilization | Draw the resonance structures. See the partial negative charge on the carbonyl carbon. |
| Confusing product stability with reactivity | Focus on the transition state of the first step. The carbonyl carbon's electrophilicity is what matters for the rate. |
| Ignoring steric hindrance | Visualize the molecule. The benzene ring is bulky and blocks the attack. |
| Giving a one-word answer | Use the "State, Reason, Contrast" structure. Always explain the why. |
Final Correct Answer: Benzaldehyde is less reactive than propanal. The carbonyl carbon in benzaldehyde is resonance-stabilized by the benzene ring, making it less electrophilic, and the bulky ring creates steric hindrance.
- AP EAPCET 2026Set ap-2026-05-20-FN1 markMCQQ.Given below are two statements Statement I: The groups −NHCOCH3 and −OCOCH3 deactivate the benzene ring for electrophilic attack when present on it Statement II: −OH and −CH2CH3 groups activate the benzene ring for electrophilic attack when present on it The Correct answer is (A) Both the statements I and II are correct (B) Both the statements I and II are not correct (C) Statements I is correct and statement II is not correct (D) Statements I is not correct but statement II is correct
›Reveal solutionSolution
Amido/acetoxy substituents are net ring-activating (not deactivating) despite the moderating carbonyl, while −OH and alkyl groups are genuinely activating — so only Statement II is correct.
Concept and Intuition
Substituent effects on the benzene ring for electrophilic aromatic substitution depend on whether the group is a net electron-donor (activating, o/p-director) or electron-withdrawer (deactivating, mostly m-director). Groups like −NH2 and −OH strongly activate the ring via resonance donation of a lone pair. When that lone pair is 'tied up' partly in an adjacent carbonyl (as in −NHCOCH3 and −OCOCH3), the donation into the ring is reduced compared to −NH2/−OH — but it is not eliminated or reversed: the net effect is still donation into the ring relative to hydrogen, so these remain (weaker) activators and o,p-directors, not deactivators. Alkyl groups like ethyl activate through hyperconjugation and the inductive (+I) effect, and −OH is one of the strongest activators known.
Step-by-Step Solution
- Statement I claims −NHCOCH3 and −OCOCH3 deactivate the ring — but both groups have a heteroatom lone pair (N or O) still available for resonance donation into the ring, making them net activators (this is exactly why anilides/acetanilide undergo EAS faster than benzene, e.g. in bromination/nitration) — Statement I is FALSE.
- Statement II claims −OH and −CH2CH3 activate the ring — −OH is a textbook strong activator (o,p-director); ethyl is a classic weak activator (alkyl groups donate electron density via hyperconjugation/+I effect) — Statement II is TRUE.
- Combining: I is incorrect, II is correct → option (D).
Common Mistakes
- Assuming that because a carbonyl is attached nearby (as in amides/esters), the group must be deactivating — this conflates the carbonyl's own character with the character of the whole substituent as seen by the ring.
- Forgetting that alkyl groups, though weak, are activating (not neutral) substituents on benzene.
✓Final answerThe correct option is (D) — Statement I is not correct but statement II is correct.
ANSWER: D
- AP EAPCET 2025Set eng-2025-05-24-FN1 markMCQQ.In compound (X), hyperconjugation is present and in (Y), resonance effect is present. What are X and Y, respectively? (A) Toluene, prop-2-en-1-ol (B) Aniline, 2-propenal (C) Toluene, nitrobenzene (D) 1-Bromopropane, phenol
›Reveal solutionSolution
Toluene is the standard example of hyperconjugation (benzylic C–H with the ring), and nitrobenzene is the standard example of resonance (NO2 conjugated with the ring) — matching X, Y to option (C).
Concept and Intuition
Hyperconjugation is the (no-bond resonance) delocalisation of σ(C−H) or σ(C−C) electrons into an adjacent empty or π orbital — it needs a C–H (or C–C) bond directly attached to an sp2/cationic centre. Resonance (mesomeric effect) needs an actual lone pair or π bond directly conjugated (in a continuous overlapping system) with another π system, such as a substituent's lone pair feeding into an aromatic ring.
Step-by-Step Solution
- Toluene has a −CH3 group attached directly to the benzene ring. The three benzylic C–H σ-bonds align with the ring's π system and delocalise into it — this is the textbook example of hyperconjugation, strengthening the ring and directing substitution ortho/para. So toluene fits X.
- Nitrobenzene has −NO2 directly bonded to the ring, and the nitrogen's p-orbital (with its formal double-bond character to oxygen) is fully conjugated with the ring's π system, giving genuine resonance structures that withdraw electron density from the ring (a strong −M group). So nitrobenzene fits Y.
- Checking (A): prop-2-en-1-ol is CH2=CH−CH2−OH; the OH-bearing carbon is sp3 and is not directly attached to the double bond (there's an intervening CH2), so the oxygen lone pair is not conjugated with the π bond — no resonance here, ruling out (A).
- Checking (B): aniline's hallmark effect is resonance (the N lone pair delocalising into the ring), not hyperconjugation, so it doesn't fit as X (which needs hyperconjugation), ruling out (B).
- Checking (D): 1-bromopropane is not the standard example used for hyperconjugation (that role usually goes to alkyl-substituted alkenes/arenes/carbocations), so it's a weaker fit than toluene, ruling out (D).
- Hence (C) Toluene (hyperconjugation), nitrobenzene (resonance) is the best-fitting pair.
Common Mistakes
- Assuming any conjugated-looking system automatically shows "resonance" and any alkyl group automatically shows "hyperconjugation" without checking that the relevant bond is actually adjacent/aligned to the π system.
- Missing that in allylic alcohols like prop-2-en-1-ol, the position of OH relative to the double bond matters — only if OH is directly on the alkene carbon (an enol) would resonance apply.
✓Final answerThe correct option is (C) — Toluene, nitrobenzene.
ANSWER: C
- AP EAPCET 2025Set eng-2025-05-24-FN1 markMCQQ.Arrange the following in decreasing order of electrophilicity of carbonyl carbon. I. Benzaldehyde (benzene ring with CHO substituent) II. Benzoic acid (benzene ring with COOH substituent) III. Acetophenone (benzene ring with COCH3 substituent) IV. CH3CH2CHO (A) IV > I > III > II (B) IV > I > II > III (C) I > IV > III > II (D) I > II > IV > III
›Reveal solutionSolution
This tests how substituents on a carbonyl carbon change its electrophilicity by resonance/inductive donation. The order is CH3CH2CHO>benzaldehyde>acetophenone>benzoic acid.
Concept and Intuition
A carbonyl carbon is electrophilic because oxygen pulls electron density away from it in the C=O bond. Anything that pumps extra electron density back into that carbon (by resonance or by a strongly electron-donating neighbour) makes it less electrophilic and less reactive toward nucleophiles. Anything that only inductively donates weakly (like a simple alkyl chain) barely changes this, so simple aliphatic aldehydes stay highly electrophilic.
Key donors to compare, from weakest to strongest electron donation into the carbonyl carbon:
- An alkyl chain (only +I effect, mild).
- One aryl ring (resonance donation of its π system into C=O).
- Two donating groups on the same carbonyl carbon (aryl and alkyl, as in a ketone).
- A directly attached −OH (a full lone pair resonates straight into the carbonyl), as in a carboxylic acid.
Step-by-Step Solution
- IV, CH3CH2CHO: an aliphatic aldehyde. The ethyl group only donates weakly by induction, so the carbonyl carbon stays strongly electron-deficient — most electrophilic.
- I, benzaldehyde (C6H5CHO): the phenyl ring conjugates with the carbonyl, delocalising electron density onto the carbonyl carbon by resonance. This is a stronger donation than a simple alkyl chain's induction, so benzaldehyde is less electrophilic than IV.
- III, acetophenone (C6H5COCH3): here the carbonyl carbon has both a phenyl ring (resonance donor) and a methyl group (inductive/hyperconjugative donor) attached — two donating influences instead of one — so it is even less electrophilic than benzaldehyde.
- II, benzoic acid (C6H5COOH): the −OH oxygen's lone pair resonates directly into the carbonyl carbon (the classic carboxylic-acid resonance structure with a C−OH+ / C=O− contributor), which is the strongest donation of the four — so benzoic acid's carbonyl carbon is the least electrophilic.
- Putting it together: IV > I > III > II.
Common Mistakes
- Assuming an aromatic ring always makes a carbonyl more reactive (in fact it stabilises/donates and lowers electrophilicity).
- Forgetting that a carboxylic acid's −OH is a stronger resonance donor than a ketone's second alkyl/aryl group, so acids end up less electrophilic than ketones, not more.
✓Final answerThe correct option is (A) — IV > I > III > II.
ANSWER: A
- AP EAPCET 2023Set ap-2023-05-22-AN1 markMCQQ.Which of the following carbonyl compound reacts with HCN at faster rate? (A) O2N-C6H4-CHO (para-nitrobenzaldehyde) (B) MeO-C6H4-CHO (para-methoxybenzaldehyde) (C) C6H5-COCH2CH3 (propiophenone) (D) O2N-C6H4-COCH3 (para-nitroacetophenone)
›Reveal solutionSolution
The fastest carbonyl toward HCN addition is the one whose carbonyl carbon is most electrophilic and least hindered — an aldehyde bearing a strong electron-withdrawing group.
Concept and Intuition
Nucleophilic addition to C=O proceeds by the nucleophile (CN−) attacking the electrophilic carbonyl carbon. Two factors control rate: (i) electronic — electron-withdrawing substituents increase the positive character on the carbonyl carbon (favouring attack), electron-donating groups decrease it;
(ii) steric/electronic from substituent type — aldehydes (H + one group) are less hindered and inherently more electrophilic than ketones (two alkyl/aryl groups, which are electron-donating and bulkier).
Step-by-Step Solution
- Compare aldehydes vs ketones: propiophenone and para-nitroacetophenone are ketones — less reactive than aldehydes in general.
- Between the two aldehydes: para-methoxybenzaldehyde has −OMe, an electron-donating group (by resonance), which decreases electrophilicity of the carbonyl carbon → slower.
- Para-nitrobenzaldehyde has −NO2, a strong electron-withdrawing group (by resonance and induction), which increases electrophilicity of the carbonyl carbon → faster.
- So among all four, the aldehyde with the EWG (para-nitrobenzaldehyde) reacts fastest with HCN.
Common Mistakes
- Forgetting that ketones are generally slower than aldehydes regardless of substituents.
- Assuming any nitro/methoxy substituent works the same way in all reactions — direction (EWG speeds up, EDG slows down) matters specifically for nucleophilic addition to carbonyls.
✓Final answerThe correct option is (A) — O2N-C6H4-CHO (para-nitrobenzaldehyde).
ANSWER: A
- AP EAPCET 2022Set ap-2022-07-12-AN1 markMCQQ.Assertion (A): The carboxylic carbon is more electrophilic than carbonyl carbon Reason (R): All the bonds attached to carboxylic carbon lie in one plane (A) Both A & R are true and R is correct explanation (B) Both A & R are correct but R is not the correct explanation for A (C) A is correct, R is wrong (D) A is wrong, R is corrent
›Reveal solutionSolution
Carboxylic-acid carbon is actually LESS electrophilic than a simple carbonyl carbon (resonance from –OH), so the assertion is false, while the planarity statement (reason) is true.
Concept and Intuition
In an aldehyde/ketone, the carbonyl carbon bears a partial positive charge from C=O polarisation, making it a good electrophile. In a carboxylic acid, the adjacent –OH group's oxygen lone pair conjugates into the carbonyl system, spreading the positive charge over both oxygens (this is exactly why carboxylic acids show resonance stabilisation, and why nucleophilic addition to the carboxyl carbon is harder than to an aldehyde/ketone carbonyl carbon). So the carboxyl carbon is less electrophilic, not more.
Step-by-Step Solution
- Evaluate assertion: "carboxylic carbon is more electrophilic than carbonyl carbon" — compare a carboxylic acid's carbonyl carbon to a plain aldehyde/ketone carbonyl carbon.
- Resonance donation from the –OH oxygen's lone pair into the C=O in a carboxylic acid partially neutralises the electron deficiency at that carbon, making it less electrophilic than an ordinary carbonyl carbon — so the assertion is FALSE.
- Evaluate reason: the carboxyl carbon is sp2 hybridised, trigonal planar, so the three groups attached (=O, –OH/–O⁻ character, and R) do lie in one plane — this statement is TRUE.
- Since A is false and R is true, and R is a fact about geometry unrelated to being the "explanation" of a false statement, the correct combination is: A incorrect, R correct.
Common Mistakes
- Assuming carboxylic acids are always "more reactive" in every sense — they are actually less reactive toward nucleophilic addition at the carbonyl carbon than aldehydes/ketones, specifically because of this resonance effect.
✓Final answerThe correct option is (D) — (A) is incorrect but (R) is correct.
ANSWER: D
- AP EAPCET 2021Set eng-2021-08-25-AN1 markMCQQ.The correct order of increasing stabilities of the following carbocations.(i) Allyl carbocation(ii) Ethyl carbocation(iii) Benzyl carbocation(iv) Isobutyl carbocation (A)(iv) <(iii) <(ii) <(i) (B)(iv) <(ii) <(i) <(iii) (C)(ii) <(iv) <(i) <(iii) (D)(ii) <(iv) <(iii) < (i)
›Reveal solutionSolution
Carbocation stability is governed mainly by resonance delocalization; benzyl and allyl cations are resonance-stabilized and far more stable than simple alkyl cations, while a branched primary cation (isobutyl) is even less stable than a plain primary (ethyl) one due to steric hindrance to solvation.
Concept and Intuition
A carbocation is stabilized by anything that spreads out its positive charge: resonance (delocalization through π systems) is the strongest stabilizing effect, followed by hyperconjugation and inductive donation from alkyl groups. Benzyl and allyl cations enjoy genuine resonance delocalization (into the aromatic ring, or across the allylic double bond), making them dramatically more stable than any simple alkyl cation. Among non-resonance-stabilized primary cations, bulky branching near the cationic centre (as in isobutyl, −CH2−CH(CH3)2+) does not add meaningful extra stabilization but does sterically hinder the approach of solvent molecules that would otherwise help stabilize the charge — so a branched primary cation like isobutyl is actually less stable than the simplest primary cation, ethyl.
Step-by-Step Solution
- Benzyl cation (iii): positive charge delocalizes into the benzene ring over multiple resonance structures — very stable.
- Allyl cation (i): positive charge delocalizes over the adjacent double bond via one resonance structure — stable, but less so than benzyl (fewer/weaker resonance contributors and no aromatic ring involvement).
- Ethyl cation (ii): a simple, unbranched primary carbocation stabilized only by weak hyperconjugation from the adjacent CH3 group.
- Isobutyl cation (iv): also primary, but the branching at the β-carbon sterically hinders solvation of the cationic centre without providing any resonance benefit, making it the least stable of the four.
- Putting these together (increasing stability): isobutyl (iv) < ethyl (ii) < allyl (i) < benzyl (iii).
Common Mistakes
- Assuming more alkyl substitution near a cationic centre always stabilizes it — branching that hinders solvation can actually destabilize a primary cation further.
- Ranking allyl above benzyl — benzylic delocalization into an aromatic ring is stronger than simple allylic delocalization.
✓Final answerThe correct option is (B) — (iv) < (ii) < (i) < (iii).
ANSWER: B
🎓Unlock everything free for 14 days
- ✓Full step-by-step solutions
- ✓Concept-first explanations
- ✓Methods, shortcuts & mistakes
- ✓PYQ mapping + timed mock tests
Full access for 14 days. No credit card required.