The Core Intuition: Who Wants to Leave, and Who Can Wait?
Imagine you're at a party where the host (the leaving group) is about to leave. The party (the reaction) happens in two stages. First, the host walks out the door — that's the slow, painful step. Then, a new guest (the nucleophile) rushes in to take the empty spot.
The SN1 reaction works exactly like this: the leaving group leaves first, forming a carbocation intermediate. The nucleophile attacks after the leaving group is gone. This means the rate of the reaction depends only on how easily the leaving group can leave — it does not depend on the nucleophile at all.
So the question becomes: What makes a carbocation form easily? The answer is stability. A carbocation that is more stable will form faster and last longer, making the SN1 reaction faster.
The Precise Statement of SN1 Reactivity Order
SN1 Reactivity Order (for alkyl halides):
Allylic≈Benzyl>3∘>2∘≫1∘≈Methyl
This is the order of how fast the SN1 reaction proceeds. Let's unpack why.
Why This Order? The Stability Ladder
A carbocation is a carbon with only six electrons in its valence shell — it's electron-deficient and positively charged. The more you can spread out (delocalize) that positive charge, the more stable the carbocation becomes.
1. Methyl and 1° Carbocations: The Unstable Ones
A methyl carbocation (CHX3X+) has no alkyl groups attached to the positive carbon. There is zero electron-donating effect to stabilize the charge. It is so unstable that it practically never forms in an SN1 reaction — the reaction simply doesn't happen.
A primary (1°) carbocation has one alkyl group attached. Alkyl groups are weakly electron-donating (through hyperconjugation and inductive effect), so it's slightly more stable than methyl — but still far too unstable to form under normal SN1 conditions.
Watch out
Never say "SN1 happens on a primary carbon" in an exam. It is essentially impossible under standard conditions because the carbocation is too unstable.
2. Secondary (2°) Carbocations: The Borderline Case
A secondary carbocation has two alkyl groups donating electron density. It is moderately stable — stable enough to form, but only under certain conditions (like a good leaving group and a polar protic solvent). SN1 reactions on secondary carbons are possible, but they are slower than on tertiary carbons.
3. Tertiary (3°) Carbocations: The Sweet Spot
Three alkyl groups donate electron density to the positive carbon. This makes the carbocation very stable. Tertiary alkyl halides undergo SN1 reactions readily — they are the classic example.
4. Allylic and Benzylic: The Champions
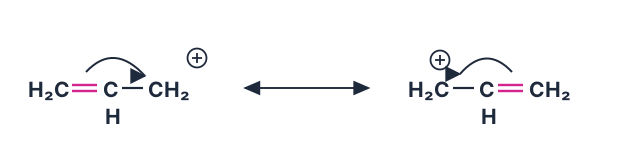
These are special cases. In an allylic carbocation, the positive charge is adjacent to a carbon-carbon double bond. The π electrons of the double bond can delocalize the positive charge onto the second carbon:
CHX2=CH−CHX2X+↔+CHX2−CH=CHX2
Two resonance structures of the allyl carbocation, CH2=CH-CH2+, with the positive charge delocalised between the two terminal carbons (Structure I and Structure II)
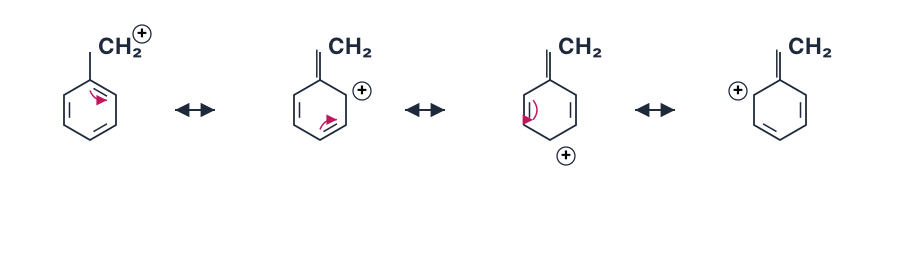
In a benzylic carbocation, the positive charge is adjacent to a benzene ring. The π system of the ring delocalizes the charge across multiple carbons:
CX6HX5−CHX2X+↔(several resonance structures)
Here are those resonance structures — the positive charge cycles from the CH₂ carbon onto the ortho and para positions of the ring:
Four resonance structures of the benzyl carbocation, C6H5-CH2+, with curved electron-pushing arrows showing the positive charge delocalising from the exocyclic CH2 carbon onto the ortho and para ring carbons
This resonance stabilization makes allylic and benzylic carbocations even more stable than tertiary ones. They form the fastest in SN1 reactions.
The Complete Picture in a Table
| Carbocation Type | Stability | SN1 Reactivity | Example |
The SN1 reaction (Substitution Nucleophilic Unimolecular) proceeds via a carbocation intermediate. The reactivity order is determined entirely by the stability of this carbocation — because the rate-determining step is its formation.
The Core Principle
The rate law for SN1 is:
Rate=k[RX]
Only the substrate appears in the rate law — the nucleophile does not participate in the slow step. The slow step is:
RXslowR++X−
Thus, anything that stabilizes the carbocation (R⁺) lowers the activation energy and increases the reaction rate.
The Reactivity Order
For alkyl halides (RX), the SN1 reactivity order is:
This is purely about hyperconjugation and inductive effect.
Tertiary carbocation: Three alkyl groups donate electron density via hyperconjugation (C–H σ bonds overlap with empty p orbital) and +I effect. This spreads the positive charge over more atoms → most stable.
Secondary: Two alkyl groups → less stabilization.
Primary: Only one alkyl group → very little stabilization.
Methyl: No alkyl groups → least stable (only inductive effect from H atoms, which is negligible).
Key formula: The number of α-hydrogens (H on carbons adjacent to the positive carbon) determines hyperconjugation. More α-H → more resonance structures → more stable.
2. Why Allylic and Benzylic Are Even Faster
These carbocations are resonance-stabilized.
Allylic carbocation: The positive charge is delocalized over two carbon atoms via π-bond conjugation:
CH2=CH−CH2+⟷CH2+−CH=CH2
Two resonance structures of the allyl carbocation with the positive charge shared between the two terminal CH2 carbons
Benzylic carbocation: The positive charge is delocalized into the aromatic ring:
C6H5−CH2+⟷several resonance forms involving the ring
Those resonance forms look like this:
Resonance structures of the benzylic carbocation C6H5-CH2+ showing the positive charge delocalised onto the ortho and para carbons of the benzene ring
This resonance stabilization is so powerful that even a primary allylic or benzylic carbocation is more stable than a tertiary alkyl carbocation.
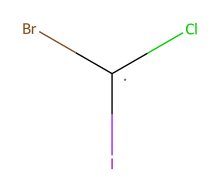
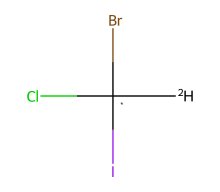
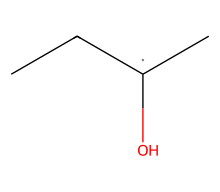
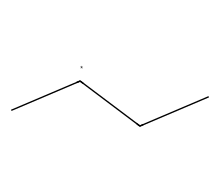
A carbon is asymmetric (chiral) only if it has four different substituents. Checking each option: (a) H, I, Br, Cl — all different → chiral; (b) D, I, Br, Cl — all different → chiral; (c) H, OH, C₂H₅, CH₃ — all different → chiral; (d) H, H, C₂H₅, CH₃ — two H’s are identical → not chiral. So the correct set is (a), (b), (c).
The core idea: An asymmetric carbon (chiral centre) is a carbon atom bonded to four different atoms or groups. If any two substituents are identical, the carbon is not asymmetric — it’s a symmetric centre. This is the single rule that decides everything here.
Let’s go through each option one by one.
Option (a): The carbon marked * is bonded to H, I, Br, and Cl.
These are four different halogen atoms plus hydrogen — all distinct. No two are the same.
→ This carbon is asymmetric.
Option (b): The carbon is bonded to D (deuterium, an isotope of hydrogen), I, Br, and Cl.
Deuterium is chemically nearly identical to hydrogen but is a different isotope — and crucially, for chirality, isotopes count as different substituents because they differ in mass and nuclear composition. So D, I, Br, Cl are four different groups.
→ This carbon is asymmetric.
Option (c): The carbon is bonded to H, OH, C₂H₅ (ethyl), and CH₃ (methyl).
These are four distinct groups: a hydrogen, a hydroxyl, a two-carbon chain, and a one-carbon chain. No two are the same.
An asymmetric carbon (chiral centre) is a carbon atom bonded to four different atoms or groups. If any two groups are identical, the carbon is not asymmetric.
Method: The Four-Group Comparison Test
Steps:
List all four substituents attached to the marked carbon (C∗).
Check for duplicates — if any two substituents are exactly the same, the carbon is not asymmetric.
If all four are different, the carbon is asymmetric.
Applying the test:
(a)C∗ bonded to H, I, Br, Cl
→ All four are different. ✓Asymmetric
(b)C∗ bonded to D (deuterium), I, Br, Cl
→ D is an isotope of H, but chemically it counts as a different group (different mass).
Common Mistakes Students Make on Asymmetric Carbon Identification
Mistake 1: Ignoring Isotopic Differences (e.g., H vs D)
The error: Students treat deuterium (D) and protium (H) as identical, thinking they are both "hydrogen." They mark option (b) as not asymmetric.
Why it's wrong: Deuterium (2H) and protium (1H) are different isotopes — they have different atomic masses. For chirality, we consider atomic number first, then mass number if atomic numbers are equal. Since D has mass 2 and H has mass 1, they are different substituents.
How to avoid: Remember: Isotopes are different groups for chirality. Always check the mass number when atomic numbers are the same.
Mistake 2: Counting Identical Atoms as Different
The error: In option (d), students see four different-looking groups (H, H, C2H5, CH3) and think the carbon is asymmetric.
Why it's wrong: The carbon has two hydrogen atoms — these are identical. A chiral carbon must have four different substituents. Two H's means it's not asymmetric.
How to avoid: Write out all four groups explicitly. If any two are the same (same element, same isotope, same structure), the carbon is not asymmetric.
Mistake 3: Confusing "Asymmetric" with "Stereocenter" in Simple Cases
The error: Students think any carbon with four different-looking groups is asymmetric, even when groups are structurally identical (like two ethyl groups).
Why it's wrong: In option (c), the groups are H, OH, C2H5, and CH3 — all four are different. This is a valid chiral carbon. But students sometimes mistakenly think C2H5 and CH3 are "similar" and reject it. …