Chemistry · Ch 6 — Haloalkanes and Haloarenes
Reactions of Haloarenes
Reactions of Haloarenes
1. Nucleophilic Substitution Reactions
Aryl halides (haloarenes) are far less reactive than alkyl halides toward nucleophilic substitution — chlorobenzene, for instance, will not hydrolyse to phenol under the mild conditions that easily convert an alkyl chloride to an alcohol. Four structural factors, taken together, explain this sluggishness.
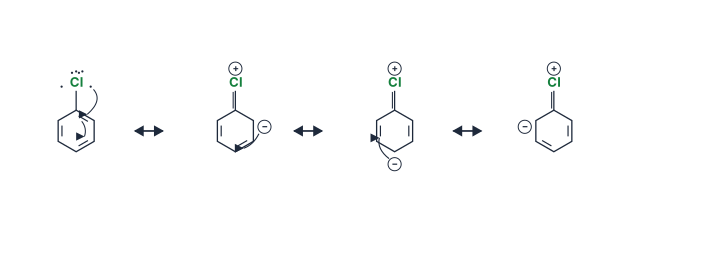
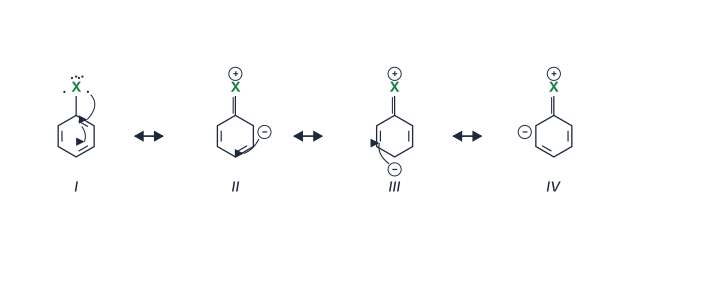
(i) Resonance effect — conjugation of the halogen lone pair with the ring. One of the lone pairs on the halogen atom overlaps with the π-system of the benzene ring. This conjugation generates additional resonance contributors in which the ring carbons ortho and para to the halogen carry a formal negative charge while the halogen itself carries a formal positive charge, with the bond gaining partial double-bond character:
Drawn by us to help you understand the concept clearly, and verified to make sure it's accurate. For exams, practice from your textbook's own diagram.
One lone pair on the halogen conjugates with the ring's system, giving three charge-separated resonance contributors alongside the neutral form: the lone pair forms a new bond while the negative charge sits at an ortho ring carbon, then at the para carbon, then at the other ortho carbon. The C–Cl bond thereby acquires partial double-bond character — shorter and stronger than a normal single bond …
Because the bond in a haloarene is effectively part-double-bond, it is shorter and stronger than a normal single bond, so it is harder to break heterolytically than the bond of an alkyl halide.
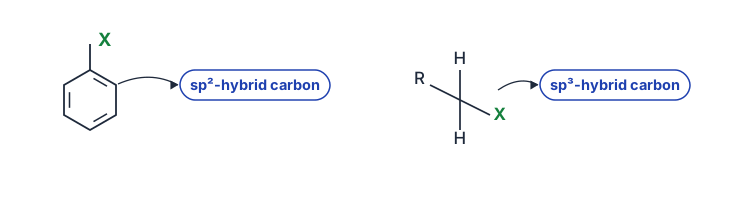
(ii) Difference in hybridisation of the carbon atom in the C–X bond. In a haloalkane, the carbon attached to is -hybridised; in a haloarene it is -hybridised. An carbon has more -character and is therefore more electronegative, so it holds the bonding electron pair of the bond more tightly. Consequently the bond length is only about 169 pm in chlorobenzene against about 177 pm in an alkyl chloride — a shorter bond is harder to cleave, reinforcing the low reactivity of haloarenes.
Drawn by us to help you understand the concept clearly, and verified to make sure it's accurate. For exams, practice from your textbook's own diagram.
The ring carbon bonded to in a haloarene is -hybridised, versus the -hybridised carbon bonded to in a haloalkane. Greater -character makes the carbon hold the C--X bonding pair more tightly, which is why the C--Cl bond is shorter in chlorobenzene (169 pm) than in an alkyl chloride (177 pm) -- and a shorter bond is harder to clea …
(iii) Instability of the phenyl cation. If a haloarene tried to ionise on its own (the way many alkyl halides do in an pathway), it would have to form a phenyl cation. This cation cannot be stabilised by resonance with the ring π-system (the empty orbital lies in the plane of the ring, not conjugated with it), so it is far too high in energy to form. This effectively rules out an -type mechanism for haloarenes.
(iv) Electronic repulsion at the ring. The benzene ring is itself electron-rich (due to the π-cloud), so an incoming electron-rich nucleophile is repelled as it approaches, making the approach of the nucleophile inherently less favourable than at the electron-poorer carbon of an alkyl halide.
Replacement by the Hydroxyl Group
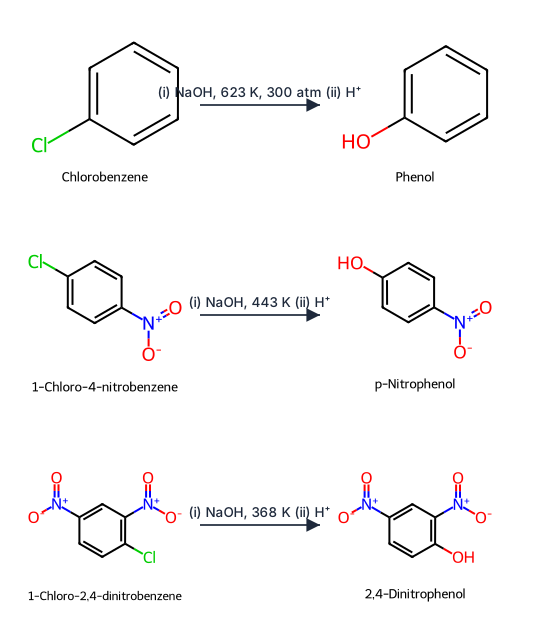
Despite this general unreactivity, chlorobenzene can be forced to give phenol if sufficiently harsh conditions are used — heating with aqueous sodium hydroxide at about 623 K under roughly 300 atmospheres of pressure, followed by acidification:
The reaction becomes dramatically easier — needing far lower temperature — when a strongly electron-withdrawing group such as is present ortho or para to the halogen:
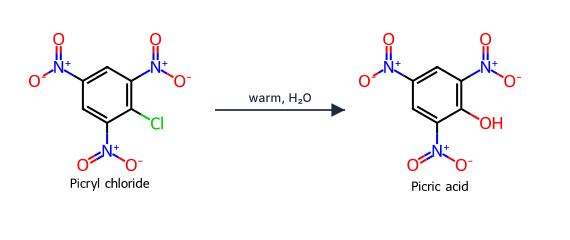
With three groups at the 2,4,6-positions, even warming the chloride with water alone is enough to displace and give the corresponding trinitrophenol. So the effect is cumulative: each additional ortho/para makes displacement of the halogen easier and lets the reaction run under progressively milder conditions. A group placed meta to the halogen, however, has essentially no effect on reactivity at all.
Drawn by us to help you understand the concept clearly, and verified to make sure it's accurate. For exams, practice from your textbook's own diagram.
Plain chlorobenzene needs harsh conditions (NaOH, 623 K, 300 atm) to hydrolyse to phenol. An electron-withdrawing group ortho or para to the chlorine stabilises the addition-elimination intermediate by resonance, so the same substitution runs at progressively milder condit …
Drawn by us to help you understand the concept clearly, and verified to make sure it's accurate. For exams, practice from your textbook's own diagram.
1-Chloro-2,4,6-trinitrobenzene (picryl chloride), reacting with just warm water, gives 2,4,6-trinitrophenol (picric acid) -- the third and most strongly activated case in the NO2-substitution series, where three electron-withdrawing nitro groups at the 2,4,6-positions make the chloride so reactive that e …
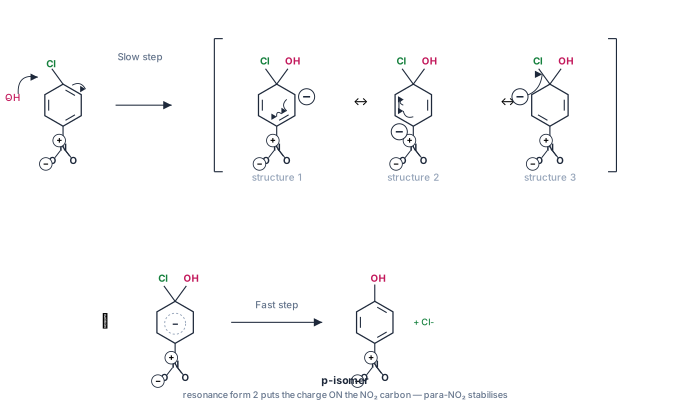
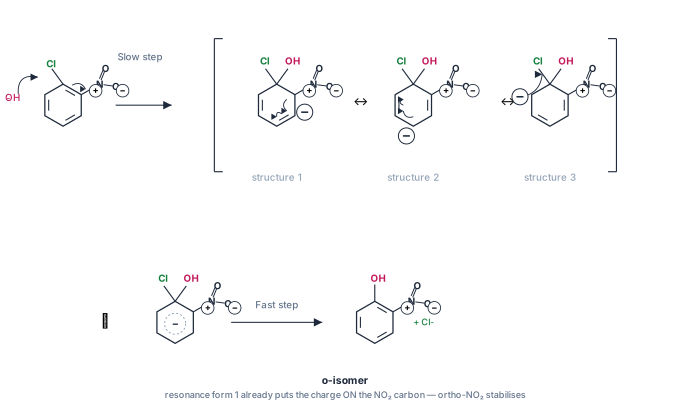
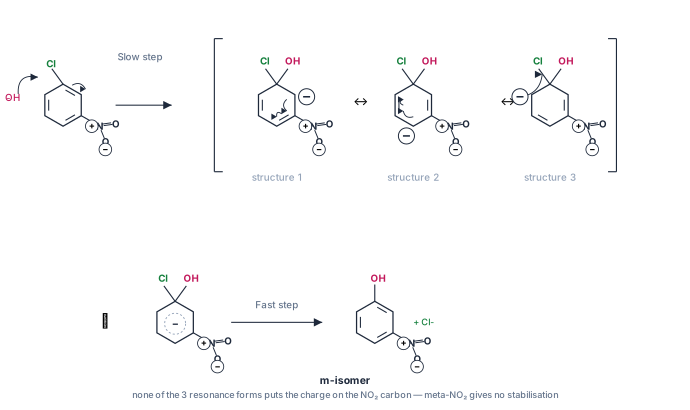
Mechanism. This activated substitution follows a two-step addition–elimination pathway (never a simple one-step displacement):
In the slow step, the hydroxide ion adds to the ring carbon bearing chlorine, momentarily destroying the aromatic sextet and generating a resonance-stabilised carbanion intermediate (this delocalised, non-aromatic intermediate is the same kind of species commonly called a Meisenheimer-type complex). When sits ortho or para to the site of attack, the negative charge of this intermediate can be pushed out onto the oxygens of the nitro group by resonance, which stabilises the intermediate strongly and speeds up the slow step. In the fast second step, the ring rearomatises as chloride ion departs.
Why only ortho/para nitro groups help, and meta does not: drawing out the resonance structures of the carbanion intermediate shows that the negative charge can be delocalised onto the ring carbon directly bearing the group only when the nucleophile has added at a position ortho or para to that group. When is meta to the point of attack, none of the resonance contributors place the negative charge on the carbon carrying , so the nitro group cannot stabilise the intermediate through resonance at all. Since the intermediate is not stabilised, a meta-nitro substituent leaves the reactivity of the haloarene essentially unchanged.
Drawn by us to help you understand the concept clearly, and verified to make sure it's accurate. For exams, practice from your textbook's own diagram.
p-Chloronitrobenzene reacting with hydroxide, OH(-): the delocalised, resonance-stabilised Meisenheimer-type intermediate CAN place negative charge directly on the carbon bearing the NO2 group, since it sits para to the site of attack -- this resonance stabilisation is exactly what activates the substitution and lets it proceed …
Drawn by us to help you understand the concept clearly, and verified to make sure it's accurate. For exams, practice from your textbook's own diagram.
o-Chloronitrobenzene reacting with hydroxide, OH(-): the same resonance activation seen in the para case, this time with the NO2 group sitting ortho to the site of nucleophilic attack -- one of the two ring positions (ortho and para) from which the intermediate's negative char …
Drawn by us to help you understand the concept clearly, and verified to make sure it's accurate. For exams, practice from your textbook's own diagram.
m-Chloronitrobenzene reacting with hydroxide, OH(-): with NO2 at the meta position, none of the resonance contributors of the addition-elimination intermediate can place negative charge on the NO2-bearing carbon, so the nitro group provides no extra activation at all -- reactivity here s …
2. Electrophilic Substitution Reactions
Haloarenes still undergo the standard electrophilic aromatic substitution reactions typical of benzene — halogenation, nitration, sulphonation and the two Friedel–Crafts reactions (alkylation and acylation). In every case the halogen substituent directs the incoming electrophile to the ortho and para positions relative to itself, even though the ring as a whole reacts more sluggishly than benzene does.
Why the halogen is ortho/para-directing. The same lone-pair conjugation described above for nucleophilic substitution is responsible here too. Resonance delocalisation of the halogen's lone pair into the ring places extra electron density preferentially at the ortho and para carbons (structures II, III and IV below all carry a formal positive charge on the halogen and enhanced electron density at these ring positions), while comparatively little extra density reaches the meta positions:
Drawn by us to help you understand the concept clearly, and verified to make sure it's accurate. For exams, practice from your textbook's own diagram.
The neutral form I and the three charge-separated contributors II–IV: in each, the halogen's lone pair has formed and the negative charge sits at an ortho (II), para (III), or the other ortho (IV) ring carbon — never meta. Electron density is therefore raised specifically at the ortho an …
So an electrophile attacking ortho or para to the halogen forms an arenium-ion (carbocation) intermediate that the halogen's lone pair can stabilise directly by resonance.
Why the ring is nevertheless deactivated overall. The halogen is strongly electronegative, so it also withdraws electron density from the ring inductively (the effect), through the sigma framework, at every position — including the position of attack. This inductive withdrawal destabilises the arenium-ion intermediate regardless of where the electrophile attacks.
The apparent contradiction — deactivating, yet ortho/para-directing. A halogen substituent exerts two opposing electronic effects at once: it withdraws electron density inductively (the effect, operating at every ring position) while it donates electron density by resonance (but only into the ortho and para positions). The inductive effect is the stronger of the two, so the net effect on the whole ring is deactivation — haloarenes react more slowly than benzene itself and need more forcing conditions for electrophilic substitution. But the resonance effect, though it cannot overcome the inductive withdrawal in absolute terms, still makes ortho/para attack considerably less disfavoured than meta attack (where no resonance stabilisation of the intermediate is possible at all). In short: the inductive effect controls how fast the ring reacts (reactivity), while the resonance effect controls where it reacts (orientation).
Drawn by us to help you understand the concept clearly, and verified to make sure it's accurate. For exams, practice from your textbook's own diagram.
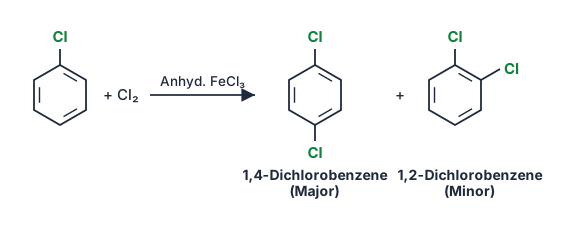
Chlorobenzene reacts with over anhydrous to give a mixture of dichlorobenzenes, exactly as the resonance argument predicts: the new lands mainly at the para position (1,4-dichlorobenzene, major) with the ortho isomer …
(i) Halogenation. Chlorobenzene reacts with in the presence of anhydrous to give a mixture of dichlorobenzenes, dominated by the para isomer:
Drawn by us to help you understand the concept clearly, and verified to make sure it's accurate. For exams, practice from your textbook's own diagram.
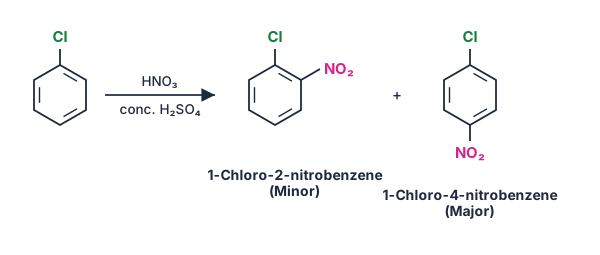
With in concentrated , chlorobenzene nitrates mainly at the para position, giving 1-chloro-4-nitrobenzene as the major product alongside the minor ortho i …
(ii) Nitration. With in concentrated , chlorobenzene gives mainly the para-nitro product:
Drawn by us to help you understand the concept clearly, and verified to make sure it's accurate. For exams, practice from your textbook's own diagram.
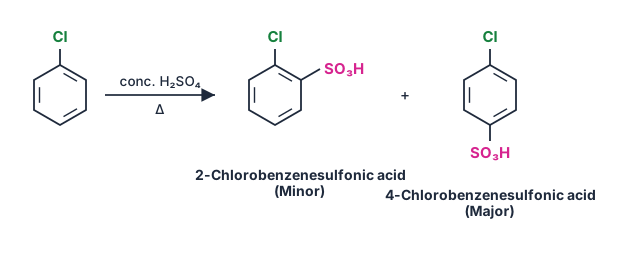
Heating chlorobenzene with concentrated gives predominantly the para-sulphonic acid (4-chlorobenzenesulfonic acid, major), with the ortho isomer (2-chlorobenzenesulf …
(iii) Sulphonation. Heating chlorobenzene with concentrated gives predominantly the para-sulphonic acid:
Drawn by us to help you understand the concept clearly, and verified to make sure it's accurate. For exams, practice from your textbook's own diagram.
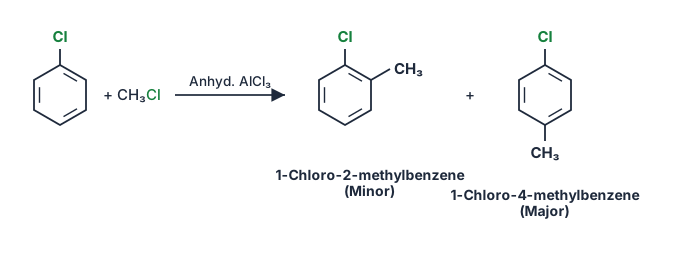
Chlorobenzene with methyl chloride over anhydrous alkylates mainly at the para position (1-chloro-4-methylbenzene, major), with 1-chloro-2-methylbenzene as the minor ortho product -- the bulkier incoming methyl group faces less steric hindrance para to the halogen than ortho to it, on top of b …
(iv) Friedel–Crafts reactions. With methyl chloride and anhydrous (alkylation): …
Drawn by us to help you understand the concept clearly, and verified to make sure it's accurate. For exams, practice from your textbook's own diagram.
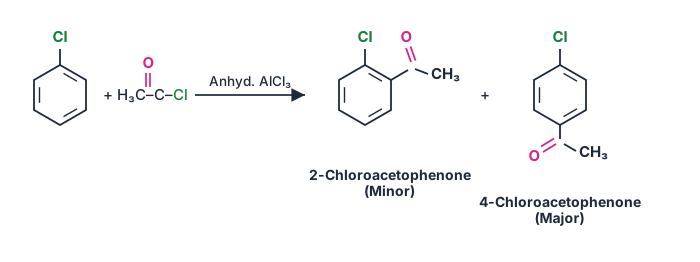
Chlorobenzene with acetyl chloride over anhydrous acylates mainly at the para position, giving 4-chloroacetophenone as the major product, with 2-chloroacetophenon …
Drawn by us to help you understand the concept clearly, and verified to make sure it's accurate. For exams, practice from your textbook's own diagram.
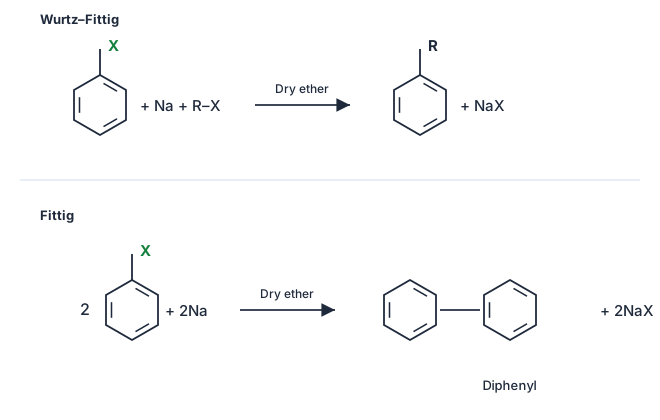
Treating a mixture of an alkyl halide and an aryl halide with sodium metal in dry ether couples the two fragments directly, giving an alkylarene plus -- the Wurtz-Fittig reaction, the aryl-alkyl analogue of the Wurtz coupling used for two alkyl halides. When only an aryl halide (no alkyl halide) is treated with sodium in dry ether, two aryl groups couple to each other instead, giving biphenyl (diphenyl) -- the * …