Chemistry · Ch 3 — Chemical Kinetics
First Order Reactions
First Order Reactions
What "first order" means
A reaction is first order when its rate is proportional to the first power of the concentration of one reactant. For a generic reaction
the rate law is
where is the concentration of the reactant remaining at time and is the rate constant.
Deriving the integrated rate equation
Separating the variables,
Integrating both sides,
where is the constant of integration. At , (the initial concentration), so
Substituting back gives the integrated rate equation
which can be rearranged into the two most useful working forms:
More generally, if the concentrations are and at two times and respectively, subtracting the integrated equation written at from the one at eliminates and gives
Taking the antilog of the rearranged equation also gives an explicit, exponential-decay form for the concentration itself:
The base-10 (log) form
Because log tables and calculators more commonly use base-10 logarithms, the same relation is usually written using :
Reading off a graph
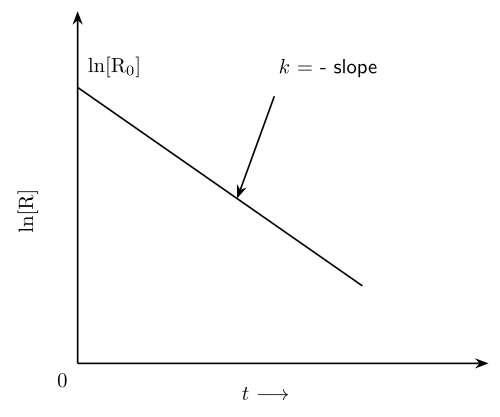
Matching against the straight-line form shows that a plot of against is a straight line with slope and intercept (see the accompanying -vs- figure).
Drawn by us to help you understand the concept clearly, and verified to make sure it's accurate. For exams, practice from your textbook's own diagram.
The figure plots the natural logarithm of the reactant concentration, , on the vertical axis (y-axis) against time, , on the horizontal axis (x-axis). The plot shows a single straight line that slopes downward from left to right.
- The y-intercept of this line is , where is the initial concentration of the reactant at .
- The slope of the line is negative and equals , where is the rate constant of the first-order reaction.
The physical idea is that for a first-order reaction, the concentration of the reactant decays exponentially with time. Taking the natural logarithm converts this exponential decay into a linear relationship, making it easy to determine the rate constant from the slope of the graph.
The key formula developed from this figure is the integrated rate law for a first-order reaction:
This is directly compared to the equation of a straight line, , where:
- (the dependent variable)
- (the independent variable)
- (the slope)
- (the y-intercept)
The textbook also derives an equivalent form using base-10 logarithms:
…
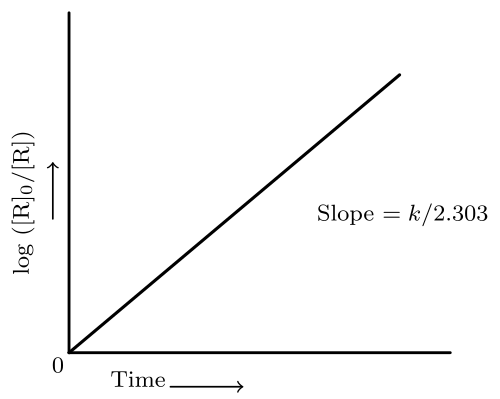
Equivalently, a plot of against is a straight line through the origin with slope (see the accompanying figure for that plot).
Drawn by us to help you understand the concept clearly, and verified to make sure it's accurate. For exams, practice from your textbook's own diagram.
Figure 3.5 is a graphical representation of the integrated rate law for a first-order reaction, plotted in a specific form that makes the rate constant easy to determine.
What the plot shows:
- The y-axis is , where is the initial concentration of the reactant and is its concentration at time .
- The x-axis is time .
- The graph is a single straight line that starts from the origin (0,0). This is because when , , so .
- The slope of this line is , where is the rate constant of the first-order reaction.
Physical idea:
For a first-order reaction, the ratio grows exponentially with time. Taking the logarithm of this ratio linearises the relationship, so a straight line confirms first-order kinetics. The steeper the line, the larger the rate constant .
Key formula developed from this figure:
The textbook derives the integrated rate law in logarithmic form:
Here:
- = initial concentration of reactant (at )
- = concentration of reactant at time
- = rate constant (units depend on reaction order; for first order, typically or )
- = time elapsed
- = logarithm base 10
Comparing this with the straight-line equation :
- slope
- intercept (line passes through origin) …
Either plot lets be extracted directly from experimental concentration-vs-time data without needing to measure an instantaneous slope at any single point.
Where first order kinetics shows up
The hydrogenation of ethene is a first order reaction:
All natural and artificial radioactive decay of unstable nuclei follows first order kinetics, e.g.
The decomposition of and of are further examples of first order gas-phase reactions.
First order kinetics from gas-phase pressure data
For a gas-phase first order reaction such as
…