Q.Write resonance structures of CH₃COO⁻ and show the movement of electrons by curved arrows.
Concept understanding — Resonance Structures Drawing
Resonance Structures: What They Are and How to Draw Them
Let's start with a simple question. When you draw a molecule like ozone (O3), you might put a double bond between the central oxygen and one of the end oxygens, and a single bond to the other. But experiments show both O–O bonds are identical — same length, same strength. So which drawing is correct?
Neither single drawing is correct. The real molecule is a hybrid of both possibilities. That's the core idea of resonance.
The Intuition: A Musical Analogy
Think of a chord played on a piano. A C major chord is made of three notes: C, E, G. No single note is the chord — the chord is the blend of all three. Similarly, a resonance hybrid is the blend of all valid Lewis structures (called resonance contributors or canonical forms) for a molecule. The real molecule is not flipping between these forms; it exists as a single, stable average.
Resonance structures are not in equilibrium. The molecule does not switch from one form to another. It is a single structure that is the weighted average of all contributors.
The Precise Definition
Resonance structures are two or more Lewis structures that differ only in the placement of electrons (pi bonds and lone pairs), never in the positions of atoms. The real molecule is described by a resonance hybrid — a superposition of all contributors.
Rules for Valid Resonance Structures
- Atoms never move. Only electrons (pi bonds, lone pairs, and sometimes sigma bonds in special cases) change positions.
- The total number of electrons stays the same. You are just redistributing them.
- Each structure must obey the octet rule (for second-period elements) and have valid formal charges.
- All structures must have the same net charge and the same number of unpaired electrons (if any).
How to Draw Resonance Structures: A Step-by-Step Method
Let's use the carbonate ion (CO32−) as our example.
Step 1: Draw the best Lewis structure
Start with the skeleton: carbon in the center, three oxygens around it. Count valence electrons: C has 4, each O has 6, plus 2 for the charge = 4+18+2=24 electrons. Place bonds and lone pairs to satisfy octets. You'll get one structure with a C=O double bond and two C–O single bonds, each single-bonded oxygen carrying a negative charge.
Step 2: Identify movable electrons
Look for pi bonds (double or triple bonds) and lone pairs that are adjacent to pi bonds or to atoms with an empty p orbital. In carbonate, the C=O pi bond and the lone pairs on the negatively charged oxygens are the movable parts.
Step 3: Push electrons using curved arrows
An arrow starts at the electron source (a pi bond or lone pair) and points to where the electrons go (to form a new pi bond or to become a lone pair). In carbonate:
- Take the pi bond from C=O and push it to become a lone pair on that oxygen.
- Simultaneously, take a lone pair from a negatively charged oxygen and push it to form a new C=O pi bond.
Step 4: Draw the new structure
After pushing, you get a second structure where the double bond is on a different oxygen. Repeat to get the third structure (all three oxygens take turns being double-bonded).
Always check that the total number of electrons and the net charge remain unchanged after each arrow push. A common mistake is to accidentally add or remove electrons.
Common Patterns to Recognize
| Pattern | Example | What moves |
|---|---|---|
| Allylic system | CH2=CH−CH2+ | Pi bond shifts, positive charge moves |
| Conjugated diene | CH2=CH−CH=CH2 | Pi bonds shift (less common in neutral molecules) |
| Carbonyl group | R2C=O | Lone pair from O forms pi bond, pi bond becomes lone pair |
| Benzene ring | C6H6 | Alternating double bonds shift around the ring |
The Most Common Mistake Beginners Make
Breaking sigma bonds. Remember: sigma bonds (single bonds between atoms) never break in resonance. Only pi bonds and lone pairs move. If you find yourself moving an atom or breaking a single bond, you are drawing a different molecule (a constitutional isomer), not a resonance structure.
If your two structures have different atom connectivity (e.g., one has C–O–C and the other has C–C–O), they are not resonance structures. They are different molecules.
How to Evaluate Which Contributor Is More Important
Not all resonance structures contribute equally. The real hybrid is weighted toward the more stable contributors. Here's the hierarchy:
- More octets satisfied — structures where all second-period atoms have 8 electrons are more stable.
- Fewer formal charges — neutral structures are better than charged ones.
- Negative charge on more electronegative atoms — a negative charge on oxygen is better than on carbon.
- Positive charge on less electronegative atoms — a positive charge on carbon is better than on oxygen.
- No like charges adjacent — avoid placing two positive or two negative charges next to each other.
The Final Answer: What You Need to Remember
Resonance structures are different electron arrangements for the same atomic skeleton. The real molecule is a hybrid of all valid contributors. To draw them, move only pi electrons and lone pairs using curved arrows, keeping atoms fixed. The most stable contributor (with the most octets, fewest charges, and charges on appropriate atoms) dominates the hybrid.
Practice with ozone, carbonate, nitrate, and benzene. After a few examples, the pattern becomes automatic — you'll see the movable electrons instantly.
Looking up "Resonance Structures Drawing: Definition, Formula & Real-World Examples" or "Resonance Structures Drawing important questions 11" is a common way students land here, and rightly so — resonance structures drawing is a core part of the Class 11 Chemistry NCERT/CBSE curriculum. Expect it to reappear, often in a slightly disguised form, across JEE Main, NEET and state CET Chemistry papers.
The key idea is that the acetate ion’s negative charge is delocalised over the two oxygen atoms via conjugation, giving two equivalent resonance structures.
Steps:
- Draw the Lewis structure of CH3COO−: a methyl group attached to a carbonyl carbon, which is double-bonded to one oxygen and single-bonded to the other oxygen (which carries the negative charge).
- Move the π bond from the C=O to the singly bonded oxygen. Use a curved arrow from the π bond towards that oxygen.
- Simultaneously, move a lone pair from the negatively charged oxygen to form a new π bond with carbon. Use a curved arrow from that oxygen’s lone pair towards the carbon.
- The result is the second structure: the carbonyl oxygen becomes the negatively charged one, and the former negative oxygen becomes doubly bonded.
The two structures are identical in energy and are separated by a double-headed resonance arrow (↔).
The resonance structures of CH3COO− are two equivalent forms with the negative charge alternating between the two oxygen atoms, shown by curved arrows moving the π bond and a lone pair.
The acetate ion (CH3COO−) has two equivalent resonance structures where the negative charge is delocalised equally over both oxygen atoms. The actual ion is a hybrid of these two forms, with each C–O bond having a bond order of 1.5.
Why Resonance Matters Here
The acetate ion is a classic example of resonance stabilisation. If you try to draw a single Lewis structure for CH3COO−, you'll face a dilemma: which oxygen gets the negative charge? The answer is that neither oxygen "owns" the charge permanently — instead, the charge is shared between them through a delocalised π-electron system.
This delocalisation makes the acetate ion much more stable than a simple Lewis structure would suggest. The two C–O bonds become identical, with a bond length somewhere between a single and a double bond.
Drawing the Resonance Structures
Step 1: Draw the basic skeleton
The acetate ion has a methyl group (CH3) attached to a carboxylate group (COO−). The carbon of the carboxylate is sp2 hybridized, forming a planar structure.
    O
    ||
H₃C — C
    |
    O⁻
This is one possible Lewis structure — but it's not the whole story.
Step 2: Identify the delocalisable electrons
The key players are:
- The π-bond (double bond) between carbon and the top oxygen
- The lone pair on the negatively charged bottom oxygen
These four electrons (2 from the π-bond, 2 from the lone pair) can move because they're in a conjugated system — the p orbitals on all three atoms (C, O₁, O₂) overlap.
Step 3: Push the electrons with curved arrows
From the structure above, the lone pair on the bottom oxygen moves to form a π-bond between that oxygen and carbon. Simultaneously, the existing π-bond between carbon and the top oxygen breaks, and those electrons move onto the top oxygen as a lone pair.
The curved arrow starts at the electron source (the lone pair) and points to where the bond forms. A second arrow starts at the π-bond and points to the oxygen that receives the electrons.
This gives the second resonance structure:
    O⁻
    |
H₃C — C
    ||
    O
Step 4: Recognise the equivalence
These two structures are identical in energy — they're just mirror images. The methyl group doesn't participate in the resonance, so it stays unchanged.
A quick way to check: count the total number of electrons in each structure. Both have 24 valence electrons (4 from C, 1 from each H, 6 from each O, plus 1 for the negative charge). The connectivity is the same; only the π-bond location changes.
Step 5: Draw the resonance hybrid
The actual acetate ion is not flipping between these two forms — it's a single, stable hybrid. The two C–O bonds are identical, each with a bond order of 1.5. The negative charge is spread equally over both oxygen atoms.
You can represent this with a dashed line between the two oxygens and the carbon, or with a δ⁻ on each oxygen.
A common mistake is to draw the curved arrow from the π-bond to the oxygen without also showing the lone pair moving to form the new π-bond. Both arrows are essential — you're moving four electrons total, not just two. If you only show one arrow, you'll end up with an impossible structure (a carbon with only 6 electrons).
The Final Structures
Here are the two resonance structures with curved arrows:
Structure 1 (left form):
    O                    O⁻
    ||                   |
H₃C — C      →      H₃C — C
    |                   ||
    O⁻                  O
The arrow from the lone pair on O− points toward the C–O bond region. The arrow from the C=O π-bond points toward the top oxygen.
Structure 2 (right form):
    O⁻                  O
    |                   ||
H₃C — C      →      H₃C — C
    ||                  |
    O                   O⁻
The arrows are reversed — the lone pair on the new O− moves to form the π-bond, and the old π-bond breaks to give a lone pair on the other oxygen.
The resonance hybrid is often written as:
H3C−C(O−O)
with a dashed line or a δ− on both oxygens to show equal charge distribution.
The acetate ion has two equivalent resonance structures, with the negative charge delocalised equally over both oxygen atoms, represented by curved arrows showing the movement of a lone pair to form a π-bond and the simultaneous breaking of the existing π-bond.
Showing the 12 most recent of 14 on this concept.
- AP EAPCET 2026Set eng-2026-05-18-FN1 markMCQQ.From the following list, identify the number of substituents which exert +R effect when present on benzene ring −Cl, −COCH3, −NHC2H5, −OCH3, −NHCOCH3, −COOCH3 (A) 5 (B) 6 (C) 3 (D) 4
›Reveal solutionSolution
A substituent shows +R effect if the atom attached to the ring has a lone pair to donate into it; of the six given groups, −Cl, −NHC2H5, −OCH3, −NHCOCH3 qualify (4 total), while −COCH3 and −COOCH3 are −R (carbonyl withdraws electron density).
Concept and Intuition
Resonance (R/mesomeric) effect on a benzene ring depends on whether the directly-attached atom can donate a lone pair into the ring (+R, activating by resonance) or whether the ring's π electrons are pulled into the substituent through a multiple bond (like C=O), which is −R (deactivating by resonance). Groups with an available lone pair on the ipso atom (halogens, −OR, −NR2, −NHCOR) are +R; groups where a π-bonded electronegative atom (as in C=O, C≡N) sits right next to the ring are −R.
Step-by-Step Solution
- −Cl: chlorine's lone pair conjugates into the ring (+R), even though its strong −I effect makes it net deactivating — it is still a +R group.
- −COCH3 (acetyl): the carbonyl carbon is attached directly to the ring; ring electrons delocalise into the C=O, pulling electron density away from the ring — this is −R, not +R.
- −NHC2H5: nitrogen's lone pair conjugates strongly into the ring, a classic strong +R donor (like −NH2).
- −OCH3: oxygen's lone pair conjugates into the ring — a classic +R donor.
- −NHCOCH3 (acetamido): despite partial delocalisation of the N lone pair into the adjacent amide carbonyl, the nitrogen still donates into the ring (it is an ortho, para-directing activator, just weaker than −NH2) — counted as +R.
- −COOCH3 (ester): like the acetyl group, the carbonyl carbon attached to the ring withdraws ring electron density by resonance — −R, not +R.
- Counting the +R groups: −Cl, −NHC2H5, −OCH3, −NHCOCH3 = 4 groups.
Common Mistakes
- Forgetting that halogens, despite being net deactivating, still show a genuine +R effect.
- Assuming any nitrogen-containing substituent is automatically a strong +R donor without checking if it is directly bonded through a carbonyl (as in an amide's carbonyl carbon) versus through the nitrogen itself.
✓Final answerThe correct option is (D) — 4.
ANSWER: D
- AP EAPCET 2026Set eng-2026-05-18-AN1 markMCQQ.Consider the following carbocations [FIGURE] (four labelled carbocation skeletal structures:(a) a secondary carbocation CH3−CH2−C+H−CH(CH3)−CH3;(b) an ether-oxygen-stabilised cation CH3−C+H−O−CH(CH3)−CH2CH3;(c) a primary carbocation C+H2−CH2−CH2−CH(CH3)−CH3;(d) an ether-oxygen-stabilised cation CH3CH2−O−C+H−CH2CH3) The correct stability order for the above carbocations is (A)(b) >(a) >(d) >(c) (B)(b) >(d) >(c) >(a) (C)(d) >(b) >(c) >(a) (D)(d) >(b) >(a) >(c)
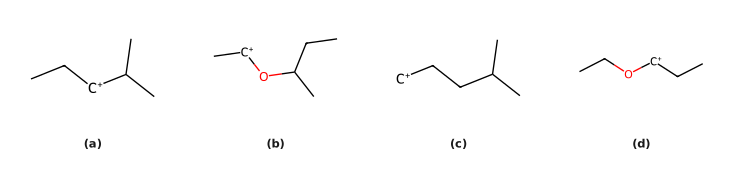
›Reveal solutionSolution
Resonance (mesomeric) donation from an adjacent ether oxygen stabilises a carbocation far more than simple alkyl hyperconjugation does, so both oxygen-adjacent cations rank above both plain alkyl cations; within the plain alkyl pair, the secondary beats the primary. Overall order: (d) > (b) > (a) > (c).
Concept and Intuition
Carbocation stability is governed, in decreasing order of strength, by: resonance/mesomeric (+M) donation > hyperconjugation/inductive (+I) donation from alkyl groups. An ether oxygen directly bonded to the electron-deficient carbon can donate a lone pair into the empty p-orbital, generating an oxocarbenium-type resonance structure (R–O+=CR2′) that delocalises the positive charge onto the (more electronegative but resonance-tolerant) oxygen. This resonance stabilisation is substantially stronger than the modest stabilisation a carbocation gets merely from being flanked by one extra alkyl group (hyperconjugation). Consequently, even a comparatively less-substituted carbocation that is directly bonded to oxygen outranks an ordinary alkyl carbocation that lacks such resonance support.
Step-by-Step Solution
- Classify each cation: (a) is a plain secondary alkyl carbocation (no heteroatom assistance); (c) is a plain primary alkyl carbocation (no heteroatom assistance); (b) and (d) both have the cationic carbon directly bonded to an ether oxygen.
- Because O's lone pair can donate by resonance into the empty orbital on both (b) and (d), both are oxocarbenium-stabilised and this resonance effect outweighs the plain hyperconjugative stabilisation available to (a) and (c). So {b,d}>{a,c}.
- Within the non-resonance-stabilised pair, ordinary carbocation stability rules apply: a secondary cation (a) is more stable than a primary cation (c) due to greater hyperconjugation/induction from two alkyl groups vs one. So a>c.
- Combining, the overall order consistent with resonance dominating hyperconjugation is: (d)>(b)>(a)>(c).
Common Mistakes
- Ranking purely by 1°/2° alkyl substitution and ignoring that resonance donation from a heteroatom lone pair is a much stronger stabilising effect than hyperconjugation, which would incorrectly place a plain secondary cation above an oxygen-stabilised primary one.
- Assuming resonance-stabilised cations must still strictly follow 1°<2°<3° ordering as if the heteroatom weren't present.
✓Final answerThe correct option is (D) — (d) > (b) > (a) > (c).
ANSWER: D
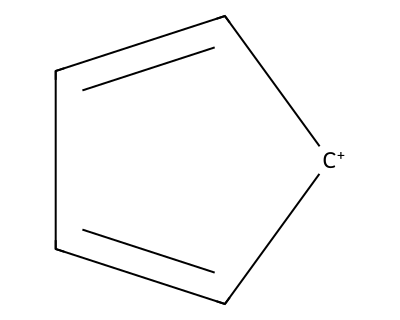
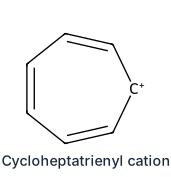
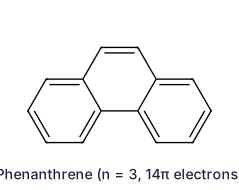
- KCET 2025Set D-41 markMCQQ.Which of the following is not an aromatic compound (A)
 (B)
 (C)
 (D)
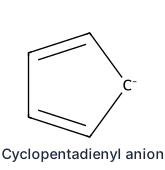
›Reveal solutionSolution
Count π electrons and apply Hückel's 4n+2 rule: the cyclopentadienyl cation has 4πe− (4n) and is anti-aromatic, not aromatic.
Step 1 — The criteria for aromaticity.
A species is aromatic if it is (i) cyclic, (ii) planar, (iii) fully conjugated (an unbroken ring of p-orbitals) and (iv) contains (4n+2) π electrons — Hückel's rule (n=0,1,2,…, i.e. 2, 6, 10, 14 …). A cyclic, planar, conjugated system with 4n π electrons (4, 8, 12 …) is anti-aromatic — actively destabilised.
Step 2 — Count the π electrons in each option.
Option Species π electrons Verdict (A) Cyclopentadienyl cation (5-ring, 2 C=C, ⊕) 2×2=4 (the positive carbon is an empty p-orbital) 4n (n=1) → anti-aromatic ✗ (B) Cycloheptatrienyl / tropylium cation (7-ring, 3 C=C, ⊕) 3×2=6 4n+2 (n=1) → aromatic ✓ (C) Phenanthrene (3 fused benzene rings, angular) 7 C=C ⇒14 4n+2 (n=3) → aromatic ✓ (D) Cyclopentadienyl anion (5-ring, 2 C=C, ⊖) 2×2+2 (the lone pair on the carbanion enters the ring) =6 4n+2 (n=1) → aromatic ✓ Step 3 — The key contrast (A) vs (D).
Both are the same five-membered ring. In the anion, the extra lone pair occupies a p-orbital and joins the cycle, giving the magic 6 π electrons — which is why cyclopentadiene is unusually acidic (pKa≈16). In the cation, that carbon instead has an empty p-orbital, leaving only 4 π electrons — a 4n count, so it is anti-aromatic and highly unstable.
✓Final answerThe correct option is (A) — the cyclopentadienyl cation (4 π electrons, anti-aromatic, hence not aromatic).
ANSWER: A
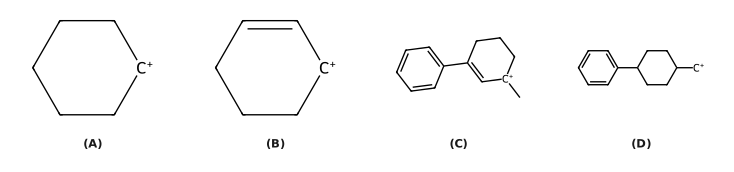
- AP EAPCET 2025Set eng-2025-05-26-AN1 markMCQQ.Identify the most stable carbocation from the following (A) [FIGURE] (a cyclohexyl cation — a six-membered ring with a positive charge on a ring carbon, no double bonds or substituents) (B) [FIGURE] (a cyclohexenyl cation — a six-membered ring with one C=C double bond, and a positive charge on the ring carbon adjacent to the double bond, i.e. allylic) (C) [FIGURE] (a cyclohexenyl ring bearing a methyl group on the cationic ring carbon and a phenyl (Ph) group on the far alkene carbon — allylic and benzylic conjugation together) (D) [FIGURE] (a cyclohexane ring with an exocyclic CH2+ group, and a phenyl (Ph) substituent on a ring carbon a few positions away — a primary benzylic-type cation)
›Reveal solutionSolution
Carbocation stability is set by how much positive charge can be delocalised by resonance (allylic/benzylic conjugation) plus hyperconjugation/induction from alkyl groups. The cation with BOTH allylic and benzylic conjugation, plus a methyl substituent, is the most stable.
Concept and Intuition
A carbocation is stabilised whenever its empty p-orbital can overlap with an adjacent π-system (resonance/conjugation) or with adjacent C–H/C–C sigma bonds (hyperconjugation), and destabilised when it sits isolated with no such support. Combining two independent resonance-donating groups (here, both an adjacent ring double bond AND a phenyl ring through that double bond) gives an extended conjugated system — much more stabilising than either alone.
Step-by-Step Solution
- Option (A): a cyclohexyl cation on a saturated ring, no adjacent π-bond, no aryl group — a simple secondary cation with only ordinary hyperconjugation. Least stabilised of the set.
- Option (B): a cyclohexenyl cation, cationic carbon directly next to the ring's C=C — this is a genuine allylic cation, delocalised over two carbons by resonance. More stable than (A), but only single-bond-worth of delocalisation.
- Option (D): the cationic carbon is an exocyclic CH2+ attached to the ring; the phenyl group sits on a different, non-adjacent ring carbon and is explicitly not conjugated with the cationic centre. So this cation behaves essentially like an isolated primary cation — very poorly stabilised despite having a phenyl group present on the molecule.
- Option (C): the cationic carbon bears a methyl group (extra hyperconjugation/induction) and is directly conjugated (allylic) to the ring's C=C, whose far carbon carries a phenyl substituent. This makes the positive charge delocalisable all the way onto the phenyl ring too (a styrene/cinnamyl-type extended conjugation: cation–C=C–Ph ↔ ring resonance forms with charge spread onto phenyl ortho/para positions). This combined allylic+benzylic delocalisation, together with the methyl group's support, makes (C) by far the most stabilised cation.
- Hence (C) is the most stable carbocation.
Common Mistakes
- Assuming any cation adjacent to a phenyl-substituted alkene is automatically 'benzylic' without checking that the phenyl is actually on a carbon conjugated to the cation (as in (D), where it explicitly is not).
- Forgetting that combining two resonance pathways (allylic + benzylic) is more stabilising than either alone, so ranking purely by 'does it touch a ring double bond' without checking for the extended phenyl conjugation.
✓Final answerThe correct option is (C) — the methyl- and phenyl-substituted cyclohexenyl cation, stabilised by combined allylic and benzylic (styrene-type) conjugation.
ANSWER: C
- MHT-CET 2024Set pcm-2024-05-10-M1 markMCQQ.Which of the following groups exhibits (+)R effect? (A) −NHR (B) −CN (C) −NO2 (D) −COOR
›Reveal solutionSolution
-NHR donates lone pair, +R
−NHR shows +R (electron donation by resonance); CN, NO2, COOR are -R groups.
✓Final answerThe correct option is (A).
ANSWER: A
- TG EAPCET 2023Set eng-2023-05-13-FN1 markMCQQ.In the following reaction ‘C’ is an aromatic compound having substituents D&E. What are D&E?
[!FORMULA] (structure)Cr2O3773K,10−20atm(A)(i) KMnO4/OH−(ii) H3O+(B)Conc HNO3+H2SO4(C)
(A) −OH, −SO3H (B) −CHO, −NO2 (C) −COOH, −NO2 (D) −SO3H, −NO2›Reveal solutionSolution
The reaction sequence starts with toluene, which is oxidised to benzoic acid, then nitrated to give 3‑nitrobenzoic acid; the substituents D and E are –COOH and –NO₂, so the correct option is (C).
Concept & Intuition
This is a classic organic synthesis puzzle. The first step uses chromia (Cr₂O₃) at high temperature and pressure—a typical condition for the dehydrogenation of an alkylbenzene to an aromatic aldehyde or acid. But here the product (A) is then treated with alkaline KMnO₄ followed by acid, which is a strong oxidation that converts any alkyl side‑chain (or aldehyde) into a carboxylic acid. So (B) must be a benzoic acid derivative. Finally, nitration with conc. HNO₃/H₂SO₄ introduces a nitro group. The key is to identify the starting material from the given options: the final compound (C) is aromatic with two substituents D and E. Working backwards, the only combination that fits the oxidation and nitration pattern is –COOH and –NO₂.
Step‑by‑Step Reasoning
-
Identify the starting material
The first arrow shows a structure (not drawn here, but typical in such problems) being passed over Cr₂O₃ at 773 K and 10–20 atm. This is the dehydrogenation of an alkylbenzene (e.g., toluene) to benzaldehyde or benzoic acid. In fact, Cr₂O₃ at high temperature often gives the aldehyde, but the exact product (A) is not yet fully oxidised.
-
Oxidation to (B)
Step (i) KMnO₄/OH⁻ followed by (ii) H₃O⁺ is a vigorous oxidation that converts any alkyl group (–CH₃) or aldehyde (–CHO) directly to a carboxylic acid (–COOH). So (B) must be benzoic acid (or a substituted benzoic acid if the starting material already had a substituent). Since the starting material is a simple aromatic hydrocarbon (likely toluene), (B) is unsubstituted benzoic acid.
-
Nitration to (C)
Conc. HNO₃ + H₂SO₄ is the standard electrophilic aromatic nitration. Benzoic acid has a –COOH group, which is meta‑directing. Therefore the nitro group (–NO₂) will be introduced at the meta position relative to –COOH. Hence (C) is 3‑nitrobenzoic acid (meta‑nitrobenzoic acid).
-
Match with the options
The substituents D and E on the aromatic ring in (C) are –COOH and –NO₂. Looking at the choices:
- (A) –OH, –SO₃H
- (B) –CHO, –NO₂
- (C) –COOH, –NO₂
- (D) –SO₃H, –NO₂ Only option (C) matches.
Watch outA common mistake is to think that Cr₂O₃ oxidation stops at the aldehyde, but the subsequent KMnO₄ step ensures full oxidation to the acid. Also, remember that –COOH is meta‑directing, so the nitro group goes to the meta position, not ortho/para.
TipIf you ever see a sequence: alkylbenzene → Cr₂O₃/high T → KMnO₄/OH⁻ → H⁺ → nitration, the final product is almost always meta‑nitrobenzoic acid. This is a standard route to introduce a meta‑directing group.
✓Final answerThe correct option is (C).
ANSWER: C
-
- TG EAPCET 2023Set eng-2023-05-13-AN1 markMCQQ.Assertion (A) : pKa of phenol is 4.19 and that of benzoic acid is 10 Reason (R) : Phenoxide ion is stabilised by non-equivalent resonance structures whereas benzoate ion by two equivalent resonance structures (A) A and R are true. R is the correct explanation of A (B) A and R are true, but R is not the correct explanation for A (C) A is true but R is false (D) A is false but R is true
›Reveal solutionSolution
The assertion is false because the pKa values are swapped (phenol ~10, benzoic acid ~4.2), but the reason about resonance stabilisation is true. So the correct choice is (D).
The key here is to understand what pKa tells us about acidity. A lower pKa means a stronger acid — the molecule more readily donates its proton. The reason given compares the stability of the conjugate bases (phenoxide vs. benzoate) via resonance. Let’s check both statements carefully.
-
Check the Assertion (A):
The problem states: pKa of phenol is 4.19 and that of benzoic acid is 10.
In reality, the pKa of phenol is about 10, and the pKa of benzoic acid is about 4.2.
So the assertion has the numbers reversed — phenol is the weaker acid, benzoic acid is the stronger one.
Therefore, Assertion (A) is false.
-
Check the Reason (R):
The reason says: Phenoxide ion is stabilised by non-equivalent resonance structures whereas benzoate ion by two equivalent resonance structures.
This is chemically correct:
- In the phenoxide ion, the negative charge can be delocalised into the ring, but the resonance structures are not all equivalent (some place the charge on carbon, which is less stable than on oxygen).
- In the benzoate ion, the two major resonance structures place the negative charge equally on the two oxygen atoms — they are equivalent, giving extra stability.
- Greater stabilisation of the conjugate base means a stronger acid. Benzoate is more stabilised than phenoxide, so benzoic acid (pKa ~4.2) is stronger than phenol (pKa ~10). Hence, Reason (R) is true.
-
Relating A and R:
Since A is false and R is true, R cannot be the correct explanation of A (nor is it needed). The correct option is the one where A is false but R is true.
Watch outA common pitfall is memorising pKa values incorrectly. Many students remember “phenol is acidic” but forget it’s much weaker than carboxylic acids. Always recall: carboxylic acids ~ pKa 4–5, phenols ~ pKa 10.
TipA quick mnemonic: “Carboxylic acids are king of acidity among organics without extra electron-withdrawing groups.” Phenol’s acidity is special but still ~10⁶ times weaker than benzoic acid.
✓Final answerThe correct option is (D).
ANSWER: D
-
- TG EAPCET 2023Set eng-2023-05-14-FN1 markMCQQ.The following molecule with the structure acts as
[!FORMULA] O2NC6H4NHCOCHCl2CH−CH−CH2OHOH
(A) Antibiotic (B) Antiseptic (C) Analgesic (D) Tranquilizer›Reveal solutionSolution
The molecule is chloramphenicol, a broad-spectrum antibiotic, so the correct answer is (A).
The key to this question is recognizing the structural features of the molecule. The given compound contains a nitrobenzene ring, a dichloroacetamide group (NHCOCHCl₂), and a chain with two hydroxyl groups and a primary alcohol. This exact arrangement is the hallmark of chloramphenicol, a well-known antibiotic.
-
Identify the functional groups: The molecule has:
- A para-nitrophenyl group (O₂N–C₆H₄–).
- An amide linkage (–NHCO–) attached to a dichloromethyl group (–CHCl₂).
- A three-carbon chain with two hydroxyl groups (–OH) and a terminal –CH₂OH.
-
Recall the known drug structure: Chloramphenicol is a natural antibiotic (originally from Streptomyces venezuelae) with the systematic name: 2,2-dichloro-N-[(1R,2R)-1,3-dihydroxy-1-(4-nitrophenyl)propan-2-yl]acetamide. Its structure matches exactly: a p-nitrophenyl ring, a dichloroacetamide, and a dihydroxypropyl side chain.
-
Eliminate other options:
- (B) Antiseptic: Antiseptics (e.g., phenol, iodine) are simpler, non-specific germicides; chloramphenicol is a specific systemic antibiotic.
- (C) Analgesic: Pain relievers like aspirin or paracetamol lack the nitro and dichloroacetamide groups.
- (D) Tranquilizer: Sedatives (e.g., diazepam) have different ring systems (benzodiazepines), not this structure.
-
Confirm the role: Chloramphenicol inhibits bacterial protein synthesis by binding to the 50S ribosomal subunit. It is used to treat serious infections like typhoid and meningitis, making it an antibiotic.
Watch outA common mistake is confusing chloramphenicol with antiseptics like chloroxylenol (which also has a chlorine and phenol) or with analgesics like paracetamol (which has an amide but no nitro or dichloro groups). Always check the full side chain.
TipMemorize the "p-nitrophenyl + dichloroacetamide + diol chain" pattern — it’s unique to chloramphenicol among common drugs.
✓Final answerThe correct option is (A).
ANSWER: A
-
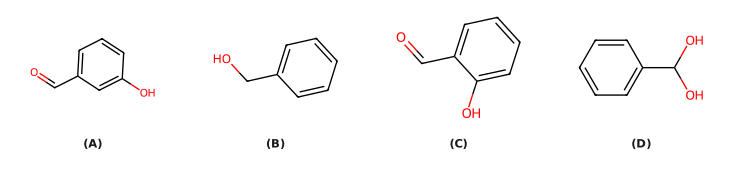
- TG EAPCET 2022Set ap-2022-07-31-AN1 markMCQQ.Identify the product, 'A' in the reaction given below C6H5CHO+Conc. NaOH⟶A+C6H5COONa (A) 3-hydroxybenzaldehyde: a benzene ring bearing −OH and −CHO in the meta positions (m-HO-C6H4-CHO) (B) Benzyl alcohol: a benzene ring bearing −CH2OH (C6H5CH2OH) (C) 2-hydroxybenzaldehyde: a benzene ring bearing −OH and −CHO in the ortho positions (o-HO-C6H4-CHO) (D) Benzaldehyde hydrate: a benzene ring attached to a carbon carrying two −OH groups and one H (C6H5-CH(OH)2)
›Reveal solutionSolution
Benzaldehyde has no α-hydrogen, so with concentrated NaOH it undergoes the Cannizzaro disproportionation: one molecule is oxidised to sodium benzoate and the other reduced to benzyl alcohol. Product 'A' is C6H5CH2OH — option (B).
The concept first
When you meet an aldehyde and a strong base, the first question is always: does it have an α-hydrogen?
- With an α-H (e.g. ethanal), the base pulls it off to make an enolate → aldol condensation.
- Without an α-H, no enolate is possible. With concentrated alkali the aldehyde has only one escape route: it oxidises one of its own molecules and reduces another. That self-oxidation–reduction is the Cannizzaro reaction.
In benzaldehyde the carbon attached to −CHO is an aromatic ring carbon carrying no hydrogen on an sp3 centre, so there is no α-hydrogen — Cannizzaro is guaranteed.
Step-by-step mechanism
Step 1 — Hydroxide attacks the carbonyl.
C6H5CHO+O−H⟶C6H5CH(O−)OH
A tetrahedral alkoxide intermediate forms.
Step 2 — Hydride transfer. This intermediate (or its doubly deprotonated dianion, which is the better hydride donor) collapses: the C−H bond breaks and the hydrogen leaves with its bonding pair as H−, attacking the carbonyl carbon of a second benzaldehyde molecule.
Step 3 — Two different fates.
- The molecule that gave away the hydride becomes benzoic acid, immediately deprotonated by the alkali to C6H5COO−Na+ — the oxidation product (printed in the question).
- The molecule that received the hydride becomes the alkoxide C6H5CH2O−, which picks up a proton on work-up to give C6H5CH2OH — the reduction product.
Step 4 — Overall equation.
2C6H5CHOconc. NaOHC6H5CH2OH+C6H5COONa
Step 5 — Rule out the others. Salicylaldehyde (ortho, option C) comes from the Reimer–Tiemann reaction of phenol with CHCl3/NaOH — a different substrate entirely; the meta isomer (A) is not formed by any of these routes; the gem-diol (D) is the transient hydrate of benzaldehyde in water and is not an isolable product.
✓Final answer'A' is benzyl alcohol, C6H5CH2OH, the reduction half of the Cannizzaro disproportionation.
ANSWER: B
- TG EAPCET 2021Set eng-2021-08-06-FN1 markMCQQ.The total number of overlapping p-orbitals present in cycloheptatrienyl cation is (A) 4 (B) 5 (C) 6 (D) 7
›Reveal solutionSolution
The cycloheptatrienyl cation is an aromatic species where all 7 carbon atoms in the ring are sp2 hybridized, each contributing a p-orbital to a continuous, delocalized π-system. Therefore, there are 7 overlapping p-orbitals.
Concept and Intuition
In organic chemistry, the concept of "overlapping p-orbitals" is central to understanding the stability and reactivity of molecules, particularly those with double bonds and cyclic structures. When we talk about overlapping p-orbitals in a cyclic system, we are referring to the formation of a continuous π-electron cloud above and below the plane of the ring. This continuous overlap is a prerequisite for aromaticity, a special stability found in certain cyclic, planar, fully conjugated systems.
For a cyclic system to have continuous overlap of p-orbitals, two main conditions must be met:
- Each atom in the ring must be sp2 or sp hybridized. This ensures that each atom has at least one unhybridized p-orbital available. In most aromatic systems, the atoms are sp2 hybridized.
- These p-orbitals must be aligned parallel to each other. This allows for effective side-by-side overlap, forming a delocalized π-system.
The number of overlapping p-orbitals is simply the count of atoms in the ring that contribute an unhybridized p-orbital to this continuous π-system. In the case of a carbocation, if the carbon bearing the positive charge is part of a conjugated system, it will be sp2 hybridized and contribute an empty p-orbital to the overall delocalization.
Step-by-Step Solution
- Identify the structure of cycloheptatrienyl cation: The name "cycloheptatrienyl cation" indicates a 7-membered carbon ring ("cyclohept-") containing three double bonds ("-triene") and a positive charge ("-yl cation"). The structure can be drawn as a 7-membered ring with three double bonds and one carbon atom bearing a positive charge.
C1=C2/\C7+C3\/C6=C5C4
(This is a simplified representation; imagine a heptagon with alternating double bonds and a positive charge on one carbon.)2. Determine the hybridization of each carbon atom in the ring:
* Carbons involved in double bonds (e.g., C1, C2, C3, C4, C5, C6) are sp2 hybridized. Each sp2 carbon has one unhybridized p-orbital.
* The carbon bearing the positive charge (C7) is a carbocation. Carbocations are typically sp2 hybridized, with the positive charge residing in an empty p-orbital. This empty p-orbital is crucial for conjugation and delocalization.
Since all 7 carbon atoms in the ring are either part of a double bond or bear a positive charge, they are all $sp^2$ hybridized.3. Count the number of p-orbitals involved in the cyclic overlap:
Because all 7 carbon atoms in the ring are sp2 hybridized, each carbon atom contributes one unhybridized p-orbital. These p-orbitals are oriented perpendicular to the plane of the ring and are parallel to each other, allowing for continuous overlap around the entire ring.
Therefore, there are 7 p-orbitals that are overlapping.4. Verify aromaticity (optional, but confirms continuous overlap):
For a system to be aromatic, it must satisfy Hückel's rules:
* Cyclic: Yes, it's a 7-membered ring.
* Planar: Yes, all carbons are sp2 hybridized, allowing for planarity.
* Fully conjugated: Yes, all 7 sp2 carbons contribute p-orbitals (6 from double bonds, 1 empty from the carbocation) that continuously overlap around the ring.
* Hückel's rule (4n+2 π electrons):
* There are three double bonds, contributing 3×2=6 π electrons.
* The positive charge does not contribute any π electrons.
* Total π electrons = 6.
* If 4n+2=6, then 4n=4, which means n=1. Since n=1 is an integer, the cycloheptatrienyl cation is aromatic.
The aromatic nature of the cycloheptatrienyl cation confirms the presence of a continuous, delocalized $\pi$-system formed by the overlap of p-orbitals from all 7 ring atoms.✓Final answerThe total number of overlapping p-orbitals present in cycloheptatrienyl cation is 7.
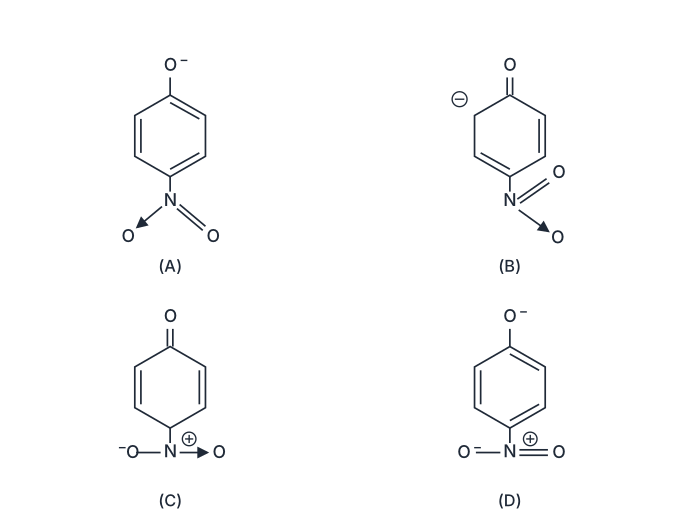
- AP EAPCET 2021Set ap-2021-10-05-FN1 markMCQQ.The most unlikely representation of resonance structure of p-nitro phenoxide is ________. (A) [FIGURE] (para-nitrophenoxide ring with the phenoxide oxygen drawn as O− at the top; the nitro group drawn with one N=O double bond and one N→O dative/coordinate bond, no formal charge shown on N) (B) [FIGURE] (a cyclohexadiene ring with a carbonyl C=O at the top and a carbanion, shown as a circled minus ⊖, on the ring carbon ortho to the carbonyl; the nitro group unchanged, drawn with a dative N→O bond) (C) [FIGURE] (a cyclohexadienone ring with a carbonyl C=O at the top; the nitro group drawn as a charge-separated form with N⊕ bonded to one O− and one O) (D) [FIGURE] (para-nitrophenoxide ring with the phenoxide oxygen drawn as O− at the top; the nitro group drawn as a charge-separated form with N⊕ bonded to two oxygen atoms)
›Reveal solutionSolution
A valid resonance structure of p-nitrophenoxide must pair an aromatic ring with an unperturbed nitro group, OR a quinonoid (C=O) ring with a charge-migrated nitro group — option (D) illegally mixes an untouched aromatic ring with an already charge-separated nitro group, which no single curved-arrow path can produce.
Concept and Intuition
In p-nitrophenoxide, the negative charge on the phenolic oxygen is stabilised by delocalising all the way to the nitro group through the ring in between ("push-pull" conjugation). Each legitimate resonance structure must differ from the next by moving exactly one pair of electrons at a time (one curved arrow, or a linked set), so the ring's bonding pattern and the nitro group's charge state must change together, in lock-step — never independently of each other.
Step-by-Step Solution
- Structure (A): aromatic ring (alternating double bonds) with O− at the top and the nitro group in its ordinary neutral-looking form (one N=O, one N→O dative bond). This is simply the reference/starting Lewis structure — a legitimate contributor.
- Pushing the phenoxide lone pair into the ring converts it to a quinonoid form: C=O appears at the ipso carbon, and the negative charge now sits as a carbanion on a ring carbon (ortho to the carbonyl), with the ring no longer aromatic — a legitimate intermediate contributor (structure B).
- Pushing that carbanion's electrons further, through the ring, into the nitro group converts the nitro group into its charge-separated form (N⊕ bonded to one O− and one O), while the ring stays quinonoid (C=O retained) — this is structure (C), a legitimate, fully-conjugated final contributor.
- Structure (D), however, shows the ring back to being fully aromatic (as if no electrons had ever moved through it) while simultaneously showing the nitro group already carrying the migrated negative charge (N⊕–O−). There is no single legal curved-arrow sequence connecting an untouched aromatic ring to an already charge-separated nitro group — the ring must pass through (and stay in) the quinonoid form for the nitro group to have received that charge.
- Hence (D) cannot be drawn from any legitimate electron-pushing sequence starting from the phenoxide — it is the "most unlikely" (in fact invalid) representation.
Common Mistakes
- Judging "likelihood" purely by apparent stability (assuming "aromatic-looking" structures are always favoured) instead of checking whether the structure is actually reachable by a continuous, valid resonance (curved-arrow) pathway.
- Confusing (A), a valid reference/starting structure, with (D), an invalid hybrid, since both superficially show an aromatic ring with O− on top — the difference is the nitro group's charge state, which is only consistent with (A)'s ring, not with (D)'s.
✓Final answerThe correct option is (D).
ANSWER: D
- AP EAPCET 2021Set eng-2021-08-23-AN1 markMCQQ.The electron transfer in the following conjugated system shows ________ [FIGURE] (four resonance structures of nitrobenzene drawn left to right with curved-arrow electron-pushing: structure 1 shows the neutral nitro group (N double-bonded to two O atoms) attached to the benzene ring with a curved arrow pushing electron density from the ring into the N=O bond; structure 2 shows one O bearing a negative charge, the N=O retained on the other oxygen, and a positive charge on the ring carbon ortho to the point of attachment, with a curved arrow moving around the ring; structure 3 is analogous but the positive charge is on the ring carbon para to the point of attachment (shown at the bottom of the ring), with a curved arrow continuing the conjugation; structure 4 shows both oxygens bearing negative charges (one shown, drawn on the left O) and a positive charge on the ring carbon ortho to the point of attachment on the other side) (A) −R effect (B) −I effect (C) +R effect (D) +I effect
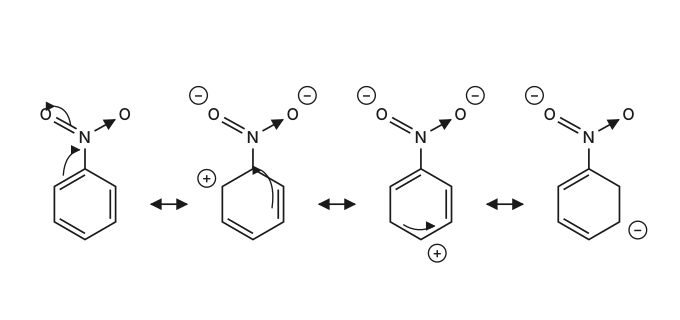
›Reveal solutionSolution
The curved arrows push π-electron density from the ring toward the nitro group, generating positive charge on ortho/para ring carbons — this is the −R (electron-withdrawing resonance) effect of −NO2.
Concept and Intuition
The resonance (mesomeric) effect describes delocalisation of π-electrons through a conjugated system. A substituent can either donate electron density into the ring by resonance (+R, e.g. −NH2, −OH, halogens) or withdraw electron density from the ring by resonance (−R, e.g. −NO2, −CHO, −COOH, −CN). The direction of electron flow in the canonical structures tells you which type of effect is operating: if the ring becomes electron-poor (positive charges appear on ring carbons) as electrons flow toward the substituent, that substituent is exerting a −R effect.
Step-by-Step Solution
- In structure 1, the nitro group is neutral; curved arrows show the ring's π-electrons beginning to shift toward the nitrogen–oxygen system.
- In structures 2 and 3, this electron shift has generated a formal positive charge on the ring carbon ortho (structure 2) and para (structure 3) to the point of attachment of −NO2, while the oxygens of the nitro group pick up negative charge.
- This is precisely the pattern of a group withdrawing electron density from the ring via conjugation/resonance — electrons flow away from the ring carbons and into the substituent, leaving positive charge behind on the ring.
- Because the resonance donates electron density out of the ring (rather than into it), this is called the −R (or −M) effect, characteristic of the nitro group, which is why −NO2 is a strong deactivator and a meta-director in electrophilic aromatic substitution.
- +R would show the opposite: electron density flowing from the substituent into the ring, generating negative charge on ortho/para ring carbons — not what is drawn here. The inductive effects (+I/−I) are unrelated to this delocalised, arrow-pushed conjugative picture.
Common Mistakes
- Confusing −R with −I: inductive effects act through σ-bonds and are not depicted by resonance structures/curved arrows through the π-system.
- Misreading the direction of charge development: positive charge appearing on the ring (not on the substituent) confirms electrons left the ring, i.e., −R, not +R.
✓Final answerThe correct option is (A) — −R effect.
ANSWER: A
🎓Unlock everything free for 14 days
- ✓Full step-by-step solutions
- ✓Concept-first explanations
- ✓Methods, shortcuts & mistakes
- ✓PYQ mapping + timed mock tests
Full access for 14 days. No credit card required.