Q.Out of C6H5CH2Cl and C6H5CHClC6H5, which is more easily hydrolysed by aqueous KOH.
🔒You're viewing a preview — the full solution, concept, methods & PYQ mapping are locked.
🔒 Start your 14-day free trial to unlock the full solution →Concept understanding — SN1 Reactivity
The Core Intuition: Who Wants to Leave, and Who Can Wait?
Imagine you're at a party where the host (the leaving group) is about to leave. The party (the reaction) happens in two stages. First, the host walks out the door — that's the slow, painful step. Then, a new guest (the nucleophile) rushes in to take the empty spot.
The SN1 reaction works exactly like this: the leaving group leaves first, forming a carbocation intermediate. The nucleophile attacks after the leaving group is gone. This means the rate of the reaction depends only on how easily the leaving group can leave — it does not depend on the nucleophile at all.
So the question becomes: What makes a carbocation form easily? The answer is stability. A carbocation that is more stable will form faster and last longer, making the SN1 reaction faster.
The Precise Statement of SN1 Reactivity Order
SN1 Reactivity Order (for alkyl halides):
Allylic≈Benzyl>3∘>2∘≫1∘≈Methyl
This is the order of how fast the SN1 reaction proceeds. Let's unpack why.
Why This Order? The Stability Ladder
A carbocation is a carbon with only six electrons in its valence shell — it's electron-deficient and positively charged. The more you can spread out (delocalize) that positive charge, the more stable the carbocation becomes.
1. Methyl and 1° Carbocations: The Unstable Ones
A methyl carbocation (CHX3X+) has no alkyl groups attached to the positive carbon. There is zero electron-donating effect to stabilize the charge. It is so unstable that it practically never forms in an SN1 reaction — the reaction simply doesn't happen.
A primary (1°) carbocation has one alkyl group attached. Alkyl groups are weakly electron-donating (through hyperconjugation and inductive effect), so it's slightly more stable than methyl — but still far too unstable to form under normal SN1 conditions.
Never say "SN1 happens on a primary carbon" in an exam. It is essentially impossible under standard conditions because the carbocation is too unstable.
2. Secondary (2°) Carbocations: The Borderline Case
A secondary carbocation has two alkyl groups donating electron density. It is moderately stable — stable enough to form, but only under certain conditions (like a good leaving group and a polar protic solvent). SN1 reactions on secondary carbons are possible, but they are slower than on tertiary carbons.
3. Tertiary (3°) Carbocations: The Sweet Spot
Three alkyl groups donate electron density to the positive carbon. This makes the carbocation very stable. Tertiary alkyl halides undergo SN1 reactions readily — they are the classic example.
4. Allylic and Benzylic: The Champions
These are special cases. In an allylic carbocation, the positive charge is adjacent to a carbon-carbon double bond. The π electrons of the double bond can delocalize the positive charge onto the second carbon:
CHX2=CH−CHX2X+↔+CHX2−CH=CHX2
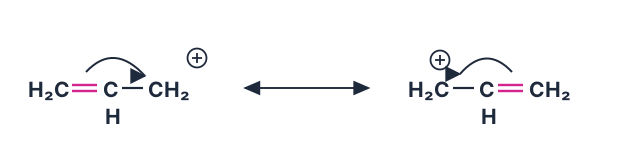
In a benzylic carbocation, the positive charge is adjacent to a benzene ring. The π system of the ring delocalizes the charge across multiple carbons:
CX6HX5−CHX2X+↔(several resonance structures)
Here are those resonance structures — the positive charge cycles from the CH₂ carbon onto the ortho and para positions of the ring:
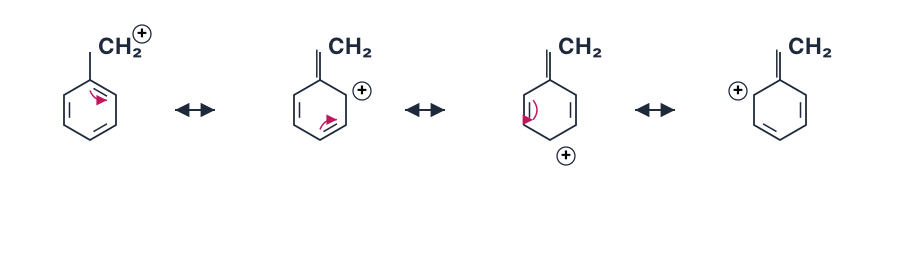
This resonance stabilization makes allylic and benzylic carbocations even more stable than tertiary ones. They form the fastest in SN1 reactions.
The Complete Picture in a Table
| Carbocation Type | Stability | SN1 Reactivity | Example |
|---|---|---|---|
| Methyl | Extremely unstable | Does not occur | CHX3Br |
Why this formula?
SN1 Reactivity: Why the Rate Law and Mechanism Hold
The Core Idea: A Two-Step, Carbocation-Mediated Process
SN1 stands for Substitution, Nucleophilic, Unimolecular. The "unimolecular" part is the key — the rate-determining step involves only one molecule (the substrate). This is fundamentally different from SN2, where both substrate and nucleophile collide.
The reaction proceeds in two distinct steps:
- Slow step: The leaving group departs, forming a carbocation intermediate.
- Fast step: The nucleophile attacks the carbocation.
Why the Rate Law is First-Order
Step 1: The Rate-Determining Step
The slow step is the heterolytic cleavage of the C–LG bond:
R–LGslowR++LG−
Since this step involves only one molecule of substrate, the rate depends only on its concentration:
Rate=k1[R–LG]
Step 2: The Fast Step
The nucleophile then attacks the carbocation:
R++Nu−fastR–Nu
Because this step is fast, it does not affect the overall rate. The nucleophile concentration does not appear in the rate law.
The Resulting Rate Law
Rate=k[R–LG]
This is first-order in substrate and zero-order in nucleophile — a hallmark of SN1.
The classic example — hydrolysis of 2-bromo-2-methylpropane — shows both steps:
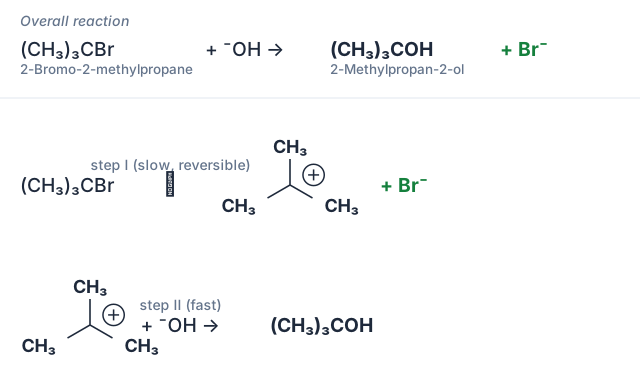
Why the Carbocation Stability Dictates Reactivity
The slow step involves breaking a bond without any help from the nucleophile. This creates a high-energy carbocation intermediate. The activation energy for this step depends entirely on how stable that carbocation is.
Carbocation Stability Order
Methyl<Primary<Secondary<Tertiary<Allylic/Benzylic
Why this order? Three factors stabilize carbocations:
- Hyperconjugation: Adjacent C–H or C–C bonds donate electron density into the empty p-orbital.
- Inductive effect: Alkyl groups are electron-donating, spreading the positive charge.
- Resonance: Allylic and benzylic carbocations delocalize the charge across multiple atoms.
For the benzylic case, that delocalisation looks like this:
The Reactivity Consequence
- Tertiary substrates form relatively stable carbocations → fast SN1.
- Primary substrates form highly unstable carbocations → SN1 is essentially impossible (the activation energy is too high).
- Methyl substrates never undergo SN1 — the carbocation is too unstable.
Why the Leaving Group Must Be Good
The slow step requires the leaving group to depart with its bonding electrons. A good leaving group:
- Is weakly basic (stable as an anion)
- Can stabilize negative charge (large, polarizable, or resonance-stabilized)
Examples: I−, Br−, Cl−, OTs−, H2O
Poor leaving groups: OH−, OR−, NH2− — these are strong bases and will not leave easily.
Why the Solvent Matters (Polar Protic Solvents)
SN1 reactions are faster in polar protic solvents (e.g., water, methanol, ethanol). Why? …
The key idea is SN1 reactivity: the rate depends on the stability of the carbocation intermediate formed after the leaving group departs.
- Both compounds are benzylic chlorides, so both can form resonance-stabilised carbocations.
- C6H5CH2Cl gives a primary benzylic carbocation (C6H5CH2+), stabilised by resonance with one phenyl ring. …
The key idea is that SN1 reactivity depends on carbocation stability. C6H5CHClC6H5 forms a more stable, resonance-delocalized carbocation than C6H5CH2Cl, so it hydrolyses faster. The answer is C6H5CHClC6H5.
This question is about SN1 hydrolysis — a reaction where the leaving group (Cl) departs first, forming a carbocation intermediate, which is then attacked by water (or OH⁻ from KOH). The rate depends entirely on how stable that carbocation is. Aqueous KOH provides a polar, protic environment that favours SN1 over SN2 for these benzylic halides.
Let’s compare the two compounds.
-
Identify the leaving group and the potential carbocation.
Both are benzylic chlorides. In C6H5CH2Cl, the carbon bearing Cl is attached to one phenyl ring and two hydrogens. In C6H5CHClC6H5, it is attached to two phenyl rings and one hydrogen.
-
Think about carbocation stability.
A carbocation is stabilised by resonance with adjacent π-systems (like phenyl rings) and by hyperconjugation from alkyl groups. The more resonance contributors you can draw, the more the positive charge is delocalised, and the more stable the carbocation.
- For C6H5CH2Cl: The carbocation formed is C6H5CH2+ (benzyl carbocation). The positive charge can be delocalised into the phenyl ring — you can draw resonance structures where the charge moves to the ortho and para positions. This is moderately stable.
- For C6H5CHClC6H5: The carbocation formed is C6H5CH+C6H5 (diphenylmethyl carbocation, also called benzhydryl carbocation). Here, the positive charge is delocalised into two phenyl rings simultaneously. That means twice as many resonance structures and much greater charge dispersal.
Carbocation stability order:
CH3+<primary<secondary<tertiary<allylic/benzylic<diphenylmethyl<triphenylmethyl
-
Relate stability to SN1 rate.
In SN1, the rate-determining step is the formation of the carbocation. The more stable the carbocation, the lower the activation energy for its formation, and the faster the reaction. So the compound that gives the more stable carbocation will hydrolyse more easily. …
Method: Carbocation Stability Analysis for SN1 Reactivity
Concept: SN1 reactions proceed via a carbocation intermediate. The rate depends on the stability of that carbocation — more stable carbocation → faster reaction.
Steps
-
Identify the leaving group
Both compounds have Cl as the leaving group. The nucleophile (OH⁻ from KOH) is the same. So the difference lies in the carbocation formed.
-
Draw the carbocation formed after Cl⁻ leaves
- C6H5CH2Cl → benzylic carbocation: C6H5CH2+
- C6H5CHClC6H5 → diphenylmethyl carbocation: (C6H5)2CH+
-
Compare carbocation stability
- Both are benzylic (resonance-stabilized by the phenyl ring).
- The diphenylmethyl carbocation has two phenyl rings donating electron density via resonance → more delocalization of positive charge. …
This is a classic trap in SN1 reactivity. Let’s break down the common mistakes and how to avoid them.
🧠 The Core Concept First
Both compounds are benzylic halides, so both can form a resonance-stabilized carbocation. The question is: which carbocation is more stable?
- C6H5CH2Cl (benzyl chloride) → forms C6H5CH2+ (primary benzylic carbocation)
- C6H5CHClC6H5 (diphenylmethyl chloride / benzhydryl chloride) → forms (C6H5)2CH+ (secondary benzylic carbocation, stabilized by two phenyl rings)
More phenyl rings = more resonance stabilization → more stable carbocation → faster SN1 reaction.
✓ Correct answer: C6H5CHClC6H5 is more easily hydrolysed.
✗ Common Mistake #1: Confusing “more easily hydrolysed” with “more reactive toward nucleophile”
The error: Students think “easily hydrolysed” means the compound reacts faster with OH⁻ directly (SN2). They compare steric hindrance and say the less hindered one (benzyl chloride) reacts faster.
Why it’s wrong: Aqueous KOH is a weak base in water (due to solvation). SN1 is favoured over SN2 for benzylic halides because:
- The carbocation is highly stabilized.
- The nucleophile (OH⁻) is weak and solvated.
How to avoid: Always check the mechanism first. For benzylic/allylic/tertiary halides, SN1 dominates in protic solvents like water/ethanol.
✗ Common Mistake #2: Forgetting that both are benzylic — then guessing based on “primary vs secondary” alone
The error: Students say “primary halide reacts faster in SN2, so benzyl chloride is more reactive.” They forget that both are benzylic and both favour SN1.
Why it’s wrong: In SN1, the rate depends only on carbocation stability, not on the leaving group’s steric environment. The diphenylmethyl carbocation is more stable because the positive charge is delocalized over two aromatic rings.
How to avoid: Draw the resonance structures of both carbocations. Count the number of resonance contributors — more = more stable.
✗ Common Mistake #3: Ignoring the role of the solvent (aqueous KOH)
The error: Students treat this as a dry reaction and assume SN2 is always faster for primary halides. …
- CBSE 2026Set 56/3/11 markMCQQ.The intermediate formed during the slowest step involved in the dehydration of alcohol is : (A) protonated alcohol (B) carbanion (C) free radical (D) carbocation
›Reveal solutionSolution
The dehydration of alcohol proceeds via an E1 mechanism, where the slowest, rate-determining step involves the departure of a water molecule from the protonated alcohol to form a carbocation intermediate. The correct option is (D).
Alcohol dehydration is a classic example of an elimination reaction, specifically an E1 mechanism when catalyzed by acid. Understanding the mechanism is key to identifying the intermediates. The core idea here is that a poor leaving group (the hydroxyl group, −OH) must be converted into a good leaving group (water, −OH2+) before it can depart. The departure of this good leaving group is the slowest step, as it involves breaking a bond and forming a high-energy, electron-deficient species.
Let's break down the process:
-
Protonation of the Alcohol:
Alcohols are not good substrates for direct elimination because the hydroxyl group (OH−) is a strong base and thus a poor leaving group. In the presence of an acid (like H2SO4 or H3PO4), the oxygen atom of the alcohol, with its lone pairs, acts as a Lewis base and gets protonated. This step is fast and reversible.
R-CH2-CH2-OH+H+⇌R-CH2-CH2-OH2+
The protonated alcohol, R-OH2+, now has a good leaving group: a neutral water molecule (H2O).
-
Formation of the Carbocation:
This is the crucial step and the rate-determining step (slowest step) of the reaction. The protonated hydroxyl group departs as a neutral water molecule, leaving behind a positively charged carbon atom. This positively charged carbon species is called a carbocation.
R-CH2-CH2-OH2+slowR-CH2-CH2++H2O
This step is slow because it involves breaking a strong carbon-oxygen bond and forming a high-energy, unstable carbocation. The activation energy for this step is high. The stability of the carbocation formed directly influences the rate of this step; more stable carbocations form faster.
The general order of carbocation stability is:
Tertiary(3∘)>Secondary(2∘)>Primary(1∘)>Methyl
This stability is primarily due to hyperconjugation and inductive effects from alkyl groups.
-
Deprotonation and Alkene Formation: …
-
- CBSE 2024Set 56/3/11 markMCQQ.(CH3)2CH−O−CH3 when treated with HI gives : (A) (CH3)2CH−I+CH3OH (B) (CH3)2CH−OH+CH3−I (C) (CH3)2CH−I+CH3−I (D) (CH3)2CH−OH+CH3OH
›Reveal solutionSolution
With HI, an ether that has no tertiary group cleaves by SN2: iodide attacks the less hindered carbon. In (CH3)2CH–O–CH3 that is the methyl carbon, so the products are CH3I and isopropyl alcohol — option (B).
When an ether reacts with a hydrogen halide, the first step is always protonation of the ether oxygen, which converts it into a good leaving group. The regiochemistry — which fragment becomes the iodide and which becomes the alcohol — is decided by the mechanism.
- Protonation. The oxygen lone pair takes a proton from HI:
(CH3)2CH–O–CH3+HI→(CH3)2CH–O+(H)–CH3+I−
-
Choose the mechanism. An SN1 (carbocation) route operates only when one alkyl group can form a stable (tertiary or benzylic/allylic) cation. Here the choices are methyl (primary) and isopropyl (secondary) — neither gives a stable enough cation, so cleavage goes by SN2.
-
SN2 attacks the less hindered carbon. Iodide approaches the carbon with least steric crowding. The methyl carbon is far less hindered than the secondary isopropyl carbon, so I− attacks the methyl group:
(CH3)2CH–O+(H)–CH3+I−→(CH3)2CH–OH+CH3–I
The isopropyl group departs as isopropyl alcohol. …
- CBSE 2024Set ANNUAL1 markQ.Why is tert-butylbromide more reactive towards SN1 reaction?
›Reveal solutionSolution
SN1 rate is governed by how easily/stably the halide can ionize; more alkyl substitution around the leaving carbon means a more stabilized carbocation and a faster SN1 reaction.
SN1 substitution proceeds through a slow, rate-determining ionization step in which the C–X bond breaks heterolytically to form a carbocation intermediate. The stability of this carbocation directly controls the rate: 3° carbocations are far more stable than 2° or 1° carbocations because the three alkyl (methyl) groups donate electron density into the empty p-orbital via both the inductive (+I) effect and hyperconjugation (C–H σ-bonds of the methyl groups overlapping with the empty p-orbital), spreading out and stabilizing the positive charge. Since tert-butyl bromide ionizes to the e …
- CBSE 2020Set 56/1/11 markQ.Predict the major product formed when 2-Bromopentane reacts with alcoholic KOH.
›Reveal solutionSolution
Alcoholic KOH favours elimination over substitution. 2-Bromopentane undergoes dehydrohalogenation via the Saytzeff rule to give the more substituted alkene as the major product — Pent-2-ene.
The key here is the reagent: alcoholic KOH. This is a classic strong base in a polar, protic solvent (ethanol). Unlike aqueous KOH (which promotes substitution), the alcoholic medium suppresses the nucleophilic character of the hydroxide ion and enhances its basicity. So the reaction follows an E2 elimination pathway — not SN1 or SN2.
Why? Because the alkoxide ion (from ethanol) is a weaker nucleophile but a strong base, and the high temperature of the reaction (usually reflux) favours elimination over substitution. The substrate is a secondary alkyl halide, which can undergo both E1 and E2, but with a strong base like KOH, E2 dominates.
Now, the molecule is 2-Bromopentane:
CH3–CHBr–CH2–CH2–CH3
The bromine is on carbon 2. In an E2 elimination, the base abstracts a proton from a β-carbon (adjacent to the carbon bearing the leaving group), while the leaving group departs simultaneously. The question is: which β-hydrogen is removed?
There are two possible β-carbons:
- β-carbon 1 (C1) — gives a terminal alkene: Pent-1-ene
- β-carbon 3 (C3) — gives an internal alkene: Pent-2-ene
The Saytzeff rule (Zaitsev’s rule) tells us that the more substituted alkene is more stable (due to hyperconjugation and inductive effects). Pent-2-ene is disubstituted (two alkyl groups on the double bond), while Pent-1-ene is monosubstituted. So Pent-2-ene is the major product.
Watch outA common mistake is to think that the less hindered β-hydrogen (on the terminal carbon) is always removed. But in E2, the more substituted alkene is favoured unless the base is very bulky (like potassium tert-butoxide). Alcoholic KOH is not bulky, so Saytzeff product dominates.
Let’s walk through the steps:
-
Identify the substrate and reagent.
2-Bromopentane is a secondary alkyl halide. Alcoholic KOH is a strong base in ethanol. The reaction conditions (heat, base, alcohol solvent) scream E2 elimination.
-
Locate the β-hydrogens.
The α-carbon (C2) has two β-carbons:
- C1 (methyl group) has 3 hydrogens.
- C3 (methylene group) has 2 hydrogens. Both are accessible to the base.
-
Apply the Saytzeff rule. …
- CBSE 2020Set ANNUAL1 markQ.In general Alkyl Halides are more reactive than Aryl halides.
›Reveal solutionSolution
True. The C–X bond in alkyl halides is a pure single bond and easier to break than the partial-double-bond C–X bond in aryl halides.
In alkyl halides, the C–X bond is a simple sigma (sp3–p) single bond, which is relatively easy to break heterolytically, so alkyl halides readily undergo nucleophilic substitution. In aryl halides, the halogen's lone pair delocalises into the benzene ring by resonance, giving the C–X bond partial double-bond character; this makes the bond shorter, stronger and harder to break. Additionally, the sp2-hybridised carbon in aryl halides holds electrons more tightly (higher effective electronegativity of sp2 vs …
- CBSE 2018Set ANNUAL1 markQ.Which one of C6H5Cl and C6H5CH2Cl will react easily with aqueous KOH ?
›Reveal solutionSolution
Benzyl chloride reacts fast because its −CH2Cl carbon is not conjugated to the ring, whereas chlorobenzene's aryl C−Cl bond is strengthened by resonance and resists substitution.
In chlorobenzene, chlorine's lone pair conjugates with the aromatic ring (resonance donation into the ring), giving the C−Cl bond partial double-bond character; this makes the bond shorter and stronger, and the carbon is sp2 hybridised (higher effective electronegativity, more resistant to nucleophilic attack) — so chlorobenzene is very unreactive towards aqueous KOH under ordinary conditions.
…
🎓Unlock everything free for 14 days
- ✓Full step-by-step solutions
- ✓Concept-first explanations
- ✓Methods, shortcuts & mistakes
- ✓PYQ mapping + timed mock tests
Full access for 14 days. No credit card required.