Q.The polarity of C-X bond of alkyl halides is responsible for their nucleophilic substitution, elimination and their reaction with metal atoms to form organometallic compounds. Alkyl halides are prepared by the free radical halogenation of alkanes, addition of halogen acids to alkenes, replacement of -OH group of alcohols with halogens using phosphorus halides, thionyl chloride or halogen acids. Aryl halides are prepared by electrophilic substitution of arenes. Nucleophilic substitution reactions are categorised into SN1 and SN2 on the basis of their kinetic properties. Chirality has a profound role in understanding the SN1 and SN2 mechanism. Answer the following questions:
🔒You're viewing a preview — the full solution, concept, methods & PYQ mapping are locked.
🔒 Start your 14-day free trial to unlock the full solution →Part (a)Concept understanding — SN1 Reactivity Order
The Core Intuition: Who Wants to Leave, and Who Can Wait?
Imagine you're at a party where the host (the leaving group) is about to leave. The party (the reaction) happens in two stages. First, the host walks out the door — that's the slow, painful step. Then, a new guest (the nucleophile) rushes in to take the empty spot.
The SN1 reaction works exactly like this: the leaving group leaves first, forming a carbocation intermediate. The nucleophile attacks after the leaving group is gone. This means the rate of the reaction depends only on how easily the leaving group can leave — it does not depend on the nucleophile at all.
So the question becomes: What makes a carbocation form easily? The answer is stability. A carbocation that is more stable will form faster and last longer, making the SN1 reaction faster.
The Precise Statement of SN1 Reactivity Order
SN1 Reactivity Order (for alkyl halides):
Allylic≈Benzyl>3∘>2∘≫1∘≈Methyl
This is the order of how fast the SN1 reaction proceeds. Let's unpack why.
Why This Order? The Stability Ladder
A carbocation is a carbon with only six electrons in its valence shell — it's electron-deficient and positively charged. The more you can spread out (delocalize) that positive charge, the more stable the carbocation becomes.
1. Methyl and 1° Carbocations: The Unstable Ones
A methyl carbocation (CHX3X+) has no alkyl groups attached to the positive carbon. There is zero electron-donating effect to stabilize the charge. It is so unstable that it practically never forms in an SN1 reaction — the reaction simply doesn't happen.
A primary (1°) carbocation has one alkyl group attached. Alkyl groups are weakly electron-donating (through hyperconjugation and inductive effect), so it's slightly more stable than methyl — but still far too unstable to form under normal SN1 conditions.
Never say "SN1 happens on a primary carbon" in an exam. It is essentially impossible under standard conditions because the carbocation is too unstable.
2. Secondary (2°) Carbocations: The Borderline Case
A secondary carbocation has two alkyl groups donating electron density. It is moderately stable — stable enough to form, but only under certain conditions (like a good leaving group and a polar protic solvent). SN1 reactions on secondary carbons are possible, but they are slower than on tertiary carbons.
3. Tertiary (3°) Carbocations: The Sweet Spot
Three alkyl groups donate electron density to the positive carbon. This makes the carbocation very stable. Tertiary alkyl halides undergo SN1 reactions readily — they are the classic example.
4. Allylic and Benzylic: The Champions
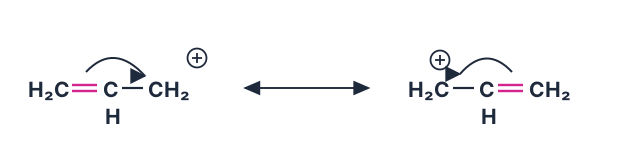
These are special cases. In an allylic carbocation, the positive charge is adjacent to a carbon-carbon double bond. The π electrons of the double bond can delocalize the positive charge onto the second carbon:
CHX2=CH−CHX2X+↔+CHX2−CH=CHX2
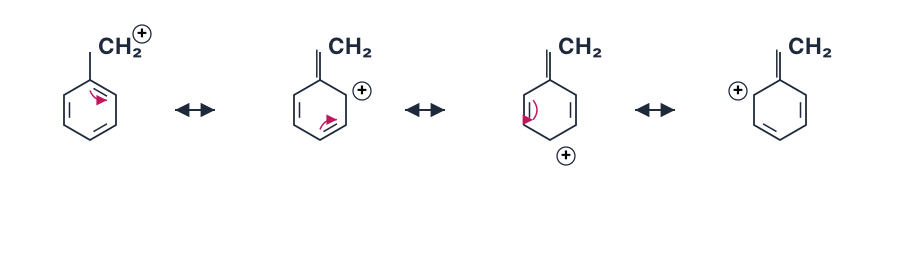
In a benzylic carbocation, the positive charge is adjacent to a benzene ring. The π system of the ring delocalizes the charge across multiple carbons:
CX6HX5−CHX2X+↔(several resonance structures)
Here are those resonance structures — the positive charge cycles from the CH₂ carbon onto the ortho and para positions of the ring:
This resonance stabilization makes allylic and benzylic carbocations even more stable than tertiary ones. They form the fastest in SN1 reactions.
The Complete Picture in a Table
| Carbocation Type | Stability | SN1 Reactivity | Example |
|------------------|-----------|----------------|---------| …
Why this formula?
SN1 Reactivity Order: Why It Holds
The SN1 reaction (Substitution Nucleophilic Unimolecular) proceeds via a carbocation intermediate. The reactivity order is determined entirely by the stability of this carbocation — because the rate-determining step is its formation.
The Core Principle
The rate law for SN1 is:
Rate=k[RX]
Only the substrate appears in the rate law — the nucleophile does not participate in the slow step. The slow step is:
RXslowR++X−
Thus, anything that stabilizes the carbocation (R⁺) lowers the activation energy and increases the reaction rate.
The Reactivity Order
For alkyl halides (RX), the SN1 reactivity order is:
Allylic>Benzyllic>Tertiary>Secondary>Primary>Methyl
Let's break down why each step holds.
1. Why Tertiary > Secondary > Primary > Methyl?
This is purely about hyperconjugation and inductive effect.
- Tertiary carbocation: Three alkyl groups donate electron density via hyperconjugation (C–H σ bonds overlap with empty p orbital) and +I effect. This spreads the positive charge over more atoms → most stable.
- Secondary: Two alkyl groups → less stabilization.
- Primary: Only one alkyl group → very little stabilization.
- Methyl: No alkyl groups → least stable (only inductive effect from H atoms, which is negligible).
Key formula: The number of α-hydrogens (H on carbons adjacent to the positive carbon) determines hyperconjugation. More α-H → more resonance structures → more stable.
2. Why Allylic and Benzylic Are Even Faster
These carbocations are resonance-stabilized.
- Allylic carbocation: The positive charge is delocalized over two carbon atoms via π-bond conjugation:
CH2=CH−CH2+⟷CH2+−CH=CH2
- Benzylic carbocation: The positive charge is delocalized into the aromatic ring:
C6H5−CH2+⟷several resonance forms involving the ring
Those resonance forms look like this:
This resonance stabilization is so powerful that even a primary allylic or benzylic carbocation is more stable than a tertiary alkyl carbocation.
3. The Complete Order (with reasoning) …
Part (b)Concept understanding — Dehydrohalogenation
Dehydrohalogenation — First Look
Imagine you have a molecule that is "unstable" in a specific way — it carries a halogen atom (like Cl, Br, I) on one carbon and a hydrogen atom on the neighbouring carbon. If you treat it with a strong base, the base can pull off that hydrogen, and simultaneously the halogen leaves as a negative ion. The two carbons that lost these atoms now form a double bond between them. That's the core idea: dehydrohalogenation is the elimination of H and X (halogen) from adjacent carbons, producing an alkene.
The name itself tells you what happens: dehydro (removal of hydrogen) + halogenation (removal of halogen). So you are literally removing a hydrogen halide (HX) from the molecule.
The Precise Reaction
A haloalkane (alkyl halide) is treated with alcoholic KOH (potassium hydroxide dissolved in ethanol). The KOH acts as a strong base. The reaction follows this general pattern:
R−CH2−CH2−XKOHalcoholicR−CH=CH2+KX+H2O
For example, bromoethane gives ethene:
CH3−CH2−BrKOHalcoholicCH2=CH2+KBr+H2O
Aqueous KOH (KOH in water) does not cause elimination — it gives substitution (an alcohol). The alcoholic medium is essential because it keeps the base strong enough to pull off the hydrogen, and it does not favour the competing substitution reaction.
Why Alcoholic KOH and Not Aqueous?
In water, the hydroxide ion (OH−) is heavily solvated — surrounded by water molecules — which reduces its basic strength. In ethanol, the solvation is weaker, so OH− is a much stronger base. A strong base is needed to abstract the β-hydrogen (the hydrogen on the carbon next to the one bearing the halogen). The reaction proceeds via a one-step concerted mechanism (E2) where the base pulls the H, the halogen leaves, and the double bond forms — all at once.
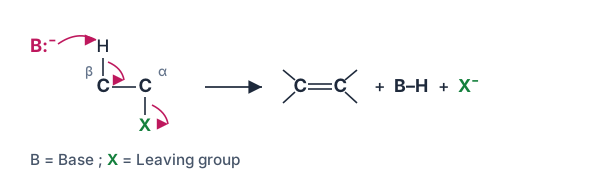
Saytzeff's Rule — Which Alkene Forms?
When the haloalkane has more than one possible β-hydrogen (i.e., the carbon next to the halogen is attached to two different sets of hydrogens), more than one alkene can form. Saytzeff's rule tells you which one is the major product:
In dehydrohalogenation, the alkene with the more substituted double bond (the one with more alkyl groups attached to the double-bonded carbons) is the major product.
Why? More substituted alkenes are more stable (hyperconjugation and inductive effects). The reaction favours the pathway that leads to the more stable alkene.
NCERT's own example uses 2-bromopentane, and the preference is just as clear there:
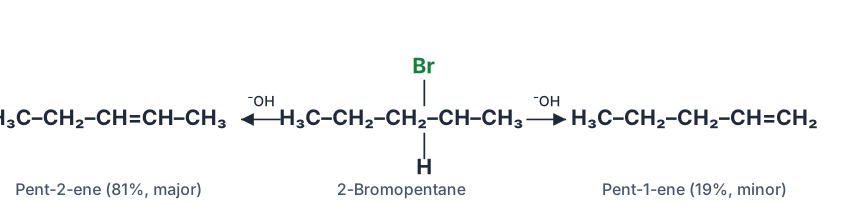
Example: 2-bromobutane has two possible β-hydrogens: …
Part (a)
(i) Bromobenzene + Mg (dry ether). Forms a Grignard reagent, phenylmagnesium bromide:
C6H5Br+MgdryetherC6H5MgBr
(ii) Faster in SN1 (governed by carbocation stability):
- CH2=CH−CH2Cl vs CH3CH2CH2Cl → allyl chloride (resonance-stabilised allyl cation) is faster.
- (CH3)3CCl vs CH3Cl → t-butyl chloride (stable 3° cation) is faster.
(iii) Preparation of 1-iodobutane
- From 1-chlorobutane — Finkelstein reaction: CH3CH2CH2CH2Cl+NaIacetoneCH3CH2CH2CH2I+NaCl↓.
- From but-1-ene — add HBr with peroxide (anti-Markovnikov) to 1-bromobutane, then Finkelstein: …
- Bromobenzene + Mg/ether → phenylmagnesium bromide; SN1 favours the stabler carbocation, so allyl chloride and t-butyl chloride react faster; 1-iodobutane comes from 1-chlorobutane (Finkelstein) or from but-1-ene (HBr/peroxide → 1-bromobutane, then Finkelstein).
- 2-bromopropane + alc. KOH → propene (elimination); benzene + CH3COCl/AlCl3 → acetophenone (Friedel–Crafts acylation).
Part (a)
(i) Bromobenzene with Mg in dry ether
An aryl halide reacts with magnesium in anhydrous ether to give a Grignard reagent:
C6H5Br+MgdryetherC6H5MgBr (phenylmagnesium bromide)
Dry conditions are essential — even traces of water destroy the reagent.
(ii) Which reacts faster in SN1
SN1 rate depends on the stability of the carbocation formed in the slow step.
- CH2=CH−CH2Cl gives a resonance-stabilised allyl cation, far stabler than the primary cation from CH3CH2CH2Cl → allyl chloride is faster.
- (CH3)3CCl gives a stable tertiary cation (hyperconjugation + +I), whereas CH3Cl would give the very unstable CH3+ → t-butyl chloride is faster.
(iii) Preparation of 1-iodobutane
(1) From 1-chlorobutane — Finkelstein reaction: halide exchange with NaI in acetone (NaCl precipitates, driving it forward):
CH3CH2CH2CH2Cl+NaIacetoneCH3CH2CH2CH2I+NaCl↓
(2) From but-1-ene: we need the terminal (anti-Markovnikov) halide, so add HBr in the presence of peroxide to get 1-bromobutane, then convert by Finkelstein:
CH3CH2CH=CH2HBrperoxideCH3CH2CH2CH2BrNaI/acetoneCH3CH2CH2CH2I …
Showing the 12 most recent of 26 on this concept.
- CBSE 2026Set 56/3/11 markMCQQ.Which of the following will be the least reactive towards nucleophilic substitution reaction ? (A) Benzyl chloride (C6H5CH2Cl) (B) 1-chloro-4-methylbenzene (4ext−CH3C6H4Cl) (C) CH3−Cl (D) Chlorocyclohexane (C6H11Cl)
›Reveal solutionSolution
The key idea is that nucleophilic substitution reactivity depends on the stability of the carbocation intermediate (for SN1) or the accessibility of the carbon (for SN2). Among the given options, 1-chloro-4-methylbenzene (4-methylchlorobenzene) is an aryl halide where the chlorine is directly attached to an aromatic ring — its lone pairs are delocalised into the ring, making the C–Cl bond extremely strong and resistant to both SN1 and SN2. The least reactive is therefore option (B).
Why this approach works
Nucleophilic substitution reactions (SN1 and SN2) both require the leaving group (here, Cl⁻) to depart. The ease of this departure depends on:
- For SN1: The stability of the carbocation formed after Cl⁻ leaves. More stable carbocations (tertiary, allylic, benzylic) react faster.
- For SN2: The steric hindrance around the carbon bearing the leaving group. Less hindered carbons (methyl > primary > secondary) react faster.
But there is a special case: when chlorine is directly bonded to an aromatic ring (an aryl halide), the C–Cl bond gains partial double-bond character due to resonance. This makes it much stronger and harder to break — so aryl halides are notoriously unreactive in typical nucleophilic substitutions unless special conditions (like very strong nucleophiles or high temperatures) are used.
Let’s examine each option.
Step-by-step reasoning
1. Option (A): Benzyl chloride (C6H5CH2Cl)
Here, chlorine is on a carbon next to the benzene ring, not directly on it. The benzylic carbocation (C6H5CH2+) is highly stabilised by resonance with the ring. So SN1 is very fast. SN2 is also possible because the benzylic carbon is primary and not too hindered. This compound is highly reactive.
2. Option (B): 1-chloro-4-methylbenzene (4-methylchlorobenzene)
Chlorine is directly attached to the benzene ring. The lone pairs on chlorine participate in resonance with the aromatic π-system, giving the C–Cl bond partial double-bond character. This bond is very strong — about 30–40 kJ/mol stronger than a typical alkyl C–Cl bond. Neither SN1 (no stable carbocation — aryl cations are extremely unstable) nor SN2 (the carbon is sp² hybridised and the backside is blocked by the ring) works under normal conditions. This is the least reactive.
Watch outA common mistake
Students often think that the methyl group on the ring makes it more reactive (like an electron-donating group activating the ring for electrophilic substitution). But for nucleophilic substitution, the methyl group does not help — the fundamental problem is the strong C–Cl bond and the sp² carbon. The methyl group is irrelevant here. …
- CBSE 2026Set V11 markMCQQ.The major product formed in the following reaction is H3C−CH2−Br∣CH−CH3Alc.KOHΔ(a) 1-butene(b) 2-butanol(c) 1-bromobutane(d) 2-butene
›Reveal solutionSolution
Alcoholic KOH removes HBr from 2-bromobutane; by Saytzeff's rule the more substituted alkene, but-2-ene, is the major product.
The reactant H3C−CH2−BrCH−CH3 is 2-bromobutane. Alcoholic KOH is a strong base that promotes β-elimination (dehydrohalogenation), not substitution.
A β-hydrogen can be removed from either the C-1 side or the C-3 side of the C-Br carbon:
- Removal from C-3 (CH2) gives CH3CH=CHCH3 (but-2-ene) — a disubstituted double bond.
- Removal from C-1 (CH3) gives CH2=CHCH2CH3 (but-1-ene) — a monosubstituted double bond. …
- CBSE 2026Set A1 markMCQQ.Ethyl bromide on boiling with alcoholic caustic potash gives(a) Ethyl alcohol(b) Ethylene(c) Acetylene(d) Ethane
›Reveal solutionSolution
Alcoholic KOH removes HBr from ethyl bromide (E2 elimination) to give ethylene.
Alcoholic potassium hydroxide furnishes the strong base ethoxide/hydroxide in an alcoholic medium, which favours elimination over substitution. Ethyl bromide loses a molecule of HBr:
CH3-CH2-Br --(alc. KOH, heat)--> CH2=CH2 + KBr + H2O
…
- CBSE 2026Set ANNUAL1 markMCQQ.SN1 reaction will be fastest in case of(a) Tertiary halide(b) Primary halide(c) Secondary halide(d) None of these
›Reveal solutionSolution
SN1 reaction rate depends on the stability of the carbocation intermediate formed in the rate-determining (ionisation) step; more substituted carbocations are more stable, so more substituted halides react faster.
The SN1 mechanism proceeds in two steps:
- Slow, rate-determining ionisation of R-X to form a carbocation R+ and X-.
- Fast attack of the nucleophile on the carbocation. …
- CBSE 2026Set ANNUAL1 markMCQQ.Assertion [A]: 1-chlorobutane on heating with alc. KOH gives mainly Bute-1-ene. Reason [R]: Bute-1-ene is more stable than Bute-2-ene.(a) Both [A] and [R] are true and [R] is the correct explanation of [A].(b) Both [A] and [R] are true, but [R] is not the correct explanation of [A].(c) [A] is true, but [R] is false.(d) [A] is false, but [R] is true.
›Reveal solutionSolution
1-Chlorobutane does give but-1-ene on heating with alcoholic KOH — but only because it is the ONLY elimination product possible (Cl is on the terminal carbon), not because but-1-ene is more stable than but-2-ene. In fact but-2-ene (especially the trans isomer) is MORE stable than but-1-ene, so the Reason is false.
Assertion: 1-Chlorobutane is CH3CH2CH2CH2Cl — the leaving group (Cl) is on C1, a terminal carbon. β-elimination (E2, Zaitsev/Hofmann considerations) can only remove a β-hydrogen from C2 (there is no carbon "before" C1 to eliminate towards). This necessarily gives:
CH3CH2CH2CH2Clalc. KOH, ΔCH3CH2CH=CH2 (but-1-ene)+KCl+H2O
So yes, but-1-ene IS the (essentially only) product — the Assertion is TRUE.
…
- CBSE 2026Set ANNUAL1 markMCQQ.C2H5Cl on heating with alcoholic KOH will produce:(a) C2H5OH(b) C2H4(c) C2H2(d) C2H6
›Reveal solutionSolution
Alcoholic KOH promotes elimination (dehydrohalogenation), so ethyl chloride gives ethene, not the substitution product ethanol.
When an alkyl halide is heated with KOH dissolved in alcohol, the hydroxide ion acts as a strong base rather than a nucleophile, abstracting a β-hydrogen and eliminating the halide to form an alkene (E2 elimination):
CH3CH2Clalc. KOHΔCH2=CH2+KCl+H2O …
- CBSE 2026Set ANNUAL1 markQ.Write the products of the following reaction: CH3−CH2−CH2−CH(Br)−CH3Alc. KOHA+B
›Reveal solutionSolution
E2 dehydrohalogenation of the secondary bromide 2-bromopentane with alcoholic KOH gives two alkenes; by Zaitsev's rule, the more-substituted one dominates.
Numbering the given chain from the bromine end for IUPAC purposes: CH3(C1)−CHBr(C2)−CH2(C3)−CH2(C4)−CH3(C5) — this is 2-bromopentane. Two sets of β-hydrogens are available for E2 elimination: on C1 (the adjacent methyl) and on C3.
Elimination toward C3 (removing a β-H from C3) gives the internal, more substituted (disubstituted) alkene:
CH3−CH=CH−CH2−CH3(pent-2-ene)
By Zaitsev's rule, the more substituted, more stable alkene (better stabilised by hyperconjugation/alkyl substitution across the double bond) is the major product (A).
…
- CBSE 2026Set ANNUAL1 markQ.Write True or False: Lower aliphatic amines are soluble in water while higher amines are essentially insoluble in water.
›Reveal solutionSolution
True - solubility falls as the amine's carbon chain lengthens.
Lower aliphatic amines (small molecules) can hydrogen-bond with water through their N-H and lone pair, so they dissolve readily in water. As the alkyl chain becomes longer (higher amines), the large hydrophobic hydrocarbon part outweighs the small polar -NH2 group, hydrogen bonding with wa …
- CBSE 2026Set ANNUAL1 markMCQQ.Which of the following is least reactive towards nucleophilic substitution (SN1) ?(a) Benzyl chloride(b) Methyl chloride(c) Chlorobenzene(d) Allyl chloride
›Reveal solutionSolution
SN1 rate follows carbocation stability; benzyl and allyl give resonance-stabilised cations and methyl a simple one, but chlorobenzene (aryl halide) cannot ionise, so it is least reactive. Answer: (c).
- Benzyl chloride and allyl chloride ionise to resonance-stabilised (benzyl / allyl) carbocations -> fast SN1.
- Methyl chloride gives a (poor but possible) methyl cation. …
- CBSE 2025Set 56/6/11 markMCQQ.Which of the following compounds would be hydrolysed by aqueous KOH most easily ? (A) CH2=CH−Br (B) CH3−CH2−Br (C) CH3−CH(Br)−CH3 (D) CH2=CH−CH2−Br
›Reveal solutionSolution
Allylic halides hydrolyse fastest because the carbocation (or transition state) is resonance-stabilized by the adjacent π-system. The correct option is (D).
Understanding SN1 Reactivity and Carbocation Stability
Hydrolysis by aqueous KOH can proceed through two pathways: SN2 (bimolecular substitution) or SN1 (unimolecular, carbocation-mediated). When we ask which compound hydrolyses "most easily," we're really asking which forms the most stable intermediate or transition state.
The key insight: carbocation stability dictates SN1 reactivity, and resonance stabilization trumps inductive effects. Let's examine each structure.
Step-by-Step Analysis
1. Identify the type of halide in each compound
- (A) CH2=CH−Br: Vinyl halide (Br directly on sp2 carbon)
- (B) CH3−CH2−Br: Primary alkyl halide
- (C) CH3−CH(Br)−CH3: Secondary alkyl halide
- (D) CH2=CH−CH2−Br: Allylic halide (Br on carbon adjacent to C=C)
2. Evaluate vinyl halide (A)
Vinyl halides are notoriously unreactive toward both SN1 and SN2. The C–Br bond has significant sp2 character (shorter, stronger), and the hypothetical vinyl cation would be extremely unstable due to the electron-withdrawing effect of the sp2 hybridized carbon. This compound is essentially inert under typical hydrolysis conditions.
Watch outNever expect a vinyl or aryl halide to undergo simple nucleophilic substitution — the carbocation would be far too high in energy.
3. Compare primary (B) vs. secondary (C) alkyl halides
- Primary carbocation: highly unstable, so (B) proceeds mainly via SN2 (slow with weak nucleophile in aqueous medium)
- Secondary carbocation: more stable than primary due to hyperconjugation from two adjacent alkyl groups, so (C) can proceed via SN1, but still not particularly fast
The order so far: (C) > (B) >> (A).
4. Recognize the allylic system in (D)
When CH2=CH−CH2−Br ionizes, it forms the allyl cation CH2=CH−CH2+. This cation is resonance-stabilized: …
- CBSE 2025Set ANNUAL1 markMCQQ.Alkyl halides react with alcoholic KOH to give:(a) Alkanes(b) Alkenes(c) Alcohols(d) Ethers
›Reveal solutionSolution
Alcoholic KOH is a strong base used for elimination; it removes H and X from adjacent carbons of an alkyl halide to give an alkene.
When an alkyl halide (R-CH2-CH2-X) is treated with KOH dissolved in alcohol, the hydroxide ion acts as a base rather than a nucleophile. It abstracts a β-hydrogen (a hydrogen on the carbon next to the one bearing the halogen), and the halide ion leaves simultaneously (E2 mechanism), forming a carbon-carbon double bond:
CH3-CH2-X + KOH(alc.) → CH2=CH2 + KX + H2O
…
- CBSE 2025Set ANNUAL1 markMCQQ.Which of the following is most reactive towards nucleophilic substitution reaction –(i) C₆H₅Cl(ii) CH₂=CHCl(iii) ClCH₂CH=CH₂(iv) CH₃-CH=CHCl
›Reveal solutionSolution
Allylic halides (like ClCH₂CH=CH₂) are the most reactive towards nucleophilic substitution because the halogen is on an sp³ carbon and ionisation gives a resonance-stabilised allylic carbocation.
Compare the four halides:
- C₆H₅Cl (aryl halide): Cl is attached to an sp² ring carbon; the lone pair on Cl delocalises into the ring (resonance), giving the C–Cl bond partial double-bond character. This makes the bond strong and short, so it strongly resists nucleophilic substitution.
- CH₂=CHCl (vinyl halide): Cl is directly on the sp² double-bond carbon; the same resonance effect (halogen lone pair conjugating with the π bond) makes it unreactive.
- CH₃-CH=CHCl: Cl is again on an sp² vinylic carbon (same reason as above) — unreactive. …
🎓Unlock everything free for 14 days
- ✓Full step-by-step solutions
- ✓Concept-first explanations
- ✓Methods, shortcuts & mistakes
- ✓PYQ mapping + timed mock tests
Full access for 14 days. No credit card required.