Q.Ortho and para nitrophenols are more acidic than phenol. Draw the resonance structures of the corresponding phenoxide ions.
🔒You're viewing a preview — the full solution, concept, methods & PYQ mapping are locked.
🔒 Start your 14-day free trial to unlock the full solution →Concept understanding — Electrophilic Aromatic Substitution
Electrophilic Aromatic Substitution – The First Meeting
Imagine you have a benzene ring — that perfect, flat hexagon of six carbons with alternating double bonds. It's stable, almost stubbornly so. You want to attach something new to it, say a bromine atom or a nitro group. But benzene doesn't react like an alkene. It doesn't just add across a double bond. Instead, it does something more elegant: it kicks out a hydrogen and keeps its aromatic ring intact.
That's the heart of Electrophilic Aromatic Substitution (EAS).
The Intuition: Why "Substitution" and Not "Addition"?
Benzene's stability comes from its delocalised π electrons — a cloud above and below the ring. This cloud is electron-rich, so it attracts electrophiles (electron-loving species). But if an electrophile simply added to a double bond, the ring would break its aromaticity, losing that huge stabilisation. That would be energetically costly.
So benzene does something smarter: it lets the electrophile attack, temporarily breaks aromaticity to form a high-energy intermediate (the arenium ion), and then loses a proton to restore the aromatic ring. The net result? A hydrogen is replaced by the electrophile. The ring is back to its stable, aromatic self.
The key trade-off: temporary loss of aromaticity is acceptable because the final product regains it. Addition reactions would permanently destroy aromaticity — benzene avoids that.
The Precise Statement
Electrophilic Aromatic Substitution is a reaction in which an electrophile (E+) replaces a hydrogen atom on an aromatic ring, proceeding through a sigma complex (arenium ion) intermediate, and restoring aromaticity after deprotonation.
The general equation:
Ar−H+EX+Ar−E+HX+
where Ar represents an aromatic ring.
The Mechanism in Three Steps
-
Generation of the electrophile – Many EAS reactions need a catalyst to create a strong enough E+. For example, bromination uses FeBrX3 to polarise BrX2 into BrX+.
-
Attack by the aromatic ring – The π electrons of benzene attack the electrophile, forming a sigma complex (also called the arenium ion or Wheland intermediate). This intermediate is non-aromatic — it has four π electrons delocalised over five carbons, and one sp3 carbon bearing the electrophile and a hydrogen.
Benzene+E+⟶Sigma complex (non-aromatic)
- Deprotonation – A base (often the counterion of the catalyst, like FeBrX4X−) removes the proton from the sp3 carbon. The pair of electrons from the C–H bond flows back into the ring, restoring the aromatic sextet.
Sigma complex+Base⟶Product+HX+
The sigma complex is not aromatic. It's a high-energy intermediate. Students often mistakenly think it's still aromatic — it isn't. That's why the step is fast and the complex is short-lived.
Why This Matters for Exams
EAS is the gateway to understanding how to put groups onto benzene rings. The rate-determining step is usually the formation of the sigma complex (step 2). The regiochemistry (where the electrophile goes) depends on whether the ring already has a substituent — that's the topic of activating/deactivating groups and ortho/para vs. meta directors.
But for now, remember this: …
Why this formula?
Electrophilic Aromatic Substitution: Why the Mechanism Holds
The Core Puzzle: Why Benzene Doesn't Just Add
Benzene (C6H6) has three double bonds — so why doesn't it undergo addition reactions like alkenes?
The answer lies in aromatic stabilisation: benzene's delocalised π-electron cloud (the "aromatic sextet") is about 150 kJ/mol more stable than a hypothetical cyclohexatriene with localised double bonds.
If benzene simply added an electrophile (like Br2), it would lose this stabilisation — a huge energy penalty.
So, nature chooses a different path: substitution instead of addition, preserving the aromatic ring.
The Key Formula: The Reaction Profile
The rate-determining step in electrophilic aromatic substitution (EAS) is the formation of the arenium ion (σ-complex):
Ar-H+E+slowAr-E+H(σ-complex)
Then, fast deprotonation restores aromaticity:
Ar-E+H+B−fastAr-E+BH
Why This Holds: The Energy Barrier Logic
-
First step (slow): The electrophile E+ attacks the electron-rich ring. The σ-complex is non-aromatic — it has only 4 π-electrons delocalised over 5 carbons (the sixth carbon is sp3 hybridised). This intermediate is higher in energy than the starting benzene.
-
Second step (fast): A base removes the proton, restoring the aromatic sextet. This step is strongly exothermic — the system regains ~150 kJ/mol of stabilisation.
The overall reaction is exothermic, but the activation energy is dominated by the destabilisation of the σ-complex.
The Rate Law: Why It's First Order in Both
From the mechanism:
Rate=k[Arene][E+]
Reasoning:
- The slow step involves one molecule of arene and one molecule of electrophile.
- No other species appear before the rate-determining step.
- Therefore, the rate law is bimolecular — first order in each reactant.
This is not derived from the overall stoichiometry — it comes directly from the molecularity of the slow step.
The Hammett Equation: Quantifying Substituent Effects
For substituted benzenes, the rate constant k relative to benzene (k0) follows:
logk0k=σρ
Why This Holds
- σ (sigma constant): Measures the electronic effect of a substituent (electron-donating or withdrawing) relative to hydrogen. It is derived from the ionisation constants of benzoic acids — a purely empirical scale.
- ρ (rho constant): Measures the sensitivity of the reaction to substituent effects. A positive ρ means the reaction is favoured by electron-withdrawing groups (rare in EAS); a negative ρ means electron-donating groups accelerate the reaction.
Why it works:
The σ-complex has a positive charge delocalised over the ring. Substituents that stabilise this positive charge (electron-donating groups like −OH, −NH2) lower the activation energy — hence σ is negative for such groups. Electron-withdrawing groups (−NO2, −CN) destabilise the σ-complex — σ is positive.
The linear free-energy relationship holds because the transition state resembles the σ-complex in charge distribution.
The Directing Effect: Why Ortho/Para vs Meta
The position of substitution is governed by the stability of the σ-complex for each possible attack site. …
The key idea is that the nitro group stabilises the phenoxide ion through resonance, but only when it is ortho or para to the oxygen — meta nitrophenol is not more acidic than phenol for this reason.
Reasoning:
- Deprotonation of phenol gives the phenoxide ion, where the negative charge is delocalised into the ring (ortho and para positions carry partial negative charge).
- A nitro group at the ortho or para position can directly accept this negative charge via resonance, forming additional stable structures where the negative charge is on the highly electronegative oxygen atoms of the nitro group. …
The higher acidity of ortho- and para-nitrophenols arises because the nitro group stabilises the conjugate base (phenoxide ion) by delocalising the negative charge through resonance -- the ortho and para positions allow direct conjugation with the nitro group, while the meta position does not.
Why this happens -- the concept
Acidity is all about the stability of the conjugate base. For phenols, the conjugate base is the phenoxide ion. Phenol itself is weakly acidic because the negative charge on oxygen can be delocalised into the benzene ring. But when a nitro group (−NO2) is present, it is a strongly electron-withdrawing group -- it pulls electron density away from the ring. This further stabilises the phenoxide ion, making the corresponding phenol more acidic.
The key is where the nitro group is attached. The nitro group withdraws electrons both by induction and by resonance. The resonance effect is especially powerful when the nitro group is at the ortho or para position because the negative charge on the phenoxide oxygen can be delocalised onto the nitro group itself. At the meta position, this direct conjugation is not possible -- the negative charge cannot reach the nitro group through resonance.
A common mistake is to think that the nitro group withdraws electrons equally from all positions. It does not -- the resonance effect is strongly position-dependent. Only ortho and para positions allow the negative charge to be delocalised onto the nitro group.
Step-by-step reasoning
-
Start with the phenoxide ion from phenol itself. The negative charge on oxygen can be delocalised into the ring, giving resonance structures where the charge appears at the ortho and para positions (relative to the −O− group). This is why phenol is more acidic than a simple alcohol -- the charge is spread out.
-
Now consider ortho-nitrophenol. The nitro group is at the ortho position relative to the −OH group. When the phenol loses a proton, the phenoxide ion forms, and its negative charge can be delocalised onto the nitro group through resonance:
- One resonance structure has the negative charge on the phenoxide oxygen, with the ring drawn in its normal alternating-bond form.
- A second resonance structure moves the negative charge onto the ring carbon that bears the nitro group (the ortho carbon), with the ring's double bonds shifted accordingly.
- A third, key resonance structure pushes that charge further onto one of the two oxygen atoms of the nitro group itself, with the nitrogen now bearing a formal positive charge (as in the nitro group's own normal resonance form, −N+(=O)(−O−)).
The result: the negative charge is spread over three oxygen atoms (the phenoxide oxygen and the two oxygens of the nitro group). This is much more stable than the phenoxide ion from phenol, where the charge is only on the ring carbons and one oxygen.
-
Now consider para-nitrophenol. Exactly the same logic applies, but now the nitro group is at the para position. The resonance delocalisation works just as well -- the negative charge travels through the ring (via the para carbon this time) and ends up on the nitro group's oxygens, again spread over three oxygen atoms in the most stabilised resonance form.
-
Now consider meta-nitrophenol (for contrast). Here, the nitro group is at the meta position. When you try to draw resonance structures that put the negative charge on the carbon bearing the nitro group, you find it is impossible -- the meta position is not connected to the oxygen by a conjugated path that allows the charge to reach the nitro group. The nitro group can still withdraw electrons inductively (through sigma bonds), but the powerful resonance stabilisation is absent. So meta-nitrophenol is less acidic than ortho- and para-nitrophenols, though still more acidic than phenol itself.
A quick way to remember: the nitro group is a resonance acceptor at ortho and para positions. If you can draw a resonance structure where the negative charge ends up on an oxygen of the nitro group, you have extra stabilisation. If you cannot (as in meta), you don't.
The resonance structures of the phenoxide ions, described …
Method: Resonance Stabilisation Analysis of Conjugate Bases
This method explains why ortho- and para-nitrophenols are more acidic than phenol by comparing the stability of their conjugate bases (phenoxide ions) through resonance.
Step 1: Draw the resonance structures of phenoxide ion (the conjugate base of phenol)
The phenoxide ion has the negative charge delocalised into the ring:
    O⁻
    |
    ══╗
   ║   ║
    ══╝
Resonance structures (only the major ones):
- Original structure with negative charge on oxygen.
- Charge delocalised to ortho position (C-2).
- Charge delocalised to para position (C-4).
- Charge delocalised to the other ortho position (C-6).
Key observation: In phenol, the negative charge is delocalised only onto ortho and para carbons — not onto the meta carbon.
Step 2: Draw the resonance structures of ortho-nitrophenoxide ion
The nitro group (−NO2) is a strong electron-withdrawing group by both inductive and resonance effects. When present at the ortho position, it can directly accept the negative charge via resonance.
Resonance structures (focus on the key extra stabilisation):
- Negative charge on oxygen delocalises into the ring as in phenol.
- Additional structure: The negative charge moves onto the nitro group oxygen:
    O⁻
    |
    ══╗
   ║   ║
    ══╝
    |
    N⁺
   / \
  O   O⁻
This structure places the negative charge on an electronegative oxygen of the nitro group — this is highly stabilising.
Step 3: Draw the resonance structures of para-nitrophenoxide ion
Exactly analogous to ortho — the nitro group at the para position can also accept the negative charge via resonance:
- Same delocalisation as phenol.
- Additional structure: Negative charge moves onto the nitro group oxygen at the para position.
Critical point: In meta-nitrophenol, the nitro group cannot accept the negative charge through resonance because the negative charge never reaches the meta position. Hence, meta-nitrophenol is less acidic than ortho- and para-nitrophenols.
Step 4: Compare stability of conjugate bases …
Here is a breakdown of the common mistakes students make on this specific Electrophilic Aromatic Substitution (EAS) concept, along with how to avoid them.
Mistake 1: Drawing the wrong resonance for the phenoxide ion
The Error: Students often draw resonance structures for the neutral phenol molecule instead of the phenoxide ion (the conjugate base). The question explicitly asks for the phenoxide ion.
Why it’s wrong: The acidity of phenol is determined by the stability of its conjugate base (the phenoxide ion). The negative charge on the oxygen can be delocalized into the ring. If you draw the neutral molecule, you miss the entire point of charge stabilization.
How to Avoid:
- Read the question carefully. Underline "phenoxide ions."
- Start with the correct structure: Draw the benzene ring with an OX− (negative charge) attached.
- Remember the rule: The negative charge moves into the ring via resonance. The oxygen atom becomes a double bond to the ring, pushing the negative charge onto a carbon atom (usually ortho or para to the oxygen).
Mistake 2: Forgetting the nitro group's role in the phenoxide ion
The Error: Students draw the same resonance structures for ortho-nitrophenoxide and para-nitrophenoxide as they do for plain phenoxide. They fail to show how the nitro group (−NOX2) specifically stabilizes the negative charge.
Why it’s wrong: The nitro group is a strong electron-withdrawing group (EWG) via both induction and resonance. Its key role is to directly accept the negative charge from the ring. If you don't show the negative charge on the nitro group's oxygen atoms, you haven't explained why ortho/para nitrophenols are more acidic.
How to Avoid:
- Identify the special resonance: When the negative charge from the phenoxide ion lands on the carbon ortho or para to the nitro group, it can be delocalized onto the nitro group itself.
- Draw the extra step: Show the double bond from the ring carbon to the nitrogen of the −NOX2 group, breaking one of the N=O bonds and placing the negative charge on an oxygen of the nitro group.
- Key insight: This creates a quinoid-like structure with the negative charge on a highly electronegative oxygen atom. This is a major contributor to stability.
Mistake 3: Not showing the complete set of resonance structures
The Error: Students draw only 2-3 resonance structures and stop. They miss the crucial structure where the negative charge is on the nitro group.
Why it’s wrong: The exam expects you to show all significant contributors. The structure with the charge on the nitro group is the most important one for explaining the increased acidity. Missing it means you lose marks and fail to demonstrate the core concept.
How to Avoid:
- Use a systematic method:
- Start with the phenoxide ion (charge on O).
- Move the charge to the ortho and para positions of the ring (3 structures for plain phenoxide).
- For ortho-nitrophenoxide: When the charge is on the carbon ortho to the nitro group, draw the extra resonance onto the nitro group.
- For para-nitrophenoxide: When the charge is on the carbon para to the nitro group, draw the extra resonance onto the nitro group.
- Count them: For para-nitrophenoxide, you should have 5 significant resonance structures. For ortho-nitrophenoxide, you should have 5 as well (including the one on the nitro group).
Mistake 4: Confusing ortho and para directing effects with acidity
The Error: Students think that because the nitro group is a meta-director in EAS, it cannot stabilize a negative charge in the ortho or para position. …
- COMEDK 2026Set 2026-A1 markMCQQ.
[!FORMULA] What is the major product [P3] formed when n - Hexane undergoes the given series of reactions: n-Hexane [P1][P2]20 atm,773kV2O5[P1]C2H5Cl, Anhyd.AlCl △[P2] Conc. HNO3+H2SO4333 K[P3] Major product
(A) (B) (C) (D)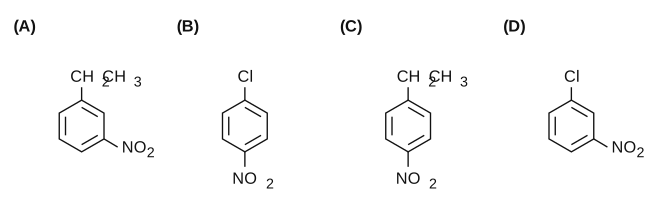
›Reveal solutionSolution
The reaction sequence converts n-hexane to benzene (via aromatization), then ethylates it to ethylbenzene, and finally nitrates it to give para-ethylnitrobenzene as the major product. The correct option is (C).
The key to this problem is recognizing that each step in the sequence is a classic organic transformation, and the final product’s structure is determined by the directing effects of substituents already on the ring. Let’s walk through it step by step.
-
First step: Aromatization of n-hexane
n-Hexane (C6H14) is treated with V2O5 at 20 atm and 773 K. This is a typical catalytic reforming or dehydrogenation condition. Vanadium pentoxide acts as a catalyst to cyclize and dehydrogenate the straight-chain alkane into an aromatic ring. The product [P1] is benzene (C6H6).
Why? Under high temperature and pressure, alkanes with six or more carbons can undergo dehydrocyclization — losing hydrogen atoms and forming a stable aromatic ring. No other functional groups are introduced here.
-
Second step: Friedel–Crafts alkylation
Benzene ([P1]) reacts with ethyl chloride (C2H5Cl) in the presence of anhydrous AlCl3 under heat (△). This is a classic Friedel–Crafts alkylation: the AlCl3 generates an ethyl carbocation (or an ethyl–AlCl3 complex) that attacks the benzene ring.
The product [P2] is ethylbenzene (C6H5CH2CH3).
Note: Alkylation gives a mono-substituted product because the ethyl group is an activating, ortho/para-directing group, but under these conditions further alkylation is minimal (or we assume mono-substitution as the intended step).
-
Third step: Nitration of ethylbenzene
Ethylbenzene ([P2]) is treated with concentrated nitric acid and sulfuric acid (a nitrating mixture) at 333 K. This is an electrophilic aromatic substitution. The nitronium ion (NO2+) is the attacking species.
The ethyl group is an ortho/para-directing group (it donates electron density via hyperconjugation and inductive effect). Therefore, the nitro group will preferentially attach at the ortho or para positions relative to the ethyl group. …
-
- COMEDK 2026Set 2026-M1 markMCQQ.
[!FORMULA] The decreasing order of reactivity towards electrophilic substitutions is:
(A) IV>I>II>IH (B) III>I>II>IV (C) I>II>IH>IV (D) I>III>IV>II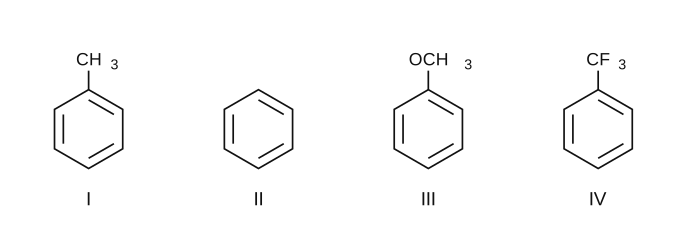
›Reveal solutionSolution
The reactivity of a benzene ring toward electrophilic substitution is governed by the electron-donating or electron-withdrawing nature of its substituent. The correct decreasing order is III (anisole) > I (toluene) > II (benzene) > IV (trifluoromethylbenzene), which corresponds to option (B).
Concept & Intuition
Electrophilic substitution on benzene proceeds via a positively charged intermediate (the arenium ion). Substituents that donate electrons (by resonance or induction) stabilize this intermediate and activate the ring, making it more reactive than benzene itself. Substituents that withdraw electrons destabilize the intermediate and deactivate the ring. The key is to compare the net electronic effect of each group:
- –OCH₃ (methoxy): Strongly activating. The oxygen lone pairs donate into the ring by resonance, overwhelming the weak inductive withdrawal.
- –CH₃ (methyl): Mildly activating. It donates electrons by hyperconjugation and the +I effect.
- –H (benzene): The reference point — no substituent effect.
- –CF₃ (trifluoromethyl): Strongly deactivating. The three fluorine atoms pull electrons inductively, and there is no resonance donation to compensate.
Thus the order is: anisole > toluene > benzene > trifluoromethylbenzene.
Step-by-step reasoning
-
Identify the substituent effects
- III (–OCH₃): The oxygen lone pairs are conjugated with the π-system, creating strong resonance donation (the methoxy group is ortho/para-directing and strongly activating).
- I (–CH₃): The methyl group donates via hyperconjugation (C–H σ bonds overlap with the ring π-orbitals) and a small +I effect. This is a mild activator.
- II (–H): No substituent — reactivity is the baseline.
- IV (–CF₃): The highly electronegative fluorines pull electron density through σ-bonds (strong –I effect). No resonance donation is possible, so the ring becomes electron-poor and deactivated.
-
Rank the activating power
Strongest activator → weakest activator → deactivator:
–OCH₃ > –CH₃ > –H > –CF₃.
-
Translate to reactivity order …
- COMEDK 2026Set 2026-M1 markMCQQ. Match reactions in Column I with the corresponding products formed as given in Column II. .tg {border-collapse:collapse;border-spacing:0;} .tg td{border-color:black;border-style:solid;border-width:1px;font-family:Arial, sans-serif;font-size:14px; overflow:hidden;padding:10px 5px;word-break:normal;} .tg th{border-color:black;border-style:solid;border-width:1px;font-family:Arial, sans-serif;font-size:14px; font-weight:normal;overflow:hidden;padding:10px 5px;word-break:normal;} .tg .tg-baqh{text-align:center;vertical-align:top} Column I Column I A C6H5 N2+Cl−+ warm H2O→ P p-Bromophenol B C6H5OH+273 KBr2 in CS2→ Q 2,4,6-Tribromophenol C C6H5OH+( ii )HCl( i )CHCl3+NaOH( aq )→ R Phenol D C6H5OH+Br2(aq)→ S 2-Hydroxybenzaldehyde (A) A−RB−PC−SD−Q (B) A−QB−PC−SD−R (C) A−SB−RC−QD−P (D) A−QB−RC−SD−P
›Reveal solutionSolution
The key is to match each reaction with its characteristic product by recalling the specific conditions: diazonium hydrolysis gives phenol, bromination in CS₂ at low temperature gives p-bromophenol, Reimer–Tiemann reaction gives salicylaldehyde, and aqueous bromination gives 2,4,6-tribromophenol. The correct matching is A→R, B→P, C→S, D→Q, which corresponds to option (A).
Concept and Intuition
This problem tests your knowledge of four classic organic reactions of benzene derivatives. Each reaction is distinguished by the reagent and conditions, which dictate the product. Instead of memorizing blindly, think about why each condition leads to a specific outcome:
- Diazonium salts are excellent leaving groups; in warm water, they simply hydrolyze to give phenol.
- Bromination of phenol can be controlled: in a non-polar solvent (CS₂) at low temperature, only one bromine substitutes at the para position (due to steric and electronic control). In aqueous solution, the reaction is much more vigorous, leading to trisubstitution.
- Reimer–Tiemann reaction uses chloroform and strong base to install a formyl group ortho to the phenol – a classic example of a carbene intermediate.
Let’s go through each match step by step.
Step-by-Step Matching
1. Reaction A: Diazonium salt with warm water
- Benzenediazonium chloride (C6H5N2+Cl−) is unstable in water. The N2+ group is a very good leaving group, and upon heating, it is replaced by a hydroxyl group from water.
- Product: Phenol (C6H5OH).
- So A → R (Phenol).
2. Reaction B: Phenol with Br2 in CS2 at 273 K
- In a non-polar solvent like carbon disulfide and at low temperature, bromination of phenol is monosubstitution. The –OH group is strongly activating and ortho/para-directing. However, the ortho positions are somewhat sterically hindered; the major product is the para isomer.
- Product: p-Bromophenol.
- So B → P (p-Bromophenol).
3. Reaction C: Phenol with CHCl3 and aqueous NaOH, then HCl
- This is the Reimer–Tiemann reaction. Chloroform in the presence of strong base generates dichlorocarbene (:CCl2), which attacks the electron-rich ortho position of the phenoxide ion. After hydrolysis with HCl, the product is an ortho-hydroxybenzaldehyde.
- Product: 2-Hydroxybenzaldehyde (salicylaldehyde). …
- KCET 2026Set D31 markMCQQ.Nitration of aniline in strong acidic medium gives significant amount of m-nitroaniline because (A) In electrophilic substitution reaction, amino group is meta directing (B) In strong acidic medium, aniline is present as anilinium ion (C) -NH2 group always directs to meta position (D) m-nitroaniline has higher molar mass than o&p nitroanilines
›Reveal solutionSolution
Protonation of aniline's amino group under strongly acidic nitrating conditions removes the usual ortho/para-directing effect of −NH2 and replaces it with the meta-directing effect of the positively charged −NH3+ group.
Step 1 — Normal behaviour of aniline
Free aniline's −NH2 group has a lone pair that donates electron density into the ring by resonance, strongly activating the ring and directing electrophiles to the ortho and para positions.
Step 2 — What happens in strongly acidic medium
Nitration uses a mixture of concentrated HNO3 and H2SO4, a highly acidic environment. Under these conditions, the nitrogen lone pair of aniline is protonated before nitration can occur, converting −NH2 into −NH3+ (the anilinium ion).
Step 3 — Why anilinium directs meta …
- KCET 2025Set D-41 markMCQQ.Which of the following reagents are suitable to differentiate Aniline and N-methylaniline chemical (A) Acetic anhydride (B) Br2 water (C) Conc. Hydrochloric acid and anhydrous zinc chloride (D) Chloroform and Alcoholic potassium hydroxide
›Reveal solutionSolution
The two compounds differ only in being a primary versus a secondary amine, so we need a test that is exclusive to primary amines — the carbylamine test (CHCl3 + alc. KOH).
Step 1 — Classify the two amines.
- Aniline, C6H5−NH2 — the nitrogen carries two hydrogens and one carbon → primary (1°) aromatic amine.
- N-methylaniline, C6H5−NH−CH3 — the nitrogen carries one hydrogen and two carbons → secondary (2°) amine.
So any reagent that distinguishes them must respond differently to 1° versus 2° amines. Anything that only probes the aromatic ring or the basicity of −NH− will react with both.
Step 2 — Test each reagent.
(A) Acetic anhydride. Acetylation needs an N–H bond. Aniline has two, N-methylaniline has one — both react:
C6H5NH2+(CH3CO)2O⟶C6H5NHCOCH3+CH3COOH
C6H5NH(CH3)+(CH3CO)2O⟶C6H5N(CH3)COCH3+CH3COOH
Both give a substituted amide, so no distinction.
(B) Br2 water. The −NH2 and −NHCH3 groups are both strongly activating, ortho/para-directing groups. Both anilines therefore undergo rapid ring bromination to give a white precipitate of the 2,4,6-tribromo derivative. No distinction.
(C) Conc. HCl + anhydrous ZnCl2. This is Lucas reagent — it is a test for alcohols (1°/2°/3°), distinguishing them by how fast a turbidity of the alkyl chloride appears. It has nothing to do with telling two amines apart; both amines simply form soluble ammonium salts with the acid. Not applicable. …
- COMEDK 2025Set 2025-M1 markMCQQ.What is the final product [Z] formed when the given reactions take place? (A) 4-Bromonitrobenzene. (B) 2-Bromonitrobenzene. (C) 2, 4, 6-tribromonitrobenzene. (D) 3-Bromonitrobenzene.
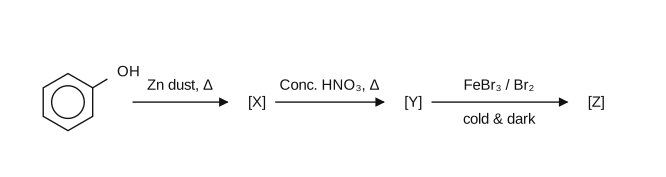
›Reveal solutionSolution
The reaction sequence converts phenol to benzene (via reduction), then nitrates it to nitrobenzene, and finally brominates it at the meta position relative to the nitro group, giving 3-bromonitrobenzene. The correct option is (D).
Concept & Intuition
This problem tests your understanding of how functional groups direct electrophilic aromatic substitution, and how a reaction sequence can completely change the directing effect. The key is to track the activating/deactivating and directing nature of the substituent at each step. Phenol’s –OH group is strongly activating and ortho/para-directing. But the first step (Zn dust, heat) removes that –OH entirely, giving benzene. Then nitration adds a meta-directing nitro group. Finally, bromination under conditions that prevent side reactions (cold & dark, FeBr₃) will place bromine meta to the nitro group. A common pitfall is to forget that the –OH is gone after step 1, and to mistakenly apply phenol’s directing effects to later steps.
Step-by-step reasoning
-
Step 1: Reduction of phenol to benzene
Phenol (CX6HX5OH) is heated with zinc dust. This is a classic reduction that removes the hydroxyl group, replacing it with hydrogen. The product [X] is benzene (CX6HX6).
Why? Zinc dust at high temperature acts as a reducing agent, cleaving the C–O bond and adding H (from trace moisture or the Zn itself). No substituent remains to direct further reactions — we now have a plain benzene ring.
-
Step 2: Nitration of benzene to nitrobenzene
Benzene [X] is treated with concentrated nitric acid and heat. This is standard electrophilic nitration. The product [Y] is nitrobenzene (CX6HX5NOX2).
Why? The nitronium ion (NOX2X+) attacks the benzene ring. Since there is no activating or deactivating group yet, the first substitution occurs at any position equally, but the product is simply nitrobenzene. The nitro group is strongly deactivating and meta-directing for further substitutions.
-
Step 3: Bromination of nitrobenzene …
-
- COMEDK 2024Set 2024-A1 markMCQQ.Arrange the following compounds in the decreasing order of reactivity towards electrophilic substitution reaction. (I) Chlorobenzene (II) Nitrobenzene (III) Benzene (IV) Isopropylbenzene (A) IV > III > I > II (B) I>II>IV>III (C) IV>I>II>III (D) III>II>IV>I
›Reveal solutionSolution
Electrophilic aromatic substitution is fastest on the ring bearing an activating group and slowest on the ring bearing a strong deactivator: isopropyl (activating) > benzene (reference) > chloro (weak deactivator) > nitro (strong deactivator), i.e. IV > III > I > II.
Rate of electrophilic substitution tracks the electron density of the ring:
- Isopropylbenzene (IV): the isopropyl group is electron-donating (+I, hyperconjugation) → activated, most reactive.
- Benzene (III): the unsubstituted reference. …
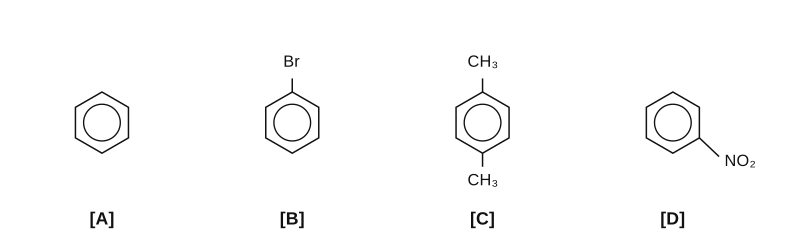
- COMEDK 2024Set 2024-E1 markMCQQ.Arrange the following compounds in the increasing order of their reactivity when each of them is reacted with chloroethane / anhydrous AlCl3. (A) D < A < B < C (B) C < D < B < A (C) D < B < A < C (D) C < B < A < D 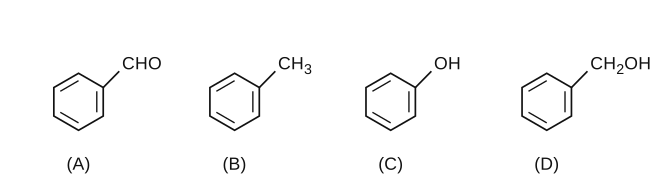
›Reveal solutionSolution
Benzene undergoes a Gattermann–Koch formylation with CO and HCl in the presence of anhydrous AlCl₃, giving benzaldehyde as the product.
The reaction you’re asking about is the Gattermann–Koch reaction — a classic method to introduce an aldehyde group directly onto an aromatic ring. The key idea is that carbon monoxide (CO) and hydrogen chloride (HCl) combine, under the Lewis acid catalysis of anhydrous AlCl₃, to generate a reactive electrophile: the formyl cation (HCO+). This electrophile then attacks the electron-rich benzene ring in an electrophilic aromatic substitution.
Why does this work? AlCl₃ coordinates with CO, polarising it, and then HCl provides the proton, forming a complex that effectively acts as HCO+. This is a strong electrophile, capable of substituting a hydrogen on benzene. The product is benzaldehyde — no further substitution occurs because the aldehyde group is deactivating and meta-directing, so the reaction stops at mono-formylation.
Let’s walk through the mechanism step by step.
- Generation of the electrophile Anhydrous AlCl₃ (a strong Lewis acid) coordinates with the oxygen of carbon monoxide, pulling electron density away from the carbon. This makes the carbon highly electrophilic. HCl then adds, forming a complex that can be thought of as HCO+ (formyl cation) stabilised by AlCl4−:
CO+HCl+AlCl3→HCO+AlCl4−
- Electrophilic attack on benzene The formyl cation attacks the benzene ring, forming a sigma complex (arenium ion). The positive charge is delocalised over the ring:
C6H6+HCO+→C6H6−CHO+
- Deprotonation to restore aromaticity The AlCl4− ion (or a chloride ion) abstracts a proton from the sigma complex, regenerating the aromatic ring and yielding benzaldehyde: …
🎓Unlock everything free for 14 days
- ✓Full step-by-step solutions
- ✓Concept-first explanations
- ✓Methods, shortcuts & mistakes
- ✓PYQ mapping + timed mock tests
Full access for 14 days. No credit card required.