Q.Write chemical equations for the following conversions:
Concept understanding — Nucleophilic Substitution Reactions
Nucleophilic Addition – From Intuition to Precision
Imagine you have a molecule with a carbon–oxygen double bond — a carbonyl group (C=O). That oxygen is greedy for electrons; it pulls them away from carbon, leaving the carbon slightly positive (δ+) and the oxygen slightly negative (δ−). Now, if you bring a species that is rich in electrons (a nucleophile, meaning "nucleus-loving"), it will naturally be attracted to that electron-deficient carbon. The nucleophile attacks the carbon, the π bond breaks, and the oxygen picks up a proton (or some other electrophile) to become stable. That, in a nutshell, is nucleophilic addition.
The key idea: a nucleophile adds across a polar multiple bond (usually C=O or C≡N), breaking the π bond and forming two new sigma bonds.
The Precise Statement
Nucleophilic addition is a reaction in which a nucleophile (an electron-rich species) forms a sigma bond with an electrophilic carbon atom of a polar multiple bond (typically a carbonyl group, C=O, or a nitrile, C≡N), while the π bond breaks. The resulting intermediate then captures a proton (or another electrophile) to give a neutral product.
In general form:
RX2C=O+NuX−HX+RX2C(OH)Nu
The nucleophile (NuX−) attacks the carbonyl carbon; the oxygen becomes negatively charged; then a proton (HX+) from the medium attaches to the oxygen, yielding an alcohol.
Why It Happens – The Driving Force
The carbonyl carbon is electrophilic because:
- Oxygen is more electronegative than carbon, so the C=O bond is polarised: CXδ+=OXδ−.
- The π bond is weaker than a σ bond, so it can break relatively easily.
A nucleophile (like OHX−, CNX−, or NHX3) has a lone pair or a negative charge. It seeks positive centres. The attack forms a new σ bond, and the π electrons move entirely to oxygen, creating an alkoxide ion (RX2C−OX−). This intermediate is then quenched by a proton.
A common mistake: thinking the nucleophile attacks the oxygen. No — oxygen is already electron-rich; the nucleophile goes to the carbon because it is electron-deficient.
A Concrete Example – Addition of HCN to a Ketone
Take acetone (CHX3COCHX3) and hydrogen cyanide (HCN). In the presence of a base, CNX− (the nucleophile) attacks the carbonyl carbon:
CHX3COCHX3+CNX−CHX3C(OX−)(CN)CHX3
The alkoxide intermediate then picks up a proton from HCN (or from water) to give a cyanohydrin:
CHX3C(OX−)(CN)CHX3+HX+CHX3C(OH)(CN)CHX3
The product is acetone cyanohydrin. Notice: two new sigma bonds formed (C−CN and O−H), and the π bond is gone.
What Makes a Good Nucleophile?
Strong nucleophiles are usually negatively charged or have lone pairs:
- OHX−, CNX−, NHX2X−, CHX3OX−, HX− (from hydride reagents like NaBHX4 or LiAlHX4)
- Neutral but polarisable: NHX3, HX2O (weaker, but can add under acidic conditions)
In exams, remember: hydride ion (HX−) is a nucleophile in reductions (e.g., NaBHX4 reduces aldehydes/ketones to alcohols via nucleophilic addition).
The Big Picture – Why This Matters
Nucleophilic addition is the fundamental reaction of carbonyl compounds. It is how:
- Aldehydes and ketones form alcohols (with NaBHX4 or LiAlHX4)
- Cyanohydrins are made (important in organic synthesis)
- Grignard reagents (RMgX) add to carbonyls to form new carbon–carbon bonds
- Hemiacetals and acetals form (key in carbohydrate chemistry)
Every time you see a C=O group, think: this carbon is a target for nucleophiles.
Final Answer
Nucleophilic addition is a reaction where an electron-rich nucleophile attacks the electrophilic carbon of a polar multiple bond (typically C=O or C≡N), breaking the π bond and forming two new sigma bonds — one to the nucleophile and one to a proton (or other electrophile). The driving force is the polarity of the C=O bond and the stability gained by forming stronger sigma bonds.
Nucleophilic addition reactions of carbonyl compounds form a central part of the NCERT Class 12 Chemistry chapter on aldehydes and ketones, and questions on cyanohydrin formation or the role of hydride nucleophiles like NaBH4 are common in CBSE boards and JEE Main. Students searching "nucleophilic addition mechanism class 12 chemistry important questions" will find this carbonyl-carbon-attack explanation is the standard NCERT approach.
Why this formula?
Nucleophilic Substitution Reactions: Why the Key Formulas Hold
Nucleophilic substitution reactions are a cornerstone of organic chemistry. Instead of just memorizing the rate laws, let's understand why they arise from the molecular events.
1. The Two Main Mechanisms: A Tale of Timing
The key formulas (rate laws) for nucleophilic substitution come directly from how many molecules are involved in the rate-determining step (RDS) — the slowest step that controls the overall reaction speed.
SN1: Unimolecular — The Leaving Group Goes First
The Rate Law:
Rate=k[RX]
Why?
The reaction happens in two steps:
- Slow step (RDS): The C–X bond breaks spontaneously, forming a carbocation intermediate. Only the substrate (RX) is involved.
RXslowR++X−
- Fast step: The nucleophile (Nu−) attacks the carbocation.
R++Nu−fastRNu
Since the slow step depends only on the concentration of RX, the rate law has no dependence on [Nu−]. The nucleophile arrives after the carbocation is formed — it cannot affect the speed of the first step.
Key insight: The rate is determined by how easily the leaving group leaves, not by how fast the nucleophile attacks.
SN2: Bimolecular — Simultaneous Attack and Departure
The Rate Law:
Rate=k[RX][Nu−]
Why?
The reaction occurs in one concerted step:
- The nucleophile attacks the carbon from one side at the same time as the leaving group departs from the opposite side.
- Both RX and Nu− must collide with the correct orientation and sufficient energy.
The rate depends on the frequency of productive collisions between the two molecules. This is directly proportional to the product of their concentrations:
Rate∝[RX]×[Nu−]
Key insight: Both partners are involved in the transition state simultaneously — if either is missing, the reaction cannot proceed.
2. The Transition State: Why the Formulas Are Not Just "Given"
For SN2, the transition state has a pentavalent carbon (five bonds partially formed/broken). The energy barrier depends on steric hindrance — bulkier groups around carbon make it harder for the nucleophile to approach, which is why SN2 is favored at primary carbons.
For SN1, the transition state of the slow step involves only bond breaking (no bond formation yet). The carbocation stability (tertiary > secondary > primary) determines the activation energy — this is why SN1 is favored at tertiary carbons.
3. The Temperature Dependence: The Arrhenius Equation
Both rate constants k follow the Arrhenius equation:
k=Ae−Ea/RT
- A = frequency factor (how often collisions occur with correct orientation)
- Ea = activation energy (the energy barrier for the RDS)
- R = gas constant
- T = temperature
This is not a separate formula — it explains why the rate constants change with temperature. Higher T increases the fraction of molecules with energy ≥Ea, speeding up the reaction.
4. Summary: The "Why" in One Table
| Mechanism | Rate Law | Why? (Molecular Reason) |
|---|---|---|
| SN1 | Rate=k[RX] | Slow step involves only the substrate breaking apart. Nucleophile waits. |
| SN2 | Rate=k[RX][Nu−] | Both molecules must collide in the single, concerted step. |
Final takeaway: The formulas are not arbitrary — they are direct consequences of which molecules are present in the slowest step. Always ask: "What is happening in the rate-determining step?" The answer gives you the rate law.
Concept: Nucleophilic Substitution Reactions (SN2) — using cyanide ion as a nucleophile to extend the carbon chain by one carbon, followed by reduction of the nitrile to a primary amine.
Reasoning:
- Both starting materials are primary alkyl halides. Treat each with alcoholic KCN (or NaCN) to perform an SN2 attack, replacing the chlorine with a cyano group (−CN). This adds one carbon atom.
- The resulting nitrile (−CN) is then reduced to a primary amine (−CH2NH2). A common reducing agent for this step is LiAlH4 in dry ether, or catalytic hydrogenation (H2/Ni or H2/Pd).
Stepwise equations:
- CH3−CH2−ClKCN(alc.)CH3−CH2−CNLiAlH4/H2OCH3−CH2−CH2−NH2
- C6H5−CH2−ClKCN(alc.)C6H5−CH2−CNLiAlH4/H2OC6H5−CH2−CH2−NH2
✓Final answer
The conversions are achieved via SN2 with KCN followed by LiAlH4 reduction.
Both conversions are one-carbon chain elongations using the cyanide ion (CN−) as a nucleophile in an SN2 reaction, followed by reduction of the nitrile (−CN) to a primary amine (−CH2NH2). The final products are propan-1-amine and 2-phenylethan-1-amine, respectively.
The Core Idea: Nucleophilic Substitution + Reduction
You have an alkyl halide (a good electrophile) and you want a product whose carbon chain is one carbon longer, ending in CH2NH2. The way to do that is to replace the halogen with a carbon nucleophile that carries the nitrogen, then reduce.
The cyanide ion (CN−) is perfect: it's a strong nucleophile, attacks the carbon bearing the halogen in an SN2 reaction, and the resulting nitrile (R–CN) can be reduced to R–CH2NH2 — exactly the product you need, with the nitrile carbon supplying the extra CH2.
A common mistake is to reach for direct amination with NH3, or for the Gabriel phthalimide synthesis. Both of those put the nitrogen onto the same carbon skeleton — from CH3CH2Cl they give ethylamine (2 carbons), not the 3-carbon target propan-1-amine. Because each target here is one carbon longer than its halide, only a chain-extending route works, and the cyanide route is the standard one. Always count carbons before picking a method.
Step-by-Step Solution
1. First conversion: CH3CH2Cl→CH3CH2CH2NH2
Step 1a: Nucleophilic substitution with KCN (or NaCN)
The chlorine atom is a good leaving group. In ethanol (the NCERT solution writes "ethanolic NaCN" — ethanol is a polar protic solvent, and the reaction works well in it), the cyanide ion attacks the electrophilic carbon.
CH3CH2Cl+KCNethanolCH3CH2CN+KCl
This is an SN2 reaction — the cyanide approaches from the back, inverting the configuration (though here the carbon is not chiral, so no stereochemical consequence). The product is propanenitrile (ethyl cyanide).
Step 1b: Reduction of the nitrile to a primary amine
The nitrile group (−CN) can be reduced to a primary amine (−CH2NH2) using a strong reducing agent. The classic choice is lithium aluminium hydride (LiAlH4) in dry ether, followed by hydrolysis. Alternatively, catalytic hydrogenation (H2/Ni) works equally well — that is the reagent NCERT itself uses in part (ii).
CH3CH2CN1. LiAlH4/ether2. H2OCH3CH2CH2NH2
The reduction adds two hydrogen atoms to the carbon and one to the nitrogen, converting the triple bond into a single bond.
You can also use H2 / Raney Ni with ammonia to avoid coupling side-products (secondary amines). But LiAlH4 or plain H2/Ni is entirely acceptable in a typical exam context.
Overall equation for (i):
CH3CH2ClKCNCH3CH2CNLiAlH4/H2OCH3CH2CH2NH2
2. Second conversion: C6H5CH2Cl→C6H5CH2CH2NH2
Step 2a: Nucleophilic substitution with KCN
Benzyl chloride (C6H5CH2Cl) is even more reactive toward SN2 than a simple primary halide — the adjacent aromatic ring stabilises the transition state, so cyanide attack is fast.
C6H5CH2Cl+KCNethanolC6H5CH2CN+KCl
The product is phenylacetonitrile (phenylethanenitrile / benzyl cyanide).
Step 2b: Reduction of the nitrile
Same reduction as before (H2/Ni, as NCERT writes, or LiAlH4):
C6H5CH2CN1. LiAlH4/ether2. H2OC6H5CH2CH2NH2
The product is 2-phenylethan-1-amine (phenethylamine).
Phenethylamine is a naturally occurring compound (found in chocolate and some brain chemistry) — a nice real-world connection.
Overall equation for (ii):
C6H5CH2ClKCNC6H5CH2CNLiAlH4/H2OC6H5CH2CH2NH2
The required conversions are:
- CH3CH2ClKCNCH3CH2CNLiAlH4/H2OCH3CH2CH2NH2
- C6H5CH2ClKCNC6H5CH2CNLiAlH4/H2OC6H5CH2CH2NH2
Method: Nucleophilic Substitution via Alkyl Cyanide (Nitrile → Amine)
This is a two-step chain elongation method using cyanide ion (CN−) as a nucleophile, followed by reduction.
General Principle
- Step 1: Alkyl halide undergoes SN2 with KCN (or NaCN, in ethanol — NCERT writes "ethanolic NaCN") to form an alkyl cyanide (nitrile).
- Step 2: The nitrile is reduced (e.g., with LiAlH4 or catalytic hydrogenation, H2/Ni) to a primary amine with one extra carbon.
(i) CH3−CH2−Cl→CH3−CH2−CH2−NH2
Step 1 — Nucleophilic substitution:
CH3−CH2−Cl+KCNethanolΔCH3−CH2−CN+KCl
Step 2 — Reduction of nitrile:
CH3−CH2−CN+4[H]LiAlH4 or H2/NiCH3−CH2−CH2−NH2
Key point: The cyanide carbon becomes the extra CH2 group next to the amine.
(ii) C6H5−CH2−Cl→C6H5−CH2−CH2−NH2
Step 1 — Nucleophilic substitution:
C6H5−CH2−Cl+KCNethanolΔC6H5−CH2−CN+KCl
Step 2 — Reduction of nitrile:
C6H5−CH2−CN+4[H]LiAlH4 or H2/NiC6H5−CH2−CH2−NH2
Key point: Benzyl chloride (C6H5CH2Cl) is especially reactive in SN2 — the adjacent aromatic ring stabilises the transition state — so the substitution proceeds smoothly.
Summary of the Method
| Step | Reaction Type | Reagent | Product |
|---|---|---|---|
| 1 | SN2 | KCN (ethanolic) | Alkyl cyanide (nitrile) |
| 2 | Reduction | LiAlH4 or H2/Ni | Primary amine (+1 carbon) |
Final result: Both conversions increase the carbon chain by one and introduce a primary amine at the terminal position.
Here are the common mistakes students make when solving these nucleophilic substitution conversions, along with how to avoid each.
Mistake 1: Choosing a Route That Doesn't Change the Carbon Count
The Mistake:
Students reach for a standard amine preparation — direct ammonolysis or the Gabriel phthalimide synthesis — without counting carbons:
- CH3CH2Cl+NH3→CH3CH2NH2 ✗ (ethylamine — only 2 carbons)
- CH3CH2Cl + potassium phthalimide → N-ethylphthalimide → hydrolysis → CH3CH2NH2 ✗ (still ethylamine)
Why it's wrong:
Both routes attach nitrogen to the existing carbon skeleton. The target of (i) is CH3CH2CH2NH2 (propan-1-amine, 3 carbons) — one carbon longer than the starting halide — so any route that doesn't add a carbon cannot give it.
How to Avoid:
Count carbons first. A one-carbon extension means the cyanide route: the CN− nucleophile supplies the extra carbon, and reduction turns −C≡N into −CH2NH2.
✓ Correct approach for (i):
- CH3CH2Cl+KCNethanolCH3CH2CN+KCl
- Reduction (LiAlH4 or H2/Ni) → CH3CH2CH2NH2
Mistake 2: Forgetting the Carbon Chain Length in (ii)
The Mistake:
Students write the product as C6H5CH2NH2 (benzylamine) instead of C6H5CH2CH2NH2 (2-phenylethan-1-amine).
Why it's wrong:
The target has two carbons between the benzene ring and the amino group. The starting material has only one carbon. You must increase the chain length by one carbon.
How to Avoid:
Always count the carbon atoms in the product vs. starting material. If the product has one more carbon, you need a cyanide ion (CN−) as the nucleophile first, then reduce.
✓ Correct approach for (ii):
- C6H5CH2Cl + KCN (ethanolic) → C6H5CH2CN (benzyl cyanide)
- Reduction: H2/Ni or LiAlH₄ → C6H5CH2CH2NH2
Mistake 3: Inventing "Better" Solvent Conditions Than the Standard Ones
The Mistake:
Insisting the substitution must be run in an anhydrous polar aprotic solvent (acetone, DMF) and marking the ethanol route wrong.
Why it's wrong:
The standard (and NCERT's own printed) condition for this reaction is ethanolic NaCN/KCN — cyanide is a strong enough nucleophile that the SN2 displacement works well in ethanol. Aprotic solvents can accelerate SN2 reactions, but they are not required here, and "correcting" the printed conditions loses marks.
How to Avoid:
Write the reagent the syllabus uses: ethanolic KCN (or NaCN), with heating. Mention SN2 as the mechanism.
Mistake 4: Choosing a Reducing Agent That Doesn't Reduce Nitriles to Primary Amines
The Mistake:
Using NaBH4 (which does not reduce nitriles), or DIBAL-H (which stops at the aldehyde stage), and expecting a primary amine.
How to Avoid:
For R−CN→R−CH2NH2, use LiAlH4 in dry ether (then water) or catalytic hydrogenation (H2/Ni) — the reagent NCERT itself uses. (H2/Raney Ni with ammonia suppresses secondary-amine coupling by-products, a useful refinement but not required.)
✓ Correct reduction:
R−CNLiAlH4/ether, then H2O (or H2/Ni)R−CH2NH2
Mistake 5: Writing Incomplete or Unbalanced Equations
The Mistake:
Writing only the organic product and forgetting byproducts (like KCl) or not balancing atoms.
Example of wrong:
CH3CH2Cl+KCN→CH3CH2CN (missing KCl)
How to Avoid:
Always write complete, balanced equations with all products. Check that the number of atoms of each element is the same on both sides.
✓ Correct:
CH3CH2Cl+KCNethanolCH3CH2CN+KCl
Quick Summary Table
| Mistake | How to Avoid |
|---|---|
| Same-carbon-count route (direct NH3, Gabriel) | Count carbons — a +1 extension needs CN− then reduction |
| Forgetting chain extension in (ii) | Use CN− then reduce |
| "Correcting" the solvent | Ethanolic KCN/NaCN is the standard condition |
| Wrong reduction reagent | Use LiAlH4 or H2/Ni (not NaBH4/DIBAL-H) |
| Unbalanced equations | Always write complete products |
Final Tip: For primary amine preparation from alkyl halides, remember two standard routes:
- No chain extension → Gabriel phthalimide
- Chain extension by one carbon → KCN followed by reduction
Both targets in this question are one carbon longer than their halides — so both must go through the cyanide route.
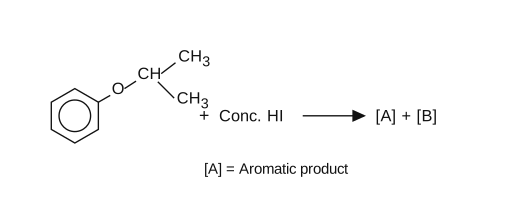
- COMEDK 2026Set 2026-A1 markMCQQ. A compound [X] undergoes reactions as given. Identify compounds [C] and [D] formed in these reactions. [A]Cr2O72−/H+[C] $$ [\text { B }] \xrightarrow[\text {(ii) } \mathrm{Na}2 \mathrm{CO}{3(\text { aq })}+\mathrm{I}_2]{\text { (i)aq. } \mathrm{KOH}} [\mathrm{D}] \quad+\mathrm{CH}_3 \mathrm{COONa}(A) \text { [C]: Benzoquinone [D]: lodoform } (B)[C]:Benzene[D]:2−iodo−propane(C)[C]:Benzoicacid[D]:lodoform(D) \text { [C]: 4-lodophenol [D]: 1-iodo-propane } $$
›Reveal solutionSolution
The compound [X] is isopropyl phenyl ether (C₆H₅–O–CH(CH₃)₂). Cleavage with concentrated HI gives phenol ([A]) and isopropyl iodide ([B]). Oxidation of phenol yields benzoquinone ([C]), and the iodoform reaction on isopropyl iodide gives iodoform ([D]) and sodium acetate. Thus the correct option is (A).
Concept & Intuition
This problem tests two classic organic reactions: ether cleavage by HI and the iodoform reaction. The key is to recognize that the ether [X] is an aryl alkyl ether (phenol derivative). When treated with concentrated HI, the C–O bond breaks selectively at the alkyl side (since the aryl–O bond is stronger due to resonance), producing phenol and an alkyl iodide. Then, phenol can be oxidized to benzoquinone, and the alkyl iodide (if it has a methyl group adjacent to the carbonyl or a secondary alcohol that can be oxidized to a methyl ketone) will undergo the iodoform test.
Let’s walk through each step.
Step-by-step reasoning
-
Identify [X] and its cleavage products
The figure shows a benzene ring with an –O–CH(CH₃)₂ group. That is isopropyl phenyl ether (C₆H₅–O–CH(CH₃)₂).
With concentrated HI, the ether bond breaks. The mechanism: HI protonates the oxygen, then iodide attacks the less hindered carbon (the isopropyl carbon, since it’s primary-like in the sense of being less sterically hindered than the aromatic ring). This gives phenol (C₆H₅OH) as the aromatic product [A] and isopropyl iodide (CH₃–CHI–CH₃) as [B].
Watch outA common mistake is to think the aromatic ring gets iodinated. But under these conditions, the C–O bond on the alkyl side breaks, not the aryl–O bond. The aromatic ring remains intact as phenol.
-
Reaction of [A] (phenol) with Cr₂O₇²⁻/H⁺ → [C]
Phenol is easily oxidized. Chromic acid (Cr₂O₇²⁻/H⁺) is a strong oxidizing agent. It oxidizes phenol to 1,4-benzoquinone (often just called benzoquinone). The reaction involves two-electron oxidation: the –OH group becomes a carbonyl, and the ring is rearranged to a quinoid structure.
So [C] = Benzoquinone.
-
Reaction of [B] (isopropyl iodide) with (i) aq. KOH, then (ii) Na₂CO₃(aq) + I₂ → [D] + CH₃COONa
- Step (i): Aqueous KOH will hydrolyze the alkyl iodide to an alcohol. Isopropyl iodide gives isopropyl alcohol (propan-2-ol, CH₃–CHOH–CH₃).
- Step (ii): The mixture of Na₂CO₃ and I₂ is the classic iodoform test reagent. It works on compounds that have a CH₃–C(=O)– group or a CH₃–CHOH– group (which gets oxidized to a methyl ketone under the basic conditions). Isopropyl alcohol (CH₃–CHOH–CH₃) is a secondary alcohol with a methyl group on the carbon bearing the –OH. Under basic I₂, it is first oxidized to acetone (CH₃–CO–CH₃). Then acetone undergoes the iodoform reaction:
CH3COCH3+3I2+4NaOH→CHI3+CH3COONa+3NaI+3H2O
The products are **iodoform** (CHI₃, a yellow precipitate) and **sodium acetate** (CH₃COONa).So [D] = Iodoform.
- Match with options
- [C] = Benzoquinone
- [D] = Iodoform This corresponds exactly to option (A).
TipThe iodoform reaction is specific to methyl ketones and secondary alcohols with a methyl group on the carbinol carbon. Isopropyl alcohol fits perfectly. If [B] had been a primary alkyl iodide (like n-propyl iodide), the product would be a carboxylic acid, not iodoform.
✓Final answerThe correct option is (A).
ANSWER: A
-
- KCET 2025Set D-41 markMCQQ.Match the compounds given in List – I with the items given in List – II. List – I (I) Benzenesulphonyl Chloride (II) Sulphanilic acid (III) Alkyl Diazonium salts (IV) Aryl Diazonium salts List – II(a) Zwitterion(b) Hinsberg reagent(c) Dyes(d) Conversion to alcohols (A) 1 – c, II – b, III – a, IV – d (B) 1 – a, II – c, III – b, IV – d (C) 1 – c, II – a, III – d, IV – b (D) 1 – b, II – a, III – d, IV – c
›Reveal solutionSolution
Benzenesulphonyl chloride = Hinsberg reagent, sulphanilic acid = zwitterion, alkyl diazonium salts → alcohols, aryl diazonium salts → azo dyes.
Step 1 — (I) Benzenesulphonyl chloride → (b) Hinsberg reagent.
C6H5SO2Cl is known as Hinsberg's reagent. It reacts with 1° amines to give a sulphonamide with an acidic N–H (soluble in alkali), with 2° amines to give a sulphonamide with no N–H (insoluble in alkali), and does not react with 3° amines — the classical test for distinguishing the three classes.
Step 2 — (II) Sulphanilic acid → (a) Zwitterion.
The −SO3H group is strongly acidic and the −NH2 group is basic, so an internal proton transfer occurs:
H2N−C6H4−SO3H⇌+H3N−C6H4−SO3−
This dipolar internal salt is a zwitterion (which is why sulphanilic acid has a high melting point and low solubility in organic solvents).
Step 3 — (III) Alkyl diazonium salts → (d) Conversion to alcohols.
Alkyl diazonium ions (R−N2+) are extremely unstable because N2 is an excellent leaving group and there is no resonance stabilisation. They decompose at once, and water traps the resulting carbocation:
R−N2+−N2R+H2OR−OH
This is why treating a 1° aliphatic amine with HNO2 gives an alcohol (plus brisk N2 evolution).
Step 4 — (IV) Aryl diazonium salts → (c) Dyes.
C6H5N2+Cl− is resonance-stabilised by the ring and is stable at 0–5 °C. It undergoes coupling with phenol or aniline to give brightly coloured azo compounds (−N=N− chromophore) — the basis of azo dyes.
Step 5 — Assemble. I–b, II–a, III–d, IV–c.
✓Final answerThe correct option is (D) — 1 – b, II – a, III – d, IV – c.
ANSWER: D
- KCET 2024Set B-21 markMCQQ.In the reaction Aniline NaNO2/dil.HCl P Phenol/NaOH Q, ‘Q’ is: (A) C6H5N2Cl (B) ortho-hydroxyazobenzene (C) para-hydroxyazobenzene (D) meta-hydroxyazobenzene
›Reveal solutionSolution
Diazotisation of aniline gives the benzenediazonium salt (P); azo-coupling of that weak electrophile with phenoxide occurs at the para position, so Q is para-hydroxyazobenzene.
1. Step 1 — Diazotisation gives P
A primary aromatic amine treated with nitrous acid (generated in situ from NaNO2+dil. HCl) at 273–278 K gives an arenediazonium salt:
C6H5NH2NaNO2/dil. HCl273−278 KC6H5N+≡N Cl−
So P= benzenediazonium chloride. (The aryl diazonium ion is stabilised by delocalisation into the ring — this is why it survives, unlike an alkyl diazonium ion.) Note that option (A) is P, not Q — a classic distractor.
2. Step 2 — Azo coupling gives Q
The diazonium ion is only a weak electrophile, so it can attack a ring only if that ring is strongly activated. Phenol in NaOH is deprotonated to the phenoxide ion, C6H5O−, whose −O− is a very powerful electron-releasing group (strong +M), pumping electron density onto the ortho and para carbons.
Electrophilic substitution therefore occurs at those positions, but coupling takes place essentially exclusively at the para position because:
- the para carbon is sterically unhindered, whereas an ortho attack would place the bulky −N=N−C6H5 group right next to the −OH;
- the para-coupled azo product is the thermodynamically favoured, fully conjugated dye.
C6H5N+≡N+C6H5O−NaOHC6H5−N=N−(p)C6H4−OH
Q=p-hydroxyazobenzene — an orange dye. (With aniline instead of phenol, the analogous product would be p-aminoazobenzene.)
3. Rejecting the others
- (A) C6H5N2Cl is the intermediate P, not the final product Q.
- (B) ortho-coupling is sterically disfavoured and is at best a minor product.
- (D) meta-coupling is impossible: −O− is an o/p-director; the meta carbons carry no extra electron density.
✓Final answerThe correct option is (C) — para-hydroxyazobenzene.
ANSWER: C
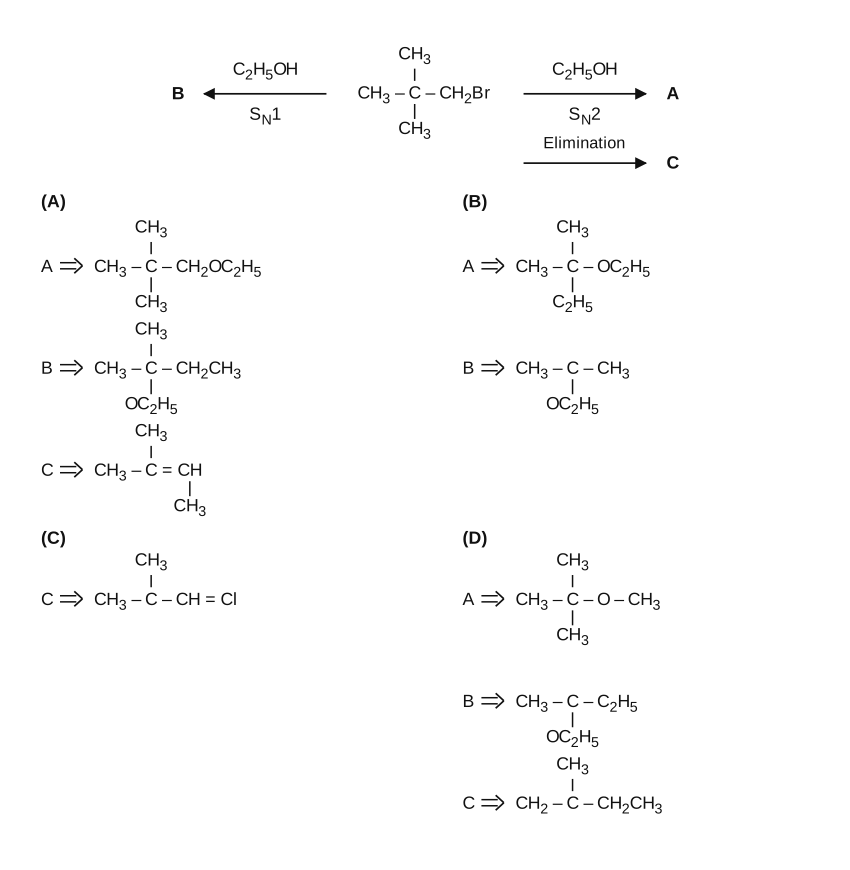
- COMEDK 2023Set 2023-M1 markMCQQ.Identify A, B and C. (A) (B) (C) (D)
›Reveal solutionSolution
The reaction scheme shows a neopentyl bromide undergoing SN1 (to B), SN2 (to A), and elimination (to C). The correct products are: A = neopentyl ethyl ether, B = 2-ethoxy-2-methylbutane (rearranged), C = 2-methyl-2-butene. Only option (A) matches all three.
Concept & Intuition
Neopentyl bromide (1-bromo-2,2-dimethylpropane) is a classic case where the substrate’s structure dictates reaction pathways. The carbon bearing bromine is primary, but it’s attached to a quaternary carbon (three methyl groups). For SN2, the backside attack is severely hindered by the bulky neopentyl group, making it very slow. For SN1, the primary carbocation would normally be unstable, but under solvolytic conditions (ethanol), the reaction proceeds via a rearranged tertiary carbocation (a methyl shift), giving a more stable intermediate. Elimination also favors the more substituted alkene (Zaitsev product). The question tests recognition of these rearrangements and the correct structures.
Step-by-step reasoning
- Identify the substrate The central structure is neopentyl bromide:
CH3–C(CH3)2–CH2Br
The bromine is on a primary carbon, but the carbon is neopentyl (tert-butylmethyl). This is crucial.
- SN2 pathway (→ A) SN2 requires a clean backside attack. The neopentyl group is extremely bulky, so SN2 is very slow. However, in ethanol (C₂H₅OH) as solvent, the ethoxide ion (from ethanol) can act as a nucleophile. The product is the unrearranged ethyl ether:
CH3–C(CH3)2–CH2–O–C2H5
This is neopentyl ethyl ether. No rearrangement occurs because SN2 is concerted.
Check options: Only option (A) shows this exact structure for A.
- SN1 pathway (→ B) SN1 proceeds via carbocation formation. The primary carbocation (CH₃–C(CH₃)₂–CH₂⁺) is very unstable. It immediately undergoes a 1,2-methyl shift to form the more stable tertiary carbocation:
CH3–C+(CH3)–CH2CH3
This tertiary carbocation is then trapped by ethanol (solvent) to give the ethyl ether:
CH3–C(OC2H5)(CH3)–CH2CH3
This is 2-ethoxy-2-methylbutane.
Check options: Only option (A) shows B as exactly this structure (with OC₂H₅ on the quaternary carbon and an ethyl group on the adjacent carbon).
- Elimination pathway (→ C) Under elimination conditions (often with a strong base, but here simply labelled “Elimination” in ethanol), the neopentyl system can undergo E2 or E1. The most stable alkene is the trisubstituted one: 2-methyl-2-butene. The carbocation rearrangement (as in SN1) leads to the tertiary carbocation, which then loses a proton to give:
CH3–C(CH3)=CH–CH3
This is 2-methyl-2-butene.
Check options: Only option (A) shows C as this alkene.
- Eliminate other options
- Option (B): Shows A as an ether on the quaternary carbon (wrong, no rearrangement in SN2) and B as a simple ether without ethyl group (missing the shift). No C given.
- Option (C): Shows C as a chloroalkene (impossible, no chlorine source). No A or B.
- Option (D): Shows A as a methyl ether (wrong nucleophile), B as an ether on a secondary carbon (wrong), and C as an unbranched alkene (wrong).
Watch outA common mistake is to assume SN1 gives the unrearranged primary ether. But neopentyl systems always rearrange under SN1 conditions because the primary carbocation is too unstable.
TipRemember: For neopentyl halides, SN2 gives unrearranged product (slow), SN1 gives rearranged product (via methyl shift), and elimination gives the most substituted alkene.
✓Final answerThe correct option is (A).
ANSWER: A
- KCET 2022Set B-31 markMCQQ.A secondary amine is (A) a compound with an NH2 group on the carbon atom in number 2 position (B) a compound in which 2 of the hydrogen of NH3 have been replaced by organic groups (C) an organic compound with two NH2 group (D) a compound with two carbon atom and an NH2 group
›Reveal solutionSolution
A secondary amine is defined by the number of alkyl/aryl groups attached to nitrogen — specifically, two organic groups replace two hydrogens of ammonia. The correct answer is (B).
The key to this question is understanding how amines are classified. Amines are derivatives of ammonia (NH3), and the classification — primary, secondary, or tertiary — depends entirely on how many of the three hydrogen atoms on nitrogen have been replaced by carbon-containing groups (alkyl or aryl). It has nothing to do with the position of a carbon atom, the number of carbon atoms in the molecule, or the count of NH2 groups.
Let’s examine each option carefully.
-
Option (A) says “a compound with an NH2 group on the carbon atom in number 2 position.” This describes a structural detail about where an amino group is attached on a carbon chain (like on C-2 of propane). That is a matter of positional isomerism, not amine classification. A primary amine can have its NH2 on carbon-2, and so can a secondary or tertiary amine if they also have other groups. This definition misses the point entirely.
-
Option (B) says “a compound in which 2 of the hydrogen of NH3 have been replaced by organic groups.” This is exactly the definition. Ammonia has three hydrogens. Replace one → primary amine (RNH2). Replace two → secondary amine (R2NH). Replace three → tertiary amine (R3N). So a secondary amine has two alkyl/aryl groups attached to nitrogen, with one hydrogen remaining.
-
Option (C) says “an organic compound with two NH2 groups.” That describes a diamine — a molecule with two amino groups, like ethylenediamine (H2NCH2CH2NH2). Each NH2 is a primary amino group, so the compound as a whole is a primary diamine, not a secondary amine. The classification is about substitution on a single nitrogen atom, not the count of amino groups in the molecule.
-
Option (D) says “a compound with two carbon atoms and an NH2 group.” That is just a specific example — ethylamine (CH3CH2NH2) has two carbons and one NH2, but it is a primary amine, not secondary. The number of carbons in the molecule is irrelevant to the classification.
Watch outA common mistake is to confuse “secondary” with “two NH2 groups” or with “two carbons.” Remember: the word “secondary” refers to the nitrogen atom’s substitution level — two organic groups on the same nitrogen — not to the count of amino groups or carbon atoms.
✓Final answerThe correct option is (B).
-
- COMEDK 2021Set 2021-B1 markMCQQ.What are the products formed when Anisole is reacted with Hydroiodic acid and heated? (A) Iodobenzene + Methane (B) Phenol + Methanol (C) Phenol + Iodomethane (D) Iodobenzene + Methanol
›Reveal solutionSolution
Anisole C6H5−O−CH3+HI→C6H5OH (phenol) +CH3I (iodomethane).
In cleavage of aryl alkyl ethers by HI, the bond broken is the O−alkyl bond, not the O−aryl bond, because forming an aryl cation/attack at the aromatic carbon is very unfavourable. I− attacks the methyl carbon (SN2), giving iodomethane, and the aromatic ring retains oxygen as phenol.
✓Final answerThe correct option is (C) — Phenol + Iodomethane
- KCET 2019Set A-11 markMCQQ.The metal nitrate that liberates NO2 on heating (A) NaNO3 (B) KNO3 (C) LiNO3 (D) RbNO3
›Reveal solutionSolution
Li+ is tiny and highly polarising, so it distorts the nitrate ion enough to break it right down to the oxide + NO2; the bigger alkali cations only take it as far as the nitrite.
Step 1 — The two possible decomposition routes
Alkali-metal nitrates decompose on heating by one of two paths:
Path 1 — to the nitrite (Na, K, Rb, Cs):
2MNO3Δ2MNO2+O2↑
Only oxygen is evolved — no brown fumes.
Path 2 — to the oxide (Li):
4LiNO3Δ2Li2O+4NO2↑+O2↑
Here the nitrate ion is destroyed completely, giving the characteristic brown NO2 gas.
Step 2 — Why lithium is the odd one out (Fajans' rules)
The polarising power of a cation scales as (radius)2charge. Among the alkali metals:
Li+(76 pm)<Na+(102)<K+(138)<Rb+(152 pm)
So Li+ is by far the smallest and therefore the most polarising. It pulls electron density out of the large, soft NO3− anion, weakening the N–O bonds so much that the anion breaks apart entirely into O2− (which stays with Li as Li2O) and NO2.
The larger cations Na+,K+,Rb+ cannot distort the nitrate that strongly. Their nitrates only shed one oxygen atom, stopping at the stable nitrite.
Step 3 — The wider pattern (worth remembering)
This is the same reason lithium shows a diagonal relationship with magnesium: like Mg(NO3)2 (an alkaline-earth nitrate), LiNO3 gives the oxide + NO2 + O2. The identical logic explains why Li2CO3 decomposes on heating while Na2CO3 and K2CO3 do not.
Step 4 — Screen the options
- (A) NaNO3 → NaNO2+O2 ✗
- (B) KNO3 → KNO2+O2 ✗
- (C) LiNO3 → Li2O+NO2+O2 ✓
- (D) RbNO3 → RbNO2+O2 ✗
✓Final answerThe correct option is (C) LiNO3 — the only alkali-metal nitrate that decomposes to the oxide and liberates NO2.
ANSWER: C
🎓Unlock everything free for 14 days
- ✓Full step-by-step solutions
- ✓Concept-first explanations
- ✓Methods, shortcuts & mistakes
- ✓PYQ mapping + timed mock tests
Full access for 14 days. No credit card required.