Q.Explain why nucleophilic substitution reactions are not very common in phenols.
🔒You're viewing a preview — the full solution, concept, methods & PYQ mapping are locked.
🔒 Start your 14-day free trial to unlock the full solution →Concept understanding — Electrophilic Aromatic Substitution
Electrophilic Aromatic Substitution – The First Meeting
Imagine you have a benzene ring — that perfect, flat hexagon of six carbons with alternating double bonds. It's stable, almost stubbornly so. You want to attach something new to it, say a bromine atom or a nitro group. But benzene doesn't react like an alkene. It doesn't just add across a double bond. Instead, it does something more elegant: it kicks out a hydrogen and keeps its aromatic ring intact.
That's the heart of Electrophilic Aromatic Substitution (EAS).
The Intuition: Why "Substitution" and Not "Addition"?
Benzene's stability comes from its delocalised π electrons — a cloud above and below the ring. This cloud is electron-rich, so it attracts electrophiles (electron-loving species). But if an electrophile simply added to a double bond, the ring would break its aromaticity, losing that huge stabilisation. That would be energetically costly.
So benzene does something smarter: it lets the electrophile attack, temporarily breaks aromaticity to form a high-energy intermediate (the arenium ion), and then loses a proton to restore the aromatic ring. The net result? A hydrogen is replaced by the electrophile. The ring is back to its stable, aromatic self.
The key trade-off: temporary loss of aromaticity is acceptable because the final product regains it. Addition reactions would permanently destroy aromaticity — benzene avoids that.
The Precise Statement
Electrophilic Aromatic Substitution is a reaction in which an electrophile (E+) replaces a hydrogen atom on an aromatic ring, proceeding through a sigma complex (arenium ion) intermediate, and restoring aromaticity after deprotonation.
The general equation:
Ar−H+EX+Ar−E+HX+
where Ar represents an aromatic ring.
The Mechanism in Three Steps
-
Generation of the electrophile – Many EAS reactions need a catalyst to create a strong enough E+. For example, bromination uses FeBrX3 to polarise BrX2 into BrX+.
-
Attack by the aromatic ring – The π electrons of benzene attack the electrophile, forming a sigma complex (also called the arenium ion or Wheland intermediate). This intermediate is non-aromatic — it has four π electrons delocalised over five carbons, and one sp3 carbon bearing the electrophile and a hydrogen.
Benzene+E+⟶Sigma complex (non-aromatic)
- Deprotonation – A base (often the counterion of the catalyst, like FeBrX4X−) removes the proton from the sp3 carbon. The pair of electrons from the C–H bond flows back into the ring, restoring the aromatic sextet.
Sigma complex+Base⟶Product+HX+
The sigma complex is not aromatic. It's a high-energy intermediate. Students often mistakenly think it's still aromatic — it isn't. That's why the step is fast and the complex is short-lived.
Why This Matters for Exams
EAS is the gateway to understanding how to put groups onto benzene rings. The rate-determining step is usually the formation of the sigma complex (step 2). The regiochemistry (where the electrophile goes) depends on whether the ring already has a substituent — that's the topic of activating/deactivating groups and ortho/para vs. meta directors.
But for now, remember this: …
Why this formula?
Electrophilic Aromatic Substitution: Why the Mechanism Holds
The Core Puzzle: Why Benzene Doesn't Just Add
Benzene (C6H6) has three double bonds — so why doesn't it undergo addition reactions like alkenes?
The answer lies in aromatic stabilisation: benzene's delocalised π-electron cloud (the "aromatic sextet") is about 150 kJ/mol more stable than a hypothetical cyclohexatriene with localised double bonds.
If benzene simply added an electrophile (like Br2), it would lose this stabilisation — a huge energy penalty.
So, nature chooses a different path: substitution instead of addition, preserving the aromatic ring.
The Key Formula: The Reaction Profile
The rate-determining step in electrophilic aromatic substitution (EAS) is the formation of the arenium ion (σ-complex):
Ar-H+E+slowAr-E+H(σ-complex)
Then, fast deprotonation restores aromaticity:
Ar-E+H+B−fastAr-E+BH
Why This Holds: The Energy Barrier Logic
-
First step (slow): The electrophile E+ attacks the electron-rich ring. The σ-complex is non-aromatic — it has only 4 π-electrons delocalised over 5 carbons (the sixth carbon is sp3 hybridised). This intermediate is higher in energy than the starting benzene.
-
Second step (fast): A base removes the proton, restoring the aromatic sextet. This step is strongly exothermic — the system regains ~150 kJ/mol of stabilisation.
The overall reaction is exothermic, but the activation energy is dominated by the destabilisation of the σ-complex.
The Rate Law: Why It's First Order in Both
From the mechanism:
Rate=k[Arene][E+]
Reasoning:
- The slow step involves one molecule of arene and one molecule of electrophile.
- No other species appear before the rate-determining step.
- Therefore, the rate law is bimolecular — first order in each reactant.
This is not derived from the overall stoichiometry — it comes directly from the molecularity of the slow step.
The Hammett Equation: Quantifying Substituent Effects
For substituted benzenes, the rate constant k relative to benzene (k0) follows:
logk0k=σρ
Why This Holds
- σ (sigma constant): Measures the electronic effect of a substituent (electron-donating or withdrawing) relative to hydrogen. It is derived from the ionisation constants of benzoic acids — a purely empirical scale.
- ρ (rho constant): Measures the sensitivity of the reaction to substituent effects. A positive ρ means the reaction is favoured by electron-withdrawing groups (rare in EAS); a negative ρ means electron-donating groups accelerate the reaction.
Why it works:
The σ-complex has a positive charge delocalised over the ring. Substituents that stabilise this positive charge (electron-donating groups like −OH, −NH2) lower the activation energy — hence σ is negative for such groups. Electron-withdrawing groups (−NO2, −CN) destabilise the σ-complex — σ is positive.
The linear free-energy relationship holds because the transition state resembles the σ-complex in charge distribution.
The Directing Effect: Why Ortho/Para vs Meta
The position of substitution is governed by the stability of the σ-complex for each possible attack site. …
The key idea is that in phenols, the oxygen lone pair is strongly delocalised into the aromatic ring via resonance. This makes the carbon–oxygen bond stronger and the ring electron-rich, both of which disfavour nucleophilic substitution.
Reasoning:
- Resonance stabilisation: The lone pair on the phenolic oxygen participates in resonance with the benzene ring, giving the C–O bond partial double-bond character. This makes it difficult for a nucleophile to break the C–O bond.
- Ring deactivation toward nucleophiles: The same resonance makes the aromatic ring electron-rich. Nucleophilic substitution requires an electron-deficient (positively polarised) carbon centre, which is absent here. …
Phenols resist nucleophilic substitution because the lone pair on oxygen is in conjugation with the ring, making the carbon–oxygen bond strong and partial double-bond in character — the C–O bond simply does not break easily under nucleophilic attack.
The core idea: why phenols are different
Nucleophilic substitution requires a good leaving group — something that can depart with its bonding electrons, usually as a stable anion. In alkyl halides, the halide ion (Cl⁻, Br⁻, I⁻) is a weak base and a fine leaving group. In phenols, the leaving group would be the hydroxide ion (OH⁻), which is a strong base and a terrible leaving group. That alone makes direct Sₙ2 or Sₙ1 impossible under normal conditions.
But there is a deeper, structural reason that makes phenols even more resistant than, say, simple alcohols.
Step-by-step reasoning
-
The oxygen lone pair is delocalised into the ring.
In phenol, the oxygen atom is sp2-hybridised (or close to it). One of its lone pairs is in a p-orbital that overlaps with the π-system of the benzene ring. This conjugation creates a partial double bond between oxygen and the ring carbon. The C–O bond order is greater than 1 — it is not a simple single bond.
-
This conjugation strengthens the C–O bond.
A stronger bond means higher bond dissociation energy. Breaking the C–O bond to release OH⁻ requires much more energy than in an aliphatic alcohol. The resonance stabilisation of the phenol molecule itself also means the ground state is lower in energy, raising the activation barrier for any reaction that breaks the conjugation.
-
The leaving group would be a very poor one.
Even if the bond could be broken, the departing species would be the hydroxide ion. Hydroxide is a strong base (pKa of water ≈ 15.7) and a very poor leaving group. In nucleophilic substitution, good leaving groups are weak bases (like halides, tosylate, etc.). Phenols cannot provide a good leaving group without prior activation.
-
The aromatic ring is electron-rich, not electron-poor.
Nucleophilic substitution on an aromatic ring (SₙAr) typically requires the ring to be strongly electron-deficient — for example, activated by nitro groups that stabilise the Meisenheimer complex. Phenol’s ring is actually electron-rich due to oxygen’s electron-donating resonance effect. This makes it even less susceptible to attack by a nucleophile.
-
Compare with alkyl halides or activated aryl halides. …
Method: Electrophilic Aromatic Substitution (EAS) Mechanism
This is the standard method for understanding how substituents (like the –OH group in phenol) direct and activate aromatic rings toward substitution.
Steps of the EAS Mechanism (for phenol as an example)
-
Generation of the electrophile
The attacking species (e.g., Br+, NO2+, SO3) is formed in the reaction medium.
-
Formation of the σ-complex (arenium ion)
The electrophile attacks the electron-rich aromatic ring. The –OH group donates electron density via resonance, making the ring highly activated.
- The lone pair on oxygen delocalises into the ring, especially to the ortho and para positions.
- This stabilises the positive charge in the σ-complex.
-
Deprotonation to restore aromaticity
A base (often the counterion or solvent) removes the proton from the carbon that bonded to the electrophile, regenerating the aromatic ring.
Key result: The –OH group is a strong activating and ortho/para-directing group. Phenol undergoes EAS much faster than benzene.
Why Nucleophilic Substitution Is Rare in Phenols
Nucleophilic substitution (e.g., SN1 or SN2) requires a good leaving group attached to a carbon that can be attacked by a nucleophile. In phenol:
- The –OH group is a very poor leaving group (OH− is a strong base and does not depart easily).
- The C–O bond in phenol is stronger than in alcohols due to partial double-bond character from resonance between the oxygen lone pair and the aromatic ring.
The key reason: Resonance stabilisation of the C–O bond
The lone pair on oxygen is delocalised into the benzene ring: …
Common Mistakes in Electrophilic Aromatic Substitution (Phenols) & How to Avoid Them
Mistake 1: Confusing Nucleophilic vs. Electrophilic Substitution
The error: Students often mix up the two reaction types and try to apply nucleophilic substitution logic to phenols.
Why it happens: Phenols have an –OH group, which looks like a good leaving group (like in alcohols). So students assume nucleophilic substitution should work.
How to avoid: Remember the key principle — in aromatic systems, the ring's π electrons make it electron-rich, so it attacks electrophiles, not nucleophiles. The –OH group in phenol activates the ring for electrophilic substitution (like nitration, halogenation), not nucleophilic.
Exam tip: If you see an aromatic ring + –OH, think EAS first, not nucleophilic substitution.
Mistake 2: Forgetting the –OH Group is a Poor Leaving Group
The error: Students think –OH can leave as OHX− in nucleophilic substitution.
Why it happens: In aliphatic alcohols, –OH can be converted to a better leaving group (e.g., via HX+ to give HX2O). But in phenols, the C−O bond is stronger due to partial double-bond character from resonance.
How to avoid: Visualise the resonance:
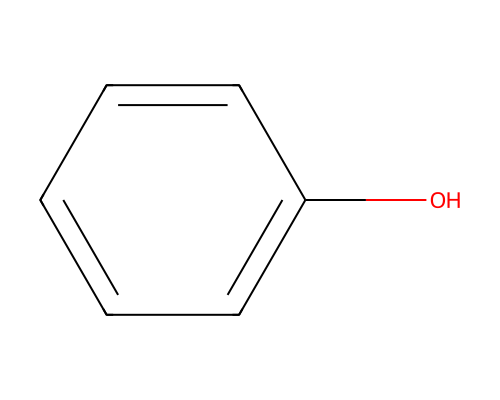
The lone pairs on oxygen delocalise into the ring, making the C−O bond shorter and stronger (about 1.36 Å vs. 1.43 Å in alcohols). This bond is not easily broken.
Key fact: CX6HX5−OH has bond dissociation energy ~460 kJ/mol — much higher than aliphatic C−OH (~380 kJ/mol).
Mistake 3: Ignoring the Aromatic Stabilisation Energy
The error: Students don't account for the energy cost of breaking aromaticity.
Why it happens: Nucleophilic substitution on an aromatic ring would require breaking the aromatic π system — which costs ~150 kJ/mol of resonance stabilisation energy.
How to avoid: Always ask: "Would this reaction break aromaticity?" If yes, it's highly unfavourable unless special conditions exist (e.g., extreme heat, strong electron-withdrawing groups).
Quick check: Benzene's resonance energy = 152 kJ/mol. Any reaction that destroys the ring must overcome this — nucleophilic substitution cannot.
Mistake 4: Confusing Phenols with Phenoxide Ions
The error: Students think that because CX6HX5OX− (phenoxide) can undergo nucleophilic substitution, phenol itself can too.
Why it happens: Phenoxide is formed in strong base, and it can undergo nucleophilic substitution at the ring (e.g., in the von Richter or Chichibabin reactions). But these are special cases.
How to avoid: Distinguish clearly:
| Species | Reactivity |
|---|---|
| Phenol (CX6HX5OH) | Electrophilic substitution only |
| Phenoxide (CX6HX5OX−) | Can undergo nucleophilic substitution (under harsh conditions) |
Exam note: Unless the question mentions strong base (NaOH, KOH) and high temperature, assume phenol — not phenoxide.
Mistake 5: Overlooking the +R Effect of –OH …
- KEAM 2026Set eng-2026-04174 marksMCQQ.p-Bromophenol is the major product formed when phenol is treated with (A) Bromine water (B) Br2 in acetic acid at 300K (C) Br2 in CCl4 at 300K (D) Br2 in CS2 at 273K (E) Br2 in acetone at 273K
›Reveal solutionSolution
Using Br2 in the non-polar solvent CS2 at low temperature (273 K) suppresses polybromination and gives p-bromophenol as the major monobrominated product.
Phenol is strongly activated, so bromine water (polar, ionising) gives 2,4,6-tribromophenol.
To stop at monobromination, a low-polarity solvent and low temperature are used, which lowers the electrophilicity/availability of Br+. …
- KEAM 2026Set eng-2026-04224 marksMCQQ.When phenol is treated with excess of bromine water, it gives (A) o-bromophenol (B) o- and p-bromophenol (C) 1,3,5-tribromophenol (D) 2,4-dibromophenol (E) 2,4,6-tribromophenol
›Reveal solutionSolution
Phenol with excess bromine water undergoes electrophilic substitution at the ortho and para positions, giving a white precipitate of 2,4,6-tribromophenol. …
- KEAM 2026Set pha-2026-0418F4 marksMCQQ.Aniline reacts with acetic anhydride in pyridine to give a product which reacts with Br2 in CH3COOH to get (A) o-bromoaniline (B) p-bromoaniline (C) p-bromoacetanilide (D) o-bromoacetanilide (E) m-bromoacetanilide
›Reveal solutionSolution
Aniline → acetanilide (acetic anhydride/pyridine) → bromination gives mainly p-bromoacetanilide.
Aniline is acetylated to acetanilide C6H5NHCOCH3. The acetamido group is an activating ortho/para director, but the bulky −NHCOCH3 hinders the ortho positions, so electrophilic bromination with Br2/CH3COOH occurs predominantly at the para po …
- KEAM 2026Set pha-2026-0420F4 marksMCQQ.Which of the following reaction yieldstarry oxidation products? (A) Sulphonation of aniline (B) Nitration of aniline (C) Firedel-Crafts alkylation aniline (D) Firedel-Crafts alkylation of aniline (E) Bromination of aniline
›Reveal solutionSolution
Aniline is readily oxidised; direct nitration with HNO3/H2SO4 oxidises it to dark tarry products, so the amino group is protected (acetylated) first.
Aniline is very easily oxidised because the ring is electron-rich. When it is subjected to direct nitration with the strongly oxidising nitrating mixture (HNO3/H2SO4), a large part of it is oxidised to dark, tarry products rather than cleanly nitrated. This is exactly why, in practice, aniline is first acetylated ( …
- KEAM 2025Set eng-2025-04264 marksMCQQ.Phenol is treated with Con.H2SO4 to gives a product 'X' which on treatment with Con.HNO3 gives compound 'Y'. The compounds 'X' and 'Y' are respectively (A) Phenol-2-sulphonic acid and 2-nitrophenol (B) Phenol-2-sulphonic acid and 4-nitrophenol (C) Phenol-2-sulphonic acid, mixture of 2-nitrophenol and 4-nitrophenol (D) Phenol-2,4-disulphonic acid, mixture of 2-nitrophenol and 4-nitrophenol (E) Phenol-2,4-disulphonic acid and picric acid
›Reveal solutionSolution
Sulphonation of phenol gives phenol-2,4-disulphonic acid; subsequent nitration replaces the –SO3H groups to yield picric acid (2,4,6-trinitrophenol).
Treating phenol with concentrated H2SO4 introduces sulphonic acid groups, giving phenol-2,4-disulphonic acid (X). On treatment with concentrated HNO3, the readily displaceable sulphonic groups are replaced by nitro groups and the ring is further nitrated, producing picric acid (2,4,6-trinitrophenol, Y). …
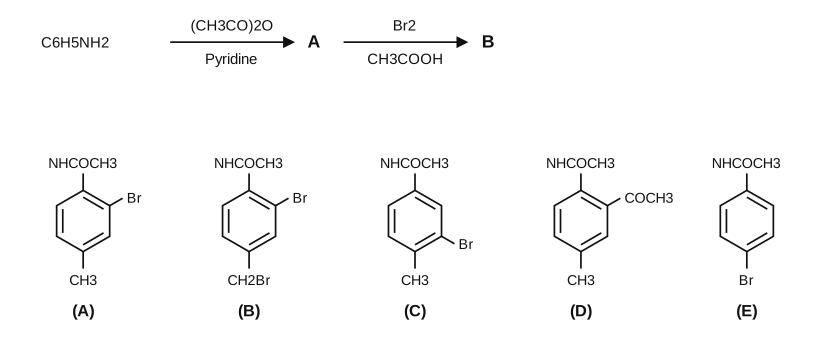
- KEAM 2025Set eng-2025-04284 marksMCQQ.In the following reaction, the final product B is C6H5NH2(CH3CO)2OPyridineABr2CH3COOHB (A) A benzene ring with NHCOCH3 at position 1, Br at position 2 (ortho), and CH3 at position 4 (para) (B) A benzene ring with NHCOCH3 at position 1, Br at position 2 (ortho), and CH2Br at position 4 (para) (C) A benzene ring with NHCOCH3 at position 1, Br at position 3 (meta), and CH3 at position 4 (D) A benzene ring with NHCOCH3 at position 1, COCH3 at position 2 (ortho), and CH3 at position 4 (para) (E) A benzene ring with NHCOCH3 at position 1 and Br at position 4 (para)
›Reveal solutionSolution
Acetylation moderates aniline; the acetamido group is an o/p-director and bromination gives mainly the para product.
C6H5NH2 + (CH3CO)2O/pyridine → acetanilide (A), C6H5NHCOCH3. The acetamido group is a strong ortho/para director; steric factors make the para product dominant. With Br2/CH3COOH the final product B is **p-bromoacetan …
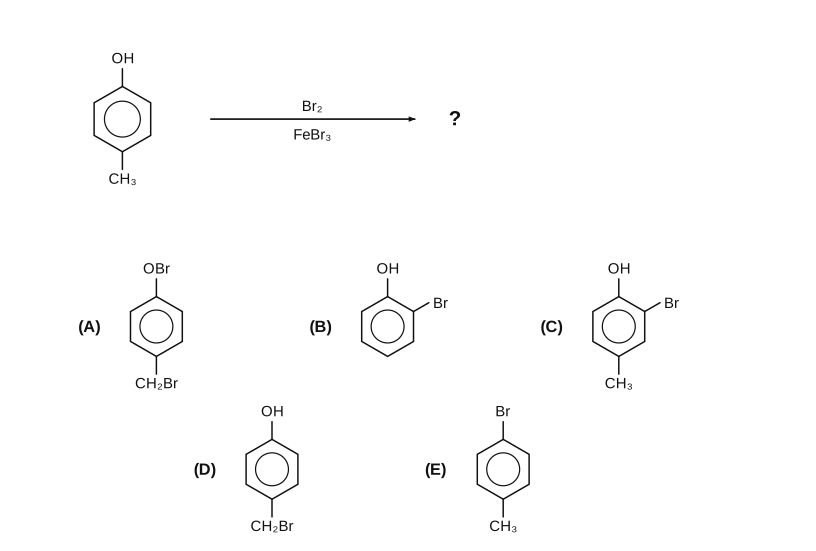
- KEAM 2025Set eng-2025-04294 marksMCQQ.What is the major product of the following reaction? 4-methylphenol (p-cresol) +Br2FeBr3 ? (A) a benzene ring with an -OBr group (para) and a -CH2Br group (B) phenol with a Br substituent ortho to the -OH (2-bromophenol) (C) a phenol (-OH) with a Br ortho to the OH and a -CH3 group para to the OH (2-bromo-4-methylphenol) (D) phenol (-OH) with a -CH2Br group at the para position (E) a benzene ring with a Br (para) and a -CH3 group (4-bromotoluene)
›Reveal solutionSolution
-OH activates and directs ortho/para. With the para position occupied by -CH3, electrophilic bromination goes ortho to the -OH, yielding 2-bromo-4-methylphenol.
p-Cresol is 4-methylphenol, with -OH and -CH3 para to each other. Both substituents are ortho/para directors, but the -OH group is a much stronger activator and controls the orientation. Its para position is already occupied by the methyl group, so electrophilic aromatic bromination (with Br2/FeBr3) occurs at the position ortho to the -OH. The major product is **2-bromo-4-methylphen …
- KEAM 2025Set pha-2025-0424A4 marksMCQQ.When chlorobenzene is treated with acetyl chloride in the presence of anhydrous AlCl3 , 4-Chloroacetophenone is formed as the major product. It is an example of (A) Nucleophilic substitution (B) Electrophilic substitution (C) Free radical substitution (D) Nucleophilic addition (E) Electrophilic addition
›Reveal solutionSolution
AlCl3 generates the acylium electrophile CH3CO+, which substitutes a ring hydrogen (para to Cl). This is electrophilic aromatic substitution.
CH3COCl+AlCl3→CH3CO++AlCl4−. The acylium ion attacks the electron-rich benzene ring of chlorobenzene at the para position (Cl is o,p-directing), for …
- KEAM 2024Set eng-2024-06084 marksMCQQ.An organic compound X (C6H6O) on reaction with zinc dust gives 'Y'. The product 'Y' reacts CH3COCl in presence of anhydrous AlCl3 to give 'Z' (C8H8O). The compounds X, Y and Z are respectively (A) benzaldehyde, benzene, methyl phenyl ketone (B) phenol, benzene, acetophenone (C) phenol, naphthalene, acetophenone (D) benzene, phenol, diphenyl ketone (E) cyclohexanol, cyclohexane, benzophenone
›Reveal solutionSolution
X = phenol, Y = benzene, Z = acetophenone: phenol is reduced by Zn dust to benzene, which undergoes Friedel–Crafts acylation to give acetophenone (C8H8O).
Identify X: C6H6O is phenol. Heating phenol with zinc dust reduces it (removes the –OH):
C6H5OH+Zn⟶C6H6+ZnO
so Y = benzene.
Benzene then undergoes Friedel–Crafts acylation with acetyl chloride and anhydrous AlCl3: …
🎓Unlock everything free for 14 days
- ✓Full step-by-step solutions
- ✓Concept-first explanations
- ✓Methods, shortcuts & mistakes
- ✓PYQ mapping + timed mock tests
Full access for 14 days. No credit card required.