Q.Write the reactions of Williamson synthesis of 2-ethoxy-3-methylpentane starting from ethanol and 3-methylpentan-2-ol.
Concept understanding — Williamson Ether Synthesis
Williamson Ether Synthesis: From Intuition to Mechanism
Imagine you want to build a simple bridge between two carbon chains — an oxygen atom linking them together. That bridge is an ether (R−O−R′). The Williamson ether synthesis is the most reliable way to build that bridge in a lab.
The Core Idea
You have two pieces: an alkoxide ion (RO−) and an alkyl halide (R′X). The alkoxide is a strong nucleophile — it loves positive charge. The alkyl halide has a carbon attached to a halogen (like Cl, Br, I) that is slightly positive because the halogen pulls electrons away.
When you mix them, the alkoxide attacks that slightly positive carbon, kicks out the halide ion, and forms a new C−O bond. The result? An ether.
R−O−+R′−X⟶R−O−R′+X−
That's the entire reaction in one line. But the devil is in the details — especially which alkyl halide you choose.
The Mechanism (SN2)
This is a classic SN2 reaction — one step, no intermediates. The alkoxide approaches the carbon from the opposite side of the halogen. As the C−O bond forms, the C−X bond breaks. The halide leaves as a stable anion.
Because it's SN2, the reaction is sensitive to steric hindrance. The carbon being attacked must be accessible.
If the alkyl halide is tertiary (3°), the reaction will not work via SN2. The bulky carbon blocks the backside attack. Instead, the alkoxide will act as a base and cause elimination (forming an alkene). You'll get no ether.
The Practical Rule
| Alkyl halide | Works? | Why |
|---|---|---|
| Methyl (CH3X) | Yes | Least hindered, fastest SN2 |
| Primary (1°) | Yes | Clean SN2 |
| Secondary (2°) | Sometimes | Works if not too bulky; elimination competes |
| Tertiary (3°) | No | Elimination dominates |
| Aryl (e.g., bromobenzene) | No | SN2 impossible on sp2 carbon |
To make an ether like R−O−R′, always use the less hindered alkyl halide and the more hindered alkoxide. For example, to make CH3CH2−O−CH(CH3)2, use CH3CH2O− (primary alkoxide) + (CH3)2CHBr (secondary halide) — not the other way around.
How to Choose the Alkoxide
You can't just buy alkoxide ions in a bottle. You make them by reacting an alcohol with a strong base like sodium hydride (NaH) or sodium metal.
ROH+NaH⟶RO−Na++H2
The alkoxide is then used immediately with the alkyl halide.
A Common Exam Trap
Students often try to make an ether by reacting two alcohols together. That doesn't work directly — you need one alcohol to become the nucleophile (alkoxide) and the other to become the electrophile (alkyl halide). The Williamson synthesis is asymmetric by design.
The Big Picture
Williamson ether synthesis is the go-to method for making unsymmetrical ethers (R−O−R′ where R=R′). It's reliable, high-yielding, and conceptually clean — as long as you respect the SN2 mechanism and avoid tertiary halides.
The one-line takeaway: An alkoxide attacks an alkyl halide in an SN2 reaction to form an ether — but only if the halide is primary or methyl.
Williamson ether synthesis is the standard method for making ethers, taught in the NCERT/CBSE Class 12 Chemistry chapter on Alcohols, Phenols and Ethers, and ‘Williamson synthesis mechanism’ or ‘Williamson ether synthesis limitations’ are frequently searched important-question topics for board exams, JEE Main and NEET. Knowing why tertiary halides fail in this SN2-based reaction is a common distinguishing question in competitive organic chemistry exams.
Why this formula?
Williamson Ether Synthesis: Why the Key Principles Hold
The Williamson Ether Synthesis is a classic method to prepare ethers. The core reaction is:
R-O−+R’-X→R-O-R’+X−
Where:
- R-O− is an alkoxide ion (strong nucleophile)
- R’-X is an alkyl halide (electrophile)
- X− is a halide ion (leaving group)
Let's break down why this works — the reasoning behind the key principles.
1. Why an Alkoxide (Not an Alcohol) is Needed
The Problem with Alcohols
Alcohols (R-OH) are weak nucleophiles. The oxygen has a partial negative charge, but the O–H bond is strong. If you mix an alcohol with an alkyl halide, the reaction is extremely slow or doesn't happen at all.
The Solution: Deprotonation
By treating the alcohol with a strong base (like NaH, Na, or KOH), you remove the proton:
R-OH+NaH→R-O−Na++H2
The alkoxide ion (R-O−) has a full negative charge on oxygen. This makes it:
- A much stronger nucleophile (higher electron density)
- More reactive toward the electrophilic carbon in the alkyl halide
Key takeaway: The alkoxide's full negative charge is what drives the reaction — it's not just about having oxygen, but about having a charged, electron-rich oxygen.
2. Why the Alkyl Halide Must Be Primary (or Methyl)
The Mechanism: SN2 is the Only Path
The Williamson synthesis proceeds exclusively via an SN2 mechanism (bimolecular nucleophilic substitution). This means:
- The nucleophile attacks the carbon from the backside
- The leaving group departs from the opposite side
- The reaction is concerted (one step, no intermediates)
Why Primary Halides Work Best
In SN2 reactions, the rate depends on steric hindrance:
| Alkyl Halide Type | Steric Hindrance | SN2 Reactivity |
|---|---|---|
| Methyl (CH3X) | Minimal | Very fast |
| Primary (RCH2X) | Low | Fast |
| Secondary (R2CHX) | Moderate | Slow |
| Tertiary (R3CX) | High | Does not occur |
Why Tertiary Halides Fail
With a tertiary halide, the bulky alkyl groups block the backside attack. Instead, the alkoxide (a strong base) will eliminate a proton from the halide, forming an alkene:
R-O−+R’3C-X→R-OH+alkene+X−
This is an E2 elimination — not the desired ether formation.
Key takeaway: The Williamson synthesis works only when the alkyl halide is primary or methyl because SN2 requires an unhindered backside.
3. Why the Leaving Group Must Be Good
The Role of the Halide
The halide (X−) must be a good leaving group — meaning it can stabilize the negative charge after departure.
| Halide | Leaving Group Ability | Reason |
|---|---|---|
| I− | Excellent | Large, polarizable, weak base |
| Br− | Good | Moderate size, weak base |
| Cl− | Fair | Smaller, stronger base |
| F− | Poor | Small, strong base, holds tightly |
Why Fluoride Fails
Fluoride is a strong base and a poor leaving group. The C–F bond is very strong, and F− does not depart easily. So alkyl fluorides are unreactive in Williamson synthesis.
Key takeaway: The leaving group must be weakly basic and polarizable — iodide and bromide are ideal.
4. Why the Alkoxide Must Be the Nucleophile (Not the Halide)
The "Wrong Way" Problem
If you try to use an alcohol as the nucleophile and an alkoxide as the leaving group, it won't work. Why?
- The alkoxide is a stronger base than the halide
- The halide is a better leaving group than the alkoxide
So the reaction is irreversible in the direction shown:
R-O−+R’-X→R-O-R’+X−
The reverse reaction (where X− attacks the ether) would require X− to be a nucleophile and R-O− to be a leaving group — but R-O− is a terrible leaving group (strong base).
Key takeaway: The reaction is driven by the difference in leaving group ability — halides leave easily, alkoxides do not.
Summary: The Three Pillars of Williamson Ether Synthesis
- Strong nucleophile (alkoxide, not alcohol) — full negative charge on oxygen
- Unhindered electrophile (primary or methyl halide) — SN2 requires backside access
- Good leaving group (iodide, bromide, or chloride) — halide must depart easily
If any of these conditions is violated, the reaction fails or gives elimination products.
Quick Exam Tip
When asked "Why does Williamson synthesis fail with tertiary halides?" — never say "because it's bulky." Say:
"Tertiary halides undergo E2 elimination instead of SN2 because the alkoxide acts as a strong base and the steric hindrance prevents backside attack."
This shows you understand the competition between substitution and elimination — a common exam trap.
The key idea is the Williamson ether synthesis: an alkoxide attacks a PRIMARY alkyl halide via SN2; using a secondary or tertiary halide instead mostly gives elimination. So the secondary alcohol (3-methylpentan-2-ol) must supply the alkoxide, and ethanol must supply the primary halide -- not the reverse.
Step 1: Convert 3-methylpentan-2-ol to its sodium alkoxide:
CH3CH2CH(CH3)CH(OH)CH3+Na→CH3CH2CH(CH3)CH(O−Na+)CH3+21H2
Step 2: Convert ethanol to the primary halide, ethyl bromide:
CH3CH2OH+HBr→CH3CH2Br+H2O
Step 3: SN2 reaction -- the alkoxide attacks the unhindered primary carbon of ethyl bromide:
CH3CH2CH(CH3)CH(O−Na+)CH3+CH3CH2Br→CH3CH2CH(CH3)CH(OC2H5)CH3+NaBr
The correct route reacts sodium 3-methylpentan-2-olate with ethyl bromide (never the reverse, which would give mostly elimination), giving CH3CH2CH(CH3)CH(OC2H5)CH3 (2-ethoxy-3-methylpentane).
Williamson ether synthesis is an SN2 reaction between an alkoxide ion and an alkyl halide. To make 2-ethoxy-3-methylpentane, the best route uses the less hindered alkoxide (from ethanol) reacting with the more hindered halide (from 3-methylpentan-2-ol), giving the ether in high yield.
The Core Idea: Williamson Ether Synthesis
Williamson ether synthesis is the most reliable laboratory method for making unsymmetrical ethers. The reaction is a straightforward SN2 substitution: an alkoxide ion (RO⁻) attacks an alkyl halide (R'X), displacing the halide and forming the ether R–O–R'.
The key constraint is that the alkyl halide must be primary (or methyl). Why? Because SN2 reactions are extremely sensitive to steric hindrance. A secondary or tertiary halide will mostly undergo elimination (forming an alkene) instead of substitution. The alkoxide, being a strong base, will deprotonate the halide's β-hydrogens rather than attack the carbon.
This gives us a critical rule: the alkoxide can be primary, secondary, or tertiary, but the alkyl halide must be primary (or methyl).
The Classic Mistake
Students often try to make the ether by using the alkoxide of the secondary alcohol and a primary halide. That works. But they also try the reverse — using a secondary halide — which fails due to elimination. Always check: is the halide primary?
Our Target: 2-ethoxy-3-methylpentane
Let's draw the structure. The name tells us:
- Parent: pentane (5-carbon chain)
- Substituents: a methyl group at carbon 3, and an ethoxy group (–O–CH₂CH₃) at carbon 2.
So the molecule is:
CH₃–CH₂–CH(CH₃)–CH(CH₃)–O–CH₂–CH₃
The ether linkage splits the molecule into two fragments:
- Fragment A (the alkoxy part): –O–CH₂CH₃ (ethoxy group)
- Fragment B (the alkyl part): the rest, which is 3-methylpentan-2-yl group
Two Possible Routes
We can make this ether in two ways, depending on which fragment becomes the alkoxide and which becomes the halide.
Route 1: Alkoxide from ethanol + halide from 3-methylpentan-2-ol
- Alkoxide: CH₃CH₂O⁻ (from ethanol)
- Halide: 3-methylpentan-2-yl halide (secondary halide)
Route 2: Alkoxide from 3-methylpentan-2-ol + halide from ethanol
- Alkoxide: 3-methylpentan-2-olate (secondary alkoxide)
- Halide: CH₃CH₂X (ethyl halide, primary)
Now apply the Williamson rule.
The Williamson Rule
The alkyl halide must be primary (or methyl) to avoid elimination. The alkoxide can be any type.
Route 1 uses a secondary halide — this is a disaster. The secondary halide will undergo E2 elimination with the strong ethoxide base, giving mostly 3-methylpent-2-ene. Very little ether forms.
Route 2 uses a primary halide (ethyl halide) — this is perfect. The secondary alkoxide attacks the unhindered primary carbon in a clean SN2 reaction. The ether forms in high yield.
The "Which Way?" Shortcut
When choosing between two routes for Williamson synthesis, always put the more hindered group as the alkoxide and the less hindered group as the halide. The alkoxide can be bulky; the halide must be small.
Step-by-Step Reactions (Route 2 — the correct one)
1. Prepare the alkoxide from 3-methylpentan-2-ol
We need to deprotonate the alcohol to make the alkoxide ion. A strong base like sodium metal or sodium hydride works well.
CH3CH2CH(CH3)CH(OH)CH3+Na⟶CH3CH2CH(CH3)CH(O−Na+)CH3+21H2
Or with NaH:
CH3CH2CH(CH3)CH(OH)CH3+NaH⟶CH3CH2CH(CH3)CH(O−Na+)CH3+H2
2. Prepare the primary alkyl halide from ethanol
Ethanol reacts with a halogenating agent like PBr₃ or HBr to give ethyl bromide.
CH3CH2OH+PBr3⟶CH3CH2Br+H3PO3
Or simply:
CH3CH2OH+HBrΔCH3CH2Br+H2O
3. Perform the Williamson ether synthesis
Now the key step: the alkoxide (from step 1) attacks the primary alkyl halide (from step 2) in an SN2 reaction.
CH3CH2CH(CH3)CH(O−Na+)CH3+BrCH2CH3⟶CH3CH2CH(CH3)CH(OCH2CH3)CH3+NaBr
The product is 2-ethoxy-3-methylpentane.
›Proof
Why Route 1 fails
If we tried Route 1, the second step would be:
CH3CH2O−Na++BrCH(CH3)CH(CH3)CH2CH3⟶elimination products (alkenes)+very little ether
The secondary halide has β-hydrogens, and the ethoxide base abstracts them preferentially. The major products are 3-methylpent-2-ene and 3-methylpent-1-ene, not the desired ether.
Summary of the Correct Reactions
| Step | Reactants | Product |
|---|---|---|
| 1 | 3-methylpentan-2-ol + Na (or NaH) | Sodium 3-methylpentan-2-olate |
| 2 | Ethanol + HBr (or PBr₃) | Ethyl bromide |
| 3 | Sodium 3-methylpentan-2-olate + ethyl bromide | 2-ethoxy-3-methylpentane + NaBr |
The correct Williamson synthesis uses sodium 3-methylpentan-2-olate (from 3-methylpentan-2-ol and Na) reacting with ethyl bromide (from ethanol and HBr) to give 2-ethoxy-3-methylpentane via SN2.
Williamson Ether Synthesis — Method & Steps
Method: Williamson Ether Synthesis (an SN2 reaction between an alkoxide ion and a primary alkyl halide / tosylate).
Core Concept
The ether oxygen comes from the alkoxide (the more acidic alcohol is deprotonated), and the alkyl group comes from the alkyl halide (the less hindered carbon is attacked).
Step-by-Step Plan for 2-ethoxy-3-methylpentane
Target ether:
CH3CH2O−CH(CH3)CH(CH3)CH2CH3
Two possible disconnections:
| Route | Alkoxide from | Alkyl halide from |
|---|---|---|
| A | Ethanol (pKa ~16) | 3-methylpentan-2-ol → 2-bromo-3-methylpentane |
| B | 3-methylpentan-2-ol (pKa ~16–18) | Ethanol → bromoethane |
Choose Route A — because the alkyl halide is secondary in Route B, which would give elimination (E2) as the major product. Williamson works best with primary alkyl halides.
Reactions (Route A)
Step 1: Form the alkoxide from ethanol
CH3CH2OH+Na⟶CH3CH2O−Na++21H2
Step 2: Convert 3-methylpentan-2-ol to a primary alkyl halide
First, convert the alcohol to a tosylate (better leaving group), then displace with bromide:
CH3CH(OH)CH(CH3)CH2CH3TsCl, pyridineCH3CH(OTs)CH(CH3)CH2CH3
CH3CH(OTs)CH(CH3)CH2CH3+NaBr⟶CH3CH(Br)CH(CH3)CH2CH3+NaOTs
Step 3: Williamson coupling
CH3CH2O−Na++CH3CH(Br)CH(CH3)CH2CH3ΔCH3CH2OCH(CH3)CH(CH3)CH2CH3+NaBr
Final product: 2-ethoxy-3-methylpentane ✓
Key Exam Point
Always use the alkoxide from the smaller alcohol and the alkyl halide from the larger alcohol — this ensures the SN2 step occurs on a primary carbon, avoiding elimination.
Here are the most common mistakes students make with this specific Williamson Ether Synthesis problem, and how to avoid each.
1. Mistake: Choosing the Wrong Alkoxide/Alkyl Halide Pair
The biggest error is not recognizing that two different ethers can form from the given alcohols, but only one is the target.
-
The Trap: Students often react ethanol with 3-methylpentan-2-ol directly, forgetting that one alcohol must be converted to an alkoxide (strong base) and the other to an alkyl halide (leaving group).
-
The Correct Logic: You have two alcohols. You must decide which one becomes the alkoxide (the nucleophile) and which one becomes the alkyl halide (the electrophile).
- Option A: Ethanol → Ethoxide + 2-bromo-3-methylpentane
- Option B: 3-methylpentan-2-ol → 3-methylpentan-2-oxide + Bromoethane
-
How to Avoid: Always check for steric hindrance. The Williamson synthesis works best when the alkyl halide is primary (or methyl). A secondary or tertiary alkyl halide will undergo elimination (E2) instead of substitution (SN2).
- In this case, 3-methylpentan-2-ol is a secondary alcohol. Converting it to an alkyl halide (2-bromo-3-methylpentane) and reacting it with ethoxide will give elimination products (alkenes), not the desired ether.
- Correct choice: Use ethanol as the alkyl halide (bromoethane, a primary halide) and 3-methylpentan-2-oxide as the alkoxide.
2. Mistake: Forgetting to Deprotonate the Alcohol First
Students often write the reaction as: Alcohol + Alkyl Halide → Ether. This is wrong.
- The Trap: Writing
CH3CH2OH + Br-CH(CH3)CH(CH3)CH2CH3 → Etherwithout a base. - The Correct Logic: The oxygen in an alcohol is a poor nucleophile. It must be converted into a strong nucleophile (alkoxide ion, RO−) by reacting with a strong base (like NaH, Na metal, or KOH).
- How to Avoid: Always write the two-step process clearly:
- Formation of alkoxide: R-OH+NaH→R-O−Na++H2
- SN2 reaction: R-O−Na++R′-X→R-O-R′+NaX
3. Mistake: Incorrect Naming of the Target Ether
The name "2-ethoxy-3-methylpentane" tells you exactly which part is the alkoxy group and which is the parent alkane.
- The Trap: Students might try to make the ether by joining the two alcohols in the wrong order, leading to a different structural isomer (e.g., 3-methylpentan-2-oxyethane, which is the same molecule but named incorrectly, or a completely different ether).
- The Correct Logic:
- Ethoxy (CH3CH2O−) is the substituent. This comes from ethanol.
- 3-methylpentane is the parent chain. This comes from 3-methylpentan-2-ol (the oxygen is on carbon #2 of the pentane chain).
- How to Avoid: Break the ether name into two parts:
- Alkoxy group: "ethoxy" → CH3CH2O−
- Parent alkane: "3-methylpentane" → CH3CH2CH(CH3)CH2−
- The oxygen is attached to carbon #2 of the parent, so the alkoxide must be derived from 3-methylpentan-2-ol.
4. Mistake: Writing the Wrong Alkyl Halide Structure
Even if you choose the correct alcohol to be the halide (ethanol), you must write the correct halide structure.
- The Trap: Writing bromoethane as CH3CH2Br is fine, but students sometimes write it as BrCH2CH3 (which is the same) or, worse, confuse it with the structure of the other alcohol.
- The Correct Logic: Ethanol (CH3CH2OH) becomes bromoethane (CH3CH2Br). The 3-methylpentan-2-ol (CH3CH2CH(CH3)CH(OH)CH3) becomes the alkoxide (CH3CH2CH(CH3)CH(O−)CH3).
- How to Avoid: Draw the full structural formula for each reactant before writing the reaction. Double-check that the carbon skeleton of the alkyl halide matches the alcohol you intend to use.
Summary: The Correct Reaction
Step 1: Formation of the alkoxide (from the secondary alcohol)
CH3CH2CH(CH3)CH(OH)CH3+NaH→CH3CH2CH(CH3)CH(O−Na+)CH3+H2
Step 2: SN2 reaction with the primary alkyl halide (from ethanol)
CH3CH2CH(CH3)CH(O−Na+)CH3+CH3CH2Br→CH3CH2CH(CH3)CH(OCH2CH3)CH3+NaBr
The final product is 2-ethoxy-3-methylpentane.
Showing the 12 most recent of 27 on this concept.
- TG EAPCET 2026Set eng-2026-05-09-AN1 markMCQQ.Which of the following statements are correct? I. Liquid sodium metal is used as coolant in fast breeder nuclear reactor II. LiCl is deliquescent III. LiF and CsI have low solubility in water (A) I, III only (B) II, III only (C) I, II only (D) I, II, III
›Reveal solutionSolution
All three statements are correct: liquid sodium is used as a coolant in fast breeder reactors, LiCl is deliquescent, and both LiF and CsI have low solubility in water. The correct option is (D).
The question asks us to identify the correct statements regarding properties of alkali metals and their compounds. We will evaluate each statement based on fundamental chemical principles.
Concept and Intuition
- Coolants in Nuclear Reactors: A good coolant needs to efficiently transfer heat, remain liquid over a wide temperature range, and ideally have a low neutron absorption cross-section in certain reactor types.
- Deliquescence: This property describes a substance that absorbs moisture from the atmosphere until it dissolves in the absorbed water to form a solution. It is related to hygroscopy (absorbing moisture) and the ability to form stable hydrates.
- Solubility in Water: The solubility of an ionic compound in water is determined by the balance between its lattice energy (energy required to break the ionic lattice) and its hydration energy (energy released when ions are surrounded by water molecules). For a compound to dissolve, the hydration energy must be sufficient to overcome the lattice energy.
Step-by-Step Evaluation
1. Statement I: Liquid sodium metal is used as coolant in fast breeder nuclear reactor
- Reasoning: Fast breeder reactors operate at very high temperatures and require a coolant that can efficiently remove heat without moderating (slowing down) the fast neutrons. Liquid sodium is an excellent choice for this purpose due to several key properties:
- High Thermal Conductivity: It can transfer heat very efficiently.
- Wide Liquid Range: Sodium has a relatively low melting point (97.8∘C) and a very high boiling point (883∘C), allowing it to remain liquid over a broad and useful temperature range for reactor operation.
- Low Neutron Absorption Cross-section: It does not significantly absorb neutrons, which is crucial for maintaining the neutron economy in a fast breeder reactor.
- Low Viscosity: This allows for easy pumping and circulation.
- Conclusion: Statement I is correct.
2. Statement II: LiCl is deliquescent
- Reasoning: Deliquescence is the property of a substance to absorb moisture from the atmosphere and dissolve in it. This property is strongly linked to the hygroscopic nature of a compound and its ability to form stable hydrates.
- Lithium chloride (LiCl) is known to be highly hygroscopic and readily forms hydrates, such as LiCl⋅H2O, LiCl⋅2H2O, and LiCl⋅3H2O.
- The small size and high charge density of the Li+ ion lead to a very high hydration energy. This strong interaction with water molecules drives the absorption of moisture from the air.
- Among the alkali metal chlorides, LiCl is the most deliquescent.
- Conclusion: Statement II is correct.
3. Statement III: LiF and CsI have low solubility in water
- Reasoning: The solubility of an ionic compound in water depends on the relative magnitudes of its lattice energy and hydration energy.
- For LiF:
- Li+ is the smallest alkali metal ion, and F− is the smallest halide ion.
- The very small sizes of both ions lead to a very short inter-ionic distance and, consequently, an exceptionally high lattice energy for LiF.
- While Li+ and F− both have high hydration energies due to their small sizes, the extremely high lattice energy of LiF dominates. The energy released upon hydration is not sufficient to overcome the energy required to break the strong ionic bonds in the crystal lattice.
- Therefore, LiF has very low solubility in water.
- For CsI:
- Cs+ is the largest alkali metal ion, and I− is the largest halide ion.
- The large sizes of both ions lead to a relatively long inter-ionic distance and thus a relatively low lattice energy compared to compounds with smaller ions.
- However, the hydration energies of very large ions like Cs+ and I− are also very low because their charge densities are low, leading to weak interactions with water molecules.
- In the case of CsI, the low hydration energy is not sufficient to overcome even the relatively low lattice energy. The energy released upon hydration is less than the energy required to break the lattice.
- Therefore, CsI also has low solubility in water.
- For LiF:
- Conclusion: Statement III is correct.
Since all three statements (I, II, and III) are correct, the option that includes all of them is the answer.
✓Final answerAll the given statements are correct, so the correct option is (D).
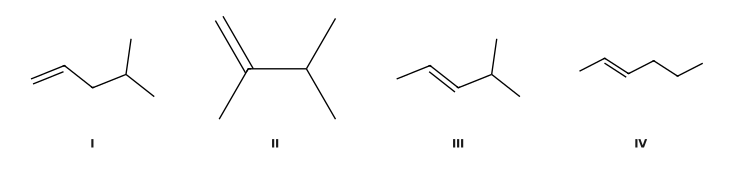
- TG EAPCET 2026Set eng-2026-05-09-AN1 markMCQQ.Which of the following are Position isomers? [FIGURE] (A) I, III (B) II, IV (C) II, III (D) I, IV
›Reveal solutionSolution
Among these C6H12 alkenes, only I (4-methylpent-1-ene) and III (4-methylpent-2-ene) have the same carbon skeleton with the double bond in different places — option (A).
The concept first
All four compounds are C6H12, so they are all isomers of one another. The question is which kind. Two structural isomers are:
- Chain (skeletal) isomers if the carbon framework itself differs (branching pattern, length of the main chain);
- Position isomers if the framework is identical and only the location of the functional group (here the C=C) changes.
So the working method is mechanical: hydrogenate each structure in your head (i.e. ignore the double bond and look only at the carbon skeleton). Any two that collapse to the same alkane are position isomers; ones that collapse to different alkanes are chain isomers.
Step-by-step
- Reduce each to its skeleton.
- I: (CH3)2CH−CH2−CH=CH2→ 2-methylpentane skeleton (5-carbon chain, methyl branch on the carbon next to one end).
- II: CH2=C(CH3)−CH(CH3)2→ 2,3-dimethylbutane skeleton (4-carbon chain, two methyl branches).
- III: (CH3)2CH−CH=CH−CH3→ 2-methylpentane skeleton — same as I.
- IV: unbranched hexene → n-hexane skeleton (no branch).
- Group by skeleton.
- 2-methylpentane skeleton: {I, III}
- 2,3-dimethylbutane skeleton: {II}
- n-hexane skeleton: {IV}
- Check the double-bond positions within the matching pair. Numbering the main chain of I and III so the double bond gets the lower locant:
- I = 4-methylpent-1-ene
- III = 4-methylpent-2-ene Same skeleton, C=C shifted from position 1 to position 2 → position isomers ✓.
- Reject the rest. II vs anything, and IV vs anything, involve a change in the carbon framework, which makes them chain isomers, not position isomers.
✓Final answerThe position-isomeric pair is I and III, so the correct option is (A).
ANSWER: A
- TG EAPCET 2026Set eng-2026-05-09-AN1 markMCQQ.What are X, Y, Z respectively in the following set of reactions? (A) [benzene ring]-Br ; [ethanol] ; [benzene ring]-O-[ethyl group]-Br (B) [benzene ring]-OH ; [ethyl bromide] ; [benzene ring]-O-[ethyl group]-Br (C) [benzene ring]-OH ; [ethyl bromide] ; [benzene ring]-(Br)2-O-[ethyl group] (D) [benzene ring]-Br ; [ethyl bromide] ; [benzene ring]-O-[ethyl group]-Br
›Reveal solutionSolution
The reaction sequence is the Williamson ether synthesis: phenol (X) reacts with ethyl bromide (Y) in the presence of a base to give phenetole, which then undergoes bromination at the para position to yield p-bromophenetole (Z). The correct option is (B).
The question presents a set of reactions where three unknown compounds X, Y, and Z are to be identified. The key is to recognize the classic Williamson ether synthesis followed by electrophilic aromatic substitution. Let's walk through the logic.
-
Identify the first reaction: X + NaOH → sodium phenoxide
The product after treatment with NaOH is a sodium salt of an aromatic alcohol. This is a characteristic reaction of phenol (C6H5OH). Phenol reacts with NaOH to form sodium phenoxide (C6H5ONa). So X must be phenol, not bromobenzene (which does not react with NaOH under these conditions). This immediately eliminates options (A) and (D), which list X as bromobenzene.
-
Identify the second reaction: sodium phenoxide + Y → an ether
Sodium phenoxide is a strong nucleophile. It reacts with an alkyl halide in an SN2 reaction to form an ether — this is the Williamson ether synthesis. The product shown is an aromatic ether with an ethyl group attached to the oxygen. Therefore Y must be ethyl bromide (C2H5Br). The product is ethyl phenyl ether, commonly called phenetole (C6H5OC2H5).
-
Identify the third reaction: phenetole + Br2 → Z
Phenetole has an activated benzene ring (the -OCH2CH3 group is strongly activating and ortho/para-directing). Bromination occurs predominantly at the para position due to steric hindrance at the ortho positions. The product Z is p-bromophenetole, which has the structure Br−C6H4−OC2H5 (with bromine para to the ethoxy group). This matches the description in option (B): [benzene ring]-O-[ethyl group]-Br.
Watch outA common mistake is to think that bromobenzene could be X because it contains bromine. But bromobenzene does not react with NaOH to form a sodium salt — it requires much harsher conditions (high pressure, high temperature) for nucleophilic substitution. The mild NaOH step here is a dead giveaway for phenol.
TipThe Williamson ether synthesis is the most reliable method for preparing unsymmetrical ethers. The alkoxide (from the alcohol/phenol) must be the nucleophile, and the alkyl halide should be primary to avoid elimination. Here, ethyl bromide is primary, so SN2 proceeds cleanly.
✓Final answerThe correct option is (B): X = phenol, Y = ethyl bromide, Z = p-bromophenetole.
-
- TG EAPCET 2026Set eng-2026-05-10-FN1 markMCQQ.Consider the following reactions Borax H2O A + B Borax H2O/HCl B + C Which of the following are the properties of A, B, C? I. Both A, B are strong bases II. B is a white crystalline solid with soapy touch III. C is water soluble (A) I, II, III (B) II, III only (C) I, III only (D) I, II only
›Reveal solutionSolution
Borax hydrolyses in water to give boric acid and a strong base; with HCl it gives boric acid and NaCl. The correct properties are II and III only — option (B).
The key is to understand what borax (sodium tetraborate decahydrate, Na2B4O7⋅10H2O) does in water and in acidic water. Borax is not a simple salt — it reacts with water to produce boric acid and a strong base. When you add HCl, the base gets neutralised, leaving a different set of products.
Let’s write the reactions clearly.
- Borax in pure water Borax dissolves and hydrolyses:
Na2B4O7+7H2O→2NaOH+4H3BO3
Here, A is NaOH (sodium hydroxide) and B is H3BO3 (boric acid).
So A is a strong base, B is a weak acid — not a base at all.
- Borax in water with HCl The same hydrolysis occurs, but the HCl neutralises the NaOH formed:
Na2B4O7+2HCl+5H2O→2NaCl+4H3BO3
Here, B is again H3BO3, and C is NaCl (sodium chloride).
Now evaluate each statement:
-
I. Both A, B are strong bases
A is NaOH (strong base), but B is boric acid — a very weak acid, not a base. So I is false.
-
II. B is a white crystalline solid with soapy touch
Boric acid (H3BO3) is indeed a white crystalline solid. It feels slippery or soapy to the touch (a property of many boron compounds). This is correct.
-
III. C is water soluble
C is NaCl, common salt — highly soluble in water. This is correct.
Only statements II and III are true.
Watch outA common mistake is to think boric acid is a base because it feels soapy. In fact, it is a weak acid (it accepts OH− from water, not donates H+ in the usual sense). Soapy touch does not imply basicity.
✓Final answerThe correct option is (B) — only II and III are true.
- TG EAPCET 2026Set eng-2026-05-10-FN1 markMCQQ.Which of the following is most reactive towards SN2 reaction? (A) (CH3)3C−Br (tert-butyl bromide) (B) CH3CH2CH2CH2−Br (1-bromobutane) (C) (CH3)2CH−CH2−Br (1-bromo-2-methylpropane) (D) CH3CH2−CH(Br)−CH3 (2-bromobutane) 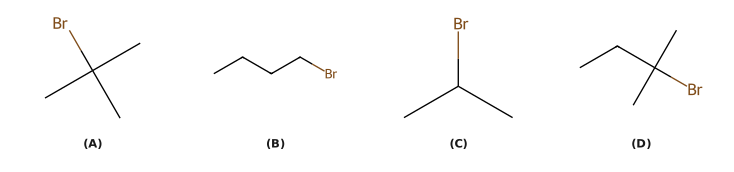
›Reveal solutionSolution
SN2 needs an unobstructed backside approach, so steric bulk kills it. The unbranched primary halide 1-bromobutane is the least hindered and therefore the most reactive — option (B).
The concept first
In the SN2 mechanism, bond-making and bond-breaking happen simultaneously in one step. The nucleophile attacks the electrophilic carbon from the side directly opposite the leaving group (backside attack), passing through a transition state in which that carbon is momentarily bonded to five groups:
Nu−+ R−Br ⟶ [Nu⋯C⋯Br]‡ ⟶ Nu−R+Br−
The carbon is sp2-like in that transition state with the three remaining groups splayed out in a plane. Every extra alkyl group attached to that carbon (or even to the carbon next to it) makes the transition state more crowded, raises its energy, and slows the reaction:
Rate: CH3X > 1∘ > 2∘ ≫ 3∘
This is the exact opposite of the SN1 order (which follows carbocation stability, 3∘>2∘>1∘). Knowing which mechanism the question names is half the battle.
Step-by-step through the options
(A) (CH3)3C−Br, tert-butyl bromide — tertiary. Three methyl groups completely block the backside of the C–Br carbon. Nucleophiles simply cannot get in; this halide reacts almost exclusively by SN1 (it forms a very stable 3∘ carbocation). Slowest for SN2. ✗
(B) CH3CH2CH2CH2−Br, 1-bromobutane — primary and unbranched. The carbon bearing Br carries only two hydrogens and one straight chain, so the backside is wide open. Fastest of the four. ✓
(C) (CH3)2CH−CH2−Br, 1-bromo-2-methylpropane (isobutyl bromide) — primary but β-branched. Although the carbon holding the Br is primary, the bulky isopropyl group on the very next carbon leans over the reaction site and hinders the incoming nucleophile. Isobutyl bromide reacts several times more slowly than n-butyl bromide in SN2. ✗
(D) CH3CH2−CH(Br)−CH3, 2-bromobutane — secondary. Two alkyl groups on the reacting carbon; distinctly slower than any primary halide. ✗
The ordering
1∘, unbranched1-bromobutane > 1∘, β-branched1-bromo-2-methylpropane > 2∘2-bromobutane ≫ 3∘tert-butyl bromide
A one-line way to remember it: SN2 hates crowds.
✓Final answerThe unhindered primary halide 1-bromobutane undergoes SN2 fastest, so the correct option is (B).
ANSWER: B
- TG EAPCET 2026Set ap-2026-05-05-FN1 markMCQQ.In the formation of photochemical smog, two oxides of nitrogen are significantly formed. The oxidation states of nitrogen in these oxides are respectively (A) +1, +4 (B) +2, +4 (C) +2, +5 (D) +1, +2
›Reveal solutionSolution
Photochemical smog forms when nitrogen oxides (NO and NO₂) from vehicle exhaust react with sunlight. The oxidation states of nitrogen in these two key oxides are +2 and +4, making option (B) correct.
The question is about the chemistry of photochemical smog — a type of air pollution that forms when sunlight triggers reactions between pollutants, especially from car engines. The two oxides of nitrogen that play the starring role here are nitric oxide (NO) and nitrogen dioxide (NO₂). They are produced in high-temperature combustion (e.g., in a car engine) when nitrogen and oxygen from the air combine.
To find the oxidation states, we apply the standard rule: oxygen almost always has an oxidation state of –2 (except in peroxides, which aren't relevant here). For a neutral molecule, the sum of oxidation states must be zero.
-
For NO: Let the oxidation state of nitrogen be x.
x+(−2)=0⟹x=+2.
So nitrogen is in the +2 oxidation state in nitric oxide.
-
For NO₂: Let the oxidation state of nitrogen be y.
y+2(−2)=0⟹y−4=0⟹y=+4.
So nitrogen is in the +4 oxidation state in nitrogen dioxide.
Watch outA common mistake is to confuse NO₂ (nitrogen dioxide, +4) with N₂O (nitrous oxide, +1) or N₂O₅ (dinitrogen pentoxide, +5). But in photochemical smog, the two oxides formed directly from combustion are NO and NO₂ — not these others.
These two oxides are the primary precursors: NO is released first, then quickly oxidised to NO₂ in the air. NO₂ absorbs sunlight and breaks down, starting a chain of reactions that produce ozone and other components of photochemical smog.
✓Final answerThe correct option is (B) +2, +4.
-
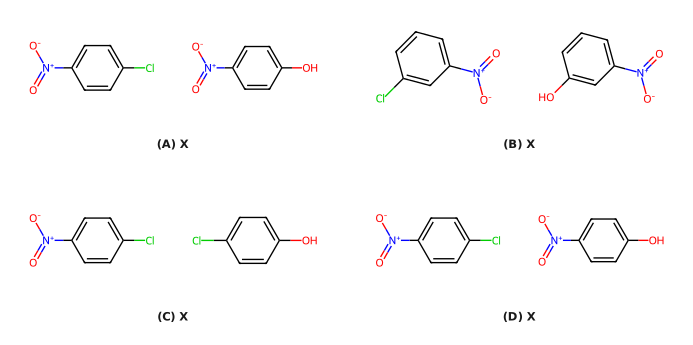
- TG EAPCET 2026Set ap-2026-05-05-FN1 markMCQQ.What are X, Y, Z respectively in the following reaction sequence? (Conc. = concentrated) Chlorobenzene HNO3conc. H2SO4XYZ (A) X = 1-chloro-4-nitrobenzene (O2N−C6H4−Cl, para) ; Y =(i) NaOH∣443 K(ii) H+ ; Z = 4-nitrophenol (O2N−C6H4−OH, para) (B) X = 1-chloro-3-nitrobenzene (Cl and NO2 meta) ; Y =(i) NaOH∣368 K(ii) H+ ; Z = 3-nitrophenol (OH and NO2 meta) (C) X = 1-chloro-4-nitrobenzene (para) ; Y =(i) NaOH∣300 K(ii) H+ ; Z = 4-chlorophenol (HO−C6H4−Cl, para) (D) X = 1-chloro-4-nitrobenzene (para) ; Y = H2O∣300 K ; Z = 4-nitrophenol (para)
›Reveal solutionSolution
Nitration of chlorobenzene gives mainly 1-chloro-4-nitrobenzene (X); the para nitro group makes the C−Cl bond susceptible to nucleophilic attack, so NaOH at 443 K followed by H+ converts it to 4-nitrophenol (Z). That is option (A).
The concept first
- Electrophilic step. Halogens are deactivating but ortho–para directing: the −I effect withdraws electron density (slowing the reaction), while the +R (lone-pair) effect places negative charge at the o and p positions, so the electrophile goes there. Steric crowding makes para the major product.
- Nucleophilic step. Aryl halides resist SN reactions (partial double-bond character of C−Cl, resonance stabilisation). Chlorobenzene itself needs 623 K/300 atm NaOH (Dow process). A strongly electron-withdrawing −NO2 at the ortho or para position changes everything: it stabilises the negatively-charged carbanion (Meisenheimer) intermediate by resonance, so the reaction now runs under much milder conditions (NaOH, 443 K). A meta nitro group gives no such stabilisation.
Step-by-step
- C6H5ClHNO3conc. H2SO4 chiefly p-nitrochlorobenzene (with a little ortho).
X=1-chloro-4-nitrobenzene
This already eliminates the meta option.
2. Treat X with aqueous NaOH at 443 K: OH− adds at the carbon bearing Cl; the resulting carbanion is delocalised onto the para −NO2 oxygen. Loss of Cl− restores aromaticity, giving sodium p-nitrophenoxide.
3. Acidify: H+ protonates the phenoxide.
Z=4-nitrophenol, O2N−C6H4−OH
- Why the other options fail.
- 300 K NaOH giving 4-chlorophenol: the NO2, not the Cl, would have to leave — nitro is a very poor leaving group here, and 300 K is far too mild for aryl-halide substitution.
- H2O at 300 K: water is far too weak a nucleophile; nothing happens.
- meta isomer: contradicts the o,p-directing power of Cl.
✓Final answerThe sequence is chlorobenzene → p-nitrochlorobenzene → (NaOH, 443 K; then H+) → p-nitrophenol, so the correct option is (A).
ANSWER: A
- TG EAPCET 2025Set eng-2025-05-02-FN1 markMCQQ.The major products P and Q from the following reactions are P(i) LiAlH4(ii) H2O C6H5CONH2 Br2 / NaOHQ (A) P=C6H5NH2 ; Q=C6H5CH2NH2 (B) P=C6H5CH2NH2 ; Q=C6H5NH2 (C) P=C6H5−OH∣CH−NH2 ; Q=C6H5COONa (D) P=C6H5CN ; Q=C6H5Br
›Reveal solutionSolution
LiAlH4 reduces benzamide to benzylamine (P), while Br2/NaOH degrades it to aniline (Q), one carbon shorter. That is option (B).
The concept first
Benzamide, C6H5CONH2, has two very different fates depending on whether you attack the carbon or the nitrogen.
- LiAlH4 delivers H− to the electrophilic carbonyl carbon. The C–N bond is never broken; the oxygen leaves and the carbon ends up as a CH2 bridging the ring and the nitrogen. Amides are the only carbonyl compounds reduced all the way to an amine rather than an alcohol — that is worth remembering.
- Br2/NaOH instead brominates the nitrogen. The resulting N-bromoamide is deprotonated, loses Br− to give a nitrene-like species, and the phenyl group migrates from carbon to nitrogen. The old carbonyl carbon ends up as an isocyanate carbon and is finally hydrolysed away as Na2CO3. Hence a one-carbon shortening.
Step-by-step
- Reduction (P):
C6H5CONH2(i) LiAlH4 (ii) H2O C6H5CH2NH2
Benzylamine, a primary amine with 7 carbons (same as benzamide).
2. Hofmann degradation (Q):
C6H5CONH2+Br2+4NaOH→C6H5NH2+Na2CO3+2NaBr+2H2O
Aniline, a primary amine with 6 carbons (one less).
3. Mechanistic checkpoints for Q: C6H5CONHBr→C6H5CONˉBr→ phenyl migration →C6H5N=C=O (phenyl isocyanate) → hydrolysis → aniline.
4. Match to the options: P=C6H5CH2NH2, Q=C6H5NH2 — exactly option (B). Option (A) swaps them; (C) invents an unstable hemiaminal; (D) has no reagent that could make a nitrile or an aryl bromide here.
✓Final answerBenzamide gives benzylamine on LiAlH4 reduction and aniline on Hofmann degradation, so the correct option is (B).
ANSWER: B
- TG EAPCET 2025Set eng-2025-05-03-FN1 markMCQQ.Identify the product 'Y' in the given sequence of reactions. Chlorobenzene HNO3Conc. H2SO4 X (Major) (i) NaOH, 443 K(ii) H+ Y (A) p-Nitrophenol (benzene ring with −OH and −NO2 para to each other) (B) o-Nitrophenol (benzene ring with −OH and −NO2 ortho to each other) (C) p-Hydroxybenzenesulphonic acid (benzene ring with −OH and −SO3H para to each other) (D) 2,4-Dinitrophenol (benzene ring with −OH and −NO2 groups at positions 2 and 4)
›Reveal solutionSolution
Chlorobenzene nitrates mainly at the para position to give p-nitrochlorobenzene (X); the para −NO2 activates the C–Cl carbon, so NaOH at 443 K followed by H+ gives p-nitrophenol — option (A).
The concept first
Chlorine on a ring plays a double game. Through its −I effect it deactivates the ring, but through lone-pair +R donation it directs an incoming electrophile to the ortho and para positions. So nitration of chlorobenzene is slower than that of benzene, yet its major product is the para isomer (ortho is crowded).
The second step is the interesting one. Aryl halides are normally inert to nucleophiles — the C–Cl bond has partial double-bond character and the ring is electron-rich. Nucleophilic aromatic substitution becomes easy only when a strong electron-withdrawing group sits ortho or para to the halogen, because then the negative charge of the intermediate carbanion (the Meisenheimer complex) can be delocalised onto that group.
Step-by-step
Step 1 — nitration (X).
C6H5Cl HNO3conc. H2SO4 p-O2N-C6H4-Cl (major) + o-isomer (minor)
The electrophile is NO2+, formed as HNO3+2H2SO4→NO2++H3O++2HSO4−. So X=p-nitrochlorobenzene. Note H2SO4 acts only as the catalyst — it does not sulphonate the ring, which removes option (C).
Step 2 — why 443 K suffices. After OH− attacks the C–Cl carbon, the resulting carbanion can push its charge onto the nitro oxygens (only possible from the ortho or para position). That extra stabilisation drops the activation energy enormously, so the harsh Dow conditions (623 K, 300 atm) are no longer needed: 443 K is enough.
Step 3 — substitution and acidification (Y).
p-O2N-C6H4-Cl NaOH443 K p-O2N-C6H4-O−Na+ H+ p-O2N-C6H4-OH
Step 4 — eliminate the rest.
- (B) o-Nitrophenol would come from the minor ortho nitration product, not the major one.
- (C) A sulphonic acid group cannot form under nitrating conditions.
- (D) 2,4-Dinitrophenol needs two nitro groups; only one nitration is shown.
✓Final answerThe sequence runs chlorobenzene → p-nitrochlorobenzene → p-nitrophenol, so the correct option is (A).
ANSWER: A
- TG EAPCET 2025Set eng-2025-05-03-FN1 markMCQQ.What is ‘Z’ in the given set of reactions?
[!FORMULA] C6H5OCH3XYHIX+YZn,Δ(benzene ring)C6H6Anhy. AlCl3Z
(A) \chemfig{*6(-=-(-CH_3)-=-)} (B) \chemfig{*6(-=-(-CH_2Cl)-=-)} (C) \chemfig{*6(-=-(-C_2H_5)-=-)} (D) \chemfig{*6(-=-(-Cl)-=-)}›Reveal solutionSolution
The reaction sequence converts anisole into iodobenzene and methanol, then iodobenzene reacts with benzene in a Friedel–Crafts alkylation to give diphenylmethane, which is toluene — so Z is methylbenzene, option (A).
Concept & Intuition
This problem tests your understanding of ether cleavage by HI and subsequent aromatic substitution. Anisole (C₆H₅OCH₃) is an aryl methyl ether. When treated with HI, the C–O bond breaks selectively because the aryl–O bond is stronger (partial double-bond character from resonance with the ring). The products are iodobenzene (X) and methanol (Y). Then, X (iodobenzene) is reduced by Zn dust to benzene. Meanwhile, Y (methanol) reacts with benzene in the presence of anhydrous AlCl₃ — a Friedel–Crafts alkylation — to give toluene (methylbenzene). The key insight: methanol generates a methyl carbocation under these conditions, which attacks benzene.
Step-by-step reasoning
- First reaction — Cleavage of anisole with HI Anisole (C₆H₅OCH₃) reacts with excess HI. The mechanism: protonation of the ether oxygen, then nucleophilic attack by I⁻ at the methyl carbon (since the methyl–O bond is weaker than the aryl–O bond). This yields iodobenzene (C₆H₅I) as X and methanol (CH₃OH) as Y.
C6H5OCH3+HI→C6H5I+CH3OH
- Second reaction — Reduction of iodobenzene X (iodobenzene) is treated with Zn dust and heated (Δ). Zinc reduces the C–I bond, replacing iodine with hydrogen. The product is benzene (C₆H₆).
C6H5I+ZnΔC6H6+ZnI2
- Third reaction — Friedel–Crafts alkylation of benzene Y is methanol (CH₃OH). In the presence of anhydrous AlCl₃ (a Lewis acid), methanol forms a complex that generates a methyl carbocation (CH₃⁺). This electrophile attacks benzene, yielding toluene (methylbenzene, C₆H₅CH₃).
CH3OH+C6H6Anhy. AlCl3C6H5CH3+H2O
The product Z is therefore toluene.
- Matching with options Option (A) shows a benzene ring with a –CH₃ substituent — that is toluene. Option (B) has –CH₂Cl, (C) has –C₂H₅, (D) has –Cl. Only (A) matches.
Watch outA common mistake is to think that HI cleaves the aryl–O bond, giving phenol and methyl iodide. But HI is a strong acid and I⁻ is a good nucleophile; it attacks the less hindered methyl carbon, not the aromatic ring. So the products are iodobenzene and methanol, not phenol.
TipRemember: For aryl alkyl ethers, HI always cleaves the alkyl–O bond, not the aryl–O bond. This is a classic “hard-soft” acid-base principle — the soft I⁻ prefers the softer alkyl carbon.
✓Final answerThe correct option is (A).
ANSWER: A
- TG EAPCET 2025Set eng-2025-05-04-AN1 markMCQQ.What are X and Y respectively in the following reactions? (A) CH3Cl, CH3COCl (B) C2H5Cl, CH3COCl (C) CH3COCl, CH3Cl (D) C2H5COCl, CH3Cl
›Reveal solutionSolution
The first step is a Friedel–Crafts alkylation (X=CH3Cl) and the second a Friedel–Crafts acylation (Y=CH3COCl). Correct option: (A).
Both steps are carried out on the aromatic ring with anhydrous AlCl3, so each reagent is a Friedel–Crafts electrophile.
- X (alkylation). An alkyl chloride substitutes an alkyl group onto the ring:
C6H6+CH3ClAlCl3C6H5CH3+HCl.
So X=CH3Cl.
- Y (acylation). An acyl chloride introduces a –COCH3 group:
ArH+CH3COClAlCl3Ar–COCH3+HCl.
So Y=CH3COCl.
Alkylation activates the ring and must precede acylation (an acyl group deactivates the ring), which fixes the order X=CH3Cl, Y=CH3COCl.
✓Final answer(A) X=CH3Cl, Y=CH3COCl.
- TG EAPCET 2025Set eng-2025-05-04-FN1 markMCQQ.In which of the following reactions, hydrogen is evolved? I. Reaction of sodium borohydride with iodine II. Oxidation of diborane III. Reaction of boron trifluoride with sodium hydride IV. Hydrolysis of diborane (A) I, II only (B) I, II, IV only (C) III, IV only (D) I, IV only
›Reveal solutionSolution
Hydrogen gas is evolved when diborane is hydrolysed and when sodium borohydride reacts with iodine — the key is to check whether the reaction produces free H2 or only hydrogen-containing byproducts. The correct set is I and IV only.
The question tests your understanding of the chemistry of boron hydrides and related hydride-transfer reactions. Hydrogen evolution means free H2 gas is released, not just that hydrogen atoms appear in a product. Many boron compounds are electron-deficient and react with protic sources or oxidising agents to liberate hydrogen, but not every reaction that involves a hydride does so.
Let’s examine each reaction carefully.
- Reaction of sodium borohydride with iodine Sodium borohydride (NaBH4) is a source of hydride ions. Iodine (I2) is a mild oxidising agent. They react to give diborane and hydrogen gas:
2NaBH4+I2→B2H6+2NaI+H2
Hydrogen is clearly evolved here. So I is correct.
- Oxidation of diborane Diborane (B2H6) burns in oxygen to form boric oxide and water:
B2H6+3O2→B2O3+3H2O
No free hydrogen is produced — the hydrogen ends up in water. So II is incorrect.
- Reaction of boron trifluoride with sodium hydride Sodium hydride (NaH) is a strong hydride donor. With BF3, it forms diborane and sodium fluoride:
2BF3+6NaH→B2H6+6NaF
All the hydrogen from NaH goes into diborane; no H2 gas is released. So III is incorrect.
- Hydrolysis of diborane Diborane reacts vigorously with water to give boric acid and hydrogen gas:
B2H6+6H2O→2H3BO3+6H2
This is a classic reaction — the electron-deficient B–H bonds are attacked by water, and each B–H unit ultimately yields one H2 molecule. So IV is correct.
Watch outA common mistake is to think that any reaction producing a boron hydride (like diborane) must evolve hydrogen. But in reaction III, the hydride from NaH is completely consumed to form B2H6 — no free H2 appears. Always check the balanced equation for H2 as a product.
TipFor boron hydride reactions, remember the pattern: if a hydride source meets a protic source (like water, alcohol, or an acid), H2 is usually evolved. If two hydride sources react (like NaH and BF3), they combine to form a higher borane without releasing H2.
Only reactions I and IV produce hydrogen gas.
✓Final answerThe correct option is (D) I, IV only.
🎓Unlock everything free for 14 days
- ✓Full step-by-step solutions
- ✓Concept-first explanations
- ✓Methods, shortcuts & mistakes
- ✓PYQ mapping + timed mock tests
Full access for 14 days. No credit card required.