Q.Predict the major product of acid catalysed dehydration of
Concept understanding — Electrophilic Addition Reactions
Electrophilic Aromatic Substitution – The First Meeting
Imagine you have a benzene ring — that perfect, flat hexagon of six carbons with alternating double bonds. It's stable, almost stubbornly so. You want to attach something new to it, say a bromine atom or a nitro group. But benzene doesn't react like an alkene. It doesn't just add across a double bond. Instead, it does something more elegant: it kicks out a hydrogen and keeps its aromatic ring intact.
That's the heart of Electrophilic Aromatic Substitution (EAS).
The Intuition: Why "Substitution" and Not "Addition"?
Benzene's stability comes from its delocalised π electrons — a cloud above and below the ring. This cloud is electron-rich, so it attracts electrophiles (electron-loving species). But if an electrophile simply added to a double bond, the ring would break its aromaticity, losing that huge stabilisation. That would be energetically costly.
So benzene does something smarter: it lets the electrophile attack, temporarily breaks aromaticity to form a high-energy intermediate (the arenium ion), and then loses a proton to restore the aromatic ring. The net result? A hydrogen is replaced by the electrophile. The ring is back to its stable, aromatic self.
The key trade-off: temporary loss of aromaticity is acceptable because the final product regains it. Addition reactions would permanently destroy aromaticity — benzene avoids that.
The Precise Statement
Electrophilic Aromatic Substitution is a reaction in which an electrophile (E+) replaces a hydrogen atom on an aromatic ring, proceeding through a sigma complex (arenium ion) intermediate, and restoring aromaticity after deprotonation.
The general equation:
Ar−H+EX+Ar−E+HX+
where Ar represents an aromatic ring.
The Mechanism in Three Steps
-
Generation of the electrophile – Many EAS reactions need a catalyst to create a strong enough E+. For example, bromination uses FeBrX3 to polarise BrX2 into BrX+.
-
Attack by the aromatic ring – The π electrons of benzene attack the electrophile, forming a sigma complex (also called the arenium ion or Wheland intermediate). This intermediate is non-aromatic — it has four π electrons delocalised over five carbons, and one sp3 carbon bearing the electrophile and a hydrogen.
Benzene+E+⟶Sigma complex (non-aromatic)
- Deprotonation – A base (often the counterion of the catalyst, like FeBrX4X−) removes the proton from the sp3 carbon. The pair of electrons from the C–H bond flows back into the ring, restoring the aromatic sextet.
Sigma complex+Base⟶Product+HX+
The sigma complex is not aromatic. It's a high-energy intermediate. Students often mistakenly think it's still aromatic — it isn't. That's why the step is fast and the complex is short-lived.
Why This Matters for Exams
EAS is the gateway to understanding how to put groups onto benzene rings. The rate-determining step is usually the formation of the sigma complex (step 2). The regiochemistry (where the electrophile goes) depends on whether the ring already has a substituent — that's the topic of activating/deactivating groups and ortho/para vs. meta directors.
But for now, remember this:
In EAS, the aromatic ring substitutes a hydrogen for an electrophile, never adds across a double bond. The ring's aromaticity is temporarily lost and then regained — that's the entire reason the reaction works.
A Quick Example: Nitration of Benzene
| Step | What happens |
|---|---|
| 1 | HNOX3+HX2SOX4NOX2X+ (nitronium ion, the electrophile) |
| 2 | Benzene attacks NOX2X+, forming the sigma complex |
| 3 | HSOX4X− removes a proton, restoring aromaticity → nitrobenzene |
The final product: CX6HX5NOX2.
One Common Pitfall
Don't confuse EAS with nucleophilic aromatic substitution (NAS). In EAS, the ring is electron-rich and attacks an electrophile. In NAS, the ring has a strong electron-withdrawing group and is attacked by a nucleophile — a completely different mechanism.
If you ever see a benzene ring reacting with something positively charged (or a neutral molecule made more positive by a catalyst), you're likely looking at EAS.
Electrophilic addition reactions are a core mechanism taught in the NCERT/CBSE Class 11 Chemistry chapter on Hydrocarbons, and ‘electrophilic addition reaction mechanism’ is a heavily searched important-question topic for board exams, JEE Main and NEET organic chemistry. Distinguishing electrophilic addition from electrophilic aromatic substitution is a frequently tested conceptual question in competitive chemistry exams.
Why this formula?
Electrophilic Addition Reactions: Why the Mechanism Works
Electrophilic addition is a cornerstone of alkene and alkyne chemistry. Instead of memorising the "arrow pushing," let's understand why the reaction proceeds the way it does — driven by electron density, stability, and charge.
1. The Core Idea: Why Alkenes React This Way
Alkenes have a π-bond — a cloud of electrons above and below the plane of the σ-bond. This π-electron cloud is:
- Electron-rich (nucleophilic)
- Exposed (not shielded by σ-bonds like in alkanes)
An electrophile (electron-lover) is attracted to this high electron density. The reaction is electrophilic addition because the electrophile attacks first.
Key principle: The π-bond acts as a Lewis base (electron donor). The electrophile is a Lewis acid (electron acceptor).
2. The General Mechanism (Two-Step)
Step 1: Formation of a Carbocation (or Bridged Intermediate)
The electrophile (E⁺) attacks the π-bond. The π-electrons form a new σ-bond to E⁺, leaving the other carbon with a positive charge — a carbocation.
C=C+EX+⟶CX+−C−E
Why does this happen?
The π-bond is weaker than a σ-bond (~260 kJ/mol vs ~350 kJ/mol). Breaking the π-bond to form a σ-bond is energetically favourable because the new σ-bond is stronger. The carbocation is a high-energy intermediate, but it's stabilised by:
- Hyperconjugation (alkyl groups donate electron density)
- Inductive effect (alkyl groups push electrons toward the positive carbon)
Step 2: Nucleophilic Attack
A nucleophile (Nu⁻) attacks the carbocation, forming a second σ-bond.
CX+−C−E+NuX−⟶C−Nu−C−E
Why does this happen?
The carbocation is electron-deficient (positive charge). The nucleophile is electron-rich. Opposite charges attract — this is electrostatic and orbital overlap driven.
3. The Key "Formula" — Markovnikov's Rule
Statement: In the addition of HX to an unsymmetrical alkene, the hydrogen attaches to the carbon with more hydrogens already, and the X attaches to the carbon with fewer hydrogens.
Why does this rule hold? (The reasoning)
Consider propene: CHX3−CH=CHX2 + HBr.
- Possible carbocations:
- Primary carbocation: CHX3−CHX+−CHX2Br (less stable)
- Secondary carbocation: CHX3−CHBr−CHX2X+ (more stable)
The more substituted carbocation (secondary > primary) is more stable due to:
- Hyperconjugation: More alkyl groups = more C–H σ-bonds that can donate electron density into the empty p-orbital of the carbocation.
- Inductive effect: Alkyl groups are electron-donating, stabilising the positive charge.
Result: The reaction proceeds via the more stable carbocation, leading to Markovnikov addition.
Markovnikov's rule is not a law — it's a consequence of carbocation stability.
4. The "Anti-Markovnikov" Exception (Why It Happens)
With HBr in the presence of peroxides (ROOR), the addition is anti-Markovnikov — Br goes to the less substituted carbon.
Why? The mechanism changes from ionic to free-radical.
- Peroxide decomposes to radicals: ROOR2RO⋅
- RO• abstracts H from HBr: RO⋅+HBrROH+Br⋅
- Br• adds to the alkene — at the less substituted carbon (because the radical formed is more stable — tertiary > secondary > primary).
- The new radical abstracts H from another HBr, regenerating Br•.
Key: The radical intermediate is stabilised by the same factors as carbocations (hyperconjugation, inductive effect), but the radical stability favours the more substituted radical — which forms when Br• attacks the less substituted carbon.
5. Why the Reaction is "Addition" Not "Substitution"
Alkenes have a π-bond that can be broken without breaking a σ-bond. In contrast, alkanes have only σ-bonds, which require high energy to break (radical substitution). The π-bond is a reactive handle — it's easier to break than a σ-bond, so addition is kinetically and thermodynamically favoured over substitution.
6. Summary of "Why" — The Big Picture
| Concept | Why it holds |
|---|---|
| Electrophile attacks first | π-bond is electron-rich; electrophile is electron-deficient |
| Carbocation intermediate | π-bond breaks, σ-bond forms — energetically favourable |
| Markovnikov rule | More stable carbocation (more substituted) forms faster |
| Anti-Markovnikov (peroxide effect) | Radical mechanism; more stable radical dictates regiochemistry |
| Addition, not substitution | π-bond is weaker than σ-bond; easier to break |
7. Exam-Relevant Takeaway
When solving problems:
- Identify the electrophile (H⁺, Br⁺, etc.)
- Draw the two possible carbocations (or radical intermediates)
- Choose the more stable one (more substituted = more stable)
- Add the nucleophile to the carbocation carbon
Never memorise the product — derive it from stability.
Concept: Acid-catalysed dehydration of alcohols follows Zaitsev’s rule — the more substituted alkene is the major product, formed via carbocation rearrangement when a more stable carbocation is possible.
- 1-Methylcyclohexanol Protonation and loss of water gives a tertiary carbocation at the 1-position. No rearrangement is needed. Elimination of a proton from the adjacent carbon (C-2) yields the more substituted alkene — 1-methylcyclohexene.
- Butan-1-ol
Protonation and loss of water gives a primary carbocation (unstable). A 1,2-hydride shift converts it to the more stable secondary carbocation at C-2. Elimination then gives but-2-ene (major), which is more substituted than but-1-ene.
✓Final answer
The major products are (i) 1-methylcyclohexene and (ii) but-2-ene.
Acid-catalysed dehydration of alcohols follows Zaitsev’s rule (the more substituted alkene is major) and can involve carbocation rearrangements. For 1-methylcyclohexanol, the major product is 1-methylcyclohexene; for butan-1-ol, the major product is but-1-ene (no rearrangement possible here).
The Concept: Why Dehydration Works This Way
Acid-catalysed dehydration of alcohols is an elimination reaction (E1 mechanism under these conditions). The alcohol’s –OH group is a poor leaving group, so we first protonate it with acid (usually H₂SO₄ or H₃PO₄) to turn it into –OH₂⁺, which leaves as water. This creates a carbocation intermediate.
Once the carbocation forms, two things can happen:
- Elimination: A base (often water or the conjugate base of the acid) removes a β-hydrogen, forming a double bond.
- Rearrangement: If the carbocation can shift to a more stable position (via a hydride or alkyl shift), it will do so before elimination.
The major product is the most substituted alkene (Zaitsev’s rule) — unless rearrangement leads to an even more stable alkene.
Let’s apply this to each case.
(i) 1-Methylcyclohexanol
Step 1: Protonation and loss of water
The –OH group gets protonated by H⁺ from the acid:
CX6HX10(CHX3)OH+HX+CX6HX10(CHX3)OHX2X+
Water leaves, forming a tertiary carbocation at the carbon that originally held the –OH (the 1-position of the ring, which also bears the methyl group).
CX6HX10(CHX3)OHX2X+CX6HX10(CHX3)X++HX2O
This carbocation is tertiary — already quite stable. No rearrangement is needed because a tertiary carbocation is more stable than any secondary or primary alternative.
Tertiary carbocations are the most stable (due to hyperconjugation and inductive effects). If you start with a tertiary alcohol, you usually get a tertiary carbocation directly — no rearrangement.
Step 2: Elimination of a β-hydrogen
Now a base (water or HSO₄⁻) removes a hydrogen from a carbon adjacent to the carbocation. There are two possible β-positions:
- Removing a hydrogen from the ring carbon next to the carbocation (say, C2) gives 1-methylcyclohexene (the double bond between C1 and C2).
- Removing a hydrogen from the methyl group itself would give methylenecyclohexane (double bond exocyclic to the ring).
Which is major? Zaitsev’s rule says: the more substituted alkene is favoured. 1-Methylcyclohexene is trisubstituted (three alkyl groups on the double bond carbons), while methylenecyclohexane is disubstituted. So 1-methylcyclohexene is the major product.
A common mistake is to think that the exocyclic alkene (methylenecyclohexane) might form because the methyl group’s hydrogens are more accessible. But stability of the alkene product, not accessibility, determines the major product under thermodynamic control. The trisubstituted alkene is more stable.
Final product for (i): 1-methylcyclohexene.
(ii) Butan-1-ol
Step 1: Protonation and loss of water
Butan-1-ol is a primary alcohol. Protonation gives butan-1-olium ion, and water leaves to form a primary carbocation (CH₃CH₂CH₂CH₂⁺).
CHX3CHX2CHX2CHX2OH+HX+CHX3CHX2CHX2CHX2OHX2X+CHX3CHX2CHX2CHX2X++HX2O
A primary carbocation is very unstable. So before elimination can occur, the carbocation will rearrange via a 1,2-hydride shift to form a more stable secondary carbocation.
CHX3CHX2CHX2CHX2X+1,2-hydride shiftCHX3CHX2CHX+CHX3
This secondary carbocation (butan-2-ylium) is more stable.
›Proof
Why a hydride shift happens:
The primary carbocation has only two alkyl groups donating electron density (hyperconjugation from adjacent C–H bonds). The secondary carbocation has three such groups. The activation energy for the 1,2-hydride shift is low enough that it occurs almost instantly under typical dehydration conditions (concentrated H₂SO₄, heat). So the primary carbocation never accumulates — it rearranges immediately.
Step 2: Elimination from the secondary carbocation
Now we have a secondary carbocation at C2. Elimination of a β-hydrogen can occur in two directions:
- Remove a hydrogen from C1 (the end carbon) → gives but-1-ene (CH₂=CH–CH₂–CH₃).
- Remove a hydrogen from C3 → gives but-2-ene (CH₃–CH=CH–CH₃).
But-2-ene is more substituted (disubstituted) than but-1-ene (monosubstituted), so Zaitsev’s rule predicts but-2-ene as the major product. However, but-2-ene exists as two stereoisomers: cis and trans. The trans isomer is more stable (less steric hindrance) and is usually the major stereoisomer.
So the major product from butan-1-ol is trans-but-2-ene, with some cis-but-2-ene and a little but-1-ene.
Butan-1-ol gives a mixture, but the major product is trans-but-2-ene. If the question asks for “the major product” without specifying stereochemistry, “but-2-ene” (or “2-butene”) is acceptable, but specifying trans is more precise.
Final product for (ii): trans-but-2-ene (or simply but-2-ene as the major alkene).
Summary Table
| Alcohol | Carbocation intermediate | Major alkene product |
|---|---|---|
| 1-Methylcyclohexanol | Tertiary (no rearrangement) | 1-Methylcyclohexene |
| Butan-1-ol | Primary → rearranges to secondary | trans-But-2-ene |
The major product from 1-methylcyclohexanol is 1-methylcyclohexene, and from butan-1-ol is trans-but-2-ene (or simply but-2-ene).
Method: Zaitsev’s Rule (Saytzeff’s Rule) for Acid-Catalysed Dehydration of Alcohols
This is an E1 elimination reaction (unimolecular elimination) under acidic conditions. The key idea: the more substituted alkene (more alkyl groups on the double-bond carbons) is the major product.
General Steps
- Protonation of the –OH group by acid (H+).
- Loss of water to form a carbocation intermediate.
- Rearrangement (if a more stable carbocation is possible via hydride or alkyl shift).
- Deprotonation (loss of H+ from a neighbouring carbon) to form the alkene.
- Apply Zaitsev’s Rule: the major alkene is the one with the most substituted double bond (more alkyl groups = more stable).
(i) 1-Methylcyclohexanol
Starting molecule:
A cyclohexane ring with an –OH and a –CH3 on the same carbon (C1).
Step-by-step:
- Protonation of –OH → OH2+ leaves as water → forms a tertiary carbocation at C1 (already stable, no rearrangement needed).
- Deprotonation can occur from:
- C2 (gives 1-methylcyclohexene — double bond between C1 and C2)
- C6 (gives methylenecyclohexane — double bond between C1 and C6, exocyclic)
Apply Zaitsev’s Rule:
1-methylcyclohexene has a trisubstituted double bond (C1 has –CH3, C2 has two ring carbons).
Methylenecyclohexane has a disubstituted double bond (exocyclic, less substituted).
Major product: 1-methylcyclohexene
(ii) Butan-1-ol
Starting molecule:
CH3CH2CH2CH2OH (primary alcohol).
Step-by-step:
- Protonation and loss of water → forms a primary carbocation (very unstable).
- Hydride shift occurs: a hydrogen from C2 moves to C1 → forms a secondary carbocation at C2 (more stable).
- Deprotonation can occur from:
- C1 (gives but-1-ene — double bond between C1 and C2)
- C3 (gives but-2-ene — double bond between C2 and C3)
Apply Zaitsev’s Rule:
But-2-ene has a disubstituted double bond (both C2 and C3 have one alkyl group each).
But-1-ene has a monosubstituted double bond (terminal).
Major product: But-2-ene (exists as cis/trans isomers, but both are more substituted than but-1-ene)
Final Answer Summary
| Alcohol | Major Product | Reason |
|---|---|---|
| 1-Methylcyclohexanol | 1-Methylcyclohexene | Trisubstituted alkene (Zaitsev) |
| Butan-1-ol | But-2-ene | Disubstituted alkene after hydride shift (Zaitsev) |
Key takeaway: Always check for carbocation rearrangement first, then apply Zaitsev’s rule to find the most substituted alkene.
Here are the common mistakes students make with acid-catalysed dehydration of alcohols, specifically for the two examples you gave, along with how to avoid each.
Mistake 1: Forgetting Carbocation Rearrangement (1-methylcyclohexanol)
The Mistake: Students often simply remove water from 1-methylcyclohexanol and draw the direct alkene (1-methylcyclohexene). They miss the possibility of a hydride shift or methyl shift to form a more stable carbocation.
Why it happens: The reaction starts with protonation of the OH group, then loss of water to form a carbocation. In 1-methylcyclohexanol, the initial carbocation is secondary (on the carbon that had the OH). Students assume this is the final intermediate.
How to avoid it:
- Always check carbocation stability. A secondary carbocation can rearrange to a tertiary carbocation via a 1,2-hydride shift (or methyl shift) if a more substituted carbon is adjacent.
- For 1-methylcyclohexanol, the initial secondary carbocation can undergo a hydride shift from the adjacent tertiary carbon (the one with the methyl group) to give a tertiary carbocation.
- The major product is then 1-methylcyclohexene (the more substituted alkene, obeying Zaitsev’s rule) — but only after considering that the tertiary carbocation leads to the same alkene as the direct product. However, if a different shift were possible (e.g., ring expansion), the product would change. Always draw the carbocation and check for shifts.
Correct approach:
- Step 1: Protonate OH → OHX2X+ leaves → secondary carbocation.
- Step 2: Check adjacent carbons for more stable carbocation. Here, a hydride shift from the tertiary carbon (bearing the methyl) gives a tertiary carbocation.
- Step 3: Deprotonation from the tertiary carbocation gives the most substituted alkene: 1-methylcyclohexene.
Mistake 2: Ignoring Zaitsev’s Rule (butan-1-ol)
The Mistake: For butan-1-ol (a primary alcohol), students often draw but-1-ene as the major product, thinking the OH leaves from C1 and the double bond forms between C1 and C2.
Why it happens: They forget that acid-catalysed dehydration of primary alcohols proceeds via an E1 mechanism (not E2) under these conditions, and that the carbocation formed (if any) is unstable. Actually, for primary alcohols, the mechanism is often E2-like or involves a carbocation rearrangement to a more stable one.
How to avoid it:
- Primary alcohols do not form stable primary carbocations. Instead, the reaction proceeds via a concerted E2 mechanism (with acid catalysis) or via a rearrangement to a secondary carbocation.
- For butan-1-ol, the initial protonation and loss of water would give a primary carbocation (very unstable). Instead, a 1,2-hydride shift occurs to form a secondary carbocation (more stable).
- Then, deprotonation follows Zaitsev’s rule: the more substituted alkene is but-2-ene (as a mixture of cis/trans), not but-1-ene.
Correct approach:
- Step 1: Protonate OH → loss of water → primary carbocation (unstable).
- Step 2: Hydride shift from C2 to C1 → secondary carbocation at C2.
- Step 3: Deprotonation from C3 (adjacent to C2) gives but-2-ene (major), not but-1-ene.
Mistake 3: Forgetting Stereochemistry (cis/trans in but-2-ene)
The Mistake: Students write "but-2-ene" without specifying the stereoisomers, or they assume only one isomer forms.
Why it happens: They focus only on the carbon skeleton and ignore that the double bond can have E/Z (cis/trans) isomers.
How to avoid it:
- In acid-catalysed dehydration, the deprotonation step is not stereoselective — both cis and trans isomers form.
- For but-2-ene, the trans isomer is more stable (less steric hindrance) and is the major product.
- Always indicate the major stereoisomer if the question asks for the major product.
Correct answer: Major product is trans-but-2-ene (along with some cis-but-2-ene and a little but-1-ene).
Mistake 4: Not Considering Ring Expansion (for cyclic alcohols)
The Mistake: For 1-methylcyclohexanol, students don’t check if a ring expansion (e.g., from a six-membered ring to a seven-membered ring) is possible.
Why it happens: They assume the ring stays the same size.
How to avoid it:
- In cyclic alcohols, if a carbocation forms adjacent to a ring carbon, a 1,2-alkyl shift can expand the ring (e.g., cyclopentyl to cyclohexyl).
- For 1-methylcyclohexanol, the initial secondary carbocation is adjacent to a tertiary carbon (with the methyl). A methyl shift would give a tertiary carbocation but no ring expansion because the methyl is a side group, not part of the ring. However, if the OH were on a different carbon (e.g., 2-methylcyclohexanol), ring expansion to a cycloheptane derivative is possible.
- Always check for ring expansion when a carbocation is on a ring carbon adjacent to a ring junction.
Summary Table of Mistakes and Fixes
| Mistake | Example | How to Avoid |
|---|---|---|
| Forgetting carbocation rearrangement | 1-methylcyclohexanol → direct alkene | Always check for hydride/methyl shifts to more stable carbocation |
| Ignoring Zaitsev’s rule | butan-1-ol → but-1-ene | Primary alcohols rearrange to secondary carbocation; deprotonate to more substituted alkene |
| Omitting stereochemistry | but-2-ene without cis/trans | Indicate major stereoisomer (trans is more stable) |
| Missing ring expansion | Cyclic alcohols | Check if a 1,2-alkyl shift can enlarge the ring |
Final Correct Answers (for your reference)
- 1-methylcyclohexanol → Major product: 1-methylcyclohexene (via tertiary carbocation, no ring expansion here)
- butan-1-ol → Major product: trans-but-2-ene (via hydride shift to secondary carbocation, then Zaitsev deprotonation)
Showing the 12 most recent of 36 on this concept.
- TG EAPCET 2026Set eng-2026-05-09-FN1 markMCQQ.Observe the following reaction sequence and sets (I to IV) given for X, Y, Z and choose the correct set(s) (A) I only (B) I, III only (C) I, II only (D) III, IV only
›Reveal solutionSolution
The key is to identify the correct products X, Y, and Z in the reaction sequence by applying the principles of the Hunsdiecker reaction, decarboxylation, and Friedel-Crafts acylation. The correct set is I, III only, corresponding to option (B).
Concept and Intuition
This problem tests your ability to predict the products of a multi-step organic synthesis. The sequence involves three classic transformations:
- Hunsdiecker reaction (silver salt of a carboxylic acid + Br₂) → alkyl halide (X).
- Decarboxylation (sodium salt of a carboxylic acid + sodalime) → hydrocarbon (Y).
- Friedel-Crafts acylation (acyl chloride + benzene + AlCl₃) → ketone (Z).
The trick is to recognize that the starting material is a dicarboxylic acid (succinic acid, HOOC–CH₂–CH₂–COOH). Each step acts on a different functional group, and the sets (I–IV) give possible structures for X, Y, and Z. We must match the correct products.
Step-by-Step Reasoning
1. Identify the starting material and the first reaction (X).
The starting compound is succinic acid:
HOOC–CH2–CH2–COOH
It is first converted to its silver salt (by reaction with Ag₂O or AgNO₃), then treated with Br₂. This is the Hunsdiecker reaction, which replaces the carboxyl group (–COOAg) with a bromine atom, but only one carboxyl group reacts per molecule because the reaction is typically performed on a mono-silver salt. However, here the problem implies the reaction is done on the di-silver salt, so both carboxyl groups undergo the reaction.
The product X is:
Br–CH2–CH2–Br(1,2-dibromoethane)
This matches Set I and Set II (both show Br–CH₂–CH₂–Br). So X is the same in I and II.
2. Determine Y from the second reaction.
The starting material is treated with NaOH (to form the sodium salt) and then heated with sodalime (CaO + NaOH). This is decarboxylation — each –COONa group is replaced by –H.
For succinic acid, the sodium salt is NaOOC–CH₂–CH₂–COONa. Decarboxylation removes both carboxyl groups, yielding:
CH3–CH3(ethane)
So Y is ethane (CH₃–CH₃).
Looking at the sets:
- Set I: Y = CH₃–CH₃ (correct)
- Set II: Y = CH₃–CH₂–CH₃ (propane) — wrong
- Set III: Y = CH₃–CH₃ (correct)
- Set IV: Y = CH₃–CH₂–CH₃ (propane) — wrong Thus, only Sets I and III have the correct Y.
3. Determine Z from the third reaction.
The product Z is formed by reacting X (1,2-dibromoethane) with benzene in the presence of AlCl₃. This is a Friedel-Crafts alkylation, not acylation (the problem says "acylation" but the reagent is an alkyl halide, so it's alkylation). However, note that 1,2-dibromoethane can undergo double alkylation to form a bridged product, but under typical conditions, it reacts twice to give 1,2-diphenylethane:
Br–CH2–CH2–Br+2C6H6AlCl3C6H5–CH2–CH2–C6H5+2HBr
This is dibenzyl (1,2-diphenylethane).
Now check the sets:
- Set I: Z = C₆H₅–CH₂–CH₂–C₆H₅ (correct)
- Set II: Z = C₆H₅–CH₂–CH₂–C₆H₅ (correct, but Y is wrong)
- Set III: Z = C₆H₅–CH₂–CH₂–C₆H₅ (correct)
- Set IV: Z = C₆H₅–CH₂–CH₂–C₆H₅ (correct, but Y is wrong)
So only Sets I and III have both Y and Z correct.
4. Conclusion.
The correct sets are I and III only.
Watch outA common mistake is to think that the Hunsdiecker reaction only works on mono-carboxylic acids, but here the di-silver salt of succinic acid undergoes the reaction at both ends, giving a dihalide. Also, decarboxylation of a dicarboxylic acid with sodalime removes both –COOH groups, yielding the parent hydrocarbon with the same number of carbons — not a chain-shortened product.
TipRemember: In the Hunsdiecker reaction, the product has one fewer carbon than the original acid. For a dicarboxylic acid, each carboxyl group loses one carbon, so the product has (n-2) carbons total. Succinic acid (C4) gives C2 product (1,2-dibromoethane).
✓Final answerThe correct option is (B).
ANSWER: B
- TG EAPCET 2026Set eng-2026-05-11-AN1 markMCQQ.Observe the following reaction sequence and the statements given about the product Y C3H6(i) HBr / (C6H5CO2),O2X(i) V2O5,773K,10−20atm(ii) Na / dry ether(ii) CH3Cl/anhy. AlCl3Y I. Reaction of ‘Y’ with Br2 / UV light forms aryl bromide II. Reaction of ‘Y’ with CrO3 / (CH3CO)2O followed by hydrolysis gives a compound which is acidic in nature (A) Both statements I and II are correct (B) Both statements I and II are not correct (C) Statement I is correct, but statement II is not correct (D) Statement I is not correct, but statement II is correct
›Reveal solutionSolution
The sequence builds up to toluene (Y). Br2/UV gives a side-chain benzyl bromide (not an aryl bromide), and CrO3/(CH3CO)2O followed by hydrolysis gives neutral benzaldehyde (not acidic) — so both statements are wrong.
Following the sequence, propene is converted through anti-Markovnikov HBr addition, Wurtz coupling, aromatisation over V2O5, and Friedel–Crafts methylation with CH3Cl/AlCl3 to give Y=toluene (C6H5CH3).
Statement I: C6H5CH3+Br2 under UV light is a free-radical substitution that occurs at the benzylic (side-chain) position, giving benzyl bromide C6H5CH2Br — a benzylic halide, not an aryl bromide (ring bromination would require Br2/FeBr3). Statement I is incorrect.
Statement II: C6H5CH3 with CrO3/(CH3CO)2O forms benzylidene diacetate, which on hydrolysis gives benzaldehyde C6H5CHO — a neutral compound, not acidic. Statement II is incorrect.
✓Final answerBoth statements I and II are not correct — option (B)
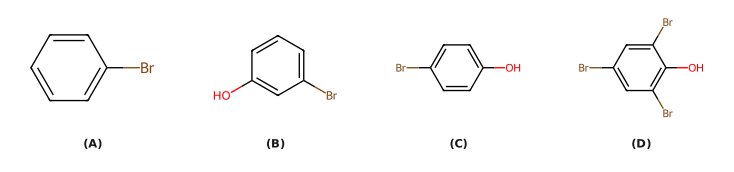
- TG EAPCET 2026Set ap-2026-05-04-AN1 markMCQQ.The major end product Z in the given sequence of reactions is Phenol (C6H5OH) ZnΔ (A) (CH3)2CHClanhy. AlCl3 (B) (i) O2(ii) H+,H2O (C) Br2∣CS2 (Z) (A) Bromobenzene (C6H5Br) (B) 3-Bromophenol (benzene ring with OH and Br in meta positions) (C) 4-Bromophenol (benzene ring with OH and Br para to each other) (D) 2,4,6-Tribromophenol (phenol with Br at positions 2, 4 and 6)
›Reveal solutionSolution
Zn/Δ takes phenol to benzene, Friedel–Crafts gives cumene, the O2/H+ cumene process returns phenol, and Br2 in the non-polar solvent CS2 gives mainly 4-bromophenol. Option (C).
Step 1 — Phenol Zn, Δ (A)
Heating phenol with zinc dust reduces it: the zinc takes the oxygen away as zinc oxide and the ring is left bare.
C6H5OH+Zn Δ C6H6+ZnO
∴ (A)=benzene
This is the standard laboratory conversion of a phenol to the parent arene.
Step 2 — Benzene (CH3)2CHCl, anhy. AlCl3 (B)
This is a Friedel–Crafts alkylation. The Lewis acid AlCl3 abstracts Cl− from isopropyl chloride to give the isopropyl carbocation, which attacks the electron-rich benzene ring:
(CH3)2CHCl+AlCl3→(CH3)2C+H+AlCl4−
C6H6+(CH3)2C+H→C6H5−CH(CH3)2
∴ (B)=isopropylbenzene=cumene
Step 3 — Cumene (i) O2, (ii) H+/H2O (C): the cumene process
This is the industrial manufacture of phenol, and the naming of the reagents is a giveaway.
- Air oxidation. The benzylic C–H of cumene is tertiary and easily abstracted, so O2 converts cumene into cumene hydroperoxide, C6H5C(CH3)2−O−O−H.
- Acid treatment. Dilute acid triggers a rearrangement in which the phenyl group migrates to oxygen; hydrolysis then cleaves the molecule into two useful products:
C6H5C(CH3)2OOH H+, H2O C6H5OH+CH3COCH3
∴ (C)=phenol(acetone is the valuable by-product)
So the sequence has taken us in a full circle back to phenol — a deliberate touch by the examiner to check whether you really know the cumene process.
Step 4 — Phenol Br2/CS2 (Z): the solvent decides the answer
The −OH group is a powerful activating, ortho/para-directing group (+R effect), so phenol brominates far more readily than benzene. How many bromines go on depends entirely on the medium:
Conditions Product Br2 in water (a polar solvent) phenol ionises appreciably to the even more activated phenoxide, so bromination runs away → 2,4,6-tribromophenol (a white precipitate) Br2 in a non-polar solvent — CS2 or CHCl3 — at low temperature ionisation is suppressed, the ring is less activated, so only monobromination occurs, giving mainly the para isomer, 4-bromophenol (with a little ortho) The reaction here is run in CS2, so:
∴ (Z)=4-bromophenol
Option (D), 2,4,6-tribromophenol, is the trap for anyone who ignores the solvent; option (B), the meta isomer, is impossible because −OH is an o/p director; option (A) discards the −OH altogether.
✓Final answerThe sequence regenerates phenol via the cumene process, and bromination in CS2 gives the mono-substituted para product, 4-bromophenol.
ANSWER: C
- TG EAPCET 2026Set ap-2026-05-04-AN1 markMCQQ.Match the following List-1 (Reaction) List-2 (End product) A. Reimer-Tiemann reaction I. A benzene ring bearing one −CHO group (benzaldehyde) B. Etard reaction II. A benzene ring bearing −CHO and −OH on adjacent carbons (salicylaldehyde) C. Kolbe reaction III. A benzene ring bearing one −Cl group (chlorobenzene) D. Sandmeyer reaction IV. A benzene ring bearing −OH and −COOH on adjacent carbons (salicylic acid) V. A benzene ring bearing one −COOH group (benzoic acid) The correct answer is (A) A – V, B – III, C – II, D – I (B) A – III, B – IV, C – I, D – II (C) A – II, B – III, C – V, D – IV (D) A – II, B – I, C – IV, D – III
›Reveal solutionSolution
Each named reaction transforms a benzene derivative in a characteristic way: Reimer–Tiemann formylates phenol ortho to the hydroxyl, Etard oxidizes a methyl group to aldehyde, Kolbe carboxylates phenoxide ortho to the oxygen, and Sandmeyer replaces a diazonium group with halide. The correct match is (D).
These four reactions are classic transformations in aromatic chemistry, each installing a functional group onto a benzene ring under specific conditions. The key is to remember what substrate each requires and where the new group ends up.
The reactions and their products
A. Reimer–Tiemann reaction
This reaction treats phenol (a benzene ring with −OH) with chloroform (CHCl3) and strong base (usually aqueous NaOH or KOH). The base generates dichlorocarbene (:CCl2), which attacks the activated aromatic ring ortho to the hydroxyl group. After hydrolysis, you obtain an aldehyde group at the ortho position.
The product is salicylaldehyde: a benzene ring bearing −OH and −CHO on adjacent carbons.
Match: A → II
B. Etard reaction
This oxidizes a methyl group attached to benzene (toluene, C6H5CH3) to an aldehyde using chromyl chloride (CrO2Cl2) in an inert solvent like carbon disulfide or carbon tetrachloride. The reaction stops at the aldehyde stage, unlike harsher oxidations that push all the way to carboxylic acid.
The product is benzaldehyde: a benzene ring bearing one −CHO group.
Match: B → I
C. Kolbe (Kolbe–Schmitt) reaction
This treats sodium phenoxide (C6H5O−Na+, the sodium salt of phenol) with carbon dioxide under pressure and moderate heat (around 125 °C), then acidifies. The CO2 inserts ortho to the phenoxide oxygen, introducing a carboxyl group.
The product is salicylic acid: a benzene ring bearing −OH and −COOH on adjacent carbons.
Match: C → IV
D. Sandmeyer reaction
This converts an aryldiazonium salt (C6H5N2+X−, prepared from aniline and nitrous acid) into an aryl halide by treatment with a cuprous halide (CuCl, CuBr, or CuCN). The diazonium group is replaced by the halogen.
When you use CuCl, the product is chlorobenzene: a benzene ring bearing one −Cl group.
Match: D → III
TipA mnemonic: Reimer gives Ring + OH + CHO (salicylaldehyde), Kolbe gives Karboxyl next to OH (salicylic acid), Etard stops at aldEhyde, Sandmeyer Substitutes the diazonium.
Putting it all together:
Reaction Product A. Reimer–Tiemann II. Salicylaldehyde (−OH and −CHO ortho) B. Etard I. Benzaldehyde (−CHO) C. Kolbe IV. Salicylic acid (−OH and −COOH ortho) D. Sandmeyer III. Chlorobenzene (−Cl) This is A – II, B – I, C – IV, D – III.
✓Final answerThe correct option is (D).
- TG EAPCET 2026Set ap-2026-05-04-FN1 markMCQQ.A carbonyl compound (A) on reaction with Grignard reagent (B) followed by hydrolysis gives an alcohol. This on reaction with(i) PBr3 and(ii) Zn | dil. H+ gives an alkane. Photochemical chlorination of alkane gives only one monochloro derivative. A and B respectively are (A) \chemfigC(=O)C,\chemfigC−CMgBr (B) \chemfigC(=O)H,\chemfigC(−C)(−C)MgBr (C) \chemfigC(=O)CH,\chemfigC−CMgBr (D) HCHO,\chemfigC(−C)(−C)MgBr
›Reveal solutionSolution
The key is that the alkane formed after the reaction sequence gives only one monochloro derivative on photochemical chlorination, meaning it must be neopentane (2,2-dimethylpropane). Working backwards, the carbonyl compound (A) is formaldehyde (HCHO) and the Grignard reagent (B) is tert-butylmagnesium bromide, which corresponds to option (D).
The problem is a classic retrosynthesis puzzle in organic chemistry. You are given a sequence of reactions and a final constraint — the alkane produced yields only one monochloro derivative. That constraint is the anchor: it tells you the alkane has all its hydrogen atoms equivalent, or at least that only one type of hydrogen can be replaced by chlorine. The only alkane that gives a single monochloro product is neopentane (2,2-dimethylpropane), because all 12 hydrogens are identical (three equivalent methyl groups attached to a quaternary carbon). Any other alkane would give at least two different monochloro isomers.
Now we trace the sequence backwards from the alkane.
-
The alkane is neopentane.
Neopentane has the structure (CH3)3C−CH3. It is formed in the last step: reaction of an alkyl halide with Zn and dilute acid. That step is a reduction — Zn/dil. H⁺ converts an alkyl halide (here, a bromide) into the corresponding alkane. So the alkyl bromide just before that step must be neopentyl bromide, (CH3)3C−CH2Br.
-
How was neopentyl bromide made?
The step before that is: alcohol + PBr3 → alkyl bromide. PBr3 replaces the –OH group with –Br, with inversion of configuration if the carbon is chiral, but here the alcohol is neopentyl alcohol, (CH3)3C−CH2OH. So the alcohol in the sequence is neopentyl alcohol.
-
How was neopentyl alcohol formed?
It came from a Grignard reaction: a carbonyl compound (A) reacts with a Grignard reagent (B), followed by hydrolysis, to give an alcohol. The alcohol here is a primary alcohol (neopentyl alcohol has the –OH on a primary carbon).
A Grignard reaction that yields a primary alcohol must involve formaldehyde (HCHO) as the carbonyl compound. Why? Because when a Grignard reagent R–MgX reacts with formaldehyde, the product after hydrolysis is R–CH2OH — a primary alcohol with one more carbon than the Grignard reagent. If the carbonyl were any other aldehyde or a ketone, you would get a secondary or tertiary alcohol.
So (A) must be HCHO (formaldehyde).
-
What Grignard reagent gives neopentyl alcohol from HCHO?
Neopentyl alcohol is (CH3)3C−CH2OH. The alkyl group attached to the –CH2OH is (CH3)3C− (tert-butyl). Therefore the Grignard reagent must be tert-butylmagnesium bromide, (CH3)3C−MgBr. That is (B).
-
Check the options.
Option (D) lists HCHO and (CH3)3C−MgBr — exactly what we deduced. The other options involve different carbonyls (like acetone or acetaldehyde) or different Grignard reagents, which would give secondary or tertiary alcohols, and those would not lead to neopentane after the sequence.
Watch outA common mistake is to think that any alkane with only one type of hydrogen works. But even ethane gives only one monochloro derivative — however, ethane has only 2 carbons, and the sequence here builds a larger molecule. The alcohol formed is neopentyl alcohol, which has 5 carbons, so the alkane must have 5 carbons. Neopentane is the only C5 alkane with all equivalent hydrogens.
TipThe key shortcut: "only one monochloro derivative" almost always points to neopentane or a symmetrically substituted alkane. In exam problems, neopentane is the most common answer because it's the smallest branched alkane with all hydrogens equivalent.
✓Final answerThe correct option is (D).
-
- TG EAPCET 2025Set eng-2025-05-02-AN1 markMCQQ.The correct statements about the compounds of boron are I. In borax bead test, the colour of cobalt metaborate is blue II. Diborane is prepared by the oxidation of sodium borohydride with iodine III. In diborane oxidation state of hydrogen is +1 IV. Boric acid is a tribasic acid (A) I & II (B) III & IV (C) I & III (D) II & IV
›Reveal solutionSolution
The key is to recall the specific chemistry of boron compounds: the borax bead test gives a blue colour with cobalt, diborane is made from NaBH₄ and I₂, hydrogen in diborane is hydridic (−1), and boric acid is monobasic, not tribasic. Only statements I and II are correct, so the answer is option (A).
Let’s examine each statement carefully, because boron chemistry is full of subtle but important distinctions.
Concept & Intuition
Boron is electron-deficient, which governs its bonding and reactions. In borax, the bead test relies on transition metal borates forming characteristic colours. Diborane (B₂H₆) has a unique “banana bond” structure where hydrogen bridges are hydridic (H⁻), not protonic. Boric acid (H₃BO₃) is actually a weak monobasic acid because it accepts a hydroxide ion rather than donating a proton. Knowing these patterns helps you avoid common traps.
-
Statement I: “In borax bead test, the colour of cobalt metaborate is blue”
- Borax (Na₂B₄O₇·10H₂O) on heating forms a glassy bead of sodium metaborate (NaBO₂) and B₂O₃. When a cobalt salt is added, cobalt metaborate (Co(BO₂)₂) forms.
- Cobalt metaborate is indeed blue (a classic test for cobalt).
- ✓ True.
-
Statement II: “Diborane is prepared by the oxidation of sodium borohydride with iodine”
- The reaction is:
2NaBH4+I2→B2H6+2NaI+H2
- Iodine oxidises BH₄⁻, coupling two boron centres to form diborane. This is a standard laboratory preparation.
- ✓ True.
-
Statement III: “In diborane oxidation state of hydrogen is +1”
- In B₂H₆, the terminal hydrogens are slightly hydridic (H⁻), and the bridging hydrogens are even more so.
- Electronegativity: B (2.04) < H (2.20), so hydrogen is slightly more electronegative. In practice, the oxidation state of H in B₂H₆ is −1 (hydride), not +1.
- ✗ False.
-
Statement IV: “Boric acid is a tribasic acid”
- Boric acid, H₃BO₃, does not donate protons directly. Instead, it acts as a Lewis acid:
B(OH)3+H2O⇌B(OH)4−+H+
- It is monobasic (only one H⁺ is released per molecule). The three hydroxyl groups are not all acidic.
- ✗ False.
Watch outA common mistake is to count the three OH groups in boric acid and assume it’s tribasic. But boron’s electron deficiency makes it accept an OH⁻, not donate H⁺. Only one proton is liberated.
Conclusion: Only statements I and II are correct.
✓Final answerThe correct option is (A).
ANSWER: A
-
- TG EAPCET 2025Set eng-2025-05-03-AN1 markMCQQ.Which of the following reactions is an example of Clemmensen reduction? (A) R−C(=O)−Cl+H2Pd/BaSO4R−C(=O)−H+HCl (B) R−C(=O)−H(i)NH2−NH2(ii)KOHCH2−CH2OHOHR−CH3 (C) R−C(=O)−OC2H51.DIBAL−H2.H2OR−C(=O)−H+C2H5OH (D) R−C(=O)−CH3Zn−Hg/HClR−CH2−CH3
›Reveal solutionSolution
Clemmensen reduction uses Zn‑Hg amalgam and concentrated HCl to reduce a carbonyl group (C=O) directly to a methylene group (CH₂). The reaction that matches this reagent set is option (D).
Concept & Intuition
The Clemmensen reduction is a classic method for converting a ketone or aldehyde into an alkane. The key idea is that the strongly acidic, reducing environment (Zn‑Hg in HCl) strips the oxygen atom from the carbonyl and replaces it with two hydrogens. This is especially useful when the molecule also contains acid‑sensitive groups that would be destroyed by the basic conditions of the alternative Wolff‑Kishner reduction.
Here, we simply need to identify which option shows a carbonyl compound being treated with Zn‑Hg / HCl to give a hydrocarbon.
Step‑by‑Step Reasoning
-
Recall the defining reagents of Clemmensen reduction.
The reaction uses zinc amalgam (Zn‑Hg) and concentrated hydrochloric acid (HCl). The substrate is typically a ketone or aldehyde, and the product is the corresponding alkane (the C=O becomes CH₂).
-
Examine each option for the correct reagent set.
- (A) Uses Pd/BaSO₄ (Lindlar’s catalyst) with H₂ — that’s a hydrogenation of an acid chloride to an aldehyde (Rosenmund reduction), not Clemmensen.
- (B) Shows hydrazine (NH₂‑NH₂) followed by KOH in ethylene glycol — that’s the Wolff‑Kishner reduction (basic conditions), not Clemmensen.
- (C) Uses DIBAL‑H then water — that’s a partial reduction of an ester to an aldehyde, not Clemmensen.
- (D) Shows a ketone (R‑CO‑CH₃) treated with Zn‑Hg / HCl and yields R‑CH₂‑CH₃ — exactly the Clemmensen reduction.
-
Confirm the transformation in (D).
The carbonyl group in the starting ketone is reduced to a methylene group:
R−C(=O)−CH3Zn−Hg/HClR−CH2−CH3
This matches the textbook definition of Clemmensen reduction.
Watch outA common mistake is to confuse Clemmensen (acidic, Zn‑Hg/HCl) with Wolff‑Kishner (basic, NH₂NH₂/KOH). Option (B) is the Wolff‑Kishner reduction, not Clemmensen.
TipIf you ever see “Zn‑Hg / conc. HCl” in a reaction scheme, you can immediately think: “This is Clemmensen reduction — the C=O is being turned into CH₂.”
✓Final answerThe correct option is (D).
ANSWER: D
-
- TG EAPCET 2025Set ap-2025-04-29-AN1 markMCQQ.An alcohol X (C4H10O) reacts with Conc.HCl|ZnCl2 to give corresponding chloride. X on dehydration forms Y, which reacts with Baeyer's reagent to give Z. What is Z? (A) \chemfig{*6(-=-=-(-[::+60]C(=[::+60]O)-[::-60]CO_2)-=)} (B) CH3COOH (C) \chemfig{CH_3-CH_2-CH_2-CHO} (D) \chemfig{CH_3-CH(OH)-CH_2-CH_2OH}
›Reveal solutionSolution
Alcohol X, identified as butan-1-ol, undergoes dehydration to form but-1-ene (Y). This alkene then reacts with Baeyer's reagent to yield butane-1,2-diol (Z). The final product is CH3−CH(OH)−CH2−CH2OH.
The problem describes a sequence of reactions starting from an alcohol X with the molecular formula C4H10O. We need to identify the final product Z. Let's break down the reactions and the properties of the compounds involved.
Concept and Intuition
-
Identifying Alcohol X: The formula C4H10O indicates a saturated monohydric alcohol. There are four possible structural isomers. The first reaction, with Conc. HCl/ZnCl2 (Lucas reagent), is a diagnostic test for the class of alcohol (primary, secondary, or tertiary). While all alcohols can react, their rates differ significantly. Tertiary alcohols react fastest, followed by secondary, and then primary alcohols. The problem states X "reacts" to give the chloride, implying the reaction occurs, which helps narrow down possibilities when combined with subsequent steps.
-
Dehydration to Alkene Y: Alcohols undergo dehydration in the presence of strong acids or heat to form alkenes. This reaction typically follows Zaitsev's rule, where the most substituted (and thus most stable) alkene is the major product.
-
Reaction with Baeyer's Reagent to Diol Z: Baeyer's reagent (cold, dilute, alkaline KMnO4) is used to test for unsaturation (carbon-carbon double or triple bonds). It causes syn-dihydroxylation of alkenes, meaning two hydroxyl groups are added to the same face of the double bond, forming a vicinal diol (a 1,2-diol). The purple color of KMnO4 disappears, and a brown precipitate of MnO2 forms.
By systematically applying these reactions to the possible isomers of X and checking against the given options for Z, we can determine the correct pathway.
Step-by-Step Solution
-
Identify possible isomers of alcohol X (C4H10O):
The molecular formula C4H10O corresponds to the following saturated alcohols:
- Butan-1-ol (1° alcohol): CH3CH2CH2CH2OH
- Butan-2-ol (2° alcohol): CH3CH2CH(OH)CH3
- 2-Methylpropan-1-ol (1° alcohol): (CH3)2CHCH2OH
- 2-Methylpropan-2-ol (3° alcohol): (CH3)3COH
-
Analyze the reaction of X with Conc. HCl/ZnCl2 (Lucas reagent):
This reaction converts an alcohol to an alkyl chloride.
R−OH+HClZnCl2R−Cl+H2O
The reactivity order is 3° > 2° > 1°. While primary alcohols react very slowly at room temperature, they do react, especially if heated or given sufficient time. The problem states X "reacts... to give corresponding chloride," which means all these alcohols are potential candidates. We will use the subsequent reactions to pinpoint X.
-
Analyze the dehydration of X to form Y:
Alcohols undergo elimination of water to form alkenes.
R−CH2−CH2−OHH2SO4/ΔR−CH=CH2+H2O
- If X is Butan-1-ol, Y would be but-1-ene (CH3CH2CH=CH2).
- If X is Butan-2-ol, Y would be a mixture of but-1-ene (minor) and but-2-ene (major, due to Zaitsev's rule).
- If X is 2-Methylpropan-1-ol, Y would be 2-methylpropene ((CH3)2C=CH2).
- If X is 2-Methylpropan-2-ol, Y would be 2-methylpropene ((CH3)2C=CH2).
-
Analyze the reaction of Y with Baeyer's reagent to form Z:
Baeyer's reagent (cold, dilute, alkaline KMnO4) performs syn-dihydroxylation on alkenes, forming vicinal diols.
R−CH=CH2KMnO4/OH−/H2OR−CH(OH)−CH2OH
-
Evaluate each possible alcohol X against the given options for Z:
Let's consider the options for Z:
(A) A complex aromatic compound - not possible from a 4-carbon aliphatic alcohol.
(B) CH3COOH (acetic acid) - a 2-carbon carboxylic acid, not a 4-carbon diol.
(C) CH3−CH2−CH2−CHO (butanal) - a 4-carbon aldehyde, not a diol.
(D) CH3−CH(OH)−CH2−CH2OH (butane-1,2-diol) - a 4-carbon vicinal diol. This is a plausible product from the dihydroxylation of a 4-carbon alkene.
Now, let's work backward or forward to see which X leads to Z as butane-1,2-diol:
-
If Z is butane-1,2-diol (CH3CH2CH(OH)CH2OH):
This diol is formed from the syn-dihydroxylation of but-1-ene (CH3CH2CH=CH2).
Therefore, Y must be but-1-ene.
-
Which alcohol X dehydrates to form but-1-ene (Y)?
- Butan-1-ol (CH3CH2CH2CH2OH): Dehydrates to form but-1-ene. This is a direct and unambiguous path.
- Butan-2-ol (CH3CH2CH(OH)CH3): Dehydrates to form a mixture of but-1-ene (minor product) and but-2-ene (major product). If Y were predominantly but-2-ene, Z would be butane-2,3-diol (CH3CH(OH)CH(OH)CH3), which is not option (D). While but-1-ene is a minor product, the question implies a specific product Z.
- 2-Methylpropan-1-ol ((CH3)2CHCH2OH): Dehydrates to form 2-methylpropene. This would lead to 2-methylpropane-1,2-diol, not butane-1,2-diol.
- 2-Methylpropan-2-ol ((CH3)3COH): Dehydrates to form 2-methylpropene. This would also lead to 2-methylpropane-1,2-diol, not butane-1,2-diol.
Considering all steps, butan-1-ol is the only alcohol X that consistently leads to butane-1,2-diol (option D) as product Z. Although butan-1-ol is a primary alcohol and reacts slowly with Lucas reagent, the problem statement only says it "reacts," which is true. The subsequent reactions confirm this choice.
The reaction sequence is:
CH3CH2CH2CH2OH (X, Butan-1-ol)
Dehydration CH3CH2CH=CH2 (Y, But-1-ene)
Baeyer’s reagent CH3CH2CH(OH)CH2OH (Z, Butane-1,2-diol)
-
The final product Z is butane-1,2-diol.
✓Final answerThe final product Z is CH3−CH(OH)−CH2−CH2OH.
-
- TG EAPCET 2025Set ap-2025-04-30-FN1 markMCQQ.What are X, Y, Z in the following reaction sequence?
[!FORMULA] CHX3−CH=CHX2HX2O/HX+CX3HX8OXYCX6HX6anhy. AlClX3Z (major)
(A) \ce{Cl2},~\ce{Cl},~\raisebox{0.5em}{\chemfig{*6(-=-=-(-[::-60]C(-[::60]CH_3)(-[::-60]CH_3))-=)}} (B) \text{conc. HCl},~\ce{Cl},~\raisebox{0.5em}{\chemfig{*6(-=-=-(-[::-60]C(-[::60]CH_2CH_3))-=)}} (C) \ce{HCl}/\ce{ZnCl2},~\ce{Cl},~\raisebox{0.5em}{\chemfig{*6(-=-=-(-[::-60]C(-[::60]CH_2CH_2CH_3))-=)}} (D) \ce{HCl}/\ce{ZnCl2},~\ce{Cl},~\raisebox{0.5em}{\chemfig{*6(-=-=-(-[::-60]C(-[::60]CH_2CH_3))-=)}}›Reveal solutionSolution
The reaction sequence involves acid-catalysed hydration of propene to isopropyl alcohol, conversion to isopropyl chloride using Lucas reagent (HCl/ZnClX2), and then Friedel–Crafts alkylation of benzene to give cumene (isopropylbenzene). The correct option is (D).
The key to this problem is recognising each transformation in the sequence and matching the reagents and products to the given options. Let’s walk through it step by step.
-
First step: hydration of propene
Propene (CHX3−CH=CHX2) reacts with water in the presence of an acid catalyst (HX2O/HX+). This is acid-catalysed hydration, which follows Markovnikov’s rule: the hydrogen adds to the less substituted carbon of the double bond, and the hydroxyl group adds to the more substituted carbon.
The product is propan-2-ol (isopropyl alcohol), CHX3−CH(OH)−CHX3, with molecular formula CX3HX8O.
So the intermediate CX3HX8O is isopropyl alcohol.
-
Second step: conversion of alcohol to alkyl halide
The alcohol is now treated with reagent X to give product Y, which is an alkyl chloride (since the next step involves benzene and AlClX3, a Friedel–Crafts alkylation).
To convert an alcohol to an alkyl chloride, common reagents include SOClX2, PClX5, PClX3, or HCl with a catalyst. Among the options, we see ClX2 (which would not directly convert an alcohol to an alkyl chloride — it would give a hypochlorite or chlorinate elsewhere), conc. HCl (which works but slowly for secondary alcohols), and HCl/ZnClX2 (Lucas reagent).
Lucas reagent (HCl/ZnClX2) is specifically used to convert secondary alcohols to alkyl chlorides rapidly at room temperature. For isopropyl alcohol, it gives isopropyl chloride, CHX3−CHCl−CHX3.
Thus X is HCl/ZnClX2 and Y is isopropyl chloride (Cl in the options refers to the isopropyl group attached to chlorine, i.e., CHX3−CHCl−CHX3).
-
Third step: Friedel–Crafts alkylation
Y (isopropyl chloride) reacts with benzene (CX6HX6) in the presence of anhydrous AlClX3, a Lewis acid catalyst. This is a classic Friedel–Crafts alkylation.
The mechanism: AlClX3 abstracts the chlorine from isopropyl chloride, generating an isopropyl carbocation (CHX3−CHX+−CHX3). This carbocation attacks the benzene ring, and after deprotonation, the product is isopropylbenzene, commonly known as cumene.
The structure of cumene is: a benzene ring with an isopropyl group (−CH(CHX3)X2) attached. In the options, this is represented as a benzene ring with a branched three-carbon chain: C(−[::60]CHX3)(−[::-60]CHX3) — that is, a carbon attached to two methyl groups.
-
Matching with the options
- Option (A): ClX2 is wrong for the first step; product is a different alkylbenzene.
- Option (B): conc. HCl is possible but less specific; the product shown has an ethyl group (−CHX2CHX3), not isopropyl.
- Option (C): HCl/ZnClX2 is correct for X, but the product shown has an n-propyl group (−CHX2CHX2CHX3), not isopropyl.
- Option (D): HCl/ZnClX2 for X, and the product is isopropylbenzene (cumene) — exactly what we expect.
Watch outA common mistake is to think that HCl alone is sufficient for the alcohol-to-chloride step. While it works, the reaction is slow for secondary alcohols; Lucas reagent (HCl/ZnClX2) is the standard reagent used in such sequences. Also, be careful with the alkyl group in the final product — Friedel–Crafts alkylation with isopropyl chloride gives a branched isopropyl group, not a straight-chain propyl group.
TipIn Friedel–Crafts alkylation, the carbocation rearranges if a more stable carbocation is possible. Here, the isopropyl carbocation is already secondary and stable, so no rearrangement occurs. If you started with n-propyl chloride, you would get rearrangement to isopropylbenzene anyway — but here we start with isopropyl chloride, so the product is directly cumene.
✓Final answerThe correct option is (D): X is HCl/ZnClX2, Y is isopropyl chloride, and Z is cumene (isopropylbenzene).
-
- TG EAPCET 2024Set eng-2024-05-09-AN1 markMCQQ.The functional groups present in the product 'X' of the reaction given below are Phenyl benzoate — a benzene ring bearing −O−C(=O)−Ph — AlCl3 X (A) −OH, −C(=O)−H (B) −OH, −C(=O)− (C) −O−ph, −C(=O)− (D) −C(=O)−OH, −OH
›Reveal solutionSolution
The reaction is a Fries rearrangement of phenyl benzoate under AlCl3, which migrates the acyl group to the ortho or para position of the phenol ring. The product 'X' contains −OH and −C(=O)− (ketone) groups, making option (B) correct.
The key here is recognising the reaction type. Phenyl benzoate is an ester of phenol and benzoic acid. When treated with a Lewis acid like AlCl3, it undergoes the Fries rearrangement — not a simple hydrolysis or acylation. The AlCl3 coordinates to the ester oxygen, weakening the C−O bond, and the acyl group (−C(=O)−Ph) migrates to the ortho or para position of the phenol ring. After work-up, you get a hydroxy ketone.
Let’s walk through it step by step.
-
Identify the starting material. Phenyl benzoate has the structure Ph−O−C(=O)−Ph. The left benzene ring is attached to oxygen (phenolic part), the right benzene ring is part of the acyl group (−C(=O)−Ph). So the molecule is an aryl ester.
-
Recall the Fries rearrangement. Under AlCl3, aryl esters rearrange to give ortho- and para-hydroxy ketones. The AlCl3 coordinates to the carbonyl oxygen, making the ester more electrophilic. The acyl group then breaks off and attacks the electron-rich ortho/para positions of the phenol ring. This is an intramolecular rearrangement — the acyl group doesn’t leave entirely; it migrates.
-
What happens to the ester linkage? The −O− bond to the original acyl group breaks. The oxygen that was part of the ester becomes a phenolic −OH after work-up (protonation). The acyl group (−C(=O)−Ph) attaches directly to the benzene ring that originally held the oxygen. So the product is HO−C6H4−C(=O)−Ph — a hydroxybenzophenone.
-
Identify the functional groups in the product. The product has:
- A phenolic hydroxyl group (−OH) attached to the benzene ring.
- A ketone carbonyl group (−C(=O)−) linking the same benzene ring to the phenyl group. This is not an aldehyde (−C(=O)−H) and not a carboxylic acid (−C(=O)−OH). It’s simply a ketone.
-
Match with the options.
- (A) −OH, −C(=O)−H — aldehyde, wrong.
- (B) −OH, −C(=O)− — correct: hydroxyl and ketone.
- (C) −O−ph, −C(=O)− — still has an ether link, not formed.
- (D) −C(=O)−OH, −OH — carboxylic acid, not formed.
Watch outA common mistake is to think the ester simply hydrolyses under AlCl3 to give phenol and benzoic acid. That would require water, which is not present. The Fries rearrangement is specific to aryl esters and Lewis acids — the product is a hydroxy ketone, not a mixture of acid and phenol.
TipThe Fries rearrangement is the reverse of the Baeyer–Villiger oxidation in a conceptual sense — here an ester becomes a hydroxy ketone, whereas Baeyer–Villiger turns a ketone into an ester. Remembering this pair can help in exams.
✓Final answerThe correct option is (B), as the product contains a phenolic −OH group and a ketone −C(=O)− group.
-
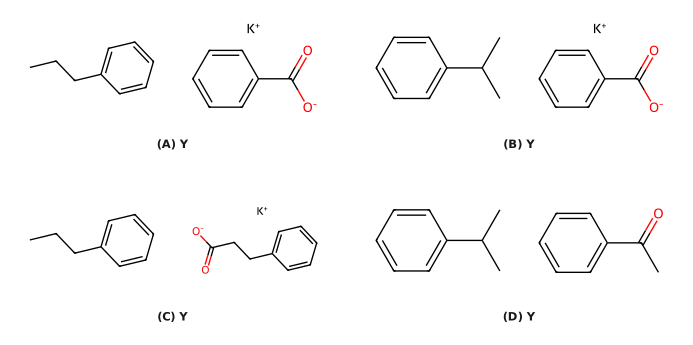
- TG EAPCET 2024Set ap-2024-05-07-AN1 markMCQQ.Hydrolysis of an alkyl halide X (C3H7Br) follows second order kinetics. Reaction of X with C6H5Cl in the presence of Na/dry ether gave Y. Oxidation of Y in the presence of KMnO4/OH gave Z. What are Y and Z respectively? (A) C6H5CH2CH2CH3 (n-propylbenzene) and C6H5COOK (potassium benzoate) (B) C6H5CH(CH3)2 (isopropylbenzene) and C6H5COOK (potassium benzoate) (C) C6H5CH2CH2CH3 (n-propylbenzene) and C6H5CH2CH2COOK (D) C6H5CH(CH3)2 (isopropylbenzene) and C6H5COCH3 (acetophenone)
›Reveal solutionSolution
Second-order hydrolysis ⇒SN2⇒ X is the primary halide, n-propyl bromide. Wurtz–Fittig with chlorobenzene gives Y = n-propylbenzene, and KMnO4/OH− chops the whole side chain to give Z = potassium benzoate. Option (A).
The concept first — three ideas, one chain
- Kinetics tells you the structure. SN1 is unimolecular (rate =k[RX], favoured by 3° > 2° > 1°), while SN2 is bimolecular (rate =k[RX][Nu], favoured by 1° > 2° > 3°, because the backside attack needs an uncrowded carbon). So "second-order kinetics" is the paper telling you X is primary.
- Wurtz–Fittig. Aryl halide + alkyl halide + 2Na in dry ether → alkylbenzene:
C6H5Cl+2Na+R−Brdry etherC6H5−R+NaCl+NaBr
- Side-chain oxidation. Hot KMnO4 attacks the benzylic carbon. Whatever the length of the alkyl chain, as long as there is at least one hydrogen on the benzylic carbon, the entire chain is degraded to a single –COOH group attached to the ring. (Only a chain with no benzylic H, like tert-butylbenzene, resists.)
Step-by-step
- Identify X. C3H7Br has two isomers. Isopropyl bromide (2°) would hydrolyse largely by SN1 (first-order). Since the observed hydrolysis is second order, the mechanism is SN2, so
X=CH3CH2CH2Br(n-propyl bromide, 1∘)
- Form Y (Wurtz–Fittig).
C6H5Cl+CH3CH2CH2Br+2Nadry etherC6H5CH2CH2CH3+NaCl+NaBr
Y=n-propylbenzene
Note the alkyl group keeps its skeleton — no rearrangement — so we get the n-propyl, not the isopropyl, product.
3. Oxidise to Z. Alkaline KMnO4 (then the alkaline work-up) oxidises the propyl side chain completely:
C6H5CH2CH2CH3KMnO4/OHΔC6H5COO−K++(2 C lost as carbonate/CO2)
Z=C6H5COOK (potassium benzoate)
Because the medium is alkaline, the product isolated is the potassium salt, not the free acid (acidification would be needed to get benzoic acid).
4. Match the options. Y = n-propylbenzene, Z = potassium benzoate → (A). Option (B) wrongly assumes X is isopropyl bromide (which would need first-order kinetics); (C) wrongly keeps two carbons of the side chain (side-chain oxidation never stops half-way); (D) is wrong on both counts.
✓Final answerX = n-propyl bromide (SN2), Y = n-propylbenzene (Wurtz–Fittig), Z = potassium benzoate (complete side-chain oxidation).
ANSWER: A
- TG EAPCET 2024Set ap-2024-05-07-FN1 markMCQQ.Phenatole can be prepared from which of the following reactants? (A) \chemfig∗6(−=−(−ONa)−=−)CH3CH2Br, (B) \chemfig∗6(−=−(−Br)−=−)CH3CH2ONa, (C) \chemfig∗6(−=−(−Br)−=−)CH3CH2CH2ONa, (D) \chemfig∗6(−=−(−ONa)−=−)CH3CH2CH2Br,
›Reveal solutionSolution
Phenatole (phenetole) is ethyl phenyl ether (C₆H₅OCH₂CH₃). It is best prepared by the Williamson ether synthesis: reacting sodium phenoxide with ethyl bromide. The correct reactants are sodium phenoxide and ethyl bromide, which corresponds to option (A).
Concept & Intuition
Phenatole (often spelled phenetole) is the common name for ethyl phenyl ether, C₆H₅–O–CH₂CH₃. The classic laboratory method for making unsymmetrical ethers like this is the Williamson ether synthesis: an alkoxide (or phenoxide) ion attacks an alkyl halide in an Sₙ2 reaction.
The key is to choose the combination that gives the desired ether in high yield. The best route uses the more nucleophilic oxygen (the phenoxide ion) and the less hindered alkyl halide (primary halide). If you reverse the roles — using a phenoxide with a secondary or tertiary halide — elimination dominates. If you use an alkoxide with an aryl halide, the reaction fails because aryl halides are unreactive toward Sₙ2.
Step-by-step reasoning
-
Identify the target molecule
Phenatole = ethyl phenyl ether = C₆H₅–O–CH₂CH₃. The oxygen is bonded to a phenyl group and an ethyl group.
-
Recall the Williamson ether synthesis
The general reaction:
RONa+R′X→ROR′+NaX
The alkoxide (or phenoxide) acts as the nucleophile; the alkyl halide must be primary (or methyl) to favor Sₙ2 over elimination.
-
Examine each option
- (A) Sodium phenoxide (C₆H₅ONa) + ethyl bromide (CH₃CH₂Br) Phenoxide is a good nucleophile; ethyl bromide is a primary halide. Sₙ2 proceeds cleanly:
C6H5ONa+CH3CH2Br→C6H5OCH2CH3+NaBr
This gives phenatole directly. ✓-
(B) Bromobenzene (C₆H₅Br) + sodium ethoxide (CH₃CH₂ONa)
Aryl halides do not undergo Sₙ2 reactions (the carbon–halogen bond has partial double‑bond character; backside attack is impossible). This would require harsh conditions (e.g., Ullmann coupling) and is not a simple Williamson synthesis. ✗
-
(C) Bromobenzene + sodium propoxide (CH₃CH₂CH₂ONa)
Same problem as (B): aryl halide is unreactive in Sₙ2. Even if it reacted, the product would be propyl phenyl ether, not ethyl phenyl ether. ✗
-
(D) Sodium phenoxide + propyl bromide (CH₃CH₂CH₂Br)
This would give propyl phenyl ether (C₆H₅OCH₂CH₂CH₃), not phenatole. ✗
- Confirm the correct choice Only option (A) pairs the correct nucleophile (phenoxide) with the correct primary alkyl halide (ethyl bromide) to yield ethyl phenyl ether.
Watch outA common mistake is to think that an aryl halide can be used in a Williamson synthesis. Aryl halides are inert to Sₙ2; you must always put the aryl group on the oxygen (as the phenoxide) and the alkyl group on the halide.
TipThe Williamson synthesis is most reliable when the alkyl halide is primary. If you need an ether like methyl phenyl ether (anisole), use methyl iodide + sodium phenoxide — never bromobenzene + sodium methoxide.
✓Final answerThe correct option is (A).
ANSWER: A
-
🎓Unlock everything free for 14 days
- ✓Full step-by-step solutions
- ✓Concept-first explanations
- ✓Methods, shortcuts & mistakes
- ✓PYQ mapping + timed mock tests
Full access for 14 days. No credit card required.