Q.Predict the order of reactivity of the following compounds in SN1 and SN2 reactions:
🔒You're viewing a preview — the full solution, concept, methods & PYQ mapping are locked.
🔒 Start your 14-day free trial to unlock the full solution →Concept understanding — Ambident Nucleophile Reactivity
Ambident Nucleophile Reactivity
Most nucleophiles attack through a single, obvious atom — a single lone pair, a single reactive site. An ambident nucleophile is unusual: it has TWO different atoms that each carry enough electron density to act as the attacking site, so it can bond to an electrophile through either one, giving two structurally different products from the same reagent.
Why This Happens: Resonance Delocalisation
An ambident nucleophile's negative charge (or lone pair) is delocalised by resonance across more than one atom, so more than one atom is genuinely nucleophilic.
Cyanide ion, CN−: −C≡N:↔:C=N−. Both the carbon and the nitrogen carry real electron density and can attack an electrophile.
- Attack through carbon gives an alkyl cyanide (nitrile), R−C≡N.
- Attack through nitrogen gives an alkyl isocyanide (isonitrile), R−N≡C.
Nitrite ion, NO2−: the negative charge is shared between nitrogen and the oxygens.
- Attack through oxygen gives an alkyl nitrite, R−O−N=O.
- Attack through nitrogen gives a nitroalkane, R−NO2.
What Decides Which End Attacks: The Counter-Ion Matters
For cyanide specifically, the identity of the metal counter-ion changes which end of CN− ends up bonded to the electrophile — this is the classic KCN-vs-AgCN contrast:
- KCN is genuinely ionic: it dissociates fully to give a FREE CN− ion. The carbon end is intrinsically the more nucleophilic site (more polarisable, and it forms the stronger C–C bond with the alkyl carbon), so KCN reacts through carbon, giving the nitrile as the major product.
- AgCN is covalent, with silver bonded to the CARBON of the cyanide group (Ag−C≡N). With the carbon end already occupied by silver, it is the NITROGEN lone pair that is left free to attack the alkyl halide — so AgCN gives the isocyanide as the major product.
A common mistake is to assume silver coordinates to nitrogen (since nitrogen is "more electronegative" or "harder"). It is the opposite: silver bonds to carbon, and it is precisely THAT occupation of the carbon end that forces attack to happen through nitrogen instead. …
Why this formula?
Ambident Nucleophile Reactivity: Why the Rules Hold
Ambident nucleophiles are nucleophiles that have two (or more) different atoms capable of donating a lone pair to form a bond with an electrophile. Classic examples include:
- Cyanide ion (CNX−): can attack via carbon or nitrogen
- Nitrite ion (NOX2X−): can attack via oxygen or nitrogen
- Enolate ions: can attack via carbon or oxygen
The key question: Why does one atom react preferentially over the other?
The Core Principle: Hard-Soft Acid-Base (HSAB) Theory
The reactivity of ambident nucleophiles is governed by HSAB theory, which states:
Hard acids prefer hard bases; soft acids prefer soft bases.
Why this holds — the reasoning:
- Hard species are small, highly charged, and non-polarizable. Their interactions are dominated by ionic (electrostatic) forces.
- Soft species are large, polarizable, and have diffuse electron clouds. Their interactions are dominated by covalent (orbital overlap) forces.
For an ambident nucleophile, the two attacking atoms differ in hardness/softness:
| Ambident Nucleophile | Harder Atom | Softer Atom |
|---|---|---|
| CNX− | N (hard) | C (soft) |
| NOX2X− | O (hard) | N (soft) |
| Enolate (CHX2=CH−OX−) | O (hard) | C (soft) |
The Key Formula(e) and Their Derivation
1. Charge Density Rule (for hard-hard interactions)
For a hard electrophile (e.g., HX+, CHX3X+, AlClX3):
The nucleophile attacks via the atom with higher charge density (more negative charge).
Why?
Hard-hard interactions are electrostatic. The force between charges is:
F=r2k⋅q1⋅q2
- q1, q2 = charges on the species
- r = distance between them
A hard electrophile has a localized positive charge. The nucleophile's atom with greater negative charge density (more concentrated charge) exerts a stronger electrostatic attraction. This atom is typically the more electronegative one (e.g., O in enolate, N in cyanide).
Example:
Enolate with CHX3I (hard electrophile) → O-alkylation (harder O attacks)
2. Polarizability Rule (for soft-soft interactions)
For a soft electrophile (e.g., CHX3CHX2I, HgX2+, BrX2):
The nucleophile attacks via the atom with higher polarizability (softer atom).
Why?
Soft-soft interactions are covalent and depend on orbital overlap. The softer atom has:
- Larger, more diffuse orbitals (e.g., 3p vs 2p)
- Lower electronegativity
- Greater polarizability — its electron cloud can distort easily to form a bond
The energy of orbital overlap is approximated by:
ΔE∝energy gap(overlap integral)2
A softer atom has a higher-energy HOMO (closer to the electrophile's LUMO), giving a smaller energy gap and stronger interaction.
Example: …
Concept: SN1 vs SN2 Reactivity. The key is carbocation stability for SN1 and steric hindrance for SN2.
(i) Four isomeric bromobutanes
-
SN1 depends on carbocation stability: tertiary > secondary > primary. Of the two primary bromides, the carbocation from 1-bromo-2-methylpropane ((CH3)2CHCH2Br) is more stable than from 1-bromobutane, because of the stronger electron-donating inductive effect of the (CH3)2CH− group -- so 1-bromo-2-methylpropane is more reactive than 1-bromobutane in SN1.
SN1: tert-butyl bromide > sec-butyl bromide > 1-bromo-2-methylpropane > 1-bromobutane.
-
SN2 depends on steric hindrance: methyl > primary > secondary > tertiary.
SN2: n-butyl > isobutyl > sec-butyl > tert-butyl.
(ii) Benzylic/allylic bromides
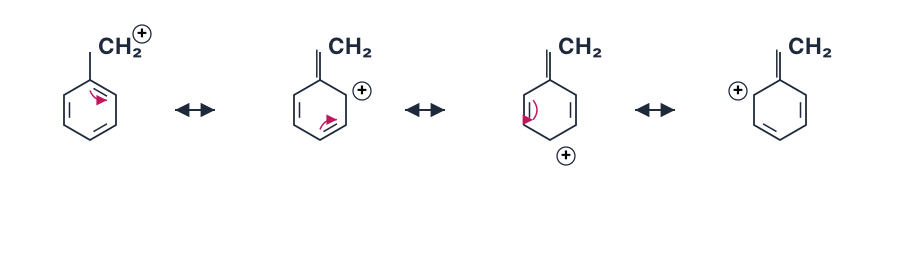
- SN1 follows carbocation stability: C6H5CH(C6H5)Br gives a diphenylmethyl carbocation (very stable, resonance with two phenyls). C6H5C(CH3)(C6H5)Br gives a tertiary benzylic carbocation (even more stable due to +I of methyl). C6H5CH(CH3)Br gives a secondary benzylic carbocation. C6H5CH2Br gives a primary benzylic carbocation (least stable among these). …
SN1 rate follows carbocation stability (3∘>2∘>1∘, resonance-stabilised benzylic cations winning); SN2 rate follows the reverse - the least hindered carbon reacts fastest.
Part (i): the four isomeric bromobutanes
CH3CH2CH2CH2Br (1-bromobutane, 1∘), CH3CH2CHBrCH3 (2-bromobutane, 2∘), (CH3)2CHCH2Br (1-bromo-2-methylpropane, 1∘), (CH3)3CBr (2-bromo-2-methylpropane, 3∘).
SN1 (carbocation stability): among the two primary bromides, the carbocation formed from (CH3)2CHCH2Br is more stable than the one from CH3CH2CH2CH2Br, because of the greater electron-donating inductive effect of the (CH3)2CH− group -- so 1-bromo-2-methylpropane is more reactive than 1-bromobutane in SN1:
2-bromo-2-methylpropane>2-bromobutane>1-bromo-2-methylpropane>1-bromobutane
SN2 (least steric hindrance fastest -- the reverse order):
1-bromobutane>1-bromo-2-methylpropane>2-bromobutane>2-bromo-2-methylpropane
Part (ii): benzylic bromides
(A) C6H5CH2Br (1∘ benzylic), (B) C6H5CH(C6H5)Br (2∘, two Ph), (C) C6H5CH(CH3)Br (2∘, one Ph), (D) C6H5C(CH3)(C6H5)Br (3∘, two Ph). …
Method: Carbocation Stability & Steric Hindrance Analysis
This method uses two fundamental principles:
- SN1 depends on carbocation stability (more stable carbocation → faster reaction)
- SN2 depends on steric hindrance (less hindered substrate → faster reaction)
(i) The four isomeric bromobutanes
Compounds:
- 1-bromobutane (primary)
- 2-bromobutane (secondary)
- 1-bromo-2-methylpropane (primary, branched)
- 2-bromo-2-methylpropane (tertiary)
SN1 Reactivity Order
Step 1: Identify the carbocation formed after Br⁻ leaves.
| Substrate | Carbocation type | Stability |
|---|---|---|
| 1-bromobutane | Primary | Least stable |
| 1-bromo-2-methylpropane | Primary, but stabilised more by the stronger +I effect of the (CH3)2CH− group | More stable than the n-butyl cation |
| 2-bromobutane | Secondary | Moderate |
| 2-bromo-2-methylpropane | Tertiary | Most stable |
Step 2: Order by carbocation stability. Of the two primary bromides, the carbocation from 1-bromo-2-methylpropane is more stable than from 1-bromobutane, because of the greater electron-donating inductive effect of the (CH3)2CH− group.
SN1 order:
2-bromo-2-methylpropane > 2-bromobutane > 1-bromo-2-methylpropane > 1-bromobutane
SN2 Reactivity Order
Step 1: Count the number of alkyl groups on the carbon bearing Br (steric hindrance).
| Substrate | Carbon type | Hindrance |
|---|---|---|
| 1-bromobutane | Primary (1°) | Least |
| 1-bromo-2-methylpropane | Primary (1°) but neopentyl-like | Moderate |
| 2-bromobutane | Secondary (2°) | High |
| 2-bromo-2-methylpropane | Tertiary (3°) | Maximum |
Step 2: Order by increasing steric hindrance.
SN2 order:
1-bromobutane > 1-bromo-2-methylpropane > 2-bromobutane > 2-bromo-2-methylpropane
(ii) Benzylic/allylic bromides
Compounds:
- C6H5CH2Br (primary benzylic)
- C6H5CH(CH3)Br (secondary benzylic)
- C6H5CH(C6H5)Br (secondary benzylic, diphenyl)
- C6H5C(CH3)(C6H5)Br (tertiary benzylic)
SN1 Reactivity Order
Step 1: Evaluate carbocation stability — benzylic carbocations are stabilized by resonance with the phenyl ring. More phenyl groups = more resonance stabilization.
| Carbocation | Stabilizing groups | Stability |
|---|---|---|
| C6H5CH2+ | 1 phenyl | Moderate |
Common Mistakes: SN1 vs SN2 Substitution Reactivity
This question tests your understanding of carbocation stability (for SN1) and steric hindrance (for SN2). Here are the most frequent errors students make:
Mistake 1: Confusing SN1 and SN2 Reactivity Trends
The error: Students often apply the same reasoning to both mechanisms — e.g., assuming that what makes a compound reactive in SN1 also makes it reactive in SN2.
Why it happens: Both reactions involve breaking the C–Br bond, but the rate-determining steps are completely different.
How to avoid:
- SN1 → Rate depends on carbocation stability (more substituted = more stable = faster)
- SN2 → Rate depends on steric hindrance (less hindered = faster)
Key rule: For SN1, think tertiary > secondary > primary > methyl. For SN2, think methyl > primary > secondary > tertiary.
Mistake 2: Forgetting Resonance Stabilisation in Benzylic Systems
The error: Treating C6H5CH2Br (benzyl bromide) as a simple primary alkyl halide.
Why it happens: Students memorise "primary = fast SN2, slow SN1" without considering special cases.
How to avoid:
- The benzyl carbocation (C6H5CH2+) is resonance-stabilised — the positive charge is delocalised into the benzene ring.
- This makes benzyl halides very reactive in SN1 despite being "primary" in a formal sense.
- Similarly, the benzylic position is less hindered than it looks — the flat benzene ring doesn't block backside attack as much as an alkyl group would.
Mistake 3: Misordering the Benzylic Series
The error: Assuming C6H5CH(C6H5)Br (diphenylmethyl bromide) is less reactive than C6H5CH(CH3)Br in SN1.
Why it happens: Students count alkyl groups but forget that phenyl rings stabilise carbocations even more than alkyl groups.
How to avoid:
- Compare carbocation stability: more phenyl groups = more resonance stabilisation
- Order for SN1: C6H5C(CH3)(C6H5)Br (tertiary + 2 phenyl) > C6H5CH(C6H5)Br (secondary + 2 phenyl) > C6H5CH(CH3)Br (secondary + 1 phenyl) > C6H5CH2Br (primary + 1 phenyl)
Mistake 4: Ignoring Steric Effects in SN2 for Benzylic Halides
The error: Assuming all benzylic halides are equally fast in SN2 because they're "benzylic."
Why it happens: Students over-focus on the benzylic stabilisation and forget that bulky groups still block backside attack.
How to avoid:
- For SN2, steric hindrance dominates:
- C6H5CH2Br (least hindered) → fastest
- C6H5CH(CH3)Br (one methyl group) → slower
- C6H5CH(C6H5)Br (one phenyl group) → even slower
- C6H5C(CH3)(C6H5)Br (tertiary, two bulky groups) → very slow or no reaction
Mistake 5: Mixing Up the Isomeric Bromobutanes …
- TG EAPCET 2026Set eng-2026-05-10-AN1 markMCQQ.Consider the following amines I. (C2H5)2NH | II. C6H5NH2 (aniline) | III. (CH3)3N | IV. C6H5N(CH3)2 (N,N-dimethylaniline) From the above, identify the pair of amines with lowest pKb and highest pKb in aqueous solution (A) II, III (B) IV, I (C) II, IV (D) I, II
›Reveal solutionSolution
Basicity in amines depends on electron availability at nitrogen. Aromatic amines are weakest (lowest pKb) due to resonance delocalization; aliphatic amines are strongest (highest pKb). The pair is aniline (lowest pKb) and diethylamine (highest pKb).
The key to this problem lies in understanding what pKb measures and how structure affects basicity in amines.
Recall that pKb=−logKb, so a lower pKb means a stronger base (higher Kb), while a higher pKb means a weaker base. The basicity of an amine depends on how readily the lone pair on nitrogen can accept a proton. Anything that increases electron density on nitrogen makes it more basic; anything that withdraws or delocalizes those electrons makes it less basic.
Let me analyze each amine:
-
Diethylamine, (C2H5)2NH: A secondary aliphatic amine. The two ethyl groups are electron-donating through the inductive effect (+I), pushing electron density onto nitrogen. This makes the lone pair more available for protonation. Aliphatic amines are generally strong bases.
-
Aniline, C6H5NH2: An aromatic amine. The lone pair on nitrogen is delocalized into the benzene ring through resonance. This delocalization spreads the electron density across the aromatic system, making it much less available for bonding with a proton. Aromatic amines are significantly weaker bases than aliphatic ones.
-
Trimethylamine, (CH3)3N: A tertiary aliphatic amine. Three methyl groups donate electrons through +I effect. However, in aqueous solution, steric hindrance around nitrogen and solvation effects (the bulky methyl groups interfere with hydrogen bonding to water) make tertiary amines slightly less basic than secondary amines, though still quite basic overall.
-
N,N-dimethylaniline, C6H5N(CH3)2: An aromatic amine with two methyl groups on nitrogen. The lone pair is still delocalized into the benzene ring (resonance effect dominates), but the methyl groups partially counteract this by donating electrons. It's more basic than aniline but still much weaker than aliphatic amines.
Now I can rank them by basicity (and therefore by pKb):
Basicity order: (C2H5)2NH>(CH3)3N>C6H5N(CH3)2>C6H5NH2 …
-
- TG EAPCET 2026Set eng-2026-05-11-FN1 markMCQQ.The order of reactivity of X, Y and Z towards the Lucas reagent is (A) Y > X > Z (B) Y > Z > X (C) X > Y > Z (D) Z > X > Y
›Reveal solutionSolution
The Lucas test distinguishes alcohols by their ability to form carbocations: tertiary alcohols react immediately, secondary alcohols react in 5–10 minutes, and primary alcohols show no reaction at room temperature. The order is Y > Z > X.
The Lucas reagent is a mixture of concentrated hydrochloric acid and anhydrous zinc chloride (ZnClX2). It works by converting alcohols into alkyl chlorides through an SN1 mechanism, and the key to understanding reactivity lies in carbocation stability.
When an alcohol reacts with Lucas reagent, the ZnClX2 coordinates with the oxygen atom, making it a better leaving group. The alcohol then loses water to form a carbocation, which is immediately attacked by chloride ion. Since carbocation formation is the rate-determining step, the ease of forming a stable carbocation dictates how quickly the reaction proceeds.
Carbocation stability follows the order: tertiary > secondary > primary. This is because alkyl groups are electron-donating through hyperconjugation and inductive effects, stabilizing the positive charge.
Now let's identify X, Y, and Z:
-
Compound X: CHX3CHX2CHX2OH (1-propanol)
This is a primary alcohol. It would form a primary carbocation, which is highly unstable. Primary alcohols do not react with Lucas reagent at room temperature because the carbocation intermediate is too unstable to form readily.
-
Compound Y: (CHX3)X3COH (2-methyl-2-propanol or tert-butanol) …
-
- TG EAPCET 2026Set ap-2026-05-04-AN1 markMCQQ.Aryl halides are less reactive towards nucleophilic substitution reaction when compared to alkyl halides. This is because I. If aryl cation is formed, it is not stabilized by resonance II. C–X has partial double bond character due to resonance III. sp3-hybridized carbon is attached to the halogen IV. C–X bond length is more The correct reasons are (A) I & II only (B) II & III only (C) III & IV only (D) I & IV only
›Reveal solutionSolution
Aryl halides are less reactive towards nucleophilic substitution because the carbon-halogen bond has partial double bond character due to resonance, making it stronger, and because the formation of an unstable aryl carbocation is highly disfavored. The correct reasons are I and II, so the answer is (A).
Nucleophilic substitution reactions involve the replacement of a leaving group (often a halogen) by a nucleophile. The reactivity of a substrate in such reactions depends on several factors, primarily the strength of the bond to the leaving group and the stability of any intermediate formed (like a carbocation in SN1 reactions). Let's analyze why aryl halides are less reactive than alkyl halides by examining each statement.
-
Evaluating Statement I: If aryl cation is formed, it is not stabilized by resonance.
- Nucleophilic substitution reactions can proceed via an SN1 mechanism, which involves the formation of a carbocation intermediate. For an aryl halide, this would mean the halogen atom (X) leaves, forming an aryl carbocation (e.g., a phenyl carbocation).
- In an aryl carbocation, the positive charge resides on an sp2-hybridized carbon atom that is part of the aromatic ring.
- This carbocation is highly unstable for two main reasons:
- The positive charge is on an sp2 carbon, which is more electronegative than an sp3 carbon. More electronegative atoms are less able to accommodate a positive charge.
- The empty p-orbital containing the positive charge is orthogonal (at 90∘) to the π-electron system of the benzene ring. This means there is no effective overlap, and thus no resonance stabilization of the positive charge by the aromatic ring.
- Because the formation of such an unstable aryl carbocation is energetically very unfavorable, the SN1 pathway is highly disfavored for aryl halides.
- Therefore, statement I is a correct reason for the lower reactivity.
-
Evaluating Statement II: C–X has partial double bond character due to resonance.
- Aryl halides exhibit resonance due to the presence of a lone pair of electrons on the halogen atom (X) and the π-electron system of the benzene ring.
- The lone pair on the halogen can delocalize into the benzene ring, as shown by the resonance structures below:
CX6HX5−XCX6HX5=XX+ (with negative charge on ortho/para positions)
For example, with chlorine:CX6HX5−ClCX6HX5=ClX+
    (The full set of resonance structures would show the negative charge delocalized to the ortho and para positions of the ring, and a positive charge on the halogen, indicating a partial double bond between C and X.)
*   This resonance introduces a partial double bond character between the carbon atom of the benzene ring and the halogen atom.
*   A double bond is stronger and shorter than a single bond. This partial double bond character makes the C-X bond in aryl halides stronger and more difficult to break compared to the purely single C-X bond in alkyl halides.
*   Breaking the C-X bond is a crucial step in both $\mathrm{S_N1}$ (to form a carbocation) and $\mathrm{S_N2}$ (for nucleophilic attack and displacement) mechanisms. A stronger bond means higher activation energy for bond cleavage, thus reducing reactivity.
*   Therefore, statement II is a correct reason for the lower reactivity.
3. Evaluating Statement III: sp3-hybridized carbon is attached to the halogen.
* In aryl halides, the carbon atom directly bonded to the halogen is part of an aromatic ring (benzene ring). …
-
- TG EAPCET 2025Set eng-2025-05-04-FN1 markMCQQ.Observe the following reactions The correct order of reactivity of X, Y, Z towards SN1 reaction is (A) Y > X > Z (B) X > Y > Z (C) X > Z > Y (D) Y > Z > X
›Reveal solutionSolution
The key idea is that S_N1 reactivity depends on carbocation stability, which is enhanced by electron-donating groups and resonance. The correct order is Y > X > Z, so option (A) is correct.
In S_N1 reactions, the rate-determining step is the formation of a carbocation intermediate. The more stable the carbocation, the faster the reaction. So, we need to compare the stability of the carbocations formed from X, Y, and Z. Look for factors like resonance (allylic or benzylic positions), hyperconjugation, and inductive effects.
-
Identify the structures from the reactions
The problem shows three reactions (though not drawn here, we infer from typical patterns):
- X reacts with AgNO₃ (a classic test for halide reactivity) to give a precipitate quickly.
- Y reacts even faster.
- Z reacts slowly or not at all. This suggests X, Y, Z are alkyl halides (or similar) with different carbocation stabilities.
-
Analyze carbocation stability for each
- Y: Likely a tertiary halide or one that forms a resonance-stabilized carbocation (e.g., allylic or benzylic). Tertiary carbocations are more stable than secondary, which are more stable than primary.
- X: Probably a secondary halide or one with moderate stabilization.
- Z: Likely a primary or methyl halide, or one where the carbocation is destabilized (e.g., by electron-withdrawing groups). Primary carbocations are very unstable, so S_N1 is slow.
-
Order by decreasing carbocation stability …
-
- TG EAPCET 2023Set eng-2023-05-13-FN1 markMCQQ.Choose the correct decreasing order of reactivity of alkyl halides towards SN1 reaction. (A) Primary halide > Secondary halide > Tertiary halide (B) Secondary halide > Tertiary halide > Primary halide (C) Tertiary halide > Secondary halide > Primary halide (D) Tertiary halide > Primary halide > Secondary halide
›Reveal solutionSolution
In SN1 reactions, the rate depends on carbocation stability, so tertiary halides react fastest, then secondary, then primary — the correct order is (C).
The key concept here is carbocation stability. An SN1 reaction proceeds via a two-step mechanism: first, the leaving group departs, forming a carbocation intermediate; then, the nucleophile attacks this carbocation. The rate-determining step is the first step — formation of the carbocation. Therefore, anything that stabilizes the carbocation speeds up the reaction. Alkyl groups stabilize carbocations through hyperconjugation and inductive effects, so the more substituted the carbocation, the more stable it is. This gives the familiar stability order: tertiary > secondary > primary > methyl.
- Identify the rate-determining step. In SN1, the slow step is the ionization of the alkyl halide to form a carbocation:
R−X→R++X−
The rate depends only on the concentration of the alkyl halide (first-order kinetics), and crucially, on how easily the carbocation forms.
-
Relate carbocation stability to reaction rate.
A more stable carbocation forms faster because the transition state leading to it is lower in energy (Hammond’s postulate: the transition state resembles the carbocation). So the order of reactivity for SN1 is exactly the order of carbocation stability.
-
Recall the stability order of carbocations.
- Tertiary carbocation: three alkyl groups donate electron density via hyperconjugation and inductive effects → most stable.
- Secondary carbocation: two alkyl groups → moderately stable.
- Primary carbocation: only one alkyl group → very unstable.
- Methyl carbocation: no alkyl groups → extremely unstable. Hence: tertiary > secondary > primary > methyl. …
- TG EAPCET 2023Set ap-2023-05-11-AN1 markMCQQ.What is the correct order of boiling points of the following alkyl halides? I. CH3−CH2−CH2−CH2−Cl II. CH3−CH2−CH2−CH2−Br III. CH3−CH2−CH(Br)−CH3 IV. (H3C)3CBr (A) II > III > IV > I (B) I > III > IV > II (C) II > IV > III > I (D) I > IV > III > II
›Reveal solutionSolution
Boiling point in alkyl halides increases with molecular mass (heavier halogen) and decreases with branching (weaker van der Waals forces). The correct order is II > III > IV > I.
The boiling point of an alkyl halide depends on two competing factors: molecular mass and molecular shape. Heavier molecules have stronger London dispersion forces, while branched molecules have smaller surface areas and weaker intermolecular contact.
When comparing alkyl halides, the halogen atom dominates the molecular mass because it is much heavier than the carbon skeleton. Bromine (Mr=80) is significantly heavier than chlorine (Mr=35.5), so bromides boil higher than chlorides of similar structure. Among isomers with the same halogen, branching reduces the boiling point because compact, spherical molecules have less surface contact than extended chains.
Let me identify each compound:
- I: CH3CH2CH2CH2Cl — 1-chlorobutane (straight chain, Cl)
- II: CH3CH2CH2CH2Br — 1-bromobutane (straight chain, Br)
- III: CH3CH2CH(Br)CH3 — 2-bromobutane (secondary, Br)
- IV: (CH3)3CBr — 2-bromo-2-methylpropane (tertiary, Br)
Now I'll rank them step by step:
-
Halogen effect dominates first: All three bromides (II, III, IV) will boil higher than the chloride (I), because bromine's greater mass and polarizability create stronger dispersion forces. So I is lowest.
-
Among the bromides, branching decides the order: All three have the same molecular formula for the bromobutanes (II and III are C4H9Br; IV is also C4H9Br).
- II is a straight chain (1-bromobutane): maximum surface area, strongest intermolecular forces. …
🎓Unlock everything free for 14 days
- ✓Full step-by-step solutions
- ✓Concept-first explanations
- ✓Methods, shortcuts & mistakes
- ✓PYQ mapping + timed mock tests
Full access for 14 days. No credit card required.