Q.Draw the possible resonance structures for CH3—O—CH2^+ (the oxygen carries two lone pairs and the terminal CH2 carbon bears a positive charge) and predict which of the structures is more stable. Give reason for your answer.
🔒You're viewing a preview — the full solution, concept, methods & PYQ mapping are locked.
🔒 Start your 14-day free trial to unlock the full solution →Concept understanding — Resonance Structures Drawing
Resonance Structures: What They Are and How to Draw Them
Let's start with a simple question. When you draw a molecule like ozone (O3), you might put a double bond between the central oxygen and one of the end oxygens, and a single bond to the other. But experiments show both O–O bonds are identical — same length, same strength. So which drawing is correct?
Neither single drawing is correct. The real molecule is a hybrid of both possibilities. That's the core idea of resonance.
The Intuition: A Musical Analogy
Think of a chord played on a piano. A C major chord is made of three notes: C, E, G. No single note is the chord — the chord is the blend of all three. Similarly, a resonance hybrid is the blend of all valid Lewis structures (called resonance contributors or canonical forms) for a molecule. The real molecule is not flipping between these forms; it exists as a single, stable average.
Resonance structures are not in equilibrium. The molecule does not switch from one form to another. It is a single structure that is the weighted average of all contributors.
The Precise Definition
Resonance structures are two or more Lewis structures that differ only in the placement of electrons (pi bonds and lone pairs), never in the positions of atoms. The real molecule is described by a resonance hybrid — a superposition of all contributors.
Rules for Valid Resonance Structures
- Atoms never move. Only electrons (pi bonds, lone pairs, and sometimes sigma bonds in special cases) change positions.
- The total number of electrons stays the same. You are just redistributing them.
- Each structure must obey the octet rule (for second-period elements) and have valid formal charges.
- All structures must have the same net charge and the same number of unpaired electrons (if any).
How to Draw Resonance Structures: A Step-by-Step Method
Let's use the carbonate ion (CO32−) as our example.
Step 1: Draw the best Lewis structure
Start with the skeleton: carbon in the center, three oxygens around it. Count valence electrons: C has 4, each O has 6, plus 2 for the charge = 4+18+2=24 electrons. Place bonds and lone pairs to satisfy octets. You'll get one structure with a C=O double bond and two C–O single bonds, each single-bonded oxygen carrying a negative charge.
Step 2: Identify movable electrons
Look for pi bonds (double or triple bonds) and lone pairs that are adjacent to pi bonds or to atoms with an empty p orbital. In carbonate, the C=O pi bond and the lone pairs on the negatively charged oxygens are the movable parts.
Step 3: Push electrons using curved arrows
An arrow starts at the electron source (a pi bond or lone pair) and points to where the electrons go (to form a new pi bond or to become a lone pair). In carbonate:
- Take the pi bond from C=O and push it to become a lone pair on that oxygen.
- Simultaneously, take a lone pair from a negatively charged oxygen and push it to form a new C=O pi bond.
Step 4: Draw the new structure
After pushing, you get a second structure where the double bond is on a different oxygen. Repeat to get the third structure (all three oxygens take turns being double-bonded).
Always check that the total number of electrons and the net charge remain unchanged after each arrow push. A common mistake is to accidentally add or remove electrons.
Common Patterns to Recognize
| Pattern | Example | What moves |
|---|---|---|
| Allylic system | CH2=CH−CH2+ | Pi bond shifts, positive charge moves |
| Conjugated diene | CH2=CH−CH=CH2 | Pi bonds shift (less common in neutral molecules) |
| Carbonyl group | R2C=O | Lone pair from O forms pi bond, pi bond becomes lone pair |
| Benzene ring | C6H6 | Alternating double bonds shift around the ring |
The Most Common Mistake Beginners Make
Breaking sigma bonds. Remember: sigma bonds (single bonds between atoms) never break in resonance. Only pi bonds and lone pairs move. If you find yourself moving an atom or breaking a single bond, you are drawing a different molecule (a constitutional isomer), not a resonance structure. …
The key idea is resonance delocalisation of the positive charge from carbon onto the
adjacent oxygen.
Structure A (given): CHX3−O⋅⋅−CHX2X+ — oxygen carries two lone
pairs and the positive charge sits on the terminal carbon (a primary carbocation, only
six electrons on that carbon).
Structure B (resonance): one oxygen lone pair forms a π bond to the CHX2X+
carbon, shifting the charge onto oxygen:
CHX3−O+=CHX2
Now every atom, including carbon, has a complete octet; the charge sits on the more
electronegative oxygen as an oxonium ion.
Stability: Structure B is the more stable (and dominant) contributor, because …
The positive charge on the oxygen-stabilised carbocation can be delocalised onto oxygen via a π-bond, giving a more stable oxonium ion structure. The resonance hybrid is dominated by the structure with a C=O double bond and a neutral oxygen.
Why resonance matters here
The given species is CH3−O−CH2+. The oxygen atom has two lone pairs, and the terminal CH2 carbon carries a positive charge. That positive carbon is directly attached to an oxygen that is rich in lone pairs — a classic setup for resonance stabilisation. The oxygen can "donate" one of its lone pairs to form a π-bond with the electron-deficient carbon, shifting the positive charge onto the oxygen itself. This delocalisation spreads the charge, making the ion more stable than a simple localised carbocation.
Drawing the resonance structures
-
Structure I (the given one)
CH3−∙∙O∙∙−CH2+
Oxygen has two lone pairs and is neutral. The terminal carbon bears a full positive charge. This is a primary carbocation — highly unstable on its own.
-
Structure II (the delocalised form)
CH3−O+=CH2
One lone pair from oxygen moves to form a π-bond between O and the CH2 carbon. Oxygen now has three bonds and a positive charge (oxonium ion). The CH2 carbon becomes neutral and has a complete octet. The methyl group remains unchanged.
-
No other significant resonance contributors
The methyl group’s C–H bonds are not in conjugation with the π-system, so hyperconjugation from the methyl group is a separate (weaker) effect, not a resonance structure. Only these two major structures matter.
Always check: does the movement of electrons create a new π-bond without exceeding the octet rule? Here, oxygen starts with two lone pairs (octet satisfied) and ends with one lone pair and a π-bond (still an octet). The carbon goes from a sextet to an octet. Both atoms obey the octet rule in both structures.
Stability comparison
Which structure is more stable? Apply the standard rules:
- Octet rule: Structure II has every atom (C, O, H) with a complete octet or duet. Structure I has a carbon with only six electrons — a major destabilising factor.
- Charge location: In Structure I, the positive charge is on a primary carbon (least stable carbocation type). In Structure II, the positive charge is on oxygen, which is more electronegative and can better accommodate a positive charge (though oxygen doesn't "like" a positive charge, it is still better than a carbon with a sextet). …
- AP EAPCET 2026Set eng-2026-05-18-FN1 markMCQQ.From the following list, identify the number of substituents which exert +R effect when present on benzene ring −Cl, −COCH3, −NHC2H5, −OCH3, −NHCOCH3, −COOCH3 (A) 5 (B) 6 (C) 3 (D) 4
›Reveal solutionSolution
A substituent shows +R effect if the atom attached to the ring has a lone pair to donate into it; of the six given groups, −Cl, −NHC2H5, −OCH3, −NHCOCH3 qualify (4 total), while −COCH3 and −COOCH3 are −R (carbonyl withdraws electron density).
Concept and Intuition
Resonance (R/mesomeric) effect on a benzene ring depends on whether the directly-attached atom can donate a lone pair into the ring (+R, activating by resonance) or whether the ring's π electrons are pulled into the substituent through a multiple bond (like C=O), which is −R (deactivating by resonance). Groups with an available lone pair on the ipso atom (halogens, −OR, −NR2, −NHCOR) are +R; groups where a π-bonded electronegative atom (as in C=O, C≡N) sits right next to the ring are −R.
Step-by-Step Solution
- −Cl: chlorine's lone pair conjugates into the ring (+R), even though its strong −I effect makes it net deactivating — it is still a +R group.
- −COCH3 (acetyl): the carbonyl carbon is attached directly to the ring; ring electrons delocalise into the C=O, pulling electron density away from the ring — this is −R, not +R.
- −NHC2H5: nitrogen's lone pair conjugates strongly into the ring, a classic strong +R donor (like −NH2).
- −OCH3: oxygen's lone pair conjugates into the ring — a classic +R donor. …
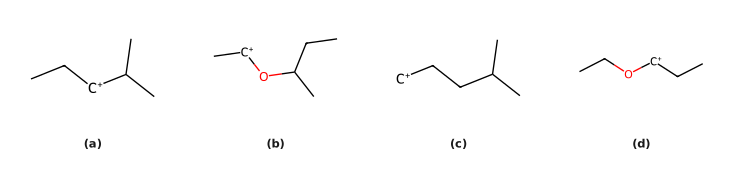
- AP EAPCET 2026Set eng-2026-05-18-AN1 markMCQQ.Consider the following carbocations [FIGURE] (four labelled carbocation skeletal structures:(a) a secondary carbocation CH3−CH2−C+H−CH(CH3)−CH3;(b) an ether-oxygen-stabilised cation CH3−C+H−O−CH(CH3)−CH2CH3;(c) a primary carbocation C+H2−CH2−CH2−CH(CH3)−CH3;(d) an ether-oxygen-stabilised cation CH3CH2−O−C+H−CH2CH3) The correct stability order for the above carbocations is (A)(b) >(a) >(d) >(c) (B)(b) >(d) >(c) >(a) (C)(d) >(b) >(c) >(a) (D)(d) >(b) >(a) >(c)
›Reveal solutionSolution
Resonance (mesomeric) donation from an adjacent ether oxygen stabilises a carbocation far more than simple alkyl hyperconjugation does, so both oxygen-adjacent cations rank above both plain alkyl cations; within the plain alkyl pair, the secondary beats the primary. Overall order: (d) > (b) > (a) > (c).
Concept and Intuition
Carbocation stability is governed, in decreasing order of strength, by: resonance/mesomeric (+M) donation > hyperconjugation/inductive (+I) donation from alkyl groups. An ether oxygen directly bonded to the electron-deficient carbon can donate a lone pair into the empty p-orbital, generating an oxocarbenium-type resonance structure (R–O+=CR2′) that delocalises the positive charge onto the (more electronegative but resonance-tolerant) oxygen. This resonance stabilisation is substantially stronger than the modest stabilisation a carbocation gets merely from being flanked by one extra alkyl group (hyperconjugation). Consequently, even a comparatively less-substituted carbocation that is directly bonded to oxygen outranks an ordinary alkyl carbocation that lacks such resonance support.
Step-by-Step Solution
- Classify each cation: (a) is a plain secondary alkyl carbocation (no heteroatom assistance); (c) is a plain primary alkyl carbocation (no heteroatom assistance); (b) and (d) both have the cationic carbon directly bonded to an ether oxygen.
- Because O's lone pair can donate by resonance into the empty orbital on both (b) and (d), both are oxocarbenium-stabilised and this resonance effect outweighs the plain hyperconjugative stabilisation available to (a) and (c). So {b,d}>{a,c}. …
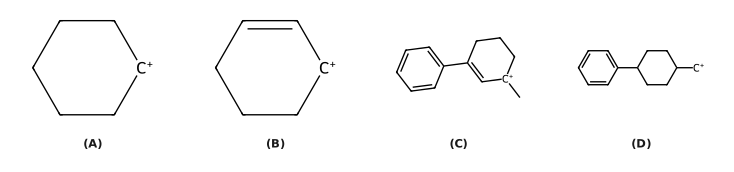
- AP EAPCET 2025Set eng-2025-05-26-AN1 markMCQQ.Identify the most stable carbocation from the following (A) [FIGURE] (a cyclohexyl cation — a six-membered ring with a positive charge on a ring carbon, no double bonds or substituents) (B) [FIGURE] (a cyclohexenyl cation — a six-membered ring with one C=C double bond, and a positive charge on the ring carbon adjacent to the double bond, i.e. allylic) (C) [FIGURE] (a cyclohexenyl ring bearing a methyl group on the cationic ring carbon and a phenyl (Ph) group on the far alkene carbon — allylic and benzylic conjugation together) (D) [FIGURE] (a cyclohexane ring with an exocyclic CH2+ group, and a phenyl (Ph) substituent on a ring carbon a few positions away — a primary benzylic-type cation)
›Reveal solutionSolution
Carbocation stability is set by how much positive charge can be delocalised by resonance (allylic/benzylic conjugation) plus hyperconjugation/induction from alkyl groups. The cation with BOTH allylic and benzylic conjugation, plus a methyl substituent, is the most stable.
Concept and Intuition
A carbocation is stabilised whenever its empty p-orbital can overlap with an adjacent π-system (resonance/conjugation) or with adjacent C–H/C–C sigma bonds (hyperconjugation), and destabilised when it sits isolated with no such support. Combining two independent resonance-donating groups (here, both an adjacent ring double bond AND a phenyl ring through that double bond) gives an extended conjugated system — much more stabilising than either alone.
Step-by-Step Solution
- Option (A): a cyclohexyl cation on a saturated ring, no adjacent π-bond, no aryl group — a simple secondary cation with only ordinary hyperconjugation. Least stabilised of the set.
- Option (B): a cyclohexenyl cation, cationic carbon directly next to the ring's C=C — this is a genuine allylic cation, delocalised over two carbons by resonance. More stable than (A), but only single-bond-worth of delocalisation.
- Option (D): the cationic carbon is an exocyclic CH2+ attached to the ring; the phenyl group sits on a different, non-adjacent ring carbon and is explicitly not conjugated with the cationic centre. So this cation behaves essentially like an isolated primary cation — very poorly stabilised despite having a phenyl group present on the molecule. …
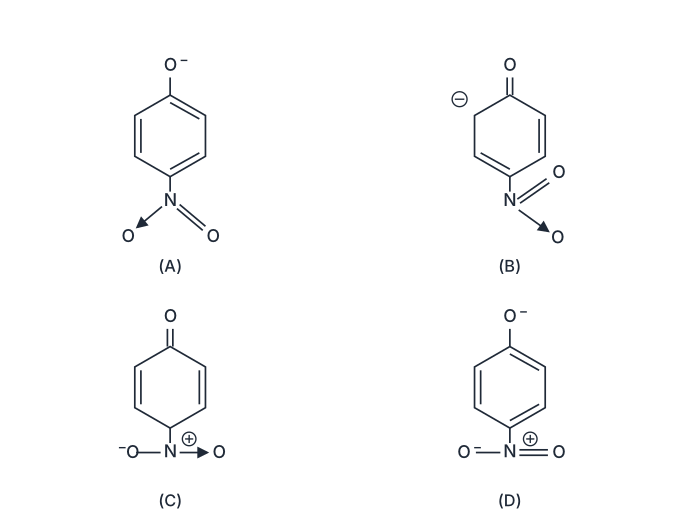
- AP EAPCET 2021Set ap-2021-10-05-FN1 markMCQQ.The most unlikely representation of resonance structure of p-nitro phenoxide is ________. (A) [FIGURE] (para-nitrophenoxide ring with the phenoxide oxygen drawn as O− at the top; the nitro group drawn with one N=O double bond and one N→O dative/coordinate bond, no formal charge shown on N) (B) [FIGURE] (a cyclohexadiene ring with a carbonyl C=O at the top and a carbanion, shown as a circled minus ⊖, on the ring carbon ortho to the carbonyl; the nitro group unchanged, drawn with a dative N→O bond) (C) [FIGURE] (a cyclohexadienone ring with a carbonyl C=O at the top; the nitro group drawn as a charge-separated form with N⊕ bonded to one O− and one O) (D) [FIGURE] (para-nitrophenoxide ring with the phenoxide oxygen drawn as O− at the top; the nitro group drawn as a charge-separated form with N⊕ bonded to two oxygen atoms)
›Reveal solutionSolution
A valid resonance structure of p-nitrophenoxide must pair an aromatic ring with an unperturbed nitro group, OR a quinonoid (C=O) ring with a charge-migrated nitro group — option (D) illegally mixes an untouched aromatic ring with an already charge-separated nitro group, which no single curved-arrow path can produce.
Concept and Intuition
In p-nitrophenoxide, the negative charge on the phenolic oxygen is stabilised by delocalising all the way to the nitro group through the ring in between ("push-pull" conjugation). Each legitimate resonance structure must differ from the next by moving exactly one pair of electrons at a time (one curved arrow, or a linked set), so the ring's bonding pattern and the nitro group's charge state must change together, in lock-step — never independently of each other.
Step-by-Step Solution
- Structure (A): aromatic ring (alternating double bonds) with O− at the top and the nitro group in its ordinary neutral-looking form (one N=O, one N→O dative bond). This is simply the reference/starting Lewis structure — a legitimate contributor.
- Pushing the phenoxide lone pair into the ring converts it to a quinonoid form: C=O appears at the ipso carbon, and the negative charge now sits as a carbanion on a ring carbon (ortho to the carbonyl), with the ring no longer aromatic — a legitimate intermediate contributor (structure B).
- Pushing that carbanion's electrons further, through the ring, into the nitro group converts the nitro group into its charge-separated form (N⊕ bonded to one O− and one O), while the ring stays quinonoid (C=O retained) — this is structure (C), a legitimate, fully-conjugated final contributor. …
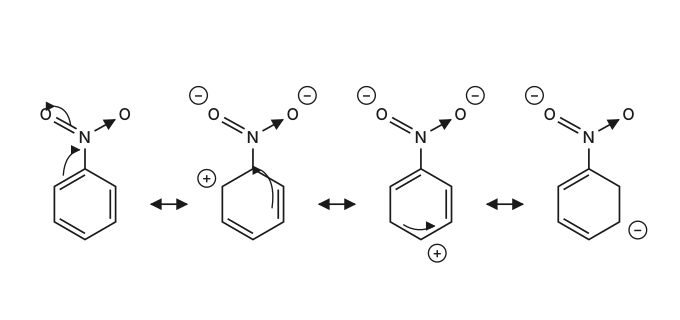
- AP EAPCET 2021Set eng-2021-08-23-AN1 markMCQQ.The electron transfer in the following conjugated system shows ________ [FIGURE] (four resonance structures of nitrobenzene drawn left to right with curved-arrow electron-pushing: structure 1 shows the neutral nitro group (N double-bonded to two O atoms) attached to the benzene ring with a curved arrow pushing electron density from the ring into the N=O bond; structure 2 shows one O bearing a negative charge, the N=O retained on the other oxygen, and a positive charge on the ring carbon ortho to the point of attachment, with a curved arrow moving around the ring; structure 3 is analogous but the positive charge is on the ring carbon para to the point of attachment (shown at the bottom of the ring), with a curved arrow continuing the conjugation; structure 4 shows both oxygens bearing negative charges (one shown, drawn on the left O) and a positive charge on the ring carbon ortho to the point of attachment on the other side) (A) −R effect (B) −I effect (C) +R effect (D) +I effect
›Reveal solutionSolution
The curved arrows push π-electron density from the ring toward the nitro group, generating positive charge on ortho/para ring carbons — this is the −R (electron-withdrawing resonance) effect of −NO2.
Concept and Intuition
The resonance (mesomeric) effect describes delocalisation of π-electrons through a conjugated system. A substituent can either donate electron density into the ring by resonance (+R, e.g. −NH2, −OH, halogens) or withdraw electron density from the ring by resonance (−R, e.g. −NO2, −CHO, −COOH, −CN). The direction of electron flow in the canonical structures tells you which type of effect is operating: if the ring becomes electron-poor (positive charges appear on ring carbons) as electrons flow toward the substituent, that substituent is exerting a −R effect.
Step-by-Step Solution
- In structure 1, the nitro group is neutral; curved arrows show the ring's π-electrons beginning to shift toward the nitrogen–oxygen system.
- In structures 2 and 3, this electron shift has generated a formal positive charge on the ring carbon ortho (structure 2) and para (structure 3) to the point of attachment of −NO2, while the oxygens of the nitro group pick up negative charge.
- This is precisely the pattern of a group withdrawing electron density from the ring via conjugation/resonance — electrons flow away from the ring carbons and into the substituent, leaving positive charge behind on the ring. …
- AP EAPCET 2021Set eng-2021-10-05-FN1 markMCQQ.The relatively more electron rich sites of the following aromatic compounds are ______ [FIGURE] Two benzene rings with numbered ring positions 1-6:(i) a ring with OH at C1 and CH3 at C3 (m-cresol/3-methylphenol), positions labelled 1(OH),2,3(CH3),4,5,6;(ii) a ring with OH at C1 and −CO−CH3 at C4 (4-hydroxyacetophenone), positions labelled 1(OH),2,3,4(COCH3),5,6 (A)(i) 2,5 &(ii) 2,3 (B)(i) 4,5 &(ii) 3,5 (C)(i) 2,4,6 &(ii) 2,6 (D)(i) 2,5,6 &(ii) 2,6
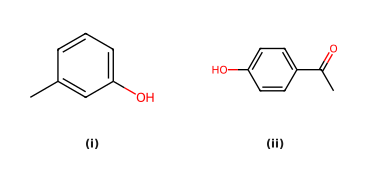
›Reveal solutionSolution
The most electron-rich sites are determined by the strongest activating substituent (OH) via resonance; for (i) the OH directs to ortho/para positions (2,4,6), and for (ii) the para-acetyl group deactivates position 4, so the OH still activates ortho/para but the para position is blocked, leaving 2 and 6 as the most electron-rich.
The key idea is that resonance effects dominate over inductive effects for strongly activating groups like –OH. The –OH group donates electron density into the ring through resonance, making the ortho and para positions (relative to itself) the most electron-rich. We must then consider whether the second substituent (CH₃ or –COCH₃) reinforces or opposes this pattern.
Step-by-step reasoning
1. Identify the activating/deactivating nature of each substituent.
- –OH (hydroxyl): Strongly activating, ortho/para-directing. Its lone pair on oxygen can delocalize into the ring, creating resonance structures that place negative charge on the ortho and para carbons.
- –CH₃ (methyl): Weakly activating, ortho/para-directing. It donates electron density via hyperconjugation and inductive effect, but much weaker than –OH.
- –COCH₃ (acetyl): Strongly deactivating, meta-directing. The carbonyl group withdraws electron density by resonance, making the ring less electron-rich overall, especially at ortho and para positions relative to itself.
2. Analyze compound (i): 3-methylphenol (OH at C1, CH₃ at C3).
- The –OH group is the dominant activator. Its resonance donation makes positions 2, 4, and 6 (ortho and para to OH) the most electron-rich.
- The –CH₃ at C3 is a weak activator and also directs ortho/para, but its effect is much smaller. It does not block any of the OH-activated positions.
- Therefore, the most electron-rich sites are C2, C4, and C6.
TipWhen two ortho/para-directors are present, the stronger one (OH) determines the primary pattern; the weaker one (CH₃) only slightly modifies the electron density.
3. Analyze compound (ii): 4-hydroxyacetophenone (OH at C1, –COCH₃ at C4).
- The –OH group again strongly activates ortho/para positions: C2, C4, C6.
- However, the –COCH₃ group at C4 is a strong deactivator. It withdraws electron density from the ring, especially from the ortho and para positions relative to itself. Since C4 is para to OH, it is also the carbon bearing the acetyl group. …
🎓Unlock everything free for 14 days
- ✓Full step-by-step solutions
- ✓Concept-first explanations
- ✓Methods, shortcuts & mistakes
- ✓PYQ mapping + timed mock tests
Full access for 14 days. No credit card required.