Q.Explain the fact that in aryl alkyl ethers
🔒You're viewing a preview — the full solution, concept, methods & PYQ mapping are locked.
🔒 Start your 14-day free trial to unlock the full solution →Concept understanding — Electrophilic Aromatic Substitution
Electrophilic Aromatic Substitution – The First Meeting
Imagine you have a benzene ring — that perfect, flat hexagon of six carbons with alternating double bonds. It's stable, almost stubbornly so. You want to attach something new to it, say a bromine atom or a nitro group. But benzene doesn't react like an alkene. It doesn't just add across a double bond. Instead, it does something more elegant: it kicks out a hydrogen and keeps its aromatic ring intact.
That's the heart of Electrophilic Aromatic Substitution (EAS).
The Intuition: Why "Substitution" and Not "Addition"?
Benzene's stability comes from its delocalised π electrons — a cloud above and below the ring. This cloud is electron-rich, so it attracts electrophiles (electron-loving species). But if an electrophile simply added to a double bond, the ring would break its aromaticity, losing that huge stabilisation. That would be energetically costly.
So benzene does something smarter: it lets the electrophile attack, temporarily breaks aromaticity to form a high-energy intermediate (the arenium ion), and then loses a proton to restore the aromatic ring. The net result? A hydrogen is replaced by the electrophile. The ring is back to its stable, aromatic self.
The key trade-off: temporary loss of aromaticity is acceptable because the final product regains it. Addition reactions would permanently destroy aromaticity — benzene avoids that.
The Precise Statement
Electrophilic Aromatic Substitution is a reaction in which an electrophile (E+) replaces a hydrogen atom on an aromatic ring, proceeding through a sigma complex (arenium ion) intermediate, and restoring aromaticity after deprotonation.
The general equation:
Ar−H+EX+Ar−E+HX+
where Ar represents an aromatic ring.
The Mechanism in Three Steps
-
Generation of the electrophile – Many EAS reactions need a catalyst to create a strong enough E+. For example, bromination uses FeBrX3 to polarise BrX2 into BrX+.
-
Attack by the aromatic ring – The π electrons of benzene attack the electrophile, forming a sigma complex (also called the arenium ion or Wheland intermediate). This intermediate is non-aromatic — it has four π electrons delocalised over five carbons, and one sp3 carbon bearing the electrophile and a hydrogen.
Benzene+E+⟶Sigma complex (non-aromatic)
- Deprotonation – A base (often the counterion of the catalyst, like FeBrX4X−) removes the proton from the sp3 carbon. The pair of electrons from the C–H bond flows back into the ring, restoring the aromatic sextet.
Sigma complex+Base⟶Product+HX+
The sigma complex is not aromatic. It's a high-energy intermediate. Students often mistakenly think it's still aromatic — it isn't. That's why the step is fast and the complex is short-lived.
Why This Matters for Exams
EAS is the gateway to understanding how to put groups onto benzene rings. The rate-determining step is usually the formation of the sigma complex (step 2). The regiochemistry (where the electrophile goes) depends on whether the ring already has a substituent — that's the topic of activating/deactivating groups and ortho/para vs. meta directors.
But for now, remember this: …
Why this formula?
Electrophilic Aromatic Substitution: Why the Mechanism Holds
The Core Puzzle: Why Benzene Doesn't Just Add
Benzene (C6H6) has three double bonds — so why doesn't it undergo addition reactions like alkenes?
The answer lies in aromatic stabilisation: benzene's delocalised π-electron cloud (the "aromatic sextet") is about 150 kJ/mol more stable than a hypothetical cyclohexatriene with localised double bonds.
If benzene simply added an electrophile (like Br2), it would lose this stabilisation — a huge energy penalty.
So, nature chooses a different path: substitution instead of addition, preserving the aromatic ring.
The Key Formula: The Reaction Profile
The rate-determining step in electrophilic aromatic substitution (EAS) is the formation of the arenium ion (σ-complex):
Ar-H+E+slowAr-E+H(σ-complex)
Then, fast deprotonation restores aromaticity:
Ar-E+H+B−fastAr-E+BH
Why This Holds: The Energy Barrier Logic
-
First step (slow): The electrophile E+ attacks the electron-rich ring. The σ-complex is non-aromatic — it has only 4 π-electrons delocalised over 5 carbons (the sixth carbon is sp3 hybridised). This intermediate is higher in energy than the starting benzene.
-
Second step (fast): A base removes the proton, restoring the aromatic sextet. This step is strongly exothermic — the system regains ~150 kJ/mol of stabilisation.
The overall reaction is exothermic, but the activation energy is dominated by the destabilisation of the σ-complex.
The Rate Law: Why It's First Order in Both
From the mechanism:
Rate=k[Arene][E+]
Reasoning:
- The slow step involves one molecule of arene and one molecule of electrophile.
- No other species appear before the rate-determining step.
- Therefore, the rate law is bimolecular — first order in each reactant.
This is not derived from the overall stoichiometry — it comes directly from the molecularity of the slow step.
The Hammett Equation: Quantifying Substituent Effects
For substituted benzenes, the rate constant k relative to benzene (k0) follows:
logk0k=σρ
Why This Holds
- σ (sigma constant): Measures the electronic effect of a substituent (electron-donating or withdrawing) relative to hydrogen. It is derived from the ionisation constants of benzoic acids — a purely empirical scale.
- ρ (rho constant): Measures the sensitivity of the reaction to substituent effects. A positive ρ means the reaction is favoured by electron-withdrawing groups (rare in EAS); a negative ρ means electron-donating groups accelerate the reaction.
Why it works:
The σ-complex has a positive charge delocalised over the ring. Substituents that stabilise this positive charge (electron-donating groups like −OH, −NH2) lower the activation energy — hence σ is negative for such groups. Electron-withdrawing groups (−NO2, −CN) destabilise the σ-complex — σ is positive.
The linear free-energy relationship holds because the transition state resembles the σ-complex in charge distribution.
The Directing Effect: Why Ortho/Para vs Meta
The position of substitution is governed by the stability of the σ-complex for each possible attack site. …
The key idea is Electrophilic Aromatic Substitution (EAS) and the resonance effect of the alkoxy (−OR) group.
Reasoning:
- The oxygen atom in the alkoxy group (−OR) has two lone pairs of electrons. These lone pairs are in conjugation with the π-electrons of the benzene ring.
- This conjugation allows the oxygen to donate electron density into the ring via resonance, creating a +R effect (resonance effect). This makes the ring more electron-rich than benzene.
- A more electron-rich ring is more attractive to an electrophile, hence the ring is activated (i) towards EAS. …
The alkoxy group (−OR) in aryl alkyl ethers is a strong activating and ortho/para-directing group because the oxygen atom donates electron density into the benzene ring through resonance, making the ring more nucleophilic at the ortho and para positions.
The Concept: Electrophilic Aromatic Substitution (EAS)
Electrophilic aromatic substitution is the fundamental reaction where an electrophile (an electron-loving species) replaces a hydrogen atom on an aromatic ring. For this to happen, the ring must be electron-rich enough to attract and stabilise the incoming positive charge.
The key question is: How does a substituent already on the ring affect this process? Substituents are classified as either activating (increase the reaction rate) or deactivating (decrease it), and as ortho/para-directing or meta-directing. The alkoxy group (−OCH3, −OC2H5, etc.) is one of the strongest activating groups and a classic ortho/para director.
Why the Alkoxy Group Activates the Ring
The oxygen atom in the −OR group has two lone pairs of electrons. These lone pairs can participate in resonance with the benzene ring. This is the single most important reason for both activation and direction.
-
Resonance donation: The lone pair on oxygen can be delocalised into the π-electron system of the benzene ring. This creates additional resonance structures where the negative charge (or rather, increased electron density) is placed specifically on the ortho and para carbon atoms.
The resonance hybrid of anisole (C6H5OCH3) shows partial negative charges on the ortho and para positions:
Resonance structures: \chemfig∗6(−=−(−OCH3)=−=)→\chemfig∗6(−[:30](−[:90]OCH3)−[:150]=−[:210](−[:270]⊖)−[:330]=−)
-
Increased electron density: Because of this resonance, the ortho and para positions become significantly more electron-rich (nucleophilic) than the meta position. The ring as a whole becomes more electron-rich than benzene itself. An electrophile is therefore more strongly attracted to the ring, and the reaction proceeds faster — hence activation.
-
Inductive effect: Oxygen is electronegative, so it pulls electron density away from the ring through the sigma bond (inductive effect). This is a deactivating effect. However, the resonance effect is much stronger and dominates, so the net result is strong activation.
A common mistake is to think that because oxygen is electronegative, the −OR group must be deactivating. This is wrong. The resonance effect (electron donation) far outweighs the inductive effect (electron withdrawal) for alkoxy groups. Always check resonance first.
Why It Directs to Ortho and Para Positions
The directing effect is a direct consequence of the resonance stabilisation of the intermediate carbocation (the Wheland intermediate or arenium ion) formed during EAS.
When an electrophile (E+) attacks the ring, a positively charged intermediate is formed. The stability of this intermediate determines which position is attacked most easily.
-
Attack at ortho or para: If the electrophile attacks at the ortho or para position, the positive charge in the intermediate can be delocalised onto the oxygen atom (via resonance). This gives an extra, highly stable resonance structure where the oxygen bears the positive charge (a tertiary oxonium ion). This is a very stable arrangement because oxygen is happy to share its lone pair.
TipThink of it this way: the oxygen atom acts like a "safety net" for the positive charge. If the electrophile hits ortho or para, the charge can be "passed" to the oxygen, which stabilises it beautifully. If it hits meta, the oxygen cannot help — the charge stays on the ring.
-
Attack at meta: If the electrophile attacks at the meta position, the positive charge cannot be delocalised onto the oxygen. The intermediate is less stable because the positive charge remains on the ring carbons without the extra resonance stabilisation from oxygen. …
Method: Resonance Effect Analysis in Electrophilic Aromatic Substitution
This method explains activation and directing ability by examining how the substituent stabilises the arenium ion intermediate through resonance.
Step 1: Identify the substituent and its lone pairs
In an aryl alkyl ether (e.g., methoxybenzene, C6H5OCH3), the alkoxy group (−OR) has two lone pairs of electrons on the oxygen atom. These are available for delocalisation into the benzene ring.
Step 2: Draw resonance structures showing electron donation
The oxygen donates electron density into the ring via resonance. The key resonance forms are:
- The lone pair on oxygen forms a double bond with the ring carbon, creating a negative charge at the ortho and para positions.
- The meta position never gets a negative charge in any resonance structure.
Result: The ring becomes electron-rich, especially at ortho and para carbons.
Step 3: Explain activation (why the ring is more reactive)
- The increased electron density on the ring makes it more nucleophilic.
- An electrophile (e.g., NO2+, Br+) is attracted more strongly to this electron-rich ring.
- The arenium ion intermediate formed during substitution is stabilised by the oxygen’s lone pair (through resonance), lowering the activation energy.
Conclusion: The alkoxy group is an activating group — it speeds up electrophilic substitution compared to benzene.
Step 4: Explain ortho/para direction (why substitution occurs at those positions)
When the electrophile attacks at the ortho or para position, the positive charge in the arenium ion can be delocalised onto the oxygen atom (a very stable contributor). This extra stabilisation is not possible if attack occurs at the meta position. …
Here is a breakdown of the common mistakes students make regarding the activating and directing effects of the alkoxy group (−OR) in aryl alkyl ethers during Electrophilic Aromatic Substitution (EAS), along with how to avoid them.
Mistake 1: Confusing the Inductive Effect (−I) with the Overall Effect
The Mistake:
Students see the electronegative oxygen atom (e.g., in methoxybenzene, anisole) and assume it must be deactivating and meta-directing because it "pulls" electron density away via the inductive effect (−I). They then get the answer wrong.
Why it’s wrong:
While the inductive effect is indeed electron-withdrawing (−I), the resonance effect (+M or +R) is much stronger and dominates the overall behavior.
How to Avoid:
- Always draw the resonance structures. The lone pair on the oxygen atom is conjugated with the π-system of the benzene ring. This delocalization puts a negative charge (high electron density) specifically on the ortho and para positions.
- Remember the hierarchy: For activating groups with lone pairs (−OH, −OR, −NH2), the +M effect > −I effect. The net result is activation of the ring.
- Key fact: The ring becomes more electron-rich than benzene itself, making it a faster target for the electrophile (E+).
Mistake 2: Forgetting the "Ortho/Para" Specificity
The Mistake:
Students correctly identify that the group is activating, but they incorrectly state it directs to meta positions, or they say it directs to "all positions equally."
Why it’s wrong:
The resonance structures that show the negative charge only place that charge on the ortho and para carbons. The meta carbon never gets a negative charge from resonance.
How to Avoid:
- Trace the resonance: Draw the curved arrows from the oxygen lone pair into the ring. You will see the π-bond shifts. The only carbons that become negatively charged (and thus most attractive to E+) are the two ortho carbons and the one para carbon.
- Memorize the pattern: Groups with a lone pair on the atom directly attached to the ring (like −OCH3, −OH, −NH2) are ortho/para directing.
- Contrast with deactivators: Groups with a positive charge or a π-bond to a more electronegative atom (like −NO2, −CN, −CHO) are meta directing because their resonance pulls electron density away from ortho/para.
Mistake 3: Ignoring Steric Hindrance at the Ortho Position
The Mistake:
Students assume that because the ortho position is activated, it will always be the major product. They forget that the alkoxy group (−OR) is bulky.
Why it’s wrong:
While both ortho and para positions are activated, the ortho position is sterically hindered by the large alkyl group (R) attached to the oxygen. A bulky electrophile will struggle to attack the ortho carbon.
How to Avoid:
- Check the size of the substituent: The −OCH3 group is not tiny. The alkyl chain (R) sticks out.
- Predict the major product: In most cases, the para product is the major product because it is both electronically favored and sterically unhindered. The ortho product is a minor product.
- Example: Nitration of anisole gives mostly p-nitroanisole and some o-nitroanisole, but very little m-nitroanisole.
Mistake 4: Confusing "Activation" with "Reactivity Order"
The Mistake: …
Showing the 12 most recent of 23 on this concept.
- AP EAPCET 2026Set eng-2026-05-12-AN1 markMCQQ.Chlorobenzene (A) on nitration gave B as major product. Reaction of B with NaOH at 443 K followed by acidification formed C. In another reaction of A with CH3COCl / anhy. AlCl3 gave D (major product). What are C and D respectively? (A) 4-chlorophenol (p-Cl-C6H4-OH) ; 4-chloroacetophenone (p-Cl-C6H4-COCH3) (B) 4-nitrophenol (p-O2N-C6H4-OH) ; 4-chloroacetophenone (p-Cl-C6H4-COCH3) (C) 4-chlorophenol (p-Cl-C6H4-OH) ; 3-chloroacetophenone (m-Cl-C6H4-COCH3) (D) 4-nitrophenol (p-O2N-C6H4-OH) ; 3-chloroacetophenone (m-Cl-C6H4-COCH3)
›Reveal solutionSolution
B (from nitration) = p-chloronitrobenzene; hydrolysing its (activated) C–Cl with hot NaOH then acidifying gives C = 4-nitrophenol; separately, Friedel–Crafts acylation of chlorobenzene gives the para product D = 4-chloroacetophenone.
Concept and Intuition
Two independent transformations of the same starting material, chlorobenzene: (i) nitration followed by an activated nucleophilic aromatic substitution of Cl by OH (only possible because the ring is now strongly deactivated/activated-for-SNAr by an ortho/para nitro group), and (ii) a direct Friedel–Crafts acylation, both controlled by the o,p-directing, weakly deactivating nature of Cl, with sterics favouring the para product in each case.
Step-by-Step Solution
- Chlorobenzene nitration: Cl is o,p-directing (weak deactivator); despite modest steric differences, the major product is the para isomer, B = 1-chloro-4-nitrobenzene.
- Ordinary (unactivated) chlorobenzene needs extremely harsh conditions (Dow process: ~623 K, 300 atm) to convert Cl to OH. But once a strongly electron-withdrawing NO2 group sits ortho/para to the Cl, it stabilises the Meisenheimer intermediate of nucleophilic aromatic substitution, so B reacts with NaOH at a much milder 443 K to displace Cl by OH, giving (after acidifying the phenoxide) C = 4-nitrophenol (p-O2N-C6H4-OH). …
- AP EAPCET 2026Set eng-2026-05-12-AN1 markMCQQ.What are X and Y respectively in the following set of reactions? Anisole Br2CH3CO2H X (major product) Toluene (i) Br2 ∣ UV Light(ii) OH− Y (major product) (A) C6H5−O−CH2CH2Br ; C6H5−CH(OH)−CH3 (B) 4-bromoanisole (p-Br-C6H4-OCH3) ; benzyl alcohol (C6H5−CH2OH) (C) C6H5−O−CH(Br)−CH3 ; benzyl alcohol (C6H5−CH2OH) (D) 2-bromoanisole (o-Br-C6H4-OCH3) ; p-cresol (p-HO-C6H4-CH3)
›Reveal solutionSolution
Anisole undergoes ring (electrophilic) bromination to give mainly para-bromoanisole; toluene under UV light undergoes side-chain (radical) bromination at the benzylic carbon, and hydrolysis of that benzylic bromide gives benzyl alcohol.
Concept and Intuition
The conditions decide whether bromination happens on the ring or on the side chain: Br2 in acetic acid (no light) with an activated ring (anisole) is classic electrophilic aromatic substitution; Br2 with UV light on an alkyl-substituted arene (toluene) instead generates bromine radicals that abstract the weak benzylic C–H, giving side-chain substitution.
Step-by-Step Solution
- Anisole: the −OCH3 group is a powerful o,p-activator via resonance donation of a lone pair into the ring. With molecular Br2 in acetic acid (ionic mechanism, electrophilic aromatic substitution), bromination occurs mainly at the position para to −OCH3 (steric hindrance disfavours ortho): X = 4-bromoanisole (p-Br-C6H4-OCH3).
- Toluene + Br2/UV light: UV light homolyses Br2 into radicals; the benzylic C–H bond of the methyl group is weak and gives a resonance-stabilised benzylic radical, so substitution happens on the side chain, not the ring, giving benzyl bromide (C6H5CH2Br). …
- AP EAPCET 2026Set eng-2026-05-14-AN1 markMCQQ.What are X and Y respectively in the following reaction sequence? (Me=−CH3) Z+Y (Benzene derivative)HBrΔAnisoleCH3Clanhy. AlCl3X (major product) (A) 3-Methylanisole (–OMe and –Me in meta positions) ; Bromobenzene (B) 2-Methylanisole (–OMe and –Me in ortho positions) ; Phenol (C) 4-Methylanisole (–OMe and –Me in para positions) ; Bromobenzene (D) 4-Methylanisole (–OMe and –Me in para positions) ; Phenol
›Reveal solutionSolution
Friedel–Crafts methylation of anisole gives predominantly the para product (X = 4-methylanisole); HBr cleaves anisole's ether linkage at the methyl carbon to give phenol (Y) and methyl bromide (Z).
Concept and Intuition
−OCH3 on benzene is a strong activating, ortho/para-directing group. In Friedel–Crafts alkylation, both ortho and para products form, but the para isomer is favoured as the major product because of steric hindrance to ortho attack from the existing −OCH3 group.
Separately, aryl alkyl ethers like anisole are cleaved by strong acids such as HBr/HI on heating. Because the Caryl−O bond cannot easily undergo SN2 (aryl carbons resist backside attack, and aryl cations are high energy), the nucleophile (Br−) instead attacks the methyl carbon (SN2 on CH3), breaking the O−CH3 bond. This releases the phenolic −OH intact on the ring (giving phenol) and generates methyl bromide.
Step-by-Step Solution
- Anisole + CH3Cl, anhydrous AlCl3 → Friedel–Crafts alkylation; −OMe directs ortho/para; steric bulk favours para as the major product → X = 4-methylanisole (OMe, Me para).
- Anisole + HBr, Δ → ether cleavage; Br− attacks the less hindered methyl carbon (SN2) → products are CH3Br (Z) and phenol (Y). …
- AP EAPCET 2025Set eng-2025-05-21-AN1 markMCQQ.Which of the following does not involve in Friedel-Craft reaction? (A) C6H5OCH3 (anisole, drawn as a benzene ring with an −OCH3 substituent) (B) C6H5NH2 (aniline, drawn as a benzene ring with an −NH2 substituent) (C) C6H5Cl (chlorobenzene, drawn as a benzene ring with a −Cl substituent) (D) C6H6 (benzene, drawn as a plain benzene ring)
›Reveal solutionSolution
Aniline is the classic exception that fails to undergo Friedel–Crafts reactions, because its −NH2 lone pair complexes with the AlCl3 catalyst, killing both the catalyst's activity and deactivating the ring.
Concept and Intuition
Friedel–Crafts alkylation/acylation needs a Lewis-acid catalyst (commonly anhydrous AlCl3) to generate the electrophile. This only works smoothly on arenes whose ring is not strongly deactivated and whose substituents don't sequester the catalyst. Aniline's amino group is a strong Lewis base — its nitrogen lone pair readily donates into AlCl3's empty orbital, forming a stable acid–base adduct. This (a) uses up the catalyst so it can't generate the required electrophile, and (b) converts the ring substituent into an electron-withdrawing −NH2→AlCl3 group (rather than the strongly activating free −NH2), strongly deactivating the ring toward electrophilic attack. As a result, aniline famously fails to give normal Friedel–Crafts products.
Step-by-Step Solution
- Anisole (C6H5OCH3): the ether oxygen is a much weaker Lewis base than an amine nitrogen and still activates the ring strongly (o,p-director); anisole undergoes Friedel–Crafts reactions normally.
- Aniline (C6H5NH2): the amine nitrogen's lone pair is a strong Lewis base, complexing irreversibly with AlCl3, disabling the catalyst and deactivating the ring — Friedel–Crafts fails. …
- AP EAPCET 2025Set eng-2025-05-23-AN1 markMCQQ.Consider the following Statement-I: In the nitration of aniline, more amount of m-nitroaniline is formed than expected. Statement-II: In the presence of a strongly acidic medium, aniline is protonated to form anilinium ion, which is meta directing. The correct answer is (A) Both statement-I and statement-II are correct (B) Both statement-I and statement-II are not correct (C) Statement-I is correct, but statement-II is not correct (D) Statement-I is not correct, but statement-II is correct
›Reveal solutionSolution
Protonation of aniline to anilinium ion under strongly acidic nitration conditions explains the unexpectedly high m-nitroaniline yield; both statements are true. Answer: (A).
Concept and Intuition
Aniline's −NH2 group is a lone-pair donor and is normally a strong ortho/para director via resonance. However, nitration is carried out in strongly acidic media (conc. HNO3/conc. H2SO4), and aniline is a reasonably basic amine — a substantial fraction of it is protonated under these conditions to the anilinium ion, C6H5NH3+. The −NH3+ group has no lone pair available to donate into the ring (it is used in the N–H bonds/protonation) and instead withdraws electron density inductively, behaving like other positively-charged substituents (−NR3+) as a deactivating, meta-directing group. So nitration of aniline under these conditions is really a mixture of two competing pathways — direct nitration of the small residual free amine (ortho/para-directing) and nitration of the dominant anilinium ion (meta-directing) — and the meta contribution pushes the overall m-nitroaniline yield well above what pure amine-directed reasoning would predict.
Step-by-Step Solution
- Aniline + conc. HNO3/conc. H2SO4: strongly acidic medium protonates much of the aniline to anilinium ion.
- Anilinium ion (−NH3+) is electron-withdrawing and meta-directing, unlike the free amine.
- Nitration therefore proceeds partly through the anilinium ion pathway, contributing meta product. …
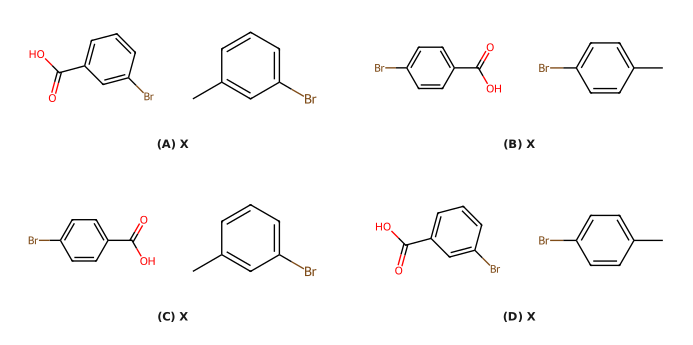
- AP EAPCET 2025Set eng-2025-05-26-AN1 markMCQQ.What are the major products X and Y respectively in the following set of reactions? (dark = dark) [FIGURE] (a benzene ring with COOH, reacting with Br2/Fe(dark) to give X) [FIGURE] (a benzene ring with CH3, reacting with Br2/Fe(dark) to give Y) (A) [FIGURE] (X = a benzene ring with COOH and Br meta to each other; Y = a benzene ring with CH3 and Br meta to each other) (B) [FIGURE] (X = a benzene ring with COOH and Br para to each other; Y = a benzene ring with CH3 and Br para to each other) (C) [FIGURE] (X = a benzene ring with COOH and Br para to each other; Y = a benzene ring with CH3 and Br meta to each other) (D) [FIGURE] (X = a benzene ring with COOH and Br meta to each other; Y = a benzene ring with CH3 and Br para to each other)
›Reveal solutionSolution
COOH (meta director) sends Br to the meta position in benzoic acid; CH3 (ortho/para director) sends Br predominantly to the para position in toluene — so X is meta-substituted and Y is para-substituted.
Concept and Intuition
Both reactions are electrophilic aromatic bromination catalysed by Fe (which generates the electrophile Br+ from Br2); "dark" conditions rule out the alternative photochemical free-radical side-chain bromination pathway, confirming these are ring substitutions. The regiochemistry is governed entirely by the directing effect of the substituent already on the ring:
- −COOH is strongly electron-withdrawing (by resonance and induction), so it deactivates the ring but directs the incoming electrophile to the meta position (the position where the resulting arenium-ion intermediate avoids placing positive charge on the carbon bearing the electron-poor COOH group).
- −CH3 is electron-donating (hyperconjugation/+I), activating the ring and directing ortho/para; because Br is a moderately bulky electrophile, steric hindrance disfavours the ortho position relative to the CH3 group, so the para isomer is the major product.
Step-by-Step Solution
- Benzoic acid + Br2/Fe: COOH is a meta director ⇒ Br enters meta to COOH, giving X = 3-bromobenzoic acid (meta relationship between COOH and Br). …
- AP EAPCET 2025Set eng-2025-05-26-AN1 markMCQQ.Which of the following will undergo methylation with CH3Cl / anhy. AlCl3 ? a) Aniline b) Chlorobenzene c) Benzoic acid d) Anisole (A) a, d (B) b, c (C) b, d (D) a, c
›Reveal solutionSolution
Friedel-Crafts alkylation with CH3Cl/AlCl3 works on chlorobenzene (weakly deactivated) and anisole (strongly activated) but fails on aniline (N lone pair complexes AlCl3) and benzoic acid (too strongly deactivated).
Concept and Intuition
Friedel-Crafts reactions need the aromatic ring to be nucleophilic enough to attack the AlCl3-generated electrophile (+CH3 here). Two situations cause the reaction to fail:
- Deactivation of the AlCl3 catalyst itself: groups with a lone pair that is basic enough to coordinate directly with the Lewis acid — most notably −NH2, −NHR, −NHCOR — tie up AlCl3 as a salt/complex, leaving none to generate the electrophile. Aniline is the textbook example of this failure mode.
- Strong deactivation of the ring itself: strongly electron-withdrawing meta-directing groups (COOH, NO2, CN, SO3H, etc.) make the ring too electron-poor for the (only moderately electrophilic) Friedel-Crafts species to attack. Benzoic acid falls in this category.
Halogens (as in chlorobenzene) are only weakly deactivating (net effect: deactivating overall by induction, but still ortho/para-directing by resonance donation of lone pairs) — the ring remains nucleophilic enough for alkylation to proceed. Ethers like anisole (−OCH3) are strongly activating (electron-donating by resonance) and undergo Friedel-Crafts alkylation readily.
Step-by-Step Solution
- Aniline (a): −NH2 lone pair coordinates with AlCl3, forming a complex that both consumes the catalyst and makes the ring effectively bear a positively-charged, strongly deactivating −NH2+AlCl3−-like group ⇒ Friedel-Crafts fails. …
- AP EAPCET 2024Set ap-2024-05-17-AN1 markMCQQ.What are X and Y respectively in the following reactions? C6H5CH3XC6H5CHONitrationY (A) X = KMnO4/OH− ; Y = 3-nitrobenzaldehyde (m-O2N-C6H4-CHO) (B) X =(i) CrO2Cl2,(ii) H3O+ ; Y = 3-nitrobenzaldehyde (m-O2N-C6H4-CHO) (C) X =(i) CrO2Cl2,(ii) H3O+ ; Y = 4-nitrobenzaldehyde (p-O2N-C6H4-CHO) (D) X = KMnO4/OH− ; Y = 4-nitrobenzaldehyde (p-O2N-C6H4-CHO)
›Reveal solutionSolution
This tests the Etard reaction (the selective route from toluene to benzaldehyde) and the meta-directing effect of the –CHO group during nitration.
Concept and Intuition
Strong oxidants like alkaline KMnO4 oxidise a toluene methyl group all the way to −COOH, overshooting the aldehyde stage. The Etard reaction (CrO2Cl2 followed by aqueous hydrolysis) selectively stops the oxidation at the aldehyde. Once formed, the electron-withdrawing −CHO group deactivates the ring and directs incoming electrophiles to the meta position.
Step-by-Step Solution
- X: converting toluene directly to benzaldehyde (not benzoic acid) requires the Etard reaction — (i) treatment with CrO2Cl2 to form a chromium complex, (ii) hydrolysis with H3O+ to release C6H5CHO. KMnO4/OH− would instead push the oxidation all the way to C6H5COOH, so it is the wrong reagent here.
- Y: nitrating benzaldehyde. The −CHO group is deactivating and meta-directing (like other carbonyl-bearing groups), so the nitro group is introduced predominantly at the meta position, giving 3-nitrobenzaldehyde (m-O2N-C6H4-CHO). …
- AP EAPCET 2024Set ap-2024-05-17-FN1 markMCQQ.What is Y in the following reaction sequence? C6H5OCH3CH3Clanhy. AlCl3X (Major)(i) KMnO4/OH−, Δ(ii) H3O+Y (A) 3-methoxybenzoic acid (OCH3 meta to COOH on the benzene ring) (B) 4-methoxybenzoic acid (OCH3 para to COOH on the benzene ring) (C) 3-hydroxybenzoic acid (OH meta to COOH on the benzene ring) (D) 4-hydroxybenzoic acid (OH para to COOH on the benzene ring)
›Reveal solutionSolution
Friedel-Crafts methylation of anisole gives the para isomer as the major product;
oxidizing its methyl side chain to -COOH gives 4-methoxybenzoic acid, keeping the
methoxy and carboxyl groups para to each other.
Concept and Intuition
The methoxy group (-OCH3) is a powerful ortho, para-director (through resonance
donation of a lone pair into the ring) in electrophilic aromatic substitution. In
Friedel-Crafts alkylation, the bulky alkyl electrophile combined with steric hindrance
near the -OCH3 group means the para product dominates as the major isomer.
Separately, hot alkaline KMnO4 is a powerful oxidant that converts any
benzylic alkyl side chain (regardless of its length) all the way down to a -COOH group
directly attached to the ring — it does not affect the ether linkage itself.
Step-by-Step Solution
- C6H5OCH3 (anisole) + CH3Cl/anhydrous AlCl3 (Friedel-Crafts alkylation): methoxy directs the incoming methyl group ortho/para; the major product (less steric strain) is X = 1-methoxy-4-methylbenzene (p-cresol methyl ether), with −OCH3 and −CH3 para to each other.
- X + KMnO4/OH−, Δ, then H3O+: vigorous oxidation …
- AP EAPCET 2023Set ap-2023-05-23-FN1 markMCQQ.The electrophile involved in sulphonation of benzene is (A) S2Cl2 (B) SO3 (C) SO2 (D) SCl4
›Reveal solutionSolution
Sulphonation of benzene is an electrophilic aromatic substitution in which the attacking electrophile is SO3 (or its protonated form), generated from fuming sulphuric acid.
Concept and Intuition
Electrophilic aromatic substitution needs a species that is electron-deficient enough to be attacked by the benzene ring's π-electron cloud. In sulphonation, benzene is treated with fuming sulphuric acid (H2SO4 containing dissolved SO3, i.e. oleum). The sulphur atom in SO3 is highly electrophilic because it is bonded to three highly electronegative oxygens, leaving it electron-poor and eager to accept electron density from the aromatic ring.
Step-by-Step Solution
- Fuming sulphuric acid supplies SO3 as the reactive species (sometimes considered as the protonated form HSO3+ in strongly acidic medium).
- The ring's π-electrons attack the electrophilic sulphur of SO3, forming a sigma complex (arenium ion) with a −SO3− group attached to the ring carbon. …
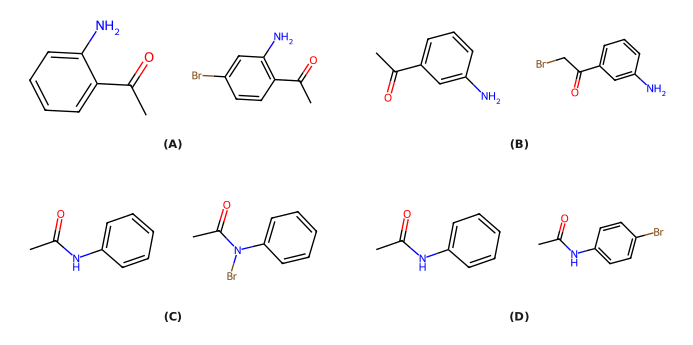
- AP EAPCET 2023Set eng-2023-05-16-FN1 markMCQQ.X and Y (major products) in the following reaction sequence are [FIGURE] (aniline - a benzene ring bearing an NH2 substituent - treated with acetic anhydride/pyridine to give X, then X treated with Br2/CH3COOH to give Y) (A) [FIGURE] X: a benzene ring with NH2 and COCH3 substituents ortho to each other; [FIGURE] Y: the same ortho NH2/COCH3-substituted ring with an added Br substituent para to the COCH3 group (B) [FIGURE] X: a benzene ring with NH2 and COCH3 substituents in a meta relationship; [FIGURE] Y: the same meta-substituted ring but with the methyl of the acetyl group brominated, i.e. COCH2Br instead of COCH3 (C) [FIGURE] X: acetanilide - a benzene ring bearing an NHCOCH3 substituent; [FIGURE] Y: N-bromo-N-phenylacetamide - the same structure but with Br on the nitrogen instead of H, i.e. Br−N(COCH3)-phenyl (D) [FIGURE] X: acetanilide - a benzene ring bearing an NHCOCH3 substituent; [FIGURE] Y: 4-bromoacetanilide - the same NHCOCH3-substituted ring with a Br substituent para to the NHCOCH3 group
›Reveal solutionSolution
Tests the classic amine-protection strategy before electrophilic aromatic substitution. Answer: X = acetanilide, Y = 4-bromoacetanilide (option D).
Concept and Intuition
Free aniline is such a strong activator (via the lone pair on N) that direct bromination is hard to control — it gives 2,4,6-tribromoaniline even with dilute bromine water, because the ring is so highly activated. To get a clean mono-bromination product, the amine is first "protected" by acetylation (acetic anhydride/pyridine), converting -NH2 to the less-activating, sterically bulkier -NHCOCH3 group (acetanilide). This still directs ortho/para (via resonance donation from N), but the bulky acetamido group sterically disfavours the ortho position, so bromination occurs predominantly at the para position.
Step-by-Step Solution
- Aniline + (CH₃CO)₂O / pyridine → acetanilide (X), C6H5NHCOCH3, via N-acetylation.
- Acetanilide + Br2/CH3COOH: the acetamido group is an ortho/para director but bulky, so bromination occurs mainly at the position para to it.
- Product Y = p-bromoacetanilide (4-bromoacetanilide), with Br para to the -NHCOCH3 group. …
- AP EAPCET 2023Set eng-2023-05-18-FN1 markMCQQ.What are 'X' and 'Y' respectively in the following reactions? Anisole (C6H5OCH3) Br2CH3COOHX (major); Anisole HIΔY (A) X = a benzene ring bearing OCH2Br (i.e. anisole's methyl hydrogen replaced by Br; ring itself unsubstituted); Y = phenol (benzene-OH) + CH3I (B) X = 4-bromoanisole (structure: benzene ring with OCH3 and Br para to each other); Y = phenol (benzene-OH) + CH3I (C) X = 2-bromoanisole (structure: benzene ring with OCH3 and Br ortho to each other); Y = iodobenzene (benzene-I) + CH3OH (D) X = 4-bromoanisole (structure: benzene ring with OCH3 and Br para to each other); Y = iodobenzene (benzene-I) + CH3I
›Reveal solutionSolution
Anisole undergoes para-selective electrophilic bromination (X = 4-bromoanisole) and, with HI, ether cleavage at the methyl carbon (not the aryl carbon), giving phenol + CH3I (Y).
Concept and Intuition
- Bromination (X): The methoxy group is a powerful activating, ortho/para-directing substituent. Electrophilic aromatic substitution with Br2 in acetic acid proceeds readily; the para product dominates over ortho because of steric crowding near the bulky OCH3 group, giving 4-bromoanisole as the major product.
- Ether cleavage (Y): Aryl alkyl ethers resist cleavage at the aryl–oxygen bond because that bond has partial double-bond character from resonance delocalisation of the oxygen lone pair into the ring — breaking it would require attacking an sp2 aromatic carbon (very unfavourable for SN2, and SN1 would give a highly unstable aryl cation). Instead, the iodide ion (from HI) performs an SN2 attack on the far less hindered, non-aromatic methyl carbon, displacing the phenoxide, which is protonated to phenol. This is the basis of the Zeisel method for estimating methoxy groups.
Step-by-Step Solution
- Identify the directing effect of −OCH3: strongly activating, o/p-director.
- Steric bulk favours para substitution as the major product ⇒ X = 4-bromoanisole. …
🎓Unlock everything free for 14 days
- ✓Full step-by-step solutions
- ✓Concept-first explanations
- ✓Methods, shortcuts & mistakes
- ✓PYQ mapping + timed mock tests
Full access for 14 days. No credit card required.