Q.When 3-methylbutan-2-ol is treated with HBr, the following reaction takes place: CH3−CH3∣CH−OH∣CH−CH3 HBr CH3−CH3∣C∣Br−CH2−CH3 Give a mechanism for this reaction. (Hint : The secondary carbocation formed in step II rearranges to a more stable tertiary carbocation by a hydride ion shift from 3rd carbon atom.
🔒You're viewing a preview — the full solution, concept, methods & PYQ mapping are locked.
🔒 Start your 14-day free trial to unlock the full solution →Concept understanding — Williamson Ether Synthesis
Williamson Ether Synthesis: From Intuition to Mechanism
Imagine you want to build a simple bridge between two carbon chains — an oxygen atom linking them together. That bridge is an ether (R−O−R′). The Williamson ether synthesis is the most reliable way to build that bridge in a lab.
The Core Idea
You have two pieces: an alkoxide ion (RO−) and an alkyl halide (R′X). The alkoxide is a strong nucleophile — it loves positive charge. The alkyl halide has a carbon attached to a halogen (like Cl, Br, I) that is slightly positive because the halogen pulls electrons away.
When you mix them, the alkoxide attacks that slightly positive carbon, kicks out the halide ion, and forms a new C−O bond. The result? An ether.
R−O−+R′−X⟶R−O−R′+X−
That's the entire reaction in one line. But the devil is in the details — especially which alkyl halide you choose.
The Mechanism (SN2)
This is a classic SN2 reaction — one step, no intermediates. The alkoxide approaches the carbon from the opposite side of the halogen. As the C−O bond forms, the C−X bond breaks. The halide leaves as a stable anion.
Because it's SN2, the reaction is sensitive to steric hindrance. The carbon being attacked must be accessible.
If the alkyl halide is tertiary (3°), the reaction will not work via SN2. The bulky carbon blocks the backside attack. Instead, the alkoxide will act as a base and cause elimination (forming an alkene). You'll get no ether.
The Practical Rule
| Alkyl halide | Works? | Why |
|---|---|---|
| Methyl (CH3X) | Yes | Least hindered, fastest SN2 |
| Primary (1°) | Yes | Clean SN2 |
| Secondary (2°) | Sometimes | Works if not too bulky; elimination competes |
| Tertiary (3°) | No | Elimination dominates |
| Aryl (e.g., bromobenzene) | No | SN2 impossible on sp2 carbon |
To make an ether like R−O−R′, always use the less hindered alkyl halide and the more hindered alkoxide. For example, to make CH3CH2−O−CH(CH3)2, use CH3CH2O− (primary alkoxide) + (CH3)2CHBr (secondary halide) — not the other way around.
How to Choose the Alkoxide
You can't just buy alkoxide ions in a bottle. You make them by reacting an alcohol with a strong base like sodium hydride (NaH) or sodium metal.
ROH+NaH⟶RO−Na++H2
The alkoxide is then used immediately with the alkyl halide.
A Common Exam Trap …
Why this formula?
Williamson Ether Synthesis: Why the Key Principles Hold
The Williamson Ether Synthesis is a classic method to prepare ethers. The core reaction is:
R-O−+R’-X→R-O-R’+X−
Where:
- R-O− is an alkoxide ion (strong nucleophile)
- R’-X is an alkyl halide (electrophile)
- X− is a halide ion (leaving group)
Let's break down why this works — the reasoning behind the key principles.
1. Why an Alkoxide (Not an Alcohol) is Needed
The Problem with Alcohols
Alcohols (R-OH) are weak nucleophiles. The oxygen has a partial negative charge, but the O–H bond is strong. If you mix an alcohol with an alkyl halide, the reaction is extremely slow or doesn't happen at all.
The Solution: Deprotonation
By treating the alcohol with a strong base (like NaH, Na, or KOH), you remove the proton:
R-OH+NaH→R-O−Na++H2
The alkoxide ion (R-O−) has a full negative charge on oxygen. This makes it:
- A much stronger nucleophile (higher electron density)
- More reactive toward the electrophilic carbon in the alkyl halide
Key takeaway: The alkoxide's full negative charge is what drives the reaction — it's not just about having oxygen, but about having a charged, electron-rich oxygen.
2. Why the Alkyl Halide Must Be Primary (or Methyl)
The Mechanism: SN2 is the Only Path
The Williamson synthesis proceeds exclusively via an SN2 mechanism (bimolecular nucleophilic substitution). This means:
- The nucleophile attacks the carbon from the backside
- The leaving group departs from the opposite side
- The reaction is concerted (one step, no intermediates)
Why Primary Halides Work Best
In SN2 reactions, the rate depends on steric hindrance:
| Alkyl Halide Type | Steric Hindrance | SN2 Reactivity |
|---|---|---|
| Methyl (CH3X) | Minimal | Very fast |
| Primary (RCH2X) | Low | Fast |
| Secondary (R2CHX) | Moderate | Slow |
| Tertiary (R3CX) | High | Does not occur |
Why Tertiary Halides Fail
With a tertiary halide, the bulky alkyl groups block the backside attack. Instead, the alkoxide (a strong base) will eliminate a proton from the halide, forming an alkene:
R-O−+R’3C-X→R-OH+alkene+X−
This is an E2 elimination — not the desired ether formation.
Key takeaway: The Williamson synthesis works only when the alkyl halide is primary or methyl because SN2 requires an unhindered backside.
3. Why the Leaving Group Must Be Good
The Role of the Halide
The halide (X−) must be a good leaving group — meaning it can stabilize the negative charge after departure.
| Halide | Leaving Group Ability | Reason |
|---|---|---|
| I− | Excellent | Large, polarizable, weak base |
| Br− | Good | Moderate size, weak base |
| Cl− | Fair | Smaller, stronger base |
| F− | Poor | Small, strong base, holds tightly |
Why Fluoride Fails
Fluoride is a strong base and a poor leaving group. The C–F bond is very strong, and F− does not depart easily. So alkyl fluorides are unreactive in Williamson synthesis.
Key takeaway: The leaving group must be weakly basic and polarizable — iodide and bromide are ideal.
--- …
The key idea is that a secondary carbocation undergoes a 1,2-hydride shift to form a more stable tertiary carbocation before the nucleophile attacks.
Step 1: Protonation – The hydroxyl group of 3-methylbutan-2-ol is protonated by HBr, forming a good leaving group (HX2O).
Step 2: Formation of secondary carbocation – Water leaves, generating a secondary carbocation at the 2-position: CHX3−CH(CHX3)−CHX+−CHX3.
Step 3: Hydride shift – A hydride ion (HX−) from the 3rd carbon (the methine carbon) shifts to the positively charged 2nd carbon. This rearranges the carbocation to a more stable tertiary carbocation at the 3-position: CHX3−CX+(CHX3)−CHX2−CHX3. …
The reaction proceeds via an Sₙ1 mechanism where the initial secondary carbocation undergoes a 1,2-hydride shift to form a more stable tertiary carbocation, which is then attacked by Br⁻ to give the rearranged product.
Why this happens — the concept
The hint tells you the key: a hydride shift. But why does the carbocation bother to rearrange at all? The answer is carbocation stability. Tertiary carbocations are more stable than secondary ones because the three alkyl groups donate electron density via hyperconjugation and inductive effects, spreading the positive charge. The reaction is driven by the thermodynamic urge to form the most stable intermediate possible.
In Williamson ether synthesis, you'd use an alkoxide and an alkyl halide — but here we have an alcohol with HBr, which is a classic Sₙ1 or Sₙ2 situation. Because the alcohol is secondary and the acid provides a good leaving group (water), the reaction favours an Sₙ1 pathway, especially when rearrangement can lead to a more stable carbocation.
Step-by-step mechanism
1. Protonation of the hydroxyl group
The lone pair on oxygen of 3-methylbutan-2-ol attacks a proton from HBr, forming an oxonium ion. This makes the OH group a much better leaving group (water instead of hydroxide).
CHX3−CH(CHX3)−CH(OH)−CHX3+HBrCHX3−CH(CHX3)−CH(OHX2X+)−CHX3+BrX−
2. Loss of water to form a secondary carbocation
The C–O bond breaks heterolytically, ejecting a water molecule and leaving behind a secondary carbocation at the 2nd carbon.
CHX3−CH(CHX3)−CH(OHX2X+)−CHX3CHX3−CH(CHX3)−CHX+ −CHX3+HX2O
This carbocation is secondary — it has two alkyl groups attached to the positive carbon. It's reasonably stable, but not as stable as it could be.
A common mistake is to stop here and attack Br⁻ directly. But if you do that, you'd get the unrearranged product (2-bromo-3-methylbutane), which is not what the question shows. The product given has the bromine on a tertiary carbon — so rearrangement must occur.
3. 1,2-Hydride shift
A hydride ion (H⁻) from the 3rd carbon (the one bearing the methyl group) shifts to the positively charged 2nd carbon. This moves the positive charge to the 3rd carbon, which is now tertiary (attached to three alkyl groups: two methyls and one ethyl group).
CHX3−CH(CHX3)−CHX+ −CHX3H− shiftCHX3−CX+(CHX3)−CHX2−CHX3
The arrow-pushing: the C–H bond at C3 breaks, and the pair of electrons moves to the empty p-orbital at C2. The result is a tertiary carbocation at C3. …
Method: Carbocation Rearrangement via Hydride Shift (SN1 Mechanism)
This reaction follows an SN1 mechanism with a 1,2-hydride shift to form a more stable tertiary carbocation.
Step-by-Step Mechanism
- Protonation of the hydroxyl group The alcohol’s –OH is protonated by HBr, turning it into a good leaving group (HX2O).
CHX3−CH(CHX3)−CH(OH)−CHX3+HBrCHX3−CH(CHX3)−CH(OHX2X+)−CHX3+BrX−
- Loss of water to form a secondary carbocation Water leaves, generating a secondary carbocation at the 2nd carbon.
CHX3−CH(CHX3)−CH(OHX2X+)−CHX3CHX3−CH(CHX3)−CHX+−CHX3+HX2O
- 1,2-Hydride shift (rearrangement) A hydride ion (HX−) from the 3rd carbon shifts to the positively charged 2nd carbon. This forms a tertiary carbocation (more stable due to hyperconjugation and inductive effects).
CHX3−CH(CHX3)−CHX+−CHX3CHX3−CX+(CHX3)−CHX2−CHX3
- Nucleophilic attack by bromide ion The bromide ion (BrX−) attacks the tertiary carbocation, giving the final product. …
Mistake 1: Forgetting that the OH group must be protonated first
What students do wrong:
They jump straight to breaking the C–O bond, showing OH⁻ as a leaving group. That’s impossible — OH⁻ is a terrible leaving group.
How to avoid:
Always remember: in acidic conditions, the first step is protonation of the –OH to make it a good leaving group (H2O).
Write:
CHX3−CH(CHX3)−CH(OH)−CHX3+HBrCHX3−CH(CHX3)−CH(OHX2X+)−CHX3+BrX−
Mistake 2: Showing the wrong carbocation after water leaves
What students do wrong:
They show the secondary carbocation as:
CHX3−CH(CHX3)−CHX+−CHX3
and then stop, or try to attach Br⁻ directly here.
How to avoid:
The hint says a hydride shift occurs. The secondary carbocation is unstable and will rearrange. Draw the shift clearly:
CHX3−CH(CHX3)−CHX+−CHX3HX− shiftCHX3−CX+(CHX3)−CHX2−CHX3
The tertiary carbocation is more stable — this is the driving force.
Mistake 3: Misidentifying which H atom shifts
What students do wrong:
They shift a hydride from the wrong carbon (e.g., from the methyl group on C2 or from C4).
How to avoid:
The hint says: hydride ion shift from the 3rd carbon atom.
Count carbons carefully:
- C1: CHX3X− (attached to C2)
- C2: −CH(CHX3)X− (the one with OH originally)
- C3: −CH− (the middle carbon)
- C4: −CHX3
The shift is from C3 to C2, turning C2 into a tertiary carbocation.
Mistake 4: Forgetting that Br⁻ attacks the carbocation
What students do wrong:
They stop after the rearrangement, or show Br⁻ attacking the wrong carbon.
How to avoid:
After the tertiary carbocation forms, the nucleophilic Br⁻ (from HBr) attacks the positively charged carbon:
CHX3−CX+(CHX3)−CHX2−CHX3+BrX−CHX3−C(Br)(CHX3)−CHX2−CHX3
This gives the final product shown in the question.
Mistake 5: Not showing arrow pushing correctly
What students do wrong:
They draw arrows that start from nowhere, or point to the wrong atom.
How to avoid:
Every arrow must start from a lone pair or a bond and point to where the electrons go. For the hydride shift:
- Arrow starts from the C–H bond on C3
- Points to the carbocation on C2
Mistake 6: Confusing this with an SN1 or E1 reaction …
- AP EAPCET 2026Set eng-2026-05-15-FN1 markMCQQ.Which one of the following is not correct? (A) (CH3)3CONa+CH3Br→(CH3)3COCH3 (B) (C2H5)2Oexcess HIΔ2C2H5I+H2O (C) (CH3)3COC2H5HIΔ(CH3)3CI+C2H5OH (D) C6H5Br+CH3ONa→C6H5OCH3+NaBr
›Reveal solutionSolution
Aryl halides do not undergo ordinary nucleophilic substitution the way alkyl halides
do, so bromobenzene + sodium methoxide will not simply hand you anisole. Answer: (D).
Concept and Intuition
- (A) (CH3)3CONa+CH3Br→(CH3)3COCH3: this is Williamson ether synthesis. The rule of thumb is that the alkyl halide should be unhindered (methyl or primary) for a clean SN2; the bulk of the alkoxide doesn't matter much since it's the nucleophile, not the electrophile. Methyl bromide is a perfect SN2 substrate, so this reaction proceeds cleanly to give methyl tert-butyl ether. Correct.
- (B) (C2H5)2Oexcess HIΔ2C2H5I+H2O: with excess hot HI, a symmetrical ether is cleaved completely, both alkyl-oxygen bonds broken, to give two equivalents of alkyl iodide. Correct textbook fact.
- (C) (CH3)3COC2H5HIΔ(CH3)3CI+C2H5OH: this is a mixed ether with one tertiary and one primary alkyl group. Cleavage proceeds via SN1 at the carbon that gives the more stable carbocation (tertiary), so the tert-butyl group leaves as the iodide and the ethyl-oxygen fragment is released as ethanol. This is the standard textbook outcome. Correct.
- (D) Aryl halides like bromobenzene have their C–X bond strengthened by resonance with the ring (partial double-bond character) and the carbon is sp2, blocking backside SN2 attack. Nucleophilic substitution on an unactivated aryl halide by a simple alkoxide under ordinary conditions simply does not happen — it requires either very forcing conditions (high temperature and pressure, as in phenol manufacture from chlorobenzene) or a strong base capable of a benzyne (elimination-addition) pathway (e.g. NaNH2). Plain CH3ONa at ordinary conditions will not convert bromobenzene to anisole. This statement is therefore NOT correct.
Step-by-Step Solution …
- AP EAPCET 2026Set ap-2026-05-20-AN1 markMCQQ.What is the IUPAC name of the product Y formed in the given sequence of reactions? Isobutane KMnO4 X (i) Na(ii) CH3−Br Y (A) 1, 1, 1-Trimethyl methoxy methane (B) Methyl, t-Butyl ether (C) 2-Methyl-2-methoxy propane (D) 2-Methoxy-2-methyl propane
›Reveal solutionSolution
Tertiary C-H oxidation gives tert-butanol, then Williamson ether synthesis gives MTBE; the only remaining subtlety is citing the substituent prefixes in the correct alphabetical order.
Concept and Intuition
Tertiary C-H bonds are the weakest and most easily oxidised C-H bonds in an alkane (the resulting radical/cation is most stabilised), so a strong oxidant like KMnO4 selectively converts isobutane's one tertiary hydrogen into a tertiary alcohol rather than attacking the primary methyl hydrogens. Converting that alcohol to its sodium alkoxide and reacting with a primary alkyl halide (Williamson ether synthesis, SN2 at the primary carbon of CH3Br) builds the ether cleanly, since SN2 works well on primary halides.
Step-by-Step Solution
- Isobutane (CH3)3CH + KMnO4 → oxidation at the sole tertiary C-H → X = tert-butyl alcohol, (CH3)3C−OH (2-methylpropan-2-ol).
- X + Na → sodium tert-butoxide, (CH3)3C−O−Na+ (Na displaces the O-H proton).
- Sodium tert-butoxide + CH3Br → Williamson ether synthesis (the alkoxide's oxygen performs SN2 on the primary carbon of methyl bromide) → Y = (CH3)3C−O−CH3, methyl tert-butyl ether. …
- AP EAPCET 2025Set ap-2025-05-20-AN1 markMCQQ.What is the major product Y in the following reaction sequence ? C6H5NH2 (aniline) (i) NaNO2/HCl, 273K(ii) H2O, Δ X (i) NaOH, CH3Br(ii) Br2/CH3COOH Y (A) C6H5−OCH2Br (benzene ring with a single −OCH2Br substituent) (B) benzene ring with Br and −OCH2Br substituents at para positions (4-Br-C6H4-OCH2Br) (C) benzene ring with Br and −OCH3 substituents at para positions (4-Br-C6H4-OCH3, 4-bromoanisole) (D) benzene ring with Br and −CH3 substituents at para positions (4-Br-C6H4-CH3, 4-bromotoluene)
›Reveal solutionSolution
Aniline → diazonium salt → phenol (X) → anisole (via Williamson ether synthesis) → 4-bromoanisole (Y, major product of electrophilic bromination directed para by the methoxy group).
Concept and Intuition
This is a multi-step synthesis chaining three classic reactions: diazotisation/hydrolysis (amine → phenol via the diazonium salt), Williamson ether synthesis (phenoxide + alkyl halide → aryl alkyl ether), and electrophilic aromatic bromination directed by a strongly activating, ortho/para-directing methoxy group. Since anisole's methoxy substituent is already present on the ring, the incoming Br+ (from Br2 in acetic acid) attacks preferentially para (and some ortho), and para is reported as the major product due to less steric crowding.
Step-by-Step Solution
- C6H5NH2NaNO2/HCl, 273KC6H5N2+Cl− (benzenediazonium chloride) — standard diazotisation of a primary aromatic amine at 0–5°C.
- C6H5N2+Cl−H2O, Δ phenol (C6H5OH) + N2 + HCl — hydrolysis of the diazonium salt on warming. So X = phenol.
- Phenol + NaOH → sodium phenoxide (C6H5O−Na+); phenoxide + CH3Br → anisole (C6H5OCH3) via SN2 (Williamson ether synthesis). …
- AP EAPCET 2024Set eng-2024-05-21-FN1 markMCQQ.The major products X and Y respectively from the following reactions are YNaOEtCH3CH2CH2CH2Br(i)Mg/dry ether(ii)H2OX (Y = major) (A) CH3CH2CH2CH3, CH3CH2CH2CH2OC2H5 (B) CH3CH2CH3, CH2=CHCH3 (C) CH3CH2CH2CH2OH, CH2=CHCH3 (D) CH3CH2CH2CH2OH, CH3CH2CH2CH2OC2H5
›Reveal solutionSolution
Grignard + water gives the alkane (not the alcohol you'd get from a carbonyl); sodium ethoxide on a primary halide gives clean SN2 substitution (an ether), not elimination.
Concept and Intuition
A Grignard reagent R-MgX behaves like a carbanion R−. Any proton source acidic enough (even water, pKa≈15.7, is far more acidic than the alkane conjugate acid) instantly protonates it: R-MgX+H2O→R-H+Mg(OH)X. This is the classic way to convert a halide into the corresponding hydrocarbon — it is not how you make an alcohol (that needs the Grignard to attack a carbonyl carbon first).
Separately, when a nucleophile/base attacks an alkyl halide, the outcome (SN2 vs E2) depends on both the base's bulk and the substrate's branching. Ethoxide (NaOEt) is a comparatively small, strong base. Against a primary substrate with an easily accessible backside carbon (n-butyl bromide), steric hindrance to backside attack is minimal, so SN2 substitution is the major pathway, not elimination.
Step-by-Step Solution
- CH3CH2CH2CH2Br+Mgdry etherCH3CH2CH2CH2MgBr (Grignard reagent formed).
- CH3CH2CH2CH2MgBr+H2O→CH3CH2CH2CH3+Mg(OH)Br — protonolysis gives butane, so X=CH3CH2CH2CH3. …
- AP EAPCET 2024Set eng-2024-05-21-FN1 markMCQQ.What are the major products X and Y respectively in the following reactions? (CH3)3CONa+CH3CH2Br→X (CH3)3CBr+CH3CH2ONa→Y (A) CH2=CH2, (CH3)3COCH2CH3 (B) (CH3)3COCH2CH3, (CH3)3COCH2CH3 (C) CH2=CH2, (CH3)2C=CH2 (D) (CH3)3COCH2CH3, (CH3)2C=CH2
›Reveal solutionSolution
SN2 vs E2 is decided by whether backside attack at the carbon bearing the leaving group is sterically possible — here that means looking at the alkyl halide's own branching, not just the base's bulk.
Concept and Intuition
The classic trap in these paired reactions is to only look at how bulky the base/nucleophile is. What actually controls the outcome is whether the nucleophile can reach the back lobe of the C–LG bond:
- If the carbon bearing the leaving group is primary and unhindered (ethyl bromide), SN2 remains fast and dominant even with a bulky base like t-butoxide, because the bulk of the base doesn't block approach to a completely open primary carbon nearly as much as branching at the substrate would.
- If the carbon bearing the leaving group is tertiary (tert-butyl bromide), backside attack is essentially impossible regardless of which base is used — the only viable pathway is E2 (proton abstraction from a β-carbon), so even a small, strong, ionic base like ethoxide gives elimination as the major product.
Step-by-Step Solution
- Reaction 1: (CH3)3CONa (base/nucleophile) + CH3CH2Br (primary substrate). The electrophilic carbon (in CH3CH2Br) is unhindered, so t-butoxide's oxygen performs SN2 substitution: X=(CH3)3C-O-CH2CH3. …
- AP EAPCET 2023Set ap-2023-05-23-FN1 markMCQQ.Better results for the preparation of ethers 'X' and 'Y' can be obtained from reactant pairs respectively CH3CH2C(CH3)2OCH2CH2CH3 (X); a benzene ring with substituent O-CH2CH2CH3 (Y) (A) (CH3CH2C(CH3)2Br+CH3CH2CH2ONa) ; (bromobenzene C6H5Br + CH3CH2CH2ONa) (B) (CH3CH2C(CH3)2ONa+CH3CH2CH2Br) ; (phenol C6H5OH + CH3CH2CH2Br) (C) (CH3CH2C(CH3)2ONa+CH3CH2CH2Br) ; (bromobenzene C6H5Br + CH3CH2CH2ONa) (D) (CH3CH2C(CH3)2Br+CH3CH2CH2ONa) ; (phenol C6H5OH + CH3CH2CH2Br)
›Reveal solutionSolution
Williamson ether synthesis needs the bulky/aryl partner as the alkoxide and the unhindered partner as the primary halide, to avoid elimination or a non-reactive aryl-halide substitution.
Concept and Intuition
Williamson synthesis is an SN2 reaction between an alkoxide and an alkyl halide. Two structural traps show up here: (1) tertiary alkyl halides are poor SN2 substrates — with a strong nucleophile/base like an alkoxide they instead undergo E2 elimination; and (2) aryl halides (like bromobenzene) cannot undergo SN2 at all because the aryl carbon is sp2, in-plane, and shielded, and the C–X bond is strengthened by ring resonance.
Step-by-Step Solution
- For X, CH3CH2C(CH3)2−O−CH2CH2CH3: the tertiary-pentyl part must come in as the alkoxide (sodium tert-alkoxide), while the primary propyl part comes in as the halide (propyl bromide) — the primary halide undergoes clean SN2 with the bulky alkoxide, avoiding elimination.
- For Y, phenyl propyl ether (C6H5−O−CH2CH2CH3): since aryl halides (bromobenzene) cannot undergo nucleophilic substitution, the correct route is phenoxide (from phenol) attacking the primary alkyl halide (propyl bromide). …
- AP EAPCET 2022Set ap-2022-07-11-AN1 markMCQQ.The major product P from the following reaction is [FIGURE] (a benzene ring bearing a −C(CH3)2Br group at one ring position and a −CH2Br group at a nearby (meta) position) Me3CONaP (A) [FIGURE] (a benzene ring bearing a −C(CH3)2−O−C(CH3)3 group, i.e. the tertiary carbon converted to a tert-butyl ether) (B) [FIGURE] (a benzene ring bearing a −C(CH3)2Br group and a −CH2OH group at the meta position) (C) [FIGURE] (a benzene ring bearing a −C(CH3)2Br group and a vinyl group −CH=CH2 at the meta position) (D) [FIGURE] (a benzene ring bearing two tertiary alcohol groups of the form −C(CH3)2OH, one directly attached to the ring and one attached via a −CH2CH2− chain)
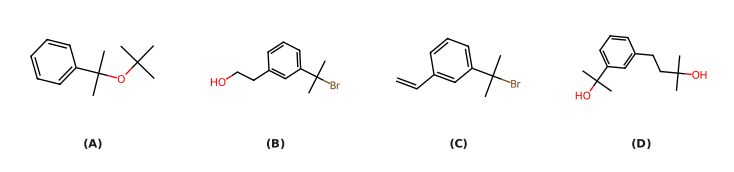
›Reveal solutionSolution
Bulky sodium tert-butoxide is a classic E2-promoting, poor-SN2 base; on the primary bromide side chain it eliminates HBr to give a terminal alkene (vinyl group), while the more hindered tertiary bromide is unaffected — matching option (C).
Concept and Intuition
The identity of a base/nucleophile controls whether an alkyl halide undergoes substitution or elimination. Sodium tert-butoxide, (CH3)3CO−Na+, is strongly basic (as a good alkoxide) but its oxygen is buried under three bulky methyl groups, making it a very poor nucleophile for SN2 (it cannot easily approach a carbon from the back side). This steric bulk is exactly why tert-butoxide is the textbook reagent of choice to force elimination (E2) rather than substitution even on primary alkyl halides, which would normally favour SN2 with a small nucleophile like hydroxide or ethoxide. For a primary bromide such as −CH2CH2Br, tert-butoxide therefore removes a β-hydrogen (E2) to give the terminal alkene −CH=CH2, releasing Br−.
Meanwhile, this same molecule also carries a tertiary, benzylic bromide, −C(CH3)2Br, directly on the ring. In problems of this kind, the answer choices make clear that the reaction is selective for the primary position over the tertiary one under the given conditions/timescale — the tertiary center is left as the unreacted starting bromide in the major product.
Step-by-Step Solution
- Identify the base: Me3CONa = sodium tert-butoxide — strong base, very weak/hindered nucleophile.
- Recognize its standard textbook behaviour: promotes E2 elimination, especially valuable for driving primary alkyl halides toward alkenes instead of the substitution product they would normally favour with a small nucleophile.
- Apply E2 to the primary bromide arm: −CH2−CH2−Br−HBr−CH=CH2 (a terminal vinyl group, the only possible elimination product from a 2-carbon primary chain). …
- AP EAPCET 2021Set eng-2021-08-25-AN1 markMCQQ.Identify the product of the following reaction: CH3CH3−C−ONaCH3+CH3Cl⟶ ? (A) C2H5−C(CH3)2−O−CH3 (B) C2H5−C(CH3)2−O−C2H5 (C) C2H5−C(CH3)2−O−Cl (D) H3C−O−C2H5
›Reveal solutionSolution
A branched sodium alkoxide plus methyl chloride is a textbook Williamson ether synthesis: the alkoxide's own carbon skeleton is retained, and a new methyl group is added onto its oxygen.
Concept and Intuition
Williamson ether synthesis works best when the alkyl halide is primary/unhindered (favours clean SN2) — methyl halides are ideal because they have no β-hydrogens at all, so competing elimination is impossible regardless of how bulky the incoming alkoxide nucleophile is. The alkoxide's own carbon skeleton is untouched by the reaction (only its oxygen attacks); the methyl group from the alkyl halide simply attaches to that oxygen.
Step-by-Step Solution
- Identify the nucleophile: the sodium alkoxide of the branched alcohol, R3C–O−Na+ (with R3C its full carbon skeleton, retained unchanged throughout the reaction).
- Identify the electrophile: CH3Cl — small, unhindered, no possibility of elimination (no β-H).
- SN2 attack: the alkoxide oxygen's lone pair displaces Cl− from the methyl carbon. …
🎓Unlock everything free for 14 days
- ✓Full step-by-step solutions
- ✓Concept-first explanations
- ✓Methods, shortcuts & mistakes
- ✓PYQ mapping + timed mock tests
Full access for 14 days. No credit card required.