Q.(a) Write reasons for the following :
🔒You're viewing a preview — the full solution, concept, methods & PYQ mapping are locked.
🔒 Start your 14-day free trial to unlock the full solution →Part (a)Concept understanding — Ortho Para Directing
The Intuition: Why Some Groups "Point" the Next Attack
Imagine you're trying to add a second substituent to a benzene ring that already has one group attached. The ring already has six hydrogens, but they aren't all equal anymore — the first group has changed the electron density at different positions. Some positions become more "attractive" to an incoming electrophile (a positive or electron-seeking species), while others become less attractive.
Ortho-para directing groups are substituents that make the next electrophile prefer to attack the positions next to the group (ortho, positions 2 and 6) or directly opposite it (para, position 4), rather than the meta position (position 3 and 5).
The terms come from Greek: ortho = straight/correct (adjacent), meta = after (one carbon away), para = beside/opposite (two carbons away, directly across).
The Precise Statement
Ortho-para directing groups are substituents that, when present on a benzene ring, cause the next electrophilic aromatic substitution (EAS) reaction to occur predominantly at the ortho and para positions relative to themselves. These groups are typically electron-donating (activating) or weakly deactivating (like halogens).
The Mechanism: How They Work
The key lies in the stability of the intermediate carbocation (the arenium ion / sigma complex) formed during the attack.
When an electrophile attacks benzene, the ring temporarily loses its aromaticity and becomes a positively charged carbocation. This intermediate is stabilised if the positive charge can be delocalised onto the substituent. Ortho-para directing groups are able to donate electron density into the ring, either through:
- Resonance effect (most important): The group has lone pairs or pi electrons that can be pushed into the ring, creating extra resonance structures where the positive charge is on the substituent (which is more stable).
- Inductive effect: The group is electron-donating through sigma bonds (e.g., alkyl groups like methyl).
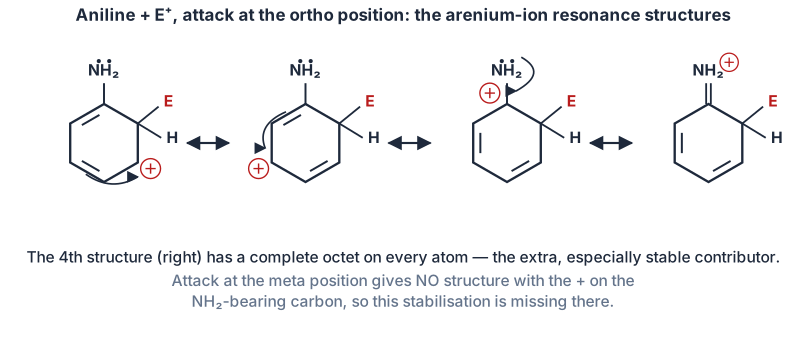
Let's see what happens when an electrophile attacks the ortho position of aniline (NH₂ group):
›Proof
Resonance stabilisation for ortho attack (aniline)
The NH₂ group donates its lone pair into the ring. When the electrophile attacks ortho, the positive charge can be delocalised onto the nitrogen atom (which is very happy to carry a positive charge because it's electronegative and has a lone pair). This gives an extra, highly stable resonance structure that is not available for meta attack.
For meta attack, the positive charge stays on the ring carbons — no extra stabilisation from the substituent. Hence ortho/para attack is favoured.
The Two Categories of Ortho-Para Directors
| Type | Examples | Effect | Why? |
|---|---|---|---|
| Strongly activating | -OH, -NH₂, -OCH₃, -NHR | Strong ortho-para directing | Strong resonance donation (lone pairs) |
| Moderately activating | -CH₃, -C₂H₅, -R (alkyl) | Ortho-para directing | Inductive electron donation (no lone pairs, but pushes electrons through sigma bonds) |
| Weakly deactivating | -F, -Cl, -Br, -I | Ortho-para directing (surprisingly!) | Halogens are electron-withdrawing inductively but electron-donating by resonance (lone pairs). The resonance effect wins for directing, but the inductive withdrawal makes the ring less reactive overall. |
Common mistake: Students think "deactivating" means "meta directing". Halogens are the exception — they deactivate the ring (slower reaction) but still direct ortho/para. The resonance donation of lone pairs is strong enough to stabilise the ortho/para intermediate, but the inductive withdrawal makes the ring less electron-rich overall. …
Why this formula?
Ortho-Para Directing: The Why Behind the Rule
Let’s build this from first principles. The question is: Why do certain groups on a benzene ring direct new substituents to the ortho and para positions, while others direct to the meta position?
The answer lies in resonance stabilization of the intermediate carbocation (the arenium ion / σ-complex) during electrophilic aromatic substitution (EAS).
1. The Core Mechanism: EAS Forms a Carbocation Intermediate
In EAS, the electrophile (E+) attacks the benzene ring. The ring temporarily loses aromaticity, forming a resonance-stabilized carbocation:
benzene+EX+[arenium ion]product
The arenium ion has three resonance forms. The stability of this intermediate determines how fast the reaction proceeds and where the electrophile attacks.
2. What Makes a Group Ortho-Para Directing?
A group is ortho-para directing if it donates electron density into the ring, especially at the ortho and para positions. This donation stabilizes the carbocation when the electrophile attacks those positions.
The Key: Resonance Structures of the Intermediate
Consider an activating group like −OH (phenol). When the electrophile attacks the ortho position, one resonance form places the positive charge directly on the carbon bearing the −OH group. The oxygen’s lone pair can then donate into that empty p-orbital, creating an extra, highly stable resonance structure:
ortho attack: ...[resonance form with C+-OH][resonance form with O+=C]
This extra resonance contributor (with a positive charge on the electronegative oxygen) is not possible for meta attack. For meta attack, the positive charge never lands on the carbon attached to the −OH group — so no extra stabilization.
The donation itself, drawn for phenol:
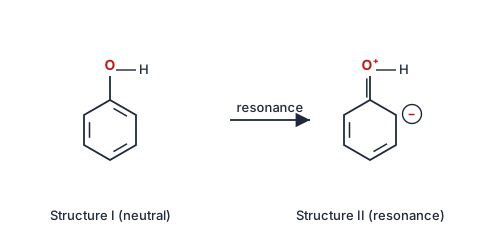
Result: The ortho/para intermediates are more stable (lower energy) than the meta intermediate. Hence, the reaction is faster at ortho/para positions.
3. The Formula: Why Ortho and Para Specifically?
The resonance structures of the arenium ion reveal the pattern:
- For ortho attack: The positive charge can be delocalized to the carbon bearing the substituent (position 1).
- For para attack: The positive charge can also be delocalized to the carbon bearing the substituent (position 1).
- For meta attack: The positive charge never reaches the carbon with the substituent.
Mathematically, if the substituent is at position 1, the positions that can stabilize the positive charge via resonance are positions 2, 4, and 6 (ortho and para). Positions 3 and 5 (meta) cannot.
4. The Deactivating Ortho-Para Directors: The Halogen Exception
Halogens (−F,−Cl,−Br,−I) are deactivating (they withdraw electron density inductively) but ortho-para directing. Why?
- Inductive effect: Halogens are electronegative → pull electron density away from the ring → deactivate (slow down EAS).
- Resonance effect: Halogens have lone pairs → can donate into the ring via resonance → stabilize the ortho/para intermediates (just like −OH).
Drawn out for chlorobenzene: …
Part (b)Concept understanding — Hofmann Bromamide Reaction
Hofmann Bromamide Degradation
Imagine you have an amide — a molecule with a carbonyl group (−CO−) attached to a nitrogen. You want to turn it into a primary amine, but you also want to chop off one carbon from the chain. That is exactly what the Hofmann bromamide reaction does: it shortens the carbon skeleton by one carbon and gives you an amine.
The Intuition
The reaction uses bromine (BrX2) in the presence of a strong alkali (like NaOH or KOH). The alkali first deprotonates the amide nitrogen, making it a strong nucleophile. This nucleophile attacks bromine, forming an N-bromoamide. Under the strongly basic conditions, this intermediate loses a bromide ion and undergoes a rearrangement — the alkyl group attached to the carbonyl carbon migrates from carbon to nitrogen. The result is an isocyanate intermediate (R−N=C=O). Finally, the isocyanate is hydrolysed by the aqueous alkali to give a primary amine and carbon dioxide.
The net effect: the carbonyl carbon is lost as COX2, and the alkyl group ends up attached to the nitrogen.
The product amine has one fewer carbon than the starting amide. The lost carbon is the carbonyl carbon.
The Precise Statement
Hofmann bromamide degradation (also called Hofmann rearrangement) is the conversion of a primary amide to a primary amine with one fewer carbon atom, using bromine and an aqueous alkali (usually NaOH or KOH).
The general reaction is:
R−CONHX2+BrX2+4NaOHR−NHX2+2NaBr+NaX2COX3+2HX2O
Or, in a more compact form:
R−CONHX2BrX2,NaOHR−NHX2+COX2
Step-by-Step Mechanism
- Deprotonation: The amide nitrogen is deprotonated by the strong base, forming an amide anion.
R−CONHX2+OHX−R−CONHX−+HX2O
- Bromination: The amide anion attacks bromine, forming an N-bromoamide.
R−CONHX−+BrX2R−CONHBr+BrX−
- Second deprotonation: The N-bromoamide is deprotonated again by the base.
R−CONHBr+OHX−R−CONBrX−+HX2O
- Rearrangement: The alkyl group migrates from the carbonyl carbon to the nitrogen, with simultaneous loss of bromide ion. This forms an isocyanate.
R−CONBrX−R−N=C=O+BrX−
- Hydrolysis: The isocyanate reacts with water to form a carbamic acid, which spontaneously decarboxylates (loses COX2) to give the primary amine.
R−N=C=O+HX2OR−NH−COOHR−NHX2+COX2
The rearrangement step (step 4) is the key. The alkyl group migrates with its bonding electrons — it is a 1,2-shift from carbon to the electron-deficient nitrogen. This is why the carbon skeleton shortens by one carbon.
Key Points for Exams
- Starting material: Primary amide (R−CONHX2) only. Secondary or tertiary amides do not undergo this reaction.
- Reagents: BrX2 and NaOH (or KOH). Sometimes ClX2 can be used instead of BrX2, but bromine is more common.
- Product: Primary amine with one fewer carbon.
- By-products: NaBr, NaX2COX3, HX2O (or COX2 if written in the simplified form). …
Part (a)
(i) Ethylamine's small –NH₂ H-bonds strongly with water and its ethyl group is small, so it dissolves; aniline's large hydrophobic benzene ring dominates (and its N lone pair is partly delocalised into the ring), so it is water-insoluble.
(ii) –NH₂ is o/p-directing by resonance, but nitration uses acidic HNO3/H2SO4 which protonates it to −N+H3 — a meta-directing, deactivating group — so a substantial amount of m-nitroaniline forms. …
Part (a): ethylamine is water-soluble (effective H-bonding, small chain) while aniline is not (bulky hydrophobic ring); in acidic nitration aniline is protonated to the meta-directing −NHX3X+, giving substantial m-nitroaniline; amines are nucleophilic because of the N lone pair. Part (b): nitrobenzene → aniline (Sn/HCl then NaOH); ethanamide → methanamine (Hofmann bromamide, one C less); ethanenitrile → ethanamine (LiAlH₄).
Part (a)
- Ethylamine soluble, aniline insoluble. Ethylamine's small –NH₂ group hydrogen-bonds strongly with water, and its short ethyl chain barely disrupts the water structure — so it is very soluble. In aniline the same –NH₂ can H-bond, but the large hydrophobic benzene ring dominates and its lone pair is partly delocalised into the ring (less available for H-bonding), so aniline is only sparingly soluble.
- o/p-directing but gives m-nitroaniline. Free –NH₂ donates its lone pair by resonance and is strongly activating, o/p-directing. Nitration, however, is done in a strongly acidic HNOX3/HX2SOX4 mixture that protonates the amine:
The −NHX3X+ group is electron-withdrawing, deactivating and meta-directing, so a substantial fraction of the product is m-nitroaniline (the o/p isomers come from the small amount of free aniline present). …
CX6HX5NHX2+HX+CX6HX5NHX3X+
Showing the 12 most recent of 27 on this concept.
- CBSE 2026Set 56/1/11 markMCQQ.At low temperature, phenol reacts with dil. HNO3 to yield (A) 2, 4, 6-Trinitrophenol (B) o-Nitrophenol only (C) p-Nitrophenol only (D) ortho-and para-nitrophenol
›Reveal solutionSolution
Phenol is highly activated toward electrophilic substitution, and dilute nitric acid at low temperature acts as a mild nitrating agent. The reaction yields a mixture of ortho- and para-nitrophenol, with the ortho isomer being the major product due to intramolecular hydrogen bonding.
The key to this question lies in understanding two things: the activating power of the phenolic –OH group, and the conditions under which the nitration is carried out.
Phenol has a hydroxyl group directly attached to the benzene ring. The lone pair on oxygen participates in resonance with the ring, pushing electron density into the ortho and para positions. This makes phenol extremely reactive toward electrophilic substitution — far more than benzene or even toluene. In fact, phenol is so activated that it can be nitrated by very mild reagents like dilute nitric acid, which would barely touch benzene.
Now, the specific conditions here are crucial: dilute HNO₃ at low temperature. This is not the concentrated nitric-sulfuric acid mixture used for benzene. Dilute nitric acid is a much weaker nitrating agent. Under these mild conditions, the reaction stops at mononitration. You do not get further substitution to dinitro or trinitro products because the nitro group, once introduced, is strongly deactivating and makes the ring much less reactive toward a second attack.
Let’s walk through the reasoning step by step.
-
Identify the directing effect of –OH.
The hydroxyl group is an ortho-para director. This means the incoming nitro group (NO2+) will attack preferentially at the positions ortho and para to the –OH group. The meta position is not favoured.
-
Consider the reaction conditions.
Dilute HNO₃ at low temperature (typically 0–5°C) is a mild nitrating system. It generates a low concentration of the nitronium ion (NO2+). This is important: a low concentration of electrophile means the reaction is selective, and the high reactivity of phenol ensures that mononitration occurs readily.
-
Predict the product mixture.
Both ortho and para positions are activated. However, the ortho position is sterically hindered by the bulky –OH group. You might expect the para product to dominate on steric grounds. But here, a special effect comes into play: intramolecular hydrogen bonding.
In ortho-nitrophenol, the –OH and –NO₂ groups are close enough to form a strong internal hydrogen bond. This stabilises the ortho isomer significantly. The para isomer cannot form such a bond. As a result, ortho-nitrophenol is actually the major product under these conditions, despite the steric hindrance.
-
Eliminate the other options. …
-
- CBSE 2026Set 56/1/11 markMCQQ.Aniline on direct nitration yields (A) 51%-ortho, 47%-para, 2%-meta derivatives (B) 51%-meta, 47%-ortho, 2%-para derivatives (C) 51%-para, 47%-meta, 2%-ortho derivatives (D) 51%-ortho, 47%-meta, 2%-para derivatives
›Reveal solutionSolution
Direct nitration of aniline is carried out in a strongly acidic medium, in which most of the aniline is protonated to the meta-directing anilinium ion. As a result a large amount of the meta isomer forms alongside para, with ortho suppressed: about 51% para, 47% meta, 2% ortho — option (C).
At first glance the −NH2 group of aniline is a strong activating, ortho/para director, because the nitrogen lone pair can be donated into the ring. That would suggest almost only ortho and para products. But nitration is special, and the answer hinges on the reaction conditions.
-
The medium is strongly acidic. Nitration uses a mixture of concentrated HNO3 and H2SO4. In this medium the basic −NH2 group is protonated, so a large fraction of aniline is present as the anilinium ion, C6H5N+H3.
-
The anilinium ion is meta-directing. Once protonated, the nitrogen carries a full positive charge and can no longer donate its lone pair; instead it withdraws electron density (–I effect) and deactivates the ring, directing the electrophile to the meta position — just like other positively charged / electron-withdrawing substituents.
-
Two directing effects operate at once. The small amount of free (unprotonated) aniline still directs ortho/para, while the anilinium ion directs meta. The net experimental result is an unusually high meta yield:
- para ≈ 51%
- meta ≈ 47%
- ortho ≈ 2% (suppressed by the bulky protonated group and steric/electronic factors)
-
Match with the options.
- (C) 51% para, 47% meta, 2% ortho — matches the experimental data. …
-
- CBSE 2026Set A1 markMCQQ.By which of the following processes, is methyl amine prepared ?(a) Wurtz reaction(b) Hoffmann Bromamide reaction(c) Friedel-Crafts reaction(d) Kolbe's reaction
›Reveal solutionSolution
The Hoffmann bromamide degradation converts an amide to a primary amine with one fewer carbon; acetamide -> methylamine.
In the Hoffmann bromamide (degradation) reaction, an amide is treated with bromine and alkali (KOH), losing the carbonyl carbon and forming a primary amine with one carbon atom fewer:
CH3-CONH2 + Br2 + 4 KOH --> CH3-NH2 + K2CO3 + 2 KBr + 2 H2O
…
- CBSE 2026Set ANNUAL1 markMCQQ.Which one of the following is used in Friedel-Crafts acylation reaction?(a) Anhy. AlCl3(b) Aq. AlCl3(c) NH2NH2(d) NH2OH
›Reveal solutionSolution
Friedel–Crafts acylation needs a genuine, water-free Lewis acid to activate the acyl halide into an electrophilic acylium ion; only anhydrous AlCl3 qualifies among the options.
Mechanism: RCOCl+AlCl3→RCO++AlCl4− The resonance-stabilised acylium ion RCO+ is the actual electrophile that attacks the benzene ring (electrophilic aromatic substitution), giving an aryl ketone after loss of H+ (which recombines with AlCl4− to regenerate HCl and the AlCl3 catalyst).
Why the other options are wrong:
- (b) Aqueous AlCl3: water reacts with and hydrolyses AlCl3 (to Al(OH)3 + HCl), destroying its Lewis-acid character; it also hydrolyses the acyl chloride reagent before it can react — the reaction cannot proceed. …
- CBSE 2026Set ANNUAL1 markMCQQ.Addition reaction of hydrogen Bromide to the unsymmetrical alkene follows(a) a) Anti Markovnikov's rule(b) b) Markovnikov's rule(c) c) Kharash effect(d) d) peroxide effect
›Reveal solutionSolution
[!TLDR]
b) Markovnikov's rule
Why
In the normal (non-peroxide) addition of HBr to an unsymmetrical alkene, H adds to the carbon already bearing more hydrogens and Br to the …
- CBSE 2025Set ANNUAL1 markMCQQ.C6H5CONH2 + Br2 + 4NaOH -----> Product, Product is(a) C6H5COOH(b) C6H5NC(c) C6H5NH2(d) C6H6
›Reveal solutionSolution
Treating a primary amide with Br2 and excess NaOH is the Hofmann bromamide degradation, which shortens the carbon chain by one and gives a primary amine.
C6H5CONH2+Br2+4NaOH→C6H5NH2+2NaBr+Na2CO3+2H2O
…
- CBSE 2025Set ANNUAL1 markMCQQ.Hofmann bromamide degradation reaction is shown by:(a) CH3CH2NH2(b) CH3CH2NO2(c) CH3CONH2(d) CH3CH2CN
›Reveal solutionSolution
The Hofmann bromamide degradation converts a primary AMIDE (RCONH2) into a primary amine with one carbon less, using Br2/KOH — only a compound with the amide (-CONH2) functional group can undergo it.
RCONH2 + Br2 + 4KOH → RNH2 + K2CO3 + 2KBr + 2H2O.
…
- CBSE 2025Set ANNUAL1 markQ.Write the structure and IUPAC name of the amine produced by the Hofmann degradation of benzamide.
›Reveal solutionSolution
Hofmann degradation of benzamide (Br2/KOH) removes the carbonyl carbon as CO2 (via K2CO3) and converts the amide nitrogen directly into the ring's amino group, giving aniline.
The Hofmann bromamide degradation converts an amide, R−CONH2, into a primary amine, R−NH2, with the loss of one carbon atom (as carbonate), when treated with bromine in aqueous/alcoholic potassium hydroxide. Mechanistically it proceeds through an N-bromoamide, then a nitrene/isocyanate intermediate, and finally hydrolysis and decarboxylation of the resulting carbamic acid.
For benzamide, R=C6H5−:
…
- CBSE 2025Set ANNUAL1 markMCQQ.The ortho/para directing group among the following is(a) -COOH(b) -CN(c) -COCH3(d) -NH2
›Reveal solutionSolution
Groups that DONATE electron density into the benzene ring (via resonance or induction) are ortho/para directors; groups that WITHDRAW electron density are meta directors. -NH2 is a strong electron donor; the other three are all electron-withdrawing (carbonyl-type or nitrile) groups.
In electrophilic aromatic substitution, substituents already on a benzene ring direct where the next incoming electrophile attacks, based on how they affect the ring's electron density:
- Ortho/para-directing, activating groups: donate electron density into the ring (by resonance and/or induction), increasing electron density preferentially at the ortho and para positions. Examples: -NH2, -OH, -OCH3, -CH3, halogens (deactivating but still o/p-directing).
- Meta-directing, deactivating groups: withdraw electron density from the ring (usually via a carbonyl-type resonance-withdrawing structure, C=O or C#N conjugated to the ring), leaving the meta position relatively more electron-rich than ortho/para. Examples: -COOH, -CHO, -COR (including -COCH3), -CN, -NO2, -SO3H.
Check each option:
- -COOH (carboxylic acid): the C=O is conjugated with the ring and withdraws electron density -> meta director …
- CBSE 2024Set B1 markQ.Write True or False: Primary amines is prepared by Hoffmann bromamide reaction.
›Reveal solutionSolution
Hofmann bromamide degradation is indeed a standard method for preparing primary amines from amides.
In the Hofmann bromamide degradation reaction, an amide is treated with bromine and concentrated aqueous/ethanolic sodium hydroxide (or KOH). This converts the amide (RCONH2) into a primary amine (RNH2) with the loss of one carbon atom (as CO2, via an isocyanate intermediate):
…
- CBSE 2024Set ANNUAL1 markMCQQ.In Hofmann-bromamide reaction an amide is converted to :(a) Primary amine(b) Secondary amine(c) Tertiary amine(d) Aldehyde
›Reveal solutionSolution
The Hofmann bromamide reaction converts an amide into a primary amine with one carbon atom less than the starting amide.
When an amide (RCONH2) is treated with bromine in aqueous or ethanolic sodium hydroxide, it is converted to a primary amine (RNH2) containing one carbon atom less than the amide. This is called the Hofmann bromamide degradation reaction.
RCONH2 + Br2 + 4NaOH -> RNH2 + Na2CO3 + 2NaBr + 2H2O
…
- CBSE 2024Set ANNUAL1 markMCQQ.Toluene reacts with a halogen in the presence of iron (III) chloride giving ortho and para halo compounds, the reaction is :(a) Electrophillic elimination reaction(b) Electrophillic substitution reaction(c) Nucleophillic substitution reaction(d) Nucleophillic addition reaction
›Reveal solutionSolution
Halogenation of an aromatic ring (toluene) using X2/FeCl3 proceeds through attack of an electrophile on the electron-rich benzene ring, replacing a ring hydrogen — a textbook electrophilic aromatic substitution.
FeCl3 is a Lewis acid catalyst that polarises the halogen molecule, generating an electrophile (X+-like species). This electrophile attacks the electron-rich aromatic ring of toluene (activated further by the ring-activating, ortho/para-directing methyl group) to form an arenium (sigma-complex) intermediate, which then loses H+ to restore aromaticity, giving ortho- and para-halotoluenes: …
🎓Unlock everything free for 14 days
- ✓Full step-by-step solutions
- ✓Concept-first explanations
- ✓Methods, shortcuts & mistakes
- ✓PYQ mapping + timed mock tests
Full access for 14 days. No credit card required.