Q.Alkenes (>C=C<) and carbonyl compounds (>C=O) both contain a π-bond but alkenes show electrophilic addition reactions whereas carbonyl compounds show nucleophilic addition reactions. Explain.
🔒You're viewing a preview — the full solution, concept, methods & PYQ mapping are locked.
🔒 Start your 14-day free trial to unlock the full solution →Concept understanding — Nucleophilic Addition
Nucleophilic Addition – From Intuition to Precision
Imagine you have a molecule with a carbon–oxygen double bond — a carbonyl group (C=O). That oxygen is greedy for electrons; it pulls them away from carbon, leaving the carbon slightly positive (δ+) and the oxygen slightly negative (δ−). Now, if you bring a species that is rich in electrons (a nucleophile, meaning "nucleus-loving"), it will naturally be attracted to that electron-deficient carbon. The nucleophile attacks the carbon, the π bond breaks, and the oxygen picks up a proton (or some other electrophile) to become stable. That, in a nutshell, is nucleophilic addition.
The key idea: a nucleophile adds across a polar multiple bond (usually C=O or C≡N), breaking the π bond and forming two new sigma bonds.
The Precise Statement
Nucleophilic addition is a reaction in which a nucleophile (an electron-rich species) forms a sigma bond with an electrophilic carbon atom of a polar multiple bond (typically a carbonyl group, C=O, or a nitrile, C≡N), while the π bond breaks. The resulting intermediate then captures a proton (or another electrophile) to give a neutral product.
In general form:
RX2C=O+NuX−HX+RX2C(OH)Nu
The nucleophile (NuX−) attacks the carbonyl carbon; the oxygen becomes negatively charged; then a proton (HX+) from the medium attaches to the oxygen, yielding an alcohol.
Why It Happens – The Driving Force
The carbonyl carbon is electrophilic because:
- Oxygen is more electronegative than carbon, so the C=O bond is polarised: CXδ+=OXδ−.
- The π bond is weaker than a σ bond, so it can break relatively easily.
A nucleophile (like OHX−, CNX−, or NHX3) has a lone pair or a negative charge. It seeks positive centres. The attack forms a new σ bond, and the π electrons move entirely to oxygen, creating an alkoxide ion (RX2C−OX−). This intermediate is then quenched by a proton.
A common mistake: thinking the nucleophile attacks the oxygen. No — oxygen is already electron-rich; the nucleophile goes to the carbon because it is electron-deficient.
A Concrete Example – Addition of HCN to a Ketone
Take acetone (CHX3COCHX3) and hydrogen cyanide (HCN). In the presence of a base, CNX− (the nucleophile) attacks the carbonyl carbon:
CHX3COCHX3+CNX−CHX3C(OX−)(CN)CHX3
The alkoxide intermediate then picks up a proton from HCN (or from water) to give a cyanohydrin:
CHX3C(OX−)(CN)CHX3+HX+CHX3C(OH)(CN)CHX3
The product is acetone cyanohydrin. Notice: two new sigma bonds formed (C−CN and O−H), and the π bond is gone.
What Makes a Good Nucleophile?
Strong nucleophiles are usually negatively charged or have lone pairs:
- OHX−, CNX−, NHX2X−, CHX3OX−, HX− (from hydride reagents like NaBHX4 or LiAlHX4)
- Neutral but polarisable: NHX3, HX2O (weaker, but can add under acidic conditions) …
Why this formula?
Nucleophilic Addition: Why the Mechanism Works the Way It Does
Let's build this from first principles — understanding why nucleophilic addition happens, not just memorising the steps.
1. The Core Problem: Why Does Addition Happen at All?
A carbonyl group (C=O) has a polarised double bond:
- Oxygen is more electronegative than carbon → it pulls electron density toward itself.
- This creates a partial positive charge on carbon (δ+) and a partial negative charge on oxygen (δ−).
CXδ+=OXδ−
Key insight: The carbon is electron-deficient — it wants electrons. A nucleophile (Nu⁻) is electron-rich — it wants to give electrons. This is a natural match.
2. The Two-Step Mechanism (Why Two Steps?)
Step 1: Nucleophilic Attack (Slow, Rate-Determining)
The nucleophile donates its lone pair to the electrophilic carbonyl carbon.
NuX−+C=O[Nu−C−O]X−
Why this happens:
- The π bond between C and O breaks — the electrons move entirely to oxygen.
- Oxygen now has a full negative charge (alkoxide ion).
- The carbon changes from sp2 (trigonal planar) to sp3 (tetrahedral).
This step is slow because the π bond must break — it requires energy.
Step 2: Protonation (Fast)
The negatively charged oxygen picks up a proton (HX+) from the solvent or acid.
[Nu−C−O]X−+HX+Nu−C−OH
Why this happens:
- The alkoxide ion is a strong base — it wants to neutralise its charge.
- Protonation gives a stable neutral alcohol product.
3. The Key Formula: Rate Law Derivation
For a general nucleophilic addition:
NuX−+RX2C=Okproducts
The rate law comes from the slow step (Step 1):
Rate=k[Nu−][RX2C=O]
Why this form?
- The reaction is bimolecular — two species must collide with correct orientation.
- Doubling either concentration doubles the rate (first order in each).
- This is second order overall.
Exam tip: This is why nucleophilic addition is often called addition-elimination when followed by loss of a leaving group (like in acyl substitution), but here it's just addition.
4. Why the Tetrahedral Intermediate Forms (And Why It's Unstable)
The intermediate is tetrahedral (sp3 hybridised carbon):
- Bond angles: ~109.5°
- Four groups around carbon: Nu, R, R', O⁻
Why it's unstable:
- The negative charge on oxygen is high-energy.
- The tetrahedral geometry is sterically crowded (especially with bulky R groups).
- The intermediate collapses quickly — either back to starting materials or forward to product. …
The key difference lies in the polarisation of the π-bond. In alkenes, the C=C bond is non-polar (or weakly polarisable), so it acts as an electron-rich centre that attracts electrophiles. In carbonyl compounds, the C=O bond is strongly polarised due to the high electronegativity of oxygen (δ−), leaving the carbon atom electron-deficient (δ+). This carbon is therefore susceptible to attack by nucleophiles.
Reasoning in steps:
- Alkene: The π-electrons are evenly shared. An electrophile (e.g., Br2, H+) is attracted to this electron density, forming a carbocation intermediate — hence, electrophilic addition. …
The key difference lies in the polarity of the π-bond. Alkenes have a non-polar C=C bond, making them electron-rich and thus reactive toward electrophiles. Carbonyl compounds have a highly polar C=O bond (oxygen is more electronegative), making the carbon electron-deficient and thus reactive toward nucleophiles.
The question asks you to explain why two seemingly similar functional groups — both containing a π-bond — undergo completely different types of addition reactions. The answer is not about the π-bond itself, but about who holds the electron density in that bond.
1. The nature of the π-bond in alkenes
In an alkene, the two carbon atoms have nearly identical electronegativity (both are about 2.5 on the Pauling scale). The π-electrons are shared almost equally between them. This means the π-bond is non-polar and the electron cloud is symmetrically distributed above and below the plane of the molecule.
Because the π-electrons are loosely held and exposed, they act as a source of electron density. An alkene is therefore nucleophilic — it "likes" to attack electron-deficient species. So when an electrophile (like Br2 or H+) approaches, it is attracted to this electron-rich region. The reaction proceeds via electrophilic addition: the electrophile attacks first, forming a carbocation, followed by attack of a nucleophile.
Alkene + Electrophile → Carbocation intermediate → Product (Electrophilic Addition)
2. The nature of the π-bond in carbonyl compounds
In a carbonyl group (C=O), oxygen is significantly more electronegative (3.5) than carbon (2.5). This creates a large dipole: the π-electrons are pulled strongly toward oxygen. The result is that the carbon becomes electron-deficient (partial positive charge, δ+) and the oxygen becomes electron-rich (partial negative charge, δ−).
So the carbonyl carbon is electrophilic — it "likes" to be attacked by electron-rich species. A nucleophile (like CN−, OH−, or NH3) is attracted to this positive centre. The reaction proceeds via nucleophilic addition: the nucleophile attacks first, forming a tetrahedral intermediate, followed by protonation.
Carbonyl compound + Nucleophile → Tetrahedral intermediate → Product (Nucleophilic Addition)
3. Why can't alkenes undergo nucleophilic addition?
If you tried to attack an alkene with a nucleophile, the nucleophile would be repelled by the electron-rich π-cloud. There is no positive centre to attract it. The alkene simply does not have an electrophilic site. So nucleophilic addition is not possible for simple alkenes.
A common mistake is to think that because both have a π-bond, they should react similarly. The π-bond is just the "reactive site" — but who controls the reaction (electrophile vs nucleophile) depends entirely on the polarity of that bond.
--- …
Method: Electronic Structure & Polarity Analysis
This method explains reaction type by comparing the electron density distribution and bond polarity of the two functional groups.
Step 1: Analyse the π-bond in alkenes (>C=C<)
- The carbon–carbon double bond is non-polar (both atoms have similar electronegativity).
- The π-electron cloud is symmetrically distributed above and below the plane.
- This makes the π-bond electron-rich — it acts as a nucleophile (attacks electron-deficient species).
Result: Alkenes attract electrophiles (positive or electron-deficient species). Hence, they undergo electrophilic addition.
Step 2: Analyse the π-bond in carbonyl compounds (>C=O)
- Oxygen is more electronegative than carbon.
- The π-bond is polarised: δ+ on carbon, δ− on oxygen.
- The carbon atom is electron-deficient (electrophilic centre).
Result: The carbonyl carbon attracts nucleophiles (electron-rich species). Hence, they undergo nucleophilic addition.
Step 3: Compare the reaction mechanisms
| Feature | Alkene (>C=C<) | Carbonyl (>C=O) |
|---|---|---|
| Polarity | Non-polar | Highly polar |
| Electron-rich centre | The π-cloud | Oxygen (lone pairs) |
Here is a breakdown of the common mistakes students make on this concept, along with how to avoid them.
Mistake 1: Forgetting the "Why" — Focusing only on the π-bond
The Mistake:
Students see both have a π-bond and assume the reaction mechanism must be the same. They memorize the answer ("alkenes are electron-rich, carbonyls are polar") without understanding why the polarity dictates the mechanism.
How to Avoid:
Always start with the electron density of the π-bond.
-
Alkenes (>C=C<): The π-bond is non-polar (or very weakly polar). The electron cloud is evenly shared between the two carbon atoms. This makes the π-bond electron-rich.
- Result: It is attracted to electrophiles (electron-loving species, like Br2 or H+). Hence, Electrophilic Addition.
-
Carbonyls (>C=O): Oxygen is much more electronegative than carbon. It pulls the π-electron cloud strongly towards itself.
- Result: The carbon becomes δ+ (electron-deficient) and the oxygen becomes δ− (electron-rich).
- Result: The carbon is now a target for nucleophiles (nucleus-loving, electron-rich species, like CN− or NH3). Hence, Nucleophilic Addition.
Key Takeaway: Don't just say "it's polar." Explain that the polarity creates an electron-deficient carbon, which is the specific site for nucleophilic attack.
Mistake 2: Confusing the Role of the Nucleophile/Electrophile
The Mistake:
Students think a nucleophile attacks the π-bond directly (like an electrophile does in alkenes). They don't realize the nucleophile attacks the carbon atom, not the bond itself.
How to Avoid:
Visualize the mechanism step-by-step.
- Alkene + Electrophile: The electrophile (E+) attacks the electron cloud of the π-bond. The π-bond breaks, and both electrons go to form a bond with E+.
- Carbonyl + Nucleophile: The nucleophile (Nu−) attacks the δ+ carbon atom directly. The π-bond breaks, and both electrons move to the oxygen (forming an alkoxide ion, O−).
Key Takeaway: The nucleophile doesn't "attack the π-bond." It attacks the partially positive carbon that the π-bond creates.
Mistake 3: Ignoring the Fate of the Leaving Group
The Mistake:
Students think nucleophilic addition is the same as nucleophilic substitution (like in alkyl halides). They forget that in addition, the π-bond breaks but no atom leaves.
How to Avoid:
Remember the name: Addition.
- Alkene: Two new σ-bonds are formed. No atoms are lost.
- Carbonyl: One π-bond is broken, and two new σ-bonds are formed (one to the nucleophile, one to the H+ that comes later). No atom leaves the carbonyl carbon.
Key Takeaway: The carbonyl carbon gains two new groups. It doesn't lose one. This is addition, not substitution.
Mistake 4: Forgetting the Second Step (Protonation) …
Showing the 12 most recent of 15 on this concept.
- TG EAPCET 2026Set eng-2026-05-09-FN1 markMCQQ.The correct statement about an SN1 reaction is (A) It is favoured by polar aprotic solvent (B) It follows second order kinetics (C) It involves racemisation (D) It involves Walden inversion
›Reveal solutionSolution
The key idea is that an SN1 reaction proceeds through a planar carbocation intermediate, which allows attack from either face, leading to racemisation. The correct statement is that it involves racemisation.
The concept behind SN1 reactions is all about the stability of the intermediate. Unlike SN2 reactions, which happen in one smooth step with backside attack, SN1 reactions are stepwise. The first and slowest step is the departure of the leaving group, forming a carbocation. This carbocation is sp²-hybridised and therefore planar — a flat, trigonal structure. Because it’s planar, the nucleophile can attack from either the top or the bottom with equal probability. If the starting material was a single enantiomer (chiral), this equal attack from both sides gives a 50:50 mixture of the two enantiomers — a racemic mixture. That’s why racemisation is the hallmark of SN1 reactions.
Now let’s examine each option carefully:
-
Option (A): “It is favoured by polar aprotic solvent”
Polar aprotic solvents (like acetone or DMF) are great for SN2 reactions because they don’t solvate the nucleophile too tightly, leaving it “naked” and reactive. But SN1 reactions need to stabilise the carbocation intermediate, and polar protic solvents (like water or ethanol) do that better by solvating both the carbocation and the leaving group. So SN1 is actually favoured by polar protic solvents, not aprotic. This statement is false.
-
Option (B): “It follows second order kinetics”
The rate of an SN1 reaction depends only on the concentration of the substrate (the alkyl halide), because the slow step is the dissociation of the leaving group. The nucleophile doesn’t appear in the rate law. So the reaction is first order, not second order. This statement is false.
-
Option (C): “It involves racemisation” …
-
- TG EAPCET 2026Set eng-2026-05-10-AN1 markMCQQ.Which of the following will be the product of Hell-Volhard-Zelinsky reaction? (A) R–CH2OH (B) R–CH–COOH∣Cl (C) R–C–NH2∣∣O (D) R–C–Cl∣∣O
›Reveal solutionSolution
The Hell-Volhard-Zelinsky (HVZ) reaction replaces the α‑hydrogen of a carboxylic acid with a halogen (usually bromine or chlorine) in the presence of a catalytic amount of PBr₃, giving an α‑halo acid. The product is an α‑chloro carboxylic acid, which matches option (B).
The Hell-Volhard-Zelinsky reaction is a classic method for selectively halogenating the carbon atom adjacent to the carboxyl group (the α‑position) of a carboxylic acid. The key insight is that without a catalyst, carboxylic acids do not easily undergo electrophilic substitution at the α‑carbon because the carboxyl group is deactivating. The HVZ reaction uses a small amount of PBr₃ (or red phosphorus + Br₂) to first form an acyl bromide, which is much more reactive toward enolization. Once the enol forms, bromination (or chlorination) occurs at the α‑position, and subsequent hydrolysis gives the α‑halo acid.
Let’s walk through the reasoning step by step.
-
Identify the reaction type.
The HVZ reaction specifically converts a carboxylic acid (R–CH₂–COOH) into an α‑halo carboxylic acid (R–CHX–COOH, where X = Cl, Br, or I). The halogen ends up on the carbon next to the –COOH group.
-
Examine each option.
- (A) R–CH₂OH is an alcohol — no carboxyl group, no halogen. Not the HVZ product.
- (B) R–CH(Cl)–COOH is an α‑chloro carboxylic acid. This matches exactly what HVZ produces.
- (C) R–CO–NH₂ is an amide. HVZ does not introduce nitrogen.
- (D) R–CO–Cl is an acyl chloride. While an acyl chloride is an intermediate in the HVZ mechanism, the final product after aqueous workup is the α‑halo acid, not the acyl chloride.
-
Confirm the mechanism (briefly).
- PBr₃ converts the carboxylic acid to an acyl bromide.
- The acyl bromide enolizes more readily, and the enol attacks Br₂ (or Cl₂) at the α‑position. …
-
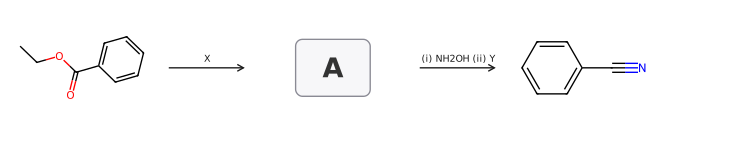
- TG EAPCET 2024Set ap-2024-05-07-FN1 markMCQQ.In the following reaction sequence, what are X and Y respectively? C6H5−COOEtXA(i) NH2OH(ii) YC6H5−CN (A) H2∣Pd; (CH3CO)2O (B) H2∣Pd; C6H5SO2Cl, Pyridine (C)(i) DIBAL-H(ii) H2O; (CH3CO)2O (D)(i) DIBAL-H(ii) H2O; C6H5SO2Cl
›Reveal solutionSolution
Since NH2OH must react with an aldehyde, A = benzaldehyde — which means X is the partial reducing agent DIBAL-H (then H2O). Y then dehydrates the aldoxime to benzonitrile: (CH3CO)2O. Option (C).
The concept first
The cleanest way through any reagent-identification question is to work backwards from the product and ask: what functional group must have been sitting there just before?
(a) The last step tells you what A is. Hydroxylamine, NH2OH, is an ammonia derivative: it condenses with a carbonyl compound to give an oxime:
R−CHO+NH2OH⟶R−CH=N−OH (aldoxime)+H2O
An aldoxime has the formula R−CH=N−OH; take one molecule of water out of it and you are left with R−C≡N — a nitrile. That is the standard aldehyde → nitrile route:
C6H5CHONH2OHC6H5CH=N−OH−H2OC6H5C≡N
A ketoxime could not do this (no H on the carbon), so A must be an aldehyde — benzaldehyde.
(b) The first step is therefore a controlled reduction. We need C6H5COOEt→C6H5CHO: the ester must be reduced only as far as the aldehyde and no further. This is exactly the job of DIBAL-H (di-isobutylaluminium hydride) at low temperature — it delivers a single hydride, giving a tetrahedral intermediate that survives until aqueous work-up releases the aldehyde. Catalytic H2/Pd is not a reagent that converts an ester into an aldehyde at all (Rosenmund's controlled hydrogenation works on an acyl chloride, not an ester, and needs the poisoned catalyst).
(c) Y is a dehydrating agent. Converting the aldoxime to the nitrile is a loss of water, so Y must dehydrate: acetic anhydride (CH3CO)2O is the standard reagent (it mops up the water as acetic acid).
Step-by-step
- Target: C6H5−C≡N (benzonitrile), reached from A by (i) NH2OH then (ii) Y.
- Deduce A. NH2OH makes an oxime; only an aldoxime can be dehydrated to a nitrile. Hence A is the aldehyde C6H5CHO (benzaldehyde). …
- TG EAPCET 2024Set eng-2024-05-10-FN1 markMCQQ.Acrolein (X) is one of the chemicals formed when O3 and NO2 react with unburnt hydrocarbons present in the polluted air. The structure of 'X' is (A) CH3−CH=CH2 (B) CH2=CH−CHO (C) CH2=CH−CN (D) CH3CO(OO)NO2
›Reveal solutionSolution
Acrolein is the simplest unsaturated aldehyde, formed from incomplete combustion or atmospheric reactions of hydrocarbons; its structure is CH2=CH−CHO, so the correct option is (B).
Acrolein is a well-known compound in atmospheric chemistry and organic chemistry. The key here is to recognize that "acrolein" is the common name for propenal — an aldehyde with a carbon‑carbon double bond. The question describes it as forming when ozone (O3) and nitrogen dioxide (NO2) react with unburnt hydrocarbons in polluted air. This is a classic photochemical smog reaction, and acrolein is a major irritant produced. So we need the structure that matches the name "acrolein."
Let’s examine each option:
-
Option (A): CH3−CH=CH2
This is propene, an alkene. It has no oxygen atom, so it cannot be an aldehyde or any oxygen-containing compound. Acrolein must contain oxygen (the “-oin” ending hints at an aldehyde or alcohol derivative). So this is incorrect.
-
Option (B): CH2=CH−CHO
This is propenal: a three-carbon chain with a double bond between C1 and C2, and an aldehyde group (−CHO) at the end. The common name for propenal is indeed acrolein. It is formed in smog from the oxidation of hydrocarbons. This matches perfectly.
-
Option (C): CH2=CH−CN
This is acrylonitrile, a nitrile (contains a cyano group, −CN). It has no aldehyde group and is not called acrolein. So this is wrong.
-
Option (D): CH3CO(OO)NO2 …
-
- TG EAPCET 2024Set eng-2024-05-11-FN1 markMCQQ.What is the IUPAC name of the product Y in the given reaction sequence?
[!FORMULA] CHO∣(CHOH)4HCNXH+/H2OY∣CH2OH
(A) 2,3,4,5,6,7-hexahydroxyheptanoic acid (B) 2,3,4,5,6-pentahydroxyhexanoic acid (C) 3,4,5-trihydroxyheptanoic acid (D) 3,4,5-trihydroxyhexanoic acid›Reveal solutionSolution
The reaction sequence involves extending an aldohexose by one carbon via cyanohydrin formation, followed by hydrolysis of the nitrile to a carboxylic acid, resulting in a 7-carbon polyhydroxy carboxylic acid named 2,3,4,5,6,7-hexahydroxyheptanoic acid.
Concept and Intuition: Chain Lengthening of Sugars
This problem demonstrates a classic method for increasing the carbon chain length of aldoses (sugars containing an aldehyde group), often referred to as the Kiliani-Fischer synthesis. The core idea is to convert the aldehyde group into a nitrile, which then extends the carbon backbone by one carbon. This new nitrile group can subsequently be transformed into other functional groups, such as a carboxylic acid, as seen in this problem.
Why this approach works:
- Aldehyde Reactivity: Aldehydes are electrophilic at their carbonyl carbon. They readily react with nucleophiles like cyanide (CN⁻).
- Cyanohydrin Formation: The addition of HCN to an aldehyde forms a cyanohydrin, which contains both a hydroxyl group and a nitrile group on the same carbon. This reaction effectively adds a new carbon atom to the chain.
- Nitrile Hydrolysis: Nitriles (-C≡N) are versatile functional groups that can be hydrolyzed under acidic or basic conditions to yield carboxylic acids (-COOH). This transformation is robust and allows for the conversion of the newly introduced carbon into a carboxylic acid functionality.
By combining these two steps, we can systematically lengthen the carbon chain of an aldose and introduce a carboxylic acid group at the new terminal end.
Let's break down the reaction sequence step by step:
1. Identify the Starting Material
The given structure is:
CHO∣(CHOH)4∣CH2OH
This represents an aldohexose. Let's count the carbons:
- One aldehyde carbon (CHO)
- Four secondary alcohol carbons (CHOH)
- One primary alcohol carbon (CH₂OH) Total carbons = 1+4+1=6 carbons. So, the starting material is a 6-carbon sugar (an aldohexose), like glucose, mannose, or galactose. For the purpose of naming the final product's carbon skeleton, the specific stereochemistry of the hydroxyl groups is not required.
2. Step 1: Reaction with HCN to form X
The first step is the reaction of the aldohexose with hydrogen cyanide (HCN). This is a cyanohydrin formation reaction.
- The aldehyde group (CHO) is the reactive site.
- The cyanide ion (CN⁻) acts as a nucleophile, attacking the electrophilic carbonyl carbon.
- The carbonyl oxygen is protonated to form a hydroxyl group.
- This reaction adds one carbon atom to the chain.
The transformation is:
R-CHO+HCN⟶R-CH(OH)-CN
Applying this to our aldohexose:
Original aldohexose: CHO−(CHOH)4−CH2OH (6 carbons)
Product X (cyanohydrin): CN−CH(OH)−(CHOH)4−CH2OH
- Carbon Count: The chain has now been extended by one carbon, so product X has 6+1=7 carbons.
- Functional Groups: Product X contains one nitrile group (-CN) and six hydroxyl groups (-OH). The carbon that was originally the aldehyde carbon (C1) is now a new chiral center (C2 in the new numbering scheme) bearing a hydroxyl group.
Cyanohydrin Formation:
R-CHO+HCN⇌R-CH(OH)-CN
This reaction is reversible and typically catalyzed by a base.
3. Step 2: Hydrolysis of X with H⁺/H₂O to form Y
The second step involves the hydrolysis of product X (the cyanohydrin) under acidic conditions (H+/H2O).
- Nitriles (-CN) are readily hydrolyzed to carboxylic acids (-COOH) in the presence of acid (or base) and water.
- The hydroxyl groups in the molecule are stable under these conditions and remain unchanged.
The transformation is:
R’-CNH+/H2OR’-COOH
Applying this to product X:
Product X: CN−CH(OH)−(CHOH)4−CH2OH
Product Y: COOH−CH(OH)−(CHOH)4−CH2OH
- Carbon Count: The carbon chain length remains 7 carbons.
- Functional Groups: Product Y contains one carboxylic acid group (-COOH) and six hydroxyl groups (-OH). …
- TG EAPCET 2023Set ap-2023-05-11-FN1 markMCQQ.The hybridisation of carbon atoms from left to right in the given compound are respectively H2C=C=CH−C≡N (A) sp3,sp,sp2,sp (B) sp2,sp2,sp2,sp (C) sp,sp2,sp,sp2 (D) sp2,sp,sp2,sp
›Reveal solutionSolution
The hybridisation of a carbon atom is determined by its steric number (number of sigma bonds + number of lone pairs). For the given compound, the hybridisations from left to right are sp2,sp,sp2,sp.
The hybridisation of a central atom in a molecule is a concept that describes the mixing of atomic orbitals to form new hybrid orbitals, which are then used to form sigma bonds and accommodate lone pairs. The type of hybridisation (e.g., sp, sp2, sp3) is directly related to the number of electron domains (sigma bonds and lone pairs) around the atom, often called the steric number.
Here's how to determine the hybridisation for each carbon atom:
- Understand the Steric Number Rule: The steric number for an atom is calculated as:
Steric Number=(Number of sigma bonds)+(Number of lone pairs)
Once the steric number is known, the hybridisation can be determined:
*   Steric Number = 2 $\implies$ $\mathrm{sp}$ hybridisation
*   Steric Number = 3 $\implies$ $\mathrm{sp^2}$ hybridisation
*   Steric Number = 4 $\implies$ $\mathrm{sp^3}$ hybridisation

> [!IMPORTANT]
> Remember that a single bond contains one sigma bond. A double bond contains one sigma bond and one pi bond. A triple bond contains one sigma bond and two pi bonds. Only the sigma bonds contribute to the steric number for hybridisation. Lone pairs also contribute to the steric number. Carbon atoms in stable organic compounds typically do not have lone pairs.
2. Draw the Lewis Structure (or interpret the condensed formula):
The given compound is H2C=C=CH−C≡N. Let's number the carbon atoms from left to right for clarity:
H2C1=C2=CH3−C4≡N
-
Determine Hybridisation for Carbon 1 (H2C=):
- This carbon is bonded to two hydrogen atoms and double-bonded to Carbon 2.
- Number of sigma bonds: 2 (to H) + 1 (to C2, from the double bond) = 3
- Number of lone pairs: 0 (carbon typically forms 4 bonds and has no lone pairs)
- Steric Number = 3 + 0 = 3
- Hybridisation = sp2
-
Determine Hybridisation for Carbon 2 (=C=):
- This carbon is double-bonded to Carbon 1 and double-bonded to Carbon 3.
- Number of sigma bonds: 1 (to C1, from the double bond) + 1 (to C3, from the double bond) = 2 …
- TG EAPCET 2022Set eng-2022-07-19-AN1 markMCQQ.A primary alcohol was reacted with pyridinium chlorochromate (PCC), which resulted in a product P. The product P on treatment with ammoniacal silver nitrate solution produces (A) Anhydride of carboxylic acid (B) Aldehyde (C) Amide (D) Carboxylate anion
›Reveal solutionSolution
PCC oxidises a primary alcohol to an aldehyde (P). An aldehyde reacts with ammoniacal silver nitrate (Tollens’ reagent) to give the carboxylate anion. The correct option is (D).
The key here is to track the oxidation state of carbon through two sequential reactions. PCC is a mild oxidising agent — it stops at the aldehyde stage for primary alcohols, unlike stronger oxidants like KX2CrX2OX7 or KMnOX4 which would push all the way to the carboxylic acid. So product P is an aldehyde.
Now, ammoniacal silver nitrate is Tollens’ reagent. It contains the diamminesilver(I) ion, [Ag(NHX3)X2]X+, in a basic medium. This reagent is famous for the silver mirror test — it oxidises an aldehyde to a carboxylate anion while reducing silver(I) to metallic silver. The reaction happens in basic solution, so the immediate product is the carboxylate salt, not the free acid.
Let’s walk through it step by step.
- First reaction: primary alcohol + PCC PCC (CX5HX5NHX+CrOX3ClX−) is a selective oxidant. It converts a primary alcohol (R−CHX2OH) to an aldehyde (R−CHO) without over-oxidising.
R−CHX2OH+PCCR−CHO
So product P is an aldehyde.
- Second reaction: aldehyde + Tollens’ reagent Tollens’ reagent is prepared by adding aqueous ammonia to silver nitrate until the initially formed AgX2O precipitate just dissolves. The active species is [Ag(NHX3)X2]X+OHX−. The aldehyde is oxidised to a carboxylate ion, and silver(I) is reduced to metallic silver (the mirror).
R−CHO+2[Ag(NHX3)X2]X++2OHX−R−COOX−+2Ag(s)+4NHX3+HX2O
Notice the product is R−COOX−, the carboxylate anion — not the free carboxylic acid, because the medium is basic.
- What about the other options?
- Anhydride of carboxylic acid: requires two carboxyl groups to couple, not formed here.
- Aldehyde: that’s P, not the final product. …
- TG EAPCET 2022Set eng-2022-07-20-AN1 markMCQQ.Ammonia is a Lewis base because it is (A) Electron pair donor (B) Electron pair acceptor (C) Proton donor (D) Proton acceptor
›Reveal solutionSolution
Ammonia donates its lone pair of electrons to form a coordinate bond, making it a Lewis base. The correct answer is (A) Electron pair donor.
The Lewis theory of acids and bases is broader than the older Brønsted–Lowry theory. A Lewis base is any species that can donate an electron pair, while a Lewis acid is any species that can accept an electron pair. This definition focuses entirely on electron pair transfer, not on protons.
Ammonia (NH3) has a nitrogen atom with five valence electrons. Three of these are used in covalent bonds with hydrogen atoms, leaving one lone pair of electrons on the nitrogen. This lone pair is available to be shared with an electron-deficient species — for example, when ammonia reacts with a proton (H+) to form the ammonium ion (NH4+), the lone pair forms a coordinate bond with the proton.
-
Identify the key feature of ammonia. The nitrogen atom in NH3 has a lone pair of electrons. This is the structural feature that determines its chemical behaviour as a base.
-
Apply the Lewis definition. A Lewis base is defined as an electron pair donor. Since ammonia can donate its lone pair, it qualifies as a Lewis base.
-
Check the other options for clarity.
- (B) Electron pair acceptor — This is the definition of a Lewis acid, not a base. Ammonia does not accept an electron pair; it donates one.
- (C) Proton donor — This is the Brønsted–Lowry definition of an acid. Ammonia does not donate a proton; it accepts one (forming NH4+). …
-
- TG EAPCET 2022Set eng-2022-07-20-AN1 markMCQQ.The process in which colloids, when subjected to DC electric field move towards an electrode is (A) Brownian movement (B) Tyndall effect (C) Peptization (D) Electrophoresis
›Reveal solutionSolution
The key idea is that the directed motion of colloidal particles under an applied DC electric field is called electrophoresis. The correct option is (D).
Colloids are mixtures where tiny particles (1–1000 nm) are dispersed in a continuous medium. These particles often carry a surface charge — either positive or negative — due to adsorption of ions from the solution. When you apply a direct current (DC) electric field across such a colloidal system, the charged particles experience a force and begin to migrate toward the electrode of opposite charge. This phenomenon is not random; it is a directed, field-driven movement.
The question asks for the specific name of this process. Let’s examine each option to see why only one fits.
-
Brownian movement is the random, zigzag motion of colloidal particles caused by collisions with solvent molecules. It has nothing to do with an electric field — it’s purely thermal and statistical. So (A) is wrong.
-
Tyndall effect is the scattering of light by colloidal particles, making a beam of light visible through the colloid. Again, no electric field is involved — it’s an optical property. So (B) is wrong.
-
Peptization is the process of converting a freshly precipitated substance back into a colloidal state by adding an electrolyte. It’s a chemical dispersion method, not an electric-field-driven motion. So (C) is wrong. …
-
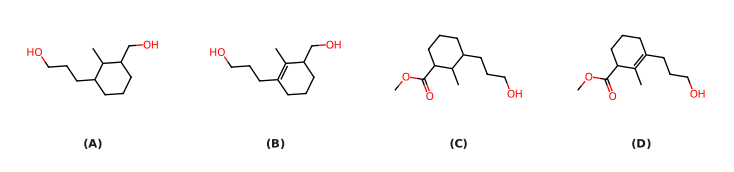
- TG EAPCET 2022Set eng-2022-07-20-AN1 markMCQQ.The major product in the following transformation is A cyclohexene ring bearing a −CH2CH2CHO side chain on one alkene carbon, a −CH3 group on the other alkene carbon, and a −CO2Me group on the adjacent ring carbon NaBH4 (A) [FIGURE] The ring double bond reduced, the side chain converted to −CH2CH2CH2OH and the −CO2Me converted to −CH2OH (B) [FIGURE] The ring double bond retained, the side chain converted to −CH2CH2CH2OH and the −CO2Me converted to −CH2OH (C) [FIGURE] The ring double bond reduced, the side chain converted to −CH2CH2CH2OH and the −CO2Me group retained (D) [FIGURE] The ring double bond retained, the side chain converted to −CH2CH2CH2OH and the −CO2Me group retained
›Reveal solutionSolution
Sodium borohydride reduces the side-chain aldehyde to a primary alcohol and nothing else — the ring double bond and the methyl ester are untouched. Option (D).
The concept: the reactivity window of NaBH4
Hydride reagents are graded by how nucleophilic the hydride is:
Reagent Reduces NaBH4 (mild) aldehydes, ketones, acid chlorides LiAlH4 (powerful) aldehydes, ketones, esters, acids, amides, nitriles H2/Pd alkenes, alkynes The B−H bond is far less polar than Al−H, so BH4− is a weak hydride donor and can attack only the most electrophilic carbonyls. An ester carbonyl is deactivated by resonance donation from its −OR group and is inert to NaBH4. An isolated alkene is a nucleophile, not an electrophile — a hydride will never attack it.
This chemoselectivity is what makes NaBH4 so valuable in synthesis, and it is exactly what the question probes.
Step 1 — Identify every functional group
- Ring C=C (trisubstituted alkene, not conjugated with the ester).
- Side-chain −CH2CH2CHO (an aldehyde).
- −CO2Me (a methyl ester).
Step 2 — Apply NaBH4
- Aldehyde: reduced.
−CH2CH2CHONaBH4−CH2CH2CH2O−work-up−CH2CH2CH2OH
- Ester: untouched (too weakly electrophilic). …
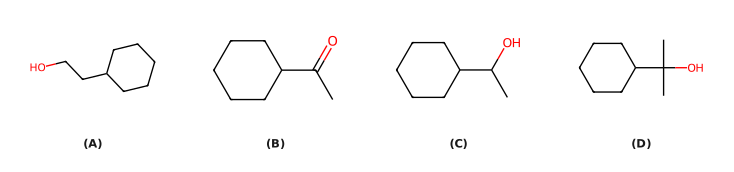
- TG EAPCET 2022Set eng-2022-07-20-AN1 markMCQQ.The major product of the following reactions is Bromocyclohexane (i) Mg; (ii) CO2; (iii) H3O+; (iv) SOCl2; (v) (CH3)2Cd (A) [FIGURE] Cyclohexane ring bearing a −CH2CH2OH group (B) [FIGURE] Cyclohexane ring bearing a −CO−CH3 group (1-cyclohexylethan-1-one) (C) [FIGURE] Cyclohexane ring bearing a −CH(OH)CH3 group (D) [FIGURE] Cyclohexane ring bearing a −C(OH)(CH3)2 group
›Reveal solutionSolution
Grignard → CO2 gives cyclohexanecarboxylic acid; SOCl2 gives the acyl chloride; dimethylcadmium then delivers just one methyl to give the ketone C6H11COCH3 — option (B).
The concept: a three-move synthesis of a ketone
Each reagent has one clean job:
1. Grignard formation
C6H11BrMg, dry etherC6H11MgBr
The carbon that carried Br is now nucleophilic — a complete polarity reversal.
2. Carboxylation with CO2
The Grignard attacks the electrophilic carbon of CO2 (usually dry ice):
C6H11MgBr+CO2→C6H11COOMgBrH3O+C6H11COOH
This is the standard way of adding exactly one carbon as a −COOH.
3. Making the acid chloride
C6H11COOH+SOCl2→C6H11COCl+SO2↑+HCl↑
4. The cadmium step — the point of the question
R2Cd has a much more covalent C−M bond than a Grignard. It is nucleophilic enough to attack an acid chloride, but not a ketone:
2C6H11COCl+(CH3)2Cd⟶2C6H11CO−CH3+CdCl2 …
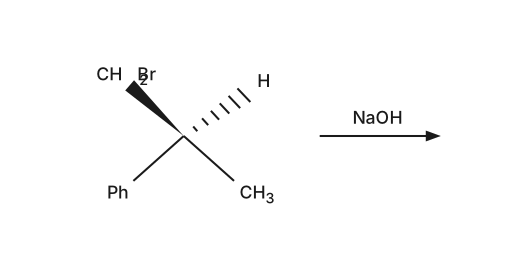
- TG EAPCET 2021Set ap-2021-08-09-AN1 markMCQQ.The correct statement about the following chemical reaction is [FIGURE] (A) It is an SN2 reaction with inversion of configuration of the reactant (B) It is an SN2 reaction with retention configuration of the reactant (C) It is an SN1 reaction with retention of configuration of the reactant (D) It is an SN1 reaction with racemisation
›Reveal solutionSolution
The Br sits on a primary −CH2− group, so OH− attacks by SN2; but the chiral carbon is the adjacent one and none of its bonds break, so the configuration is retained. The answer is (B).
The concept first — inversion is a statement about the attacked carbon. Students memorise "SN2 = inversion" and then apply it to the whole molecule. That is the trap. In SN2 the nucleophile attacks the carbon bearing the leaving group from the side opposite the leaving group; the three other bonds on that carbon flip through like an umbrella (Walden inversion). If the molecule's stereocentre is somewhere else, nothing at all happens to it — its four bonds are untouched, so its spatial arrangement (and its R/S label) is preserved.
Step 1 — Identify the substrate. From the drawing, one carbon carries four different groups: CH2Br (bold wedge), H (dashed), Ph and CH3. So the compound is
Ph−C∗H(CH3)−CH2Br
where ∗ marks the stereocentre.
Step 2 — Locate the leaving group. The bromine is on the −CH2− carbon. That carbon carries two hydrogens, one Br and one carbon — it is a primary carbon and it is not a stereocentre.
Step 3 — Decide the mechanism. A primary carbon cannot form a stable carbocation, so an SN1 path is out. With aqueous/alcoholic NaOH the strong nucleophile OH− attacks directly:
HO−+Ph−CH(CH3)−CH2Br⟶Ph−CH(CH3)−CH2OH+Br−
This is a one-step, bimolecular SN2 reaction (rate =k[RBr][OH−]). …
🎓Unlock everything free for 14 days
- ✓Full step-by-step solutions
- ✓Concept-first explanations
- ✓Methods, shortcuts & mistakes
- ✓PYQ mapping + timed mock tests
Full access for 14 days. No credit card required.