The Core Intuition: Who Wants to Leave, and Who Can Wait?
Imagine you're at a party where the host (the leaving group) is about to leave. The party (the reaction) happens in two stages. First, the host walks out the door — that's the slow, painful step. Then, a new guest (the nucleophile) rushes in to take the empty spot.
The SN1 reaction works exactly like this: the leaving group leaves first, forming a carbocation intermediate. The nucleophile attacks after the leaving group is gone. This means the rate of the reaction depends only on how easily the leaving group can leave — it does not depend on the nucleophile at all.
So the question becomes: What makes a carbocation form easily? The answer is stability. A carbocation that is more stable will form faster and last longer, making the SN1 reaction faster.
The Precise Statement of SN1 Reactivity Order
SN1 Reactivity Order (for alkyl halides):
Allylic≈Benzyl>3∘>2∘≫1∘≈Methyl
This is the order of how fast the SN1 reaction proceeds. Let's unpack why.
Why This Order? The Stability Ladder
A carbocation is a carbon with only six electrons in its valence shell — it's electron-deficient and positively charged. The more you can spread out (delocalize) that positive charge, the more stable the carbocation becomes.
1. Methyl and 1° Carbocations: The Unstable Ones
A methyl carbocation (CHX3X+) has no alkyl groups attached to the positive carbon. There is zero electron-donating effect to stabilize the charge. It is so unstable that it practically never forms in an SN1 reaction — the reaction simply doesn't happen.
A primary (1°) carbocation has one alkyl group attached. Alkyl groups are weakly electron-donating (through hyperconjugation and inductive effect), so it's slightly more stable than methyl — but still far too unstable to form under normal SN1 conditions.
Watch out
Never say "SN1 happens on a primary carbon" in an exam. It is essentially impossible under standard conditions because the carbocation is too unstable.
2. Secondary (2°) Carbocations: The Borderline Case
A secondary carbocation has two alkyl groups donating electron density. It is moderately stable — stable enough to form, but only under certain conditions (like a good leaving group and a polar protic solvent). SN1 reactions on secondary carbons are possible, but they are slower than on tertiary carbons.
3. Tertiary (3°) Carbocations: The Sweet Spot
Three alkyl groups donate electron density to the positive carbon. This makes the carbocation very stable. Tertiary alkyl halides undergo SN1 reactions readily — they are the classic example.
4. Allylic and Benzylic: The Champions
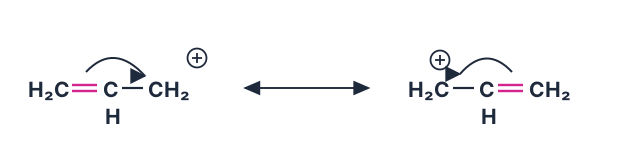
These are special cases. In an allylic carbocation, the positive charge is adjacent to a carbon-carbon double bond. The π electrons of the double bond can delocalize the positive charge onto the second carbon:
CHX2=CH−CHX2X+↔+CHX2−CH=CHX2
Two resonance structures of the allyl carbocation, CH2=CH-CH2+, with the positive charge delocalised between the two terminal carbons (Structure I and Structure II)
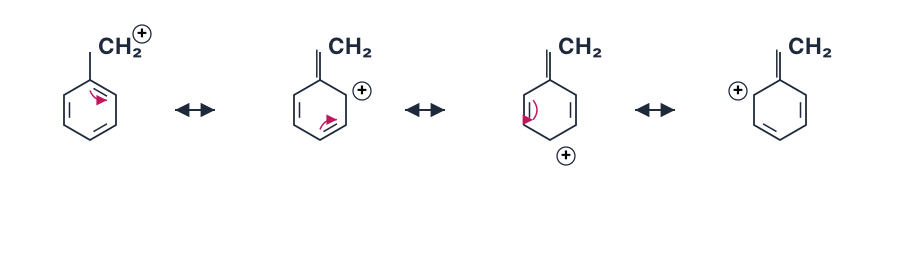
In a benzylic carbocation, the positive charge is adjacent to a benzene ring. The π system of the ring delocalizes the charge across multiple carbons:
CX6HX5−CHX2X+↔(several resonance structures)
Here are those resonance structures — the positive charge cycles from the CH₂ carbon onto the ortho and para positions of the ring:
Four resonance structures of the benzyl carbocation, C6H5-CH2+, with curved electron-pushing arrows showing the positive charge delocalising from the exocyclic CH2 carbon onto the ortho and para ring carbons
This resonance stabilization makes allylic and benzylic carbocations even more stable than tertiary ones. They form the fastest in SN1 reactions.
The Complete Picture in a Table
| Carbocation Type | Stability | SN1 Reactivity | Example |
The SN1 reaction (Substitution Nucleophilic Unimolecular) proceeds via a carbocation intermediate. The reactivity order is determined entirely by the stability of this carbocation — because the rate-determining step is its formation.
The Core Principle
The rate law for SN1 is:
Rate=k[RX]
Only the substrate appears in the rate law — the nucleophile does not participate in the slow step. The slow step is:
RXslowR++X−
Thus, anything that stabilizes the carbocation (R⁺) lowers the activation energy and increases the reaction rate.
The Reactivity Order
For alkyl halides (RX), the SN1 reactivity order is:
This is purely about hyperconjugation and inductive effect.
Tertiary carbocation: Three alkyl groups donate electron density via hyperconjugation (C–H σ bonds overlap with empty p orbital) and +I effect. This spreads the positive charge over more atoms → most stable.
Secondary: Two alkyl groups → less stabilization.
Primary: Only one alkyl group → very little stabilization.
Methyl: No alkyl groups → least stable (only inductive effect from H atoms, which is negligible).
Key formula: The number of α-hydrogens (H on carbons adjacent to the positive carbon) determines hyperconjugation. More α-H → more resonance structures → more stable.
2. Why Allylic and Benzylic Are Even Faster
These carbocations are resonance-stabilized.
Allylic carbocation: The positive charge is delocalized over two carbon atoms via π-bond conjugation:
CH2=CH−CH2+⟷CH2+−CH=CH2
Two resonance structures of the allyl carbocation with the positive charge shared between the two terminal CH2 carbons
Benzylic carbocation: The positive charge is delocalized into the aromatic ring:
C6H5−CH2+⟷several resonance forms involving the ring
Those resonance forms look like this:
Resonance structures of the benzylic carbocation C6H5-CH2+ showing the positive charge delocalised onto the ortho and para carbons of the benzene ring
This resonance stabilization is so powerful that even a primary allylic or benzylic carbocation is more stable than a tertiary alkyl carbocation.
The target, 1-phenylethanamine (C6H5CH(NH2)CH3), has ONE FEWER carbon than the starting amide, 2-phenylpropanamide (C6H5CH(CH3)CONH2) - that carbon loss is the signature of the Hofmann bromamide degradation, not a simple reduction. Br2 in aqueous NaOH degrades the amide with loss of the carbonyl carbon as CO2, giving exactly the target amine. …
1-Phenylethanamine has one fewer carbon than the starting amide, 2-phenylpropanamide - so this conversion needs the Hofmann bromamide degradation (Br2 in aqueous NaOH), which expels the carbonyl carbon as CO2, not a hydride reduction (which keeps all three carbons). The correct reagent is Br2 in aqueous NaOH, option (ii).
Compare the starting material and the target carbon-by-carbon. 2-Phenylpropanamide is C6H5-CH(CH3)-CONH2: a three-carbon amide chain (the carbonyl carbon, the CH bearing the phenyl group, and the terminal methyl). The target, 1-phenylethanamine, is C6H5-CH(NH2)-CH3: only two carbons remain, with the amino group on the carbon that used to bear the phenyl substituent - the carbonyl carbon is gone entirely.
Losing a carbon while converting the amide group to an amine is exactly what the Hofmann bromamide degradation does: treating a primary amide with Br2 in aqueous/alcoholic NaOH forms an N-bromoamide, which rearranges (via an isocyanate intermediate) with loss of the carbonyl carbon as CO2, leaving the remaining group bonded directly to NH2. Applied here, 2-phenylpropanamide converts straight to 1-phenylethanamine.
Now check the other options:
(i) excess H2/Pt does not reduce an amide carbonyl under ordinary catalytic hydrogenation conditions.
(iii) NaBH4/methanol is too mild to reduce an amide. …
This reaction converts a primary amide into a primary amine with one fewer carbon atom in the chain. The reagent used is bromine in aqueous sodium hydroxide (Br2/NaOH).
Method: Hofmann Rearrangement
Step 1 — Identify the starting material and target
Concept: Hofmann Rearrangement vs. Reduction of Amides
The reaction converts an amide (2-phenylpropanamide) into a primary amine (1-phenylethanamine). The key observation: the carbon chain loses one carbon atom — the amide carbon is lost as CO2.
✗ Mistake 1: Choosing LiAlH4 (Option D) or NaBH4 (Option C)
Why students do this:
They see "amide → amine" and immediately think of reduction. LiAlH4 is a strong reducing agent that converts amides to amines.
Why it's wrong here:
LiAlH4 reduces amides to amines without changing the carbon skeleton.
2-phenylpropanamide (C6H5CH(CH3)CONH2) would give 2-phenylpropanamine (C6H5CH(CH3)CH2NH2).
But the product asked is 1-phenylethanamine (C6H5CH(NH2)CH3) — one carbon fewer.
NaBH4 is even weaker and does not reduce amides at all under normal conditions.
How to avoid:
Always count the carbons in the reactant and product. If the chain shortens, reduction is not the answer — look for a rearrangement or degradation reaction.
✗ Mistake 2: Choosing H2/Pt (Option A)
Why students do this:
They think "hydrogenation" or "catalytic reduction" will convert the amide to an amine.
Why it's wrong:
H2/Pt reduces alkenes, alkynes, nitro groups, and nitriles — but not amides. Amides are very stable toward catalytic hydrogenation.
How to avoid:
Memorise the functional groups that H2/catalyst reduces: