Q.Assertion: Rate constants determined from Arrhenius equation are fairly accurate for simple as well as complex molecules.
Reason: Reactant molecules undergo chemical change irrespective of their orientation during collision.
🔒You're viewing a preview — the full solution, concept, methods & PYQ mapping are locked.
🔒 Start your 14-day free trial to unlock the full solution →Concept understanding — Arrhenius Equation Plot
The Arrhenius Equation Plot: Why Temperature Changes Reaction Speed
You already know that heating things up makes reactions go faster. A cold chai takes forever to dissolve sugar; hot chai does it in seconds. But how much faster? And is there a pattern that holds for every reaction?
That pattern is the Arrhenius equation, and plotting it in a clever way reveals something fundamental about how molecules need to collide to react.
The core idea: an energy barrier
Imagine a ball sitting in a valley. To get to the next valley, it must first be pushed up over a hill. That hill is the activation energy (Ea) — the minimum energy two molecules need to have when they collide, for the reaction to happen.
At a low temperature, most molecules move slowly. Only a tiny fraction have enough energy to climb that hill. Raise the temperature, and suddenly many more molecules have the required energy. The fraction of molecules with energy ≥Ea is given by the Boltzmann distribution:
fraction=e−Ea/RT
where R is the gas constant and T is the absolute temperature (in Kelvin). This exponential is the heart of the story.
The Arrhenius equation (precise statement)
The rate constant k of a reaction depends on temperature as:
k=Ae−Ea/RT
- k = rate constant (how fast the reaction proceeds)
- A = pre-exponential factor (frequency of collisions, times a steric factor — how often molecules hit in the right orientation)
- Ea = activation energy (J/mol or kJ/mol)
- R = 8.314 J/(mol·K)
- T = temperature in Kelvin
k is not the reaction rate itself — it's the proportionality constant in the rate law. But for a fixed concentration, a larger k means a faster reaction.
Why plot it? The linear trick
The equation k=Ae−Ea/RT is exponential in 1/T. That's hard to eyeball. But take the natural logarithm of both sides:
lnk=lnA−REa⋅T1
This is the equation of a straight line:
y=c+mx
where:
- y=lnk
- x=1/T
- slope m=−Ea/R
- intercept c=lnA
So if you measure k at several temperatures and plot lnk versus 1/T, you get a straight line — provided the reaction follows Arrhenius behaviour (most do, over moderate temperature ranges).
Always use Kelvin for T. Celsius will give you a curved mess because 1/T is not linear in Celsius.
What the plot tells you
From the slope, you get Ea:
Ea=−(slope)×R
A steep negative slope means a large Ea — the reaction is very sensitive to temperature. A shallow slope means a small Ea — temperature doesn't affect it much.
From the intercept, you get A:
A=eintercept
This tells you about the collision frequency and orientation factor. A high A means molecules are colliding often and in the right geometry.
A typical Arrhenius plot looks like this
| T (K) | k (s⁻¹) | 1/T (K⁻¹) | lnk |
|---|---|---|---|
| 300 | 0.0012 | 0.00333 | -6.72 |
| 310 | 0.0028 | 0.00323 | -5.88 |
| 320 | 0.0061 | 0.00313 | -5.10 |
| 330 | 0.0125 | 0.00303 | -4.38 |
Plot lnk (y-axis) vs 1/T (x-axis). The points fall on a straight line. Draw the best-fit line, measure its slope, and compute Ea. …
Why this formula?
Arrhenius Equation Plot: Why It Holds
The Arrhenius equation is not a guess — it emerges from a deep physical picture of how molecules react. Let's build that understanding step by step.
The Core Idea: Molecules Need Energy to React
For a reaction to occur, molecules must collide with enough energy to break existing bonds and form new ones. This minimum energy is called the activation energy (Ea).
But not all collisions succeed — only those with kinetic energy ≥Ea lead to a reaction.
The Key Formula
The Arrhenius equation is:
k=Ae−Ea/(RT)
Where:
- k = rate constant
- A = pre-exponential factor (frequency of collisions with correct orientation)
- Ea = activation energy (J/mol)
- R = gas constant (8.314 J/mol·K)
- T = absolute temperature (K)
Why the Exponential Term Appears
Step 1: The Boltzmann Distribution
Molecules in a gas or liquid have a distribution of kinetic energies. The fraction of molecules with energy ≥E is given by the Boltzmann factor:
Fraction=e−E/(kBT)
For molar quantities, replace kB with R:
Fraction=e−Ea/(RT)
This is not arbitrary — it comes from statistical mechanics. The exponential arises because the probability of a molecule having energy E decreases exponentially as E increases.
Step 2: Rate Depends on This Fraction
The rate constant k is proportional to:
- The collision frequency (how often molecules meet)
- The fraction of collisions with energy ≥Ea
Thus:
k∝(collision frequency)×e−Ea/(RT)
The collision frequency is captured by A, giving:
k=Ae−Ea/(RT)
Why the Plot is Linear
Take the natural logarithm of both sides:
lnk=lnA−REa⋅T1
This is of the form y=mx+c, where:
- y=lnk
- x=1/T
- Slope m=−Ea/R
- Intercept c=lnA
Thus, plotting lnk vs 1/T gives a straight line — this is the Arrhenius plot.
What the Slope Tells Us
From the slope: …
The key idea is that the Arrhenius equation ignores molecular orientation, so it is reliable only for simple molecules/collisions — not for complex ones.
Step 1: The Arrhenius equation k=Ae−Ea/RT assumes every sufficiently energetic collision reacts. This holds reasonably well for simple, near-spherical molecules, but for complex molecules the calculated k does not agree well with the observed value, because the equation takes no account of orientation. So the assertion ("fairly accurate for simple as well as complex molecules") is false. …
The Arrhenius equation gives accurate rate constants for simple reactions, but for complex molecules, steric factors and orientation matter — so the assertion is false. The reason is also false because molecules must have proper orientation for a reaction to occur. Both statements are incorrect.
The key here is to understand what the Arrhenius equation actually models and where it falls short. The equation k=Ae−Ea/RT assumes that every collision with sufficient energy leads to a reaction — but that's only true for simple, small molecules. For complex molecules, the orientation during collision becomes critical, and the Arrhenius equation overestimates the rate unless corrected by a steric factor.
Let's break down each statement.
-
Assertion: "Rate constants determined from Arrhenius equation are fairly accurate for simple as well as complex molecules."
This is incorrect. For simple molecules (like two atoms colliding), the Arrhenius equation works well because almost every energetic collision leads to reaction. But for complex molecules (large organic compounds, for instance), the molecule must hit the reactive site in the correct orientation. The Arrhenius equation ignores this — it assumes all collisions with enough energy are effective. That's why we introduce the steric factor P in collision theory: k=PZe−Ea/RT, where P is often much less than 1 for complex molecules. So the assertion is false.
-
Reason: "Reactant molecules undergo chemical change irrespective of their orientation during collision." …
Method: Conceptual Evaluation of Assertion-Reason Statements
This is not a calculation problem — it tests your understanding of the Arrhenius equation and collision theory. Evaluate each statement independently, then check if the reason correctly explains the assertion.
Step 1: Evaluate the Assertion
"Rate constants determined from Arrhenius equation are fairly accurate for simple as well as complex molecules."
- The Arrhenius equation, k=Ae−Ea/RT, implicitly assumes that every collision with sufficient energy is effective.
- For simple reactions (small, near-spherical molecules — e.g. atoms or diatomic species), this assumption holds well, and calculated k values agree closely with experiment.
- For complex molecules, the reacting groups must also be correctly oriented at the moment of collision; the plain Arrhenius equation takes no account of this, so calculated and observed rate constants do not agree well.
- Conclusion: The assertion is incorrect — Arrhenius-equation rate constants are reliable for simple molecules but NOT for complex ones.
Step 2: Evaluate the Reason
"Reactant molecules undergo chemical change irrespective of their orientation during collision." …
Here’s a breakdown of the common mistakes students make on this specific assertion-reason question about the Arrhenius equation, along with how to avoid each.
Mistake 1: Confusing “Accuracy” with “Universality”
- The Error: Students think the Arrhenius equation works perfectly for all reactions, including complex ones. They then mark the assertion as correct without checking the nuance.
- Why It’s Wrong: The Arrhenius equation is empirical and works well for simple, elementary reactions. For complex molecules or multi-step reactions, the rate constant often deviates because the equation assumes a single activation energy barrier, which isn’t true for complex pathways.
- How to Avoid: Remember: Arrhenius is accurate for simple, single-step reactions. For complex reactions, the plot of lnk vs. 1/T may be curved, not linear. The assertion says “fairly accurate for simple as well as complex molecules” — this is incorrect because it overstates the equation’s range.
Mistake 2: Misinterpreting the Reason (Orientation Factor)
- The Error: Students think the reason sounds scientific and matches the topic of collision theory, so they assume it must be correct.
- Why It’s Wrong: The reason states: “Reactant molecules undergo chemical change irrespective of their orientation during collision.” This is false. In reality, proper orientation is critical for effective collisions (the steric factor in collision theory). If orientation is wrong, no reaction occurs even if energy is sufficient.
- How to Avoid: Link the reason to collision theory:
- Effective collision = sufficient energy (Arrhenius) + proper orientation.
- The reason denies the orientation requirement, which is a classic mistake. Always check if a statement contradicts basic collision theory.
Mistake 3: Assuming “Both Correct” Without Checking the Link
- The Error: Students see two statements that both sound plausible and pick option (i) or (ii) without verifying if the reason actually explains the assertion.
- Why It’s Wrong: Even if both were correct (they aren’t here), the reason (orientation is irrelevant) does not explain why the Arrhenius equation is accurate. The accuracy of Arrhenius depends on the reaction being elementary, not on orientation.
- How to Avoid: For assertion-reason questions, always ask: “Does the reason directly cause or justify the assertion?” Here, the reason is about orientation during collision, while the assertion is about the equation’s accuracy — they are unrelated concepts.
Mistake 4: Forgetting the “Complex Molecules” Trap …
- COMEDK 2026Set 2026-M1 markMCQQ.Identify the correct values to be plotted in the graph, the slope of which can be used to determine the activation energy of a reaction. (A) lnk vs T1 (B) lnk vs T (C) Tlnk vs T (D) lnkT vs T1
›Reveal solutionSolution
The Arrhenius equation gives a linear relationship between lnk and 1/T, with slope −Ea/R, so plotting lnk vs 1/T yields the activation energy. The correct option is (A).
The key idea is the Arrhenius equation, which describes how the rate constant k depends on temperature T and activation energy Ea:
k=Ae−Ea/(RT)
Taking the natural logarithm gives:
lnk=lnA−REa⋅T1
This is in the form y=mx+c, where y=lnk, x=1/T, the slope m=−Ea/R, and the intercept c=lnA. So a plot of lnk vs 1/T gives a straight line whose slope directly yields Ea.
Now let’s examine each option:
-
Option (A): lnk vs 1/T
This matches the linear form above. The slope is −Ea/R, so Ea=−(slope)×R. This is the standard method.
-
Option (B): lnk vs T
The Arrhenius equation is exponential in T, not linear. Plotting lnk vs T gives a curve, not a straight line, so the slope is not constant and cannot be used to find Ea directly.
-
Option (C): Tlnk vs T
Substituting lnk=lnA−Ea/(RT) gives Tlnk=TlnA−RT2Ea. This is not linear in T; it’s a more complicated function. No simple slope gives Ea.
-
Option (D): lnkT vs T1 …
-
- KCET 2026Set D31 markMCQQ.The activation energy for the reaction X → Y is 150 kJ mol−1. The change in enthalpy for the above reaction is -135 kJ mol−1. Then the activation energy for Y → X is (A) 280 kJ mol−1 (B) 285 kJ mol−1 (C) 270 kJ mol−1 (D) 15 kJ mol−1
›Reveal solutionSolution
Use the energy-diagram relation between enthalpy change and the forward/reverse activation energies: ΔH=Ea(forward)−Ea(reverse).
Step 1 — Recall the relation between ΔH and activation energies
On a reaction energy profile, the enthalpy change of the reaction equals the difference between the forward and reverse activation energies:
ΔH=Ea(forward)−Ea(reverse)
Step 2 — Substitute known values
Here, Ea(forward,X→Y)=150 kJ mol−1 and ΔH=−135 kJ mol−1: …
- COMEDK 2024Set 2024-A1 markMCQQ.The temperature (T) and rate constant (k) for a first order reaction R→P, was found to follow the equation logk=−(2000) T1+8.0. The pre-exponential factor 'A' and activation energy Ea, respectively are: [Given: R=8.314 J K−1 mol−1] (A) 8.0 s−1 and 16.6 kJ mol−1 (B) 6.0 s−1 and 108 kJ mol−1 (C) 1×10⁸ s−1 and 38.3 kJ mol−1 (D) 1×10⁻⁸ s−1 and 16.6 kJ mol−1
›Reveal solutionSolution
Matching the given line to the logarithmic Arrhenius equation: the intercept 8.0 gives A=108 s−1, and the slope −2000 gives Ea=2000×2.303×R=38.3 kJ mol−1.
The Arrhenius equation in logarithmic form is
logk=logA−2.303RTEa
Comparing with the experimental fit logk=−(2000)T1+8.0:
Intercept: logA=8.0⇒A=1×10⁸ s−1. …
- COMEDK 2023Set 2023-E1 markMCQQ.Given below are graphs showing the variation in velocity constant with temperature on Kelvin scale. Identify the graph which represents Arrhenius equation. (A) D (B) C (C) A (D) B
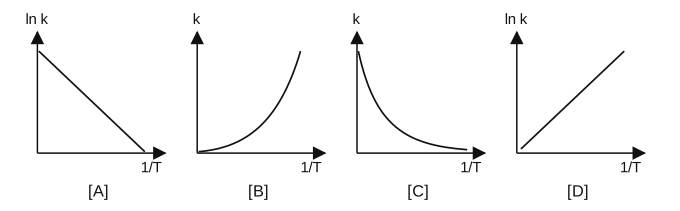
›Reveal solutionSolution
[!TLDR]
Taking logs of the Arrhenius equation gives a straight line of negative slope on lnk vs 1/T axes, which is graph [A]; the option letter for graph A is (C).
Concept
This is the standard CBSE/NCERT Class-12 Chemical Kinetics result. The Arrhenius equation relates the rate (velocity) constant to temperature as k=Ae−Ea/RT, where A is the frequency factor and Ea the activation energy. Its usefulness comes from its linearised (logarithmic) form.
Solution
Start from
k=Ae−Ea/RT.
Take the natural logarithm of both sides:
lnk=lnA−REa(T1).
Comparing with y=c+mx (with y=lnk, x=1/T), this is a straight line whose slope is m=−Ea/R (negative, since Ea>0) and whose intercept is lnA.
Examining the four graphs:
- [A] lnk vs 1/T, straight line of negative slope — matches the linearised Arrhenius form exactly.
- [B] k vs 1/T, rising exponential curve — wrong (not a lnk axis, wrong trend).
- [C] k vs 1/T, decaying curve — this is k itself vs 1/T, not the linear Arrhenius plot. …
- KCET 2022Set B-31 markMCQQ.A first order reaction is half completed in 45 min. How long does it need 99.9% of the reaction to be completed? (A) 10 Hours (B) 20 Hours (C) 5 Hours (D) 7.5 Hours
›Reveal solutionSolution
For a first-order reaction, 99.9% completion is a fall to 1/1000 of the initial concentration — almost exactly 10 half-lives, i.e. 10×45 min=7.5 hours.
Step 1 — The first-order rate law.
For a first-order reaction,
t=k2.303log[A][A]0
and the half-life is independent of concentration:
t1/2=k0.693
Step 2 — Find k from the given half-life.
k=t1/20.693=45 min0.693=0.0154 min−1
Step 3 — Translate "99.9% complete" into a concentration ratio.
If 99.9% of A has reacted, only 0.1% remains:
[A]=0.001[A]0⟹[A][A]0=1000
Step 4 — Substitute.
t99.9%=0.01542.303log(1000)=0.01542.303×3
t99.9%=149.5×3=448.6 min≈450 min
Step 5 — Convert to hours.
t=60450=7.5 hours
Step 6 — The elegant shortcut (worth knowing). …
- KCET 2022Set B-31 markMCQQ.Half-life of a reaction is found to be inversely proportional to the fifth power its initial concentration, the order of reaction is (A) 5 (B) 6 (C) 3 (D) 4
›Reveal solutionSolution
Use the general half-life–concentration relation for an nth order reaction, t1/2∝[A]01−n, and match the given exponent.
Step 1 — Why half-life depends on the initial concentration at all.
For a reaction of order n in a single reactant A, the rate law is
−dtd[A]=k[A]n
Integrating this from [A]0 at t=0 to [A] at time t (for n=1) gives
n−11([A]n−11−[A]0n−11)=kt
Step 2 — Put [A]=[A]0/2 to get the half-life.
By definition t=t1/2 when half the reactant is consumed:
kt1/2=n−11([A]0n−12n−1−[A]0n−11)=(n−1)[A]0n−12n−1−1
t1/2=k(n−1)2n−1−1⋅[A]0n−11
Everything in front of the concentration term is a constant, so the whole concentration dependence is
t1/2∝[A]0n−11i.e.t1/2∝[A]01−n
Step 3 — Match the exponent given in the question.
The problem states the half-life is inversely proportional to the fifth power of the initial concentration:
t1/2∝[A]051
Comparing exponents of [A]0 in the denominator:
n−1=5⟹n=6 …
- KCET 2021Set B-21 markMCQQ.For a reaction A+2B →Products, when concentration of B alone is increased half life remains the same. If concentration of A alone is doubled, rate remains the same. The unit of rate constant for the reaction is (A) s−1 (B) Lmol−1s−1 (C) molL−1s−1 (D) atm−1
›Reveal solutionSolution
Deduce the order in each reactant from the two clues, add them to get the overall order, then read off the units from rate=k[]n.
Step 1 — Order with respect to B.
The half-life of a reaction of order n scales as
t1/2∝[]n−11.
It is independent of concentration only when n−1=0, i.e. n=1. Since "when concentration of B alone is increased, half-life remains the same",
order in B=1.
(For a first-order reaction t1/2=0.693/k — no concentration in it, which is exactly the observation.)
Step 2 — Order with respect to A.
"If concentration of A alone is doubled, rate remains the same." If the rate depended on [A]a, doubling would multiply the rate by 2a. Rate unchanged ⇒ 2a=1 ⇒
a=0(zero order in A).
Step 3 — Write the rate law and overall order.
rate=k[A]0[B]1=k[B]
overall order=0+1=1. …
- KCET 2021Set B-21 markMCQQ.Higher order (>3) reactions are rare due to (A) Shifting of equilibrium towards reactants due to elastic collisions (B) Loss of active species on collision (C) Low probability of simultaneous collision of all reacting species (D) Increase in entropy as more molecules are involved
›Reveal solutionSolution
Higher-order steps demand a simultaneous many-body collision, whose probability falls away steeply as the number of colliding molecules rises.
Step 1 — Link order to molecularity for an elementary step.
For an elementary reaction, the order equals the molecularity — the number of species that must collide simultaneously in one act for the reaction to occur. So an order-4 elementary step requires a four-body simultaneous collision.
Step 2 — Apply collision theory.
Collision theory says a collision is fruitful only if the molecules:
- meet at the same point at the same instant,
- carry energy ≥Ea (the threshold energy), and
- arrive with the correct mutual orientation.
A binary (two-body) collision is common. A termolecular (three-body) collision is already rare — three particles must converge on one point at one instant. Requiring four or more to do so drives the probability down catastrophically: each extra body multiplies in another small factor for the coincidence of position, timing and orientation.
Step 3 — The consequence.
Because the collision frequency Z for such an encounter is effectively negligible, and the rate is proportional to it,
rate=Z⋅p⋅e−Ea/RT …
- KCET 2020Set A-11 markMCQQ.For an elementary reaction 2A+3B→4C+D the rate of appearance of C at time 't' is 2.8×10⁻³ mol L−1S−1. Rate of disappearance of B at 't', t will be (A) 41(2.8×10⁻³)mol L−1S−1 (B) 34(2.8×10⁻³)mol L−1S−1 (C) 43(2.8×10⁻³)mol L−1S−1 (D) 2(2.8×10⁻³ mol L−1S−1)
›Reveal solutionSolution
For an elementary reaction, the rate of disappearance of a reactant is related to the rate of appearance of a product by the stoichiometric coefficients. The rate of disappearance of B is 43×2.8×10−3 mol L−1 s−1, which matches option (C).
The key idea here is that for an elementary reaction, the rate can be written in terms of any reactant or product using the stoichiometric coefficients. The rate of the reaction itself is a single, unique value — but the rate of disappearance of a reactant or appearance of a product is scaled by its coefficient.
For the reaction 2A+3B→4C+D, the rate of reaction r is defined as:
r=−21dtd[A]=−31dtd[B]=41dtd[C]=dtd[D]
The negative signs indicate disappearance (decrease in concentration), and the positive signs indicate appearance (increase in concentration). The fractions ensure that no matter which species you track, you get the same numerical value for r.
We are given the rate of appearance of C: dtd[C]=2.8×10⁻³ mol L−1 s−1.
From the definition above:
41dtd[C]=−31dtd[B]
So:
−dtd[B]=43⋅dtd[C]
The left side −dtd[B] is the rate of disappearance of B (a positive quantity). Substituting the given value:
Rate of disappearance of B=43×(2.8×10−3) mol L−1s−1 …
- KCET 2019Set A-11 markMCQQ.1 L of 2 M CH3COOH is mixed with 1 L of 3M C2H5OH to form an ester. The rate of the reaction with respect to the initial rate when each solution is diluted with an equal volume of water will be (A) 0.25 times (B) 0.5 times (C) 2 times (D) 4 times
›Reveal solutionSolution
The rate law is first order in each reactant; halving both concentrations by dilution multiplies the rate by (21)2=41.
Step 1 — Write the rate law. The acid-catalysed esterification
CH3COOH+C2H5OH⇌CH3COOC2H5+H2O
is first order in each reactant (overall second order):
Rate=k[CH3COOH][C2H5OH].
Step 2 — Initial concentrations (after the original mixing).
Moles: acid =2 M×1 L=2 mol; alcohol =3 M×1 L=3 mol. Total volume =2 L.
[acid]0=22=1 M,[alcohol]0=23=1.5 M
r0=k(1)(1.5)=1.5k.
Step 3 — After each solution is diluted with an equal volume of water.
Each 1 L solution becomes 2 L, so the total volume is now 2+2=4 L, while the moles are unchanged (water only dilutes): …
- KCET 2018Set A-11 markMCQQ.The value of rate constant of a pseudo first order reaction (A) Depends only on temperature (B) Depends on the concentration of reactants present in small amounts (C) Depends on the concentration of reactants present in excess (D) Is independent of the concentration of reactants
›Reveal solutionSolution
In k′=k[B]excess, the excess reactant's (effectively constant) concentration is folded INTO the observed rate constant — so k′ carries its value.
Step 1 — What "pseudo first order" means.
Consider the classic acid-catalysed hydrolysis of an ester:
CH3COOC2H5+H2OH+CH3COOH+C2H5OH
The true rate law is second order:
rate=k[ester][H2O]
But water is the solvent — it is present in enormous excess, so [H2O] is essentially unchanged as the reaction proceeds.
Step 2 — Fold the constant term into the rate constant.
Because [H2O] is effectively constant, define
k′=k[H2O]⟹rate=k′[ester]
The reaction now behaves as first order (hence "pseudo"). The key point: k′ literally contains [H2O] as a factor. Change the amount of the excess reactant and k′ changes proportionally.
Step 3 — Eliminate the other options.
- (A) "only on temperature" — true for a genuine rate constant k (Arrhenius, k=Ae−Ea/RT), but k′ is a composite constant that also carries [B]excess. So "only" is wrong. …
🎓Unlock everything free for 14 days
- ✓Full step-by-step solutions
- ✓Concept-first explanations
- ✓Methods, shortcuts & mistakes
- ✓PYQ mapping + timed mock tests
Full access for 14 days. No credit card required.