Q.A primary amine, RNH2 can be reacted with CH3−X to get secondary amine, R−NHCH3 but the only disadvantage is that 3° amine and quaternary ammonium salts are also obtained as side products. Can you suggest a method where RNH2 forms only 2° amine?
🔒You're viewing a preview — the full solution, concept, methods & PYQ mapping are locked.
🔒 Start your 14-day free trial to unlock the full solution →Concept understanding — Nucleophilic Substitution Reactions
Nucleophilic Addition – From Intuition to Precision
Imagine you have a molecule with a carbon–oxygen double bond — a carbonyl group (C=O). That oxygen is greedy for electrons; it pulls them away from carbon, leaving the carbon slightly positive (δ+) and the oxygen slightly negative (δ−). Now, if you bring a species that is rich in electrons (a nucleophile, meaning "nucleus-loving"), it will naturally be attracted to that electron-deficient carbon. The nucleophile attacks the carbon, the π bond breaks, and the oxygen picks up a proton (or some other electrophile) to become stable. That, in a nutshell, is nucleophilic addition.
The key idea: a nucleophile adds across a polar multiple bond (usually C=O or C≡N), breaking the π bond and forming two new sigma bonds.
The Precise Statement
Nucleophilic addition is a reaction in which a nucleophile (an electron-rich species) forms a sigma bond with an electrophilic carbon atom of a polar multiple bond (typically a carbonyl group, C=O, or a nitrile, C≡N), while the π bond breaks. The resulting intermediate then captures a proton (or another electrophile) to give a neutral product.
In general form:
RX2C=O+NuX−HX+RX2C(OH)Nu
The nucleophile (NuX−) attacks the carbonyl carbon; the oxygen becomes negatively charged; then a proton (HX+) from the medium attaches to the oxygen, yielding an alcohol.
Why It Happens – The Driving Force
The carbonyl carbon is electrophilic because:
- Oxygen is more electronegative than carbon, so the C=O bond is polarised: CXδ+=OXδ−.
- The π bond is weaker than a σ bond, so it can break relatively easily.
A nucleophile (like OHX−, CNX−, or NHX3) has a lone pair or a negative charge. It seeks positive centres. The attack forms a new σ bond, and the π electrons move entirely to oxygen, creating an alkoxide ion (RX2C−OX−). This intermediate is then quenched by a proton.
A common mistake: thinking the nucleophile attacks the oxygen. No — oxygen is already electron-rich; the nucleophile goes to the carbon because it is electron-deficient.
A Concrete Example – Addition of HCN to a Ketone
Take acetone (CHX3COCHX3) and hydrogen cyanide (HCN). In the presence of a base, CNX− (the nucleophile) attacks the carbonyl carbon:
CHX3COCHX3+CNX−CHX3C(OX−)(CN)CHX3
The alkoxide intermediate then picks up a proton from HCN (or from water) to give a cyanohydrin:
CHX3C(OX−)(CN)CHX3+HX+CHX3C(OH)(CN)CHX3
The product is acetone cyanohydrin. Notice: two new sigma bonds formed (C−CN and O−H), and the π bond is gone.
What Makes a Good Nucleophile?
Strong nucleophiles are usually negatively charged or have lone pairs:
- OHX−, CNX−, NHX2X−, CHX3OX−, HX− (from hydride reagents like NaBHX4 or LiAlHX4)
- Neutral but polarisable: NHX3, HX2O (weaker, but can add under acidic conditions) …
Why this formula?
Nucleophilic Substitution Reactions: Why the Key Formulas Hold
Nucleophilic substitution reactions are a cornerstone of organic chemistry. Instead of just memorizing the rate laws, let's understand why they arise from the molecular events.
1. The Two Main Mechanisms: A Tale of Timing
The key formulas (rate laws) for nucleophilic substitution come directly from how many molecules are involved in the rate-determining step (RDS) — the slowest step that controls the overall reaction speed.
SN1: Unimolecular — The Leaving Group Goes First
The Rate Law:
Rate=k[RX]
Why?
The reaction happens in two steps:
- Slow step (RDS): The C–X bond breaks spontaneously, forming a carbocation intermediate. Only the substrate (RX) is involved.
RXslowR++X−
- Fast step: The nucleophile (Nu−) attacks the carbocation.
R++Nu−fastRNu
Since the slow step depends only on the concentration of RX, the rate law has no dependence on [Nu−]. The nucleophile arrives after the carbocation is formed — it cannot affect the speed of the first step.
Key insight: The rate is determined by how easily the leaving group leaves, not by how fast the nucleophile attacks.
SN2: Bimolecular — Simultaneous Attack and Departure
The Rate Law:
Rate=k[RX][Nu−]
Why?
The reaction occurs in one concerted step:
- The nucleophile attacks the carbon from one side at the same time as the leaving group departs from the opposite side.
- Both RX and Nu− must collide with the correct orientation and sufficient energy.
The rate depends on the frequency of productive collisions between the two molecules. This is directly proportional to the product of their concentrations:
Rate∝[RX]×[Nu−]
Key insight: Both partners are involved in the transition state simultaneously — if either is missing, the reaction cannot proceed.
2. The Transition State: Why the Formulas Are Not Just "Given"
For SN2, the transition state has a pentavalent carbon (five bonds partially formed/broken). The energy barrier depends on steric hindrance — bulkier groups around carbon make it harder for the nucleophile to approach, which is why SN2 is favored at primary carbons. …
The key idea is to use acylation followed by reduction instead of direct alkylation. Direct alkylation with CH3X is an uncontrolled SN2 process — the product RNHCH3 is more nucleophilic than RNH2, so it reacts further, giving tertiary amine and quaternary salt.
To install exactly one methyl group on nitrogen, use formylation (a one-carbon acyl group) followed by reduction:
Steps:
-
Formylate the primary amine with formic acid (HCOOH) or methyl formate (HCOOCH3) to form the N-substituted formamide:
RNH2+HCOOH→RNHCHO+H2O
-
Reduce the formamide with a strong reducing agent like LiAlH4. This cleaves the C=O bond and adds two hydrogens, converting the formyl group (−CHO) into a methyl group and giving only the secondary amine:
RNHCHOLiAlH4RNHCH3
(Acetylation with CH3COCl would instead introduce an ethyl group on reduction, giving RNHCH2CH3 — so the one-carbon formyl group is the right choice for an N-methyl product.) …
The key idea is to mask nitrogen's nucleophilicity while installing exactly one extra carbon, so a second alkylation can't happen. Formylating RNH2 with formic acid gives the formamide RNHCHO - its nitrogen lone pair is delocalised into the carbonyl, so the formamide cannot react further with CH3X. Reducing that one C=O with LiAlH4 then converts the formyl group into a methyl group, delivering R−NHCH3 as the only product.
The problem you've described is a classic headache in organic chemistry: direct alkylation of a primary amine with an alkyl halide. When you treat RNH2 with CH3X, the product R−NHCH3 is itself a better nucleophile than the starting amine. So it reacts further - first to the tertiary amine R−N(CH3)2, then to the quaternary ammonium salt R−N+(CH3)3X−. You end up with a messy mixture.
The trick is to install the extra carbon as a one-carbon acyl group first, and only then reduce it down to a methyl group - never by direct alkylation with CH3X.
Here's the step-by-step method:
- Formylate the primary amine React RNH2 with formic acid (HCOOH) or methyl formate (HCOOCH3). This gives the N-substituted formamide:
RNH2+HCOOH→RNHCHO+H2O
The nitrogen is now part of an amide bond - its lone pair is tied up in resonance with the C=O, so this formamide nitrogen is no longer a good nucleophile, and no over-substitution can happen at this stage.
- Reduce the formamide with LiAlH4 LiAlH4 reduces the carbonyl of the formamide all the way to a methylene, turning the one-carbon −CHO group into a −CH3 group: RNHCHOLiAlH4RNHCH3 …
Concept: Selective Monoalkylation of Primary Amines
The problem is that direct alkylation of a primary amine with an alkyl halide is not selective — the product (secondary amine) is itself more nucleophilic than the starting material, so it reacts further to give tertiary amine and quaternary ammonium salt.
Method: Gabriel Phthalimide Synthesis (modified for secondary amines)
This method avoids over-alkylation by using a protected nitrogen that can only be alkylated once.
Steps:
- Form the phthalimide salt Phthalimide (CX6HX4(CO)X2NH) is treated with alcoholic KOH to give potassium phthalimide.
CX6HX4(CO)X2NH+KOHCX6HX4(CO)X2NX−KX++HX2O
- Alkylate with the desired alkyl halide The potassium salt reacts with CHX3−X via SN2 to give N-alkylphthalimide.
CX6HX4(CO)X2NX−KX++CHX3−XCX6HX4(CO)X2N−CHX3+KX
- Hydrolyse to release the pure secondary amine The N-alkylphthalimide is hydrolysed (usually with aqueous NaOH or hydrazine) to give only the secondary amine and phthalic acid.
CX6HX4(CO)X2N−CHX3+2HX2OOHX−CX6HX4(COOH)X2+CHX3NHX2
Note: The product here is actually methylamine (CHX3NHX2), which is a primary amine. To get a secondary amine RNHCHX3, you must start with an N-alkylphthalimide where the alkyl group is R (from step 2 using R−X), then alkylate again? No — that would give tertiary.
Correction for your exact case:
You want only RNHCHX3 (secondary) from RNHX2 and CHX3X.
The Gabriel method as described above gives primary amine after hydrolysis.
To get a secondary amine selectively, use:
Modified Gabriel — Alkylation of a pre-formed N-alkylphthalimide
- First make N-alkylphthalimide from R−X (not CHX3X) and potassium phthalimide.
- Then alkylate that with CHX3−X — but this gives a tertiary product after hydrolysis.
So the correct method for your exact need is:
Hinsberg Test / Separation Method (not a synthesis, but a purification) …
Here’s a breakdown of the common mistakes students make on this concept — alkylation of amines — and how to avoid each.
Common Mistake 1: Forgetting that amines are nucleophilic and will keep reacting
The error:
Students often think that once the secondary amine (R−NHCH3) forms, the reaction stops. In reality, the secondary amine is more nucleophilic than the primary amine, so it reacts further with CH3X to give tertiary amine and quaternary ammonium salt.
How to avoid:
Always remember: each alkylation makes the amine more electron-rich (more alkyl groups = more +I effect), so it becomes a better nucleophile. The reaction does not self-limit — you must actively prevent further alkylation.
Common Mistake 2: Suggesting “use excess RNH2” without understanding the real problem
The error:
Students say “just take a large excess of primary amine” — but this only reduces the relative amount of side products, it does not eliminate them. Some secondary, tertiary, and quaternary products will still form.
How to avoid:
Understand that excess RNH2 is a practical trick to favour monoalkylation, but it is not a perfect method. The question asks for a method that gives only secondary amine — so excess amine is not the answer here.
Common Mistake 3: Confusing the Hinsberg test with a synthetic method
The error:
Students recall that benzenesulfonyl chloride (C6H5SO2Cl) can distinguish primary, secondary, and tertiary amines, and think it can be used to synthesise pure secondary amine.
How to avoid:
The Hinsberg test is an analytical (identification) tool, not a preparative method. You cannot use it to make a secondary amine from a primary one — it forms sulfonamides, not the free amine.
Correct Method (for reference)
The standard exam answer is:
Use the carbylamine reaction (isocyanide formation) followed by reduction.
- React RNH2 with CHCl3 and alcoholic KOH to form an isocyanide (RNC). …
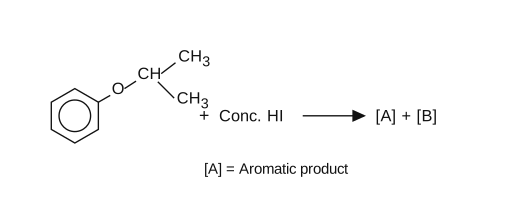
- COMEDK 2026Set 2026-A1 markMCQQ. A compound [X] undergoes reactions as given. Identify compounds [C] and [D] formed in these reactions. [A]Cr2O72−/H+[C] $$ [\text { B }] \xrightarrow[\text {(ii) } \mathrm{Na}2 \mathrm{CO}{3(\text { aq })}+\mathrm{I}_2]{\text { (i)aq. } \mathrm{KOH}} [\mathrm{D}] \quad+\mathrm{CH}_3 \mathrm{COONa}(A) \text { [C]: Benzoquinone [D]: lodoform } (B)[C]:Benzene[D]:2−iodo−propane(C)[C]:Benzoicacid[D]:lodoform(D) \text { [C]: 4-lodophenol [D]: 1-iodo-propane } $$
›Reveal solutionSolution
The compound [X] is isopropyl phenyl ether (C₆H₅–O–CH(CH₃)₂). Cleavage with concentrated HI gives phenol ([A]) and isopropyl iodide ([B]). Oxidation of phenol yields benzoquinone ([C]), and the iodoform reaction on isopropyl iodide gives iodoform ([D]) and sodium acetate. Thus the correct option is (A).
Concept & Intuition
This problem tests two classic organic reactions: ether cleavage by HI and the iodoform reaction. The key is to recognize that the ether [X] is an aryl alkyl ether (phenol derivative). When treated with concentrated HI, the C–O bond breaks selectively at the alkyl side (since the aryl–O bond is stronger due to resonance), producing phenol and an alkyl iodide. Then, phenol can be oxidized to benzoquinone, and the alkyl iodide (if it has a methyl group adjacent to the carbonyl or a secondary alcohol that can be oxidized to a methyl ketone) will undergo the iodoform test.
Let’s walk through each step.
Step-by-step reasoning
-
Identify [X] and its cleavage products
The figure shows a benzene ring with an –O–CH(CH₃)₂ group. That is isopropyl phenyl ether (C₆H₅–O–CH(CH₃)₂).
With concentrated HI, the ether bond breaks. The mechanism: HI protonates the oxygen, then iodide attacks the less hindered carbon (the isopropyl carbon, since it’s primary-like in the sense of being less sterically hindered than the aromatic ring). This gives phenol (C₆H₅OH) as the aromatic product [A] and isopropyl iodide (CH₃–CHI–CH₃) as [B].
Watch outA common mistake is to think the aromatic ring gets iodinated. But under these conditions, the C–O bond on the alkyl side breaks, not the aryl–O bond. The aromatic ring remains intact as phenol.
-
Reaction of [A] (phenol) with Cr₂O₇²⁻/H⁺ → [C]
Phenol is easily oxidized. Chromic acid (Cr₂O₇²⁻/H⁺) is a strong oxidizing agent. It oxidizes phenol to 1,4-benzoquinone (often just called benzoquinone). The reaction involves two-electron oxidation: the –OH group becomes a carbonyl, and the ring is rearranged to a quinoid structure.
So [C] = Benzoquinone.
-
Reaction of [B] (isopropyl iodide) with (i) aq. KOH, then (ii) Na₂CO₃(aq) + I₂ → [D] + CH₃COONa
- Step (i): Aqueous KOH will hydrolyze the alkyl iodide to an alcohol. Isopropyl iodide gives isopropyl alcohol (propan-2-ol, CH₃–CHOH–CH₃). …
-
- KCET 2025Set D-41 markMCQQ.Match the compounds given in List – I with the items given in List – II. List – I (I) Benzenesulphonyl Chloride (II) Sulphanilic acid (III) Alkyl Diazonium salts (IV) Aryl Diazonium salts List – II(a) Zwitterion(b) Hinsberg reagent(c) Dyes(d) Conversion to alcohols (A) 1 – c, II – b, III – a, IV – d (B) 1 – a, II – c, III – b, IV – d (C) 1 – c, II – a, III – d, IV – b (D) 1 – b, II – a, III – d, IV – c
›Reveal solutionSolution
Benzenesulphonyl chloride = Hinsberg reagent, sulphanilic acid = zwitterion, alkyl diazonium salts → alcohols, aryl diazonium salts → azo dyes.
Step 1 — (I) Benzenesulphonyl chloride → (b) Hinsberg reagent.
C6H5SO2Cl is known as Hinsberg's reagent. It reacts with 1° amines to give a sulphonamide with an acidic N–H (soluble in alkali), with 2° amines to give a sulphonamide with no N–H (insoluble in alkali), and does not react with 3° amines — the classical test for distinguishing the three classes.
Step 2 — (II) Sulphanilic acid → (a) Zwitterion.
The −SO3H group is strongly acidic and the −NH2 group is basic, so an internal proton transfer occurs:
H2N−C6H4−SO3H⇌+H3N−C6H4−SO3−
This dipolar internal salt is a zwitterion (which is why sulphanilic acid has a high melting point and low solubility in organic solvents).
Step 3 — (III) Alkyl diazonium salts → (d) Conversion to alcohols.
Alkyl diazonium ions (R−N2+) are extremely unstable because N2 is an excellent leaving group and there is no resonance stabilisation. They decompose at once, and water traps the resulting carbocation: …
- KCET 2024Set B-21 markMCQQ.In the reaction Aniline NaNO2/dil.HCl P Phenol/NaOH Q, ‘Q’ is: (A) C6H5N2Cl (B) ortho-hydroxyazobenzene (C) para-hydroxyazobenzene (D) meta-hydroxyazobenzene
›Reveal solutionSolution
Diazotisation of aniline gives the benzenediazonium salt (P); azo-coupling of that weak electrophile with phenoxide occurs at the para position, so Q is para-hydroxyazobenzene.
1. Step 1 — Diazotisation gives P
A primary aromatic amine treated with nitrous acid (generated in situ from NaNO2+dil. HCl) at 273–278 K gives an arenediazonium salt:
C6H5NH2NaNO2/dil. HCl273−278 KC6H5N+≡N Cl−
So P= benzenediazonium chloride. (The aryl diazonium ion is stabilised by delocalisation into the ring — this is why it survives, unlike an alkyl diazonium ion.) Note that option (A) is P, not Q — a classic distractor.
2. Step 2 — Azo coupling gives Q
The diazonium ion is only a weak electrophile, so it can attack a ring only if that ring is strongly activated. Phenol in NaOH is deprotonated to the phenoxide ion, C6H5O−, whose −O− is a very powerful electron-releasing group (strong +M), pumping electron density onto the ortho and para carbons.
Electrophilic substitution therefore occurs at those positions, but coupling takes place essentially exclusively at the para position because:
- the para carbon is sterically unhindered, whereas an ortho attack would place the bulky −N=N−C6H5 group right next to the −OH;
- the para-coupled azo product is the thermodynamically favoured, fully conjugated dye. …
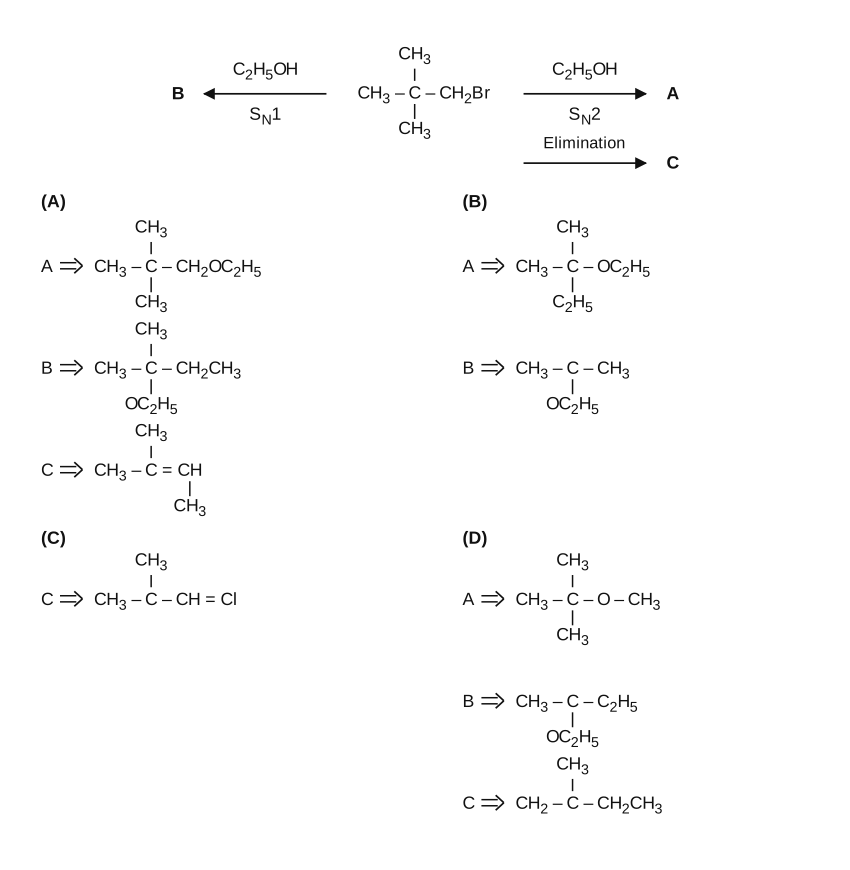
- COMEDK 2023Set 2023-M1 markMCQQ.Identify A, B and C. (A) (B) (C) (D)
›Reveal solutionSolution
The reaction scheme shows a neopentyl bromide undergoing SN1 (to B), SN2 (to A), and elimination (to C). The correct products are: A = neopentyl ethyl ether, B = 2-ethoxy-2-methylbutane (rearranged), C = 2-methyl-2-butene. Only option (A) matches all three.
Concept & Intuition
Neopentyl bromide (1-bromo-2,2-dimethylpropane) is a classic case where the substrate’s structure dictates reaction pathways. The carbon bearing bromine is primary, but it’s attached to a quaternary carbon (three methyl groups). For SN2, the backside attack is severely hindered by the bulky neopentyl group, making it very slow. For SN1, the primary carbocation would normally be unstable, but under solvolytic conditions (ethanol), the reaction proceeds via a rearranged tertiary carbocation (a methyl shift), giving a more stable intermediate. Elimination also favors the more substituted alkene (Zaitsev product). The question tests recognition of these rearrangements and the correct structures.
Step-by-step reasoning
- Identify the substrate The central structure is neopentyl bromide:
CH3–C(CH3)2–CH2Br
The bromine is on a primary carbon, but the carbon is neopentyl (tert-butylmethyl). This is crucial.
- SN2 pathway (→ A) SN2 requires a clean backside attack. The neopentyl group is extremely bulky, so SN2 is very slow. However, in ethanol (C₂H₅OH) as solvent, the ethoxide ion (from ethanol) can act as a nucleophile. The product is the unrearranged ethyl ether:
CH3–C(CH3)2–CH2–O–C2H5
This is neopentyl ethyl ether. No rearrangement occurs because SN2 is concerted.
Check options: Only option (A) shows this exact structure for A.
- SN1 pathway (→ B) SN1 proceeds via carbocation formation. The primary carbocation (CH₃–C(CH₃)₂–CH₂⁺) is very unstable. It immediately undergoes a 1,2-methyl shift to form the more stable tertiary carbocation:
CH3–C+(CH3)–CH2CH3
This tertiary carbocation is then trapped by ethanol (solvent) to give the ethyl ether:
CH3–C(OC2H5)(CH3)–CH2CH3
This is 2-ethoxy-2-methylbutane.
Check options: Only option (A) shows B as exactly this structure (with OC₂H₅ on the quaternary carbon and an ethyl group on the adjacent carbon).
- Elimination pathway (→ C) …
- KCET 2022Set B-31 markMCQQ.A secondary amine is (A) a compound with an NH2 group on the carbon atom in number 2 position (B) a compound in which 2 of the hydrogen of NH3 have been replaced by organic groups (C) an organic compound with two NH2 group (D) a compound with two carbon atom and an NH2 group
›Reveal solutionSolution
A secondary amine is defined by the number of alkyl/aryl groups attached to nitrogen — specifically, two organic groups replace two hydrogens of ammonia. The correct answer is (B).
The key to this question is understanding how amines are classified. Amines are derivatives of ammonia (NH3), and the classification — primary, secondary, or tertiary — depends entirely on how many of the three hydrogen atoms on nitrogen have been replaced by carbon-containing groups (alkyl or aryl). It has nothing to do with the position of a carbon atom, the number of carbon atoms in the molecule, or the count of NH2 groups.
Let’s examine each option carefully.
-
Option (A) says “a compound with an NH2 group on the carbon atom in number 2 position.” This describes a structural detail about where an amino group is attached on a carbon chain (like on C-2 of propane). That is a matter of positional isomerism, not amine classification. A primary amine can have its NH2 on carbon-2, and so can a secondary or tertiary amine if they also have other groups. This definition misses the point entirely.
-
Option (B) says “a compound in which 2 of the hydrogen of NH3 have been replaced by organic groups.” This is exactly the definition. Ammonia has three hydrogens. Replace one → primary amine (RNH2). Replace two → secondary amine (R2NH). Replace three → tertiary amine (R3N). So a secondary amine has two alkyl/aryl groups attached to nitrogen, with one hydrogen remaining. …
-
- COMEDK 2021Set 2021-B1 markMCQQ.What are the products formed when Anisole is reacted with Hydroiodic acid and heated? (A) Iodobenzene + Methane (B) Phenol + Methanol (C) Phenol + Iodomethane (D) Iodobenzene + Methanol
›Reveal solutionSolution
Anisole C6H5−O−CH3+HI→C6H5OH (phenol) +CH3I (iodomethane).
In cleavage of aryl alkyl ethers by HI, the bond broken is the O−alkyl bond, not the O−aryl bond, because forming an aryl cation/attack at the aromatic carbon is very unfavourable. I− attacks the methyl carbon (SN2), …
- KCET 2019Set A-11 markMCQQ.The metal nitrate that liberates NO2 on heating (A) NaNO3 (B) KNO3 (C) LiNO3 (D) RbNO3
›Reveal solutionSolution
Li+ is tiny and highly polarising, so it distorts the nitrate ion enough to break it right down to the oxide + NO2; the bigger alkali cations only take it as far as the nitrite.
Step 1 — The two possible decomposition routes
Alkali-metal nitrates decompose on heating by one of two paths:
Path 1 — to the nitrite (Na, K, Rb, Cs):
2MNO3Δ2MNO2+O2↑
Only oxygen is evolved — no brown fumes.
Path 2 — to the oxide (Li):
4LiNO3Δ2Li2O+4NO2↑+O2↑
Here the nitrate ion is destroyed completely, giving the characteristic brown NO2 gas.
Step 2 — Why lithium is the odd one out (Fajans' rules)
The polarising power of a cation scales as (radius)2charge. Among the alkali metals:
Li+(76 pm)<Na+(102)<K+(138)<Rb+(152 pm)
So Li+ is by far the smallest and therefore the most polarising. It pulls electron density out of the large, soft NO3− anion, weakening the N–O bonds so much that the anion breaks apart entirely into O2− (which stays with Li as Li2O) and NO2.
The larger cations Na+,K+,Rb+ cannot distort the nitrate that strongly. Their nitrates only shed one oxygen atom, stopping at the stable nitrite.
Step 3 — The wider pattern (worth remembering) …
🎓Unlock everything free for 14 days
- ✓Full step-by-step solutions
- ✓Concept-first explanations
- ✓Methods, shortcuts & mistakes
- ✓PYQ mapping + timed mock tests
Full access for 14 days. No credit card required.