Q.tert-Butylbromide reacts with aq. NaOH by SN1 mechanism while n-butylbromide reacts by SN2 mechanism. Why?
🔒You're viewing a preview — the full solution, concept, methods & PYQ mapping are locked.
🔒 Start your 14-day free trial to unlock the full solution →Concept understanding — SN1 Reactivity Order
The Core Intuition: Who Wants to Leave, and Who Can Wait?
Imagine you're at a party where the host (the leaving group) is about to leave. The party (the reaction) happens in two stages. First, the host walks out the door — that's the slow, painful step. Then, a new guest (the nucleophile) rushes in to take the empty spot.
The SN1 reaction works exactly like this: the leaving group leaves first, forming a carbocation intermediate. The nucleophile attacks after the leaving group is gone. This means the rate of the reaction depends only on how easily the leaving group can leave — it does not depend on the nucleophile at all.
So the question becomes: What makes a carbocation form easily? The answer is stability. A carbocation that is more stable will form faster and last longer, making the SN1 reaction faster.
The Precise Statement of SN1 Reactivity Order
SN1 Reactivity Order (for alkyl halides):
Allylic≈Benzyl>3∘>2∘≫1∘≈Methyl
This is the order of how fast the SN1 reaction proceeds. Let's unpack why.
Why This Order? The Stability Ladder
A carbocation is a carbon with only six electrons in its valence shell — it's electron-deficient and positively charged. The more you can spread out (delocalize) that positive charge, the more stable the carbocation becomes.
1. Methyl and 1° Carbocations: The Unstable Ones
A methyl carbocation (CHX3X+) has no alkyl groups attached to the positive carbon. There is zero electron-donating effect to stabilize the charge. It is so unstable that it practically never forms in an SN1 reaction — the reaction simply doesn't happen.
A primary (1°) carbocation has one alkyl group attached. Alkyl groups are weakly electron-donating (through hyperconjugation and inductive effect), so it's slightly more stable than methyl — but still far too unstable to form under normal SN1 conditions.
Never say "SN1 happens on a primary carbon" in an exam. It is essentially impossible under standard conditions because the carbocation is too unstable.
2. Secondary (2°) Carbocations: The Borderline Case
A secondary carbocation has two alkyl groups donating electron density. It is moderately stable — stable enough to form, but only under certain conditions (like a good leaving group and a polar protic solvent). SN1 reactions on secondary carbons are possible, but they are slower than on tertiary carbons.
3. Tertiary (3°) Carbocations: The Sweet Spot
Three alkyl groups donate electron density to the positive carbon. This makes the carbocation very stable. Tertiary alkyl halides undergo SN1 reactions readily — they are the classic example.
4. Allylic and Benzylic: The Champions
These are special cases. In an allylic carbocation, the positive charge is adjacent to a carbon-carbon double bond. The π electrons of the double bond can delocalize the positive charge onto the second carbon:
CHX2=CH−CHX2X+↔+CHX2−CH=CHX2
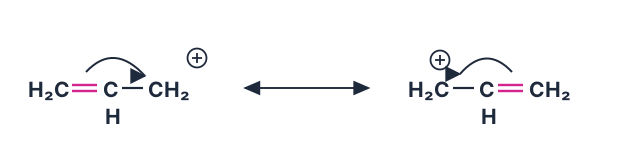
In a benzylic carbocation, the positive charge is adjacent to a benzene ring. The π system of the ring delocalizes the charge across multiple carbons:
CX6HX5−CHX2X+↔(several resonance structures)
Here are those resonance structures — the positive charge cycles from the CH₂ carbon onto the ortho and para positions of the ring:
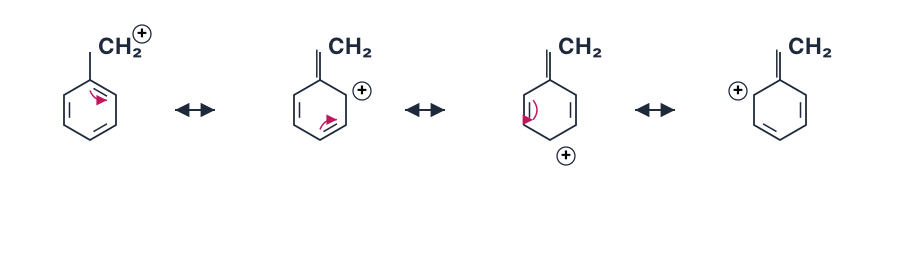
This resonance stabilization makes allylic and benzylic carbocations even more stable than tertiary ones. They form the fastest in SN1 reactions.
The Complete Picture in a Table
| Carbocation Type | Stability | SN1 Reactivity | Example |
|------------------|-----------|----------------|---------| …
Why this formula?
SN1 Reactivity Order: Why It Holds
The SN1 reaction (Substitution Nucleophilic Unimolecular) proceeds via a carbocation intermediate. The reactivity order is determined entirely by the stability of this carbocation — because the rate-determining step is its formation.
The Core Principle
The rate law for SN1 is:
Rate=k[RX]
Only the substrate appears in the rate law — the nucleophile does not participate in the slow step. The slow step is:
RXslowR++X−
Thus, anything that stabilizes the carbocation (R⁺) lowers the activation energy and increases the reaction rate.
The Reactivity Order
For alkyl halides (RX), the SN1 reactivity order is:
Allylic>Benzyllic>Tertiary>Secondary>Primary>Methyl
Let's break down why each step holds.
1. Why Tertiary > Secondary > Primary > Methyl?
This is purely about hyperconjugation and inductive effect.
- Tertiary carbocation: Three alkyl groups donate electron density via hyperconjugation (C–H σ bonds overlap with empty p orbital) and +I effect. This spreads the positive charge over more atoms → most stable.
- Secondary: Two alkyl groups → less stabilization.
- Primary: Only one alkyl group → very little stabilization.
- Methyl: No alkyl groups → least stable (only inductive effect from H atoms, which is negligible).
Key formula: The number of α-hydrogens (H on carbons adjacent to the positive carbon) determines hyperconjugation. More α-H → more resonance structures → more stable.
2. Why Allylic and Benzylic Are Even Faster
These carbocations are resonance-stabilized.
- Allylic carbocation: The positive charge is delocalized over two carbon atoms via π-bond conjugation:
CH2=CH−CH2+⟷CH2+−CH=CH2
- Benzylic carbocation: The positive charge is delocalized into the aromatic ring:
C6H5−CH2+⟷several resonance forms involving the ring
Those resonance forms look like this:
This resonance stabilization is so powerful that even a primary allylic or benzylic carbocation is more stable than a tertiary alkyl carbocation.
3. The Complete Order (with reasoning) …
The key idea is that SN1 reactivity depends on carbocation stability, while SN2 reactivity depends on steric hindrance.
- For tert-butyl bromide: The carbon attached to bromine is tertiary. In an SN1 mechanism, the C–Br bond breaks first to form a carbocation. A tertiary carbocation is highly stabilised by three alkyl groups (hyperconjugation and inductive effect), so this step is fast and favourable. The bulky tert-butyl group also strongly hinders any backside attack, making SN2 impossible. …
The reactivity difference arises from carbocation stability: tert-butyl bromide forms a stable tertiary carbocation (favouring SN1), while n-butyl bromide cannot form a stable carbocation and instead undergoes SN2 via a back-side attack.
The key to understanding this lies in the stability of the intermediate formed in each mechanism. SN1 reactions proceed through a carbocation intermediate, while SN2 reactions occur in a single step with no intermediate. The structure of the alkyl halide dictates which pathway is feasible.
- Carbocation stability determines SN1 feasibility. For an SN1 reaction, the rate-determining step is the departure of the leaving group (bromide) to form a carbocation. The more stable the carbocation, the faster this step occurs. Carbocation stability follows the order:
tertiary>secondary>primary>methyl
This is because alkyl groups are electron-donating via hyperconjugation and inductive effects, which delocalize the positive charge.
-
tert-Butyl bromide forms a stable tertiary carbocation.
The structure is (CH3)3C-Br. When bromide leaves, the resulting carbocation is (CH3)3C+, a tertiary carbocation. This is highly stabilized by three methyl groups donating electron density. The activation energy for this step is low, making SN1 the dominant mechanism.
TipA quick way to remember: tertiary alkyl halides almost always react via SN1 (or E1) because the carbocation is stable enough to form.
-
n-Butyl bromide cannot form a stable carbocation.
The structure is CH3CH2CH2CH2-Br. If bromide were to leave, the resulting carbocation would be primary (CH3CH2CH2CH2+). Primary carbocations are extremely unstable — they lack sufficient alkyl groups to stabilize the positive charge. The energy required to form such a high-energy intermediate is prohibitively high.
Watch outA common mistake is to think that n-butyl bromide could undergo a hydride shift to form a more stable carbocation. In practice, such rearrangements are possible only under strongly acidic conditions or with good leaving groups in polar solvents — but here, with aqueous NaOH, the SN2 pathway is much faster than any rearrangement.
-
Steric hindrance also plays a role. …
Concept: Steric Hindrance in Nucleophilic Substitution
The key concept is steric hindrance — the physical bulk of alkyl groups around the reaction centre (the carbon attached to the leaving group). This determines whether the mechanism is SN1 or SN2.
Method: Steric Accessibility Analysis
Steps
- Identify the substrate structure
- tert-Butylbromide: The central carbon is bonded to three methyl groups and one bromine. Structure:
(CH3)3C—Br
- n-Butylbromide: The bromine is on a primary carbon (end of a straight chain). Structure:
CH3CH2CH2CH2—Br
-
Analyse steric hindrance at the reaction centre
- In SN2, the nucleophile must attack from the backside of the carbon–bromine bond.
- tert-Butylbromide: Three bulky methyl groups block the backside approach — high steric hindrance.
- n-Butylbromide: Only a hydrogen and an alkyl chain are behind the carbon — low steric hindrance.
-
Match mechanism to steric environment
- High steric hindrance → SN2 is impossible → reaction proceeds via SN1 (two-step, carbocation intermediate).
- Low steric hindrance → SN2 is favoured (one-step, direct attack).
-
Check carbocation stability (for SN1)
- tert-Butyl carbocation (CH3)3C+ is tertiary — highly stabilised by hyperconjugation and inductive effects. …
This is a classic comparison that tests your understanding of substrate structure and reaction mechanisms in organic chemistry. Let's break down the common mistakes and how to avoid them.
🧠 The Core Concept
The key difference lies in the stability of the carbocation intermediate and steric hindrance around the carbon bearing the leaving group.
- tert-Butyl bromide (3° alkyl halide) → forms a stable tertiary carbocation → favours SN1 (two-step, via carbocation).
- n-Butyl bromide (1° alkyl halide) → forms a highly unstable primary carbocation → cannot use SN1 → instead uses SN2 (one-step, backside attack).
✗ Common Mistake #1: "Both react by the same mechanism because both have Br as leaving group"
Why it's wrong:
The leaving group is the same, but the alkyl group structure determines the mechanism. The rate-determining step in SN1 depends only on the substrate (not the nucleophile), while SN2 depends on both.
How to avoid:
Always check the degree of substitution at the carbon attached to the leaving group:
- 1° → almost always SN2 (unless special conditions)
- 3° → almost always SN1 (or E1)
- 2° → can go either way depending on solvent, nucleophile, etc.
✗ Common Mistake #2: "tert-Butyl bromide reacts faster because it has more alkyl groups"
Why it's wrong:
More alkyl groups slow down SN2 (steric hindrance) but speed up SN1 (carbocation stabilisation). So the reason for the mechanism choice is opposite for the two pathways.
How to avoid:
Remember the steric vs electronic trade-off:
- SN2: more alkyl groups = more steric hindrance → slower
- SN1: more alkyl groups = more hyperconjugation/inductive effect → more stable carbocation → faster
✗ Common Mistake #3: "n-Butyl bromide reacts by SN1 because it has a primary carbocation"
Why it's wrong:
A primary carbocation is extremely unstable — it does not form under normal conditions. The activation energy is too high.
How to avoid:
Memorise the carbocation stability order:
3∘>2∘>1∘>methyl
Only 3° and some 2° carbocations are stable enough for SN1 in typical exam problems. If you see a primary alkyl halide, assume SN2 unless told otherwise (e.g., with a very good leaving group and weak nucleophile in a polar protic solvent — but that's rare).
✗ Common Mistake #4: "The nucleophile (OH⁻) is strong, so both should be SN2"
Why it's wrong: …
Showing the 12 most recent of 17 on this concept.
- AP EAPCET 2026Set eng-2026-05-13-FN1 markMCQQ.Consider the following reactions I, II and III. The correct order of reactivity of X, Y and Z towards SN1 reaction is (I) CH2=CH−CH2−CH3HBr(X) (major) (II) CH2=CH−CH2−CH3HBr(C6H5CO)2O2(Y) (major) (III) (CH3)2C=CH2HBr(Z) (A) Z>Y>X (B) X>Z>Y (C) X>Y>Z (D) Z>X>Y
›Reveal solutionSolution
X is a secondary bromide, Y a primary bromide, Z a tertiary bromide. Since SN1 rate depends on carbocation stability (3° > 2° > 1°), the reactivity order is Z > X > Y.
Concept and Intuition
SN1 reactions proceed through a carbocation intermediate in the rate-determining step, so their rate is governed almost entirely by how stable that carbocation is — the more stable the carbocation (more substituted = more hyperconjugation/inductive electron donation from alkyl groups), the faster the SN1 reaction. This immediately tells us the reactivity order among alkyl halides towards SN1: tertiary > secondary >> primary (primary halides essentially never go by SN1 due to the very unstable primary carbocation).
So the actual chemistry of the reactions is secondary to correctly identifying the type of halide (1°, 2°, or 3°) each one produces.
Step-by-Step Solution
- Reaction I: CH2=CH−CH2−CH3 (but-1-ene) + HBr, normal ionic addition → Markovnikov's rule: H+ adds to the terminal (less substituted) carbon, generating the more stable secondary carbocation at C2, which Br− then attacks → X = 2-bromobutane (a secondary halide).
- Reaction II: same alkene + HBr in presence of a peroxide, (C6H5CO)2O2 → this triggers the free-radical (peroxide/Kharasch) mechanism, which is anti-Markovnikov: Br∙ adds first to the terminal carbon (forming the more stable secondary radical at C2), and the halogen ends up on the terminal carbon → Y = 1-bromobutane (a primary halide). …
- AP EAPCET 2026Set eng-2026-05-14-FN1 markMCQQ.An organic compound C4H9Br (A) on reaction with Na/dry ether gave B. Photochemical chlorination of B gave two monochlorides. Correct statement regarding A is (A) It is a chiral molecule (B) It undergoes nucleophilic substitution by SN1 mechanism (C) Reaction of it with NaOC2H5 gave predominantly substitution product (D) It can be obtained by the addition of HBr to but-2-ene
›Reveal solutionSolution
A is a primary C4H9Br isomer: its Wurtz coupling product B is a fairly symmetric C8H18 alkane giving only a small number of monochloro products on photochlorination. Being primary, A reacts with a moderate base/nucleophile like sodium ethoxide mainly by substitution (SN2), not elimination. Answer: predominantly substitution with NaOC2H5.
Concept and Intuition
The Wurtz reaction couples two identical alkyl halide molecules via loss of the halide (as NaX) to form a symmetric alkane with double the carbon count. Photochemical (free-radical) chlorination of that alkane substitutes different types of hydrogen atoms, and the number of distinct monochloro products equals the number of chemically distinct hydrogen environments in the molecule — fewer distinct environments (a more symmetric alkane) means fewer monochloro products, which is a strong clue about how symmetric/branched the parent alkyl halide is.
Among the four possible reactivity/behaviour statements, we test each candidate structure:
- A primary halide (e.g. n-butyl or isobutyl bromide) is not chiral, does not favor SN1 (primary carbocations are unstable), reacts with alkoxide bases predominantly via SN2 substitution (only mildly reduced by beta-branching), and cannot be made by HX addition to an internal alkene like but-2-ene (which would give a secondary halide instead).
- A secondary halide (2-bromobutane) is chiral AND can be made from but-2-ene + HBr — giving two simultaneously 'true' statements, which is inconsistent with a single-answer MCQ, ruling it out as the intended compound.
- A tertiary halide (tert-butyl bromide) reacts via SN1/E1 and gives (famously) only ONE type of monochloro product from its highly symmetric coupling product — its defining reactivity statement is the SN1 one, not the substitution-with-ethoxide one.
Given the clue of only a small number (few, not many) monochloro products from B, and that this must correspond to an answer choice that is uniquely and unambiguously true, the primary-halide identification (giving statement C) is the internally consistent choice.
Step-by-Step Solution
- A is a primary C4H9Br isomer; Wurtz coupling with Na/dry ether gives a symmetric C8H18 alkane B.
- Photochemical chlorination of B substitutes at its few distinct types of C–H bonds, consistent with a small number of monochloro products for a primary-derived, moderately symmetric alkane. …
- AP EAPCET 2026Set eng-2026-05-18-FN1 markMCQQ.Among the following, identify the compound which will form resonance stabilized carbocations after the leaving group is lost I. Chlorobenzene (C6H5Cl, Cl attached directly to the benzene ring) II. Benzyl chloride (C6H5CH2Cl) III. (E)-1-bromoprop-1-ene (CH3−CH=CH−Br, Br attached directly to the alkene carbon) IV. Allyl bromide (CH2=CH−CH2Br) The correct answer is (A) I & II only (B) I & III only (C) II & III only (D) II & IV only
›Reveal solutionSolution
This tests the distinction between allylic/benzylic positions (which give resonance-stabilized carbocations on ionization) and aryl/vinylic positions (which do not); only benzyl chloride (II) and allyl bromide (IV) qualify.
Concept and Intuition
Resonance stabilization of a carbocation requires the empty p-orbital on the cationic carbon to overlap with an adjacent π system so the positive charge can delocalize. This overlap geometry exists when the leaving group sits on an sp3 carbon next to a ring (benzylic) or a double bond (allylic) — ionization then produces a cation whose empty orbital is perfectly aligned with the neighbouring π system. It does not exist when the leaving group is itself on the sp2 carbon of the ring or double bond (aryl/vinylic), because then there is no adjacent, differently-oriented π system to delocalize into — the resulting aryl or vinyl cation is high-energy and not resonance stabilized, and in practice these C–X bonds barely ionize at all due to partial double-bond character.
Step-by-Step Solution
- I. Chlorobenzene: Cl is bonded directly to an aromatic ring carbon. Loss of Cl− would need to break a bond with significant double-bond character (from ring conjugation with the halogen lone pair) and would give a phenyl cation in the plane of the ring, orthogonal to the ring's π system — not stabilized by resonance. Excluded.
- II. Benzyl chloride: Cl is on the sp3 CH2 carbon attached to the ring. Loss of Cl− gives the benzylic cation C6H5−C+H2, whose empty p-orbital aligns with the ring π system, delocalizing the charge onto the ortho/para ring carbons. Resonance stabilized. Included. …
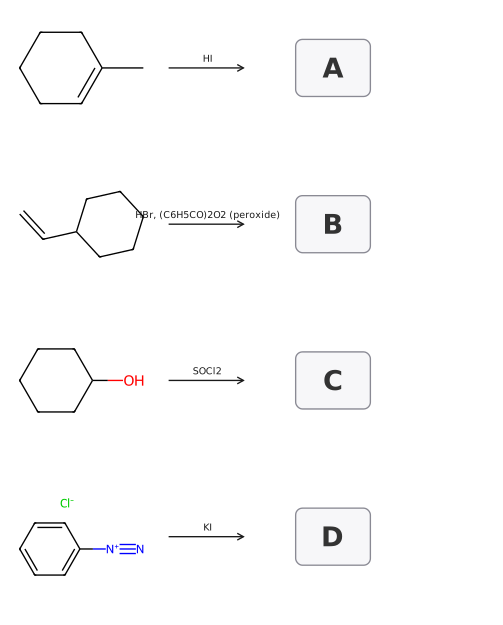
- AP EAPCET 2026Set ap-2026-05-20-FN1 markMCQQ.The correct order of reactivity of the products A, B, C and D towards SN1 reaction is [FIGURE] (four reaction schemes:(1) a methylcyclohexene ring reacting with HI to give A;(2) a vinylcyclohexane ring reacting with HBr in the presence of (C6H5CO)2O2 peroxide to give B;(3) cyclohexanol, a cyclohexane ring bearing an OH group, reacting with SOCl2 to give C;(4) a benzene ring bearing a diazonium salt group N2+Cl− reacting with KI to give D) (A) A>B>C>D (B) D>B>A>C (C) A>C>B>D (D) B>C>A>D
›Reveal solutionSolution
Identify each product's carbon type (3°, 1°, 2°, aryl respectively for A, B, C, D) and rank by SN1-relevant carbocation stability: A (3°) > C (2°) > B (1°) > D (aryl, unreactive).
Concept and Intuition
SN1 reaction rate depends entirely on how stable the intermediate carbocation is, since the rate-determining step is ionisation of the C–X bond. More substituted carbons (tertiary > secondary > primary) stabilise positive charge better through hyperconjugation and inductive electron donation. Aryl (and vinyl) halides essentially never undergo SN1: the C–X bond has partial double-bond character from ring conjugation, and the resulting aryl cation would be extremely unstable (sp² cation in the ring plane, no adjacent orbital stabilisation) — so aryl halides always rank last regardless of the specific reagent used to make them.
Step-by-Step Solution
- Product A: A methyl-substituted cyclohexene reacts with HI. Markovnikov addition places I− on the more substituted (tertiary) carbon of the double bond (bearing the methyl group) — giving a tertiary alkyl iodide.
- Product B: Vinylcyclohexane + HBr in the presence of a peroxide follows the peroxide (anti-Markovnikov) effect — a free-radical mechanism places Br on the terminal, less-hindered carbon of the vinyl group — giving a primary alkyl bromide.
- Product C: Cyclohexanol (a secondary alcohol) + SOCl2 replaces –OH with –Cl (via the chlorosulfite intermediate, SNi-type) — giving a secondary alkyl chloride, with no change in the carbon's substitution level. …
- AP EAPCET 2026Set ap-2026-05-20-AN1 markMCQQ.Observe the following sets of orders with respect to reactivity of halides against the reactions mentioned as in I and II given below I. SN1: Isobutyl iodide < sec. butyl iodide < t-butyl bromide II. SN2: n-Butylbromide > Isobutylbromide > Sec. butyl bromide correct answer is (A) Both I, II are correct (B) Both I, II are NOT correct (C) I is correct but II is NOT correct (D) I is NOT correct but II is correct
›Reveal solutionSolution
Order I correctly reflects the SN1 trend driven by carbocation stability (1° < 2° < 3°), and order II correctly reflects the SN2 trend driven by steric hindrance (unbranched primary > branched primary > secondary) — both are correct.
Concept and Intuition
SN1 reactions proceed through a carbocation intermediate, so their rate is controlled almost entirely by how stable that carbocation is: tertiary carbocations (stabilised by three alkyl/hyperconjugative groups) form fastest, followed by secondary, then primary (least stable, slowest SN1).
SN2 reactions, in contrast, proceed through a single concerted backside-attack transition state, so their rate is controlled by steric accessibility of the carbon under attack: more crowding around (or even beta to) the reacting carbon slows the reaction down. Straight-chain primary halides react fastest, branched (e.g., isobutyl, which has a beta-branch) primary halides are slower due to steric hindrance from the nearby branch, and secondary halides are slower still because the reacting carbon itself bears an extra alkyl group.
Step-by-Step Solution
- Order I (SN1): Isobutyl iodide is a primary halide (least stable carbocation if it ionised, so slowest SN1); sec-butyl iodide is secondary (more stable carbocation, faster than primary); t-butyl bromide is tertiary (most stable carbocation, fastest SN1 despite bromide being a slightly poorer leaving group than iodide, the carbocation-stability effect dominates). This gives isobutyl iodide < sec-butyl iodide < t-butyl bromide — matches the trend 1° < 2° < 3°, so order I is correct. …
- AP EAPCET 2025Set eng-2025-05-21-FN1 markMCQQ.Observe the following reactions Cyclohexene HBrX 1-Methylcyclohex-1-ene HClY (a cyclohexene ring bearing a methyl substituent on the double-bond carbon) Cyclohexanol SOCl2Z The order of reactivity of x, y, z towards SN1 reaction is (A) x > z > y (B) x > y > z (C) y > x > z (D) y > z > x
›Reveal solutionSolution
Tests SN1 reactivity trends based on carbocation stability and leaving-group ability; the answer is (C) y > x > z.
Concept and Intuition
SN1 reactions proceed through a carbocation intermediate, so their rate is governed chiefly by:
- Carbocation stability: 3° > 2° > 1° (more alkyl substitution stabilises the positive charge by hyperconjugation/induction).
- Leaving group ability: for the same carbon skeleton type, a better leaving group (weaker conjugate base) ionises faster — Br⁻ is a better leaving group than Cl⁻ since HBr is a stronger acid than HCl, so C-Br bonds ionise more readily than C-Cl bonds.
First identify what X, Y, Z actually are:
- X: Cyclohexene + HBr → Markovnikov addition of HBr across the double bond gives bromocyclohexane, a secondary alkyl bromide.
- Y: 1-Methylcyclohex-1-ene + HCl → Markovnikov addition puts H on the carbon that gives the more stable (more substituted) carbocation; since C1 already bears the methyl group and two ring bonds, the carbocation forms at C1 (tertiary), so Cl ends up at C1 too, giving 1-chloro-1-methylcyclohexane, a tertiary alkyl chloride.
- Z: Cyclohexanol + SOCl2 → gives chlorocyclohexane, a secondary alkyl chloride (via the chlorosulfite intermediate).
Step-by-Step Solution
- Y is a tertiary halide → forms the most stable (tertiary) carbocation → fastest SN1 reactant of the three. Y is highest. …
- AP EAPCET 2025Set ap-2025-05-20-FN1 markMCQQ.The SN1 reactivity of the following halides will be in the order I) C6H5CH2Br II) (C6H5)2CHBr III) (C6H5)2C(CH3)Br IV) (CH3)2CHBr (A) III > II > I > IV (B) III > I > II > IV (C) IV > I > II > III (D) II > III > I > IV
›Reveal solutionSolution
SN1 reactivity tracks carbocation stability, and stability here is set by how many phenyl rings (resonance donors) and alkyl groups (hyperconjugation) stabilize the intermediate cation: III > II > I > IV.
Concept and Intuition
The rate-determining step of an SN1 reaction is the formation of a carbocation. Any substituent that stabilizes this positive charge — through resonance (as aryl/phenyl groups do, by delocalizing the charge into the ring) or through hyperconjugation/induction (as alkyl groups do) — speeds up SN1. Aromatic (benzylic-type) stabilization is generally very strong, and stacking multiple phenyl groups (plus alkyl groups) compounds this stabilization.
Step-by-Step Solution
- I, C6H5CH2Br: ionisation gives the benzyl cation, stabilized by resonance delocalization into one phenyl ring — moderately stabilized (primary carbon, one aryl group).
- II, (C6H5)2CHBr: ionisation gives the benzhydryl (diphenylmethyl) cation, stabilized by resonance into two phenyl rings simultaneously — more stable than I.
- III, (C6H5)2C(CH3)Br: ionisation gives a tertiary cation stabilized by two phenyl rings and hyperconjugation/induction from the extra methyl group — the most stabilized cation of the four. …
- AP EAPCET 2025Set eng-2025-05-24-FN1 markMCQQ.Which one of the following halogen compounds is least reactive towards hydrolysis by SN1 mechanism? (A) Tertiary butylchloride (B) Isopropyl chloride (C) Allyl chloride (D) Ethyl chloride
›Reveal solutionSolution
This tests the link between carbocation stability and SN1 reactivity; ethyl chloride, forming an unstabilised primary carbocation, is the least reactive.
Concept and Intuition
SN1 reactions proceed through a carbocation intermediate in the rate-determining step, so anything that stabilises that carbocation speeds up the reaction. Stability order: tertiary > allylic/benzylic (resonance-stabilised) > secondary > primary. A primary carbocation, lacking both hyperconjugative stabilisation (from multiple alkyl groups) and resonance stabilisation, is highly unstable, so alkyl halides that would have to form one (like ethyl chloride) react very slowly, if at all, by the SN1 pathway — they instead prefer SN2.
Step-by-Step Solution
- Tertiary butyl chloride → forms a tertiary carbocation (most stable due to hyperconjugation/inductive donation from three methyl groups) → highly reactive in SN1.
- Allyl chloride → forms an allylic carbocation stabilised by resonance with the adjacent double bond → quite reactive in SN1, despite being formally 'primary' in connectivity.
- Isopropyl chloride → forms a secondary carbocation → moderately reactive in SN1. …
- AP EAPCET 2025Set eng-2025-05-26-FN1 markMCQQ.Identify the compound which is least reactive towards nucleophilic substitution reactions. (A) C6H5CH2Cl (benzyl chloride: benzene ring with a −CH2Cl substituent) (B) 1-chloro-4-nitrobenzene: benzene ring with Cl and NO2 (drawn as O2N−) in the para positions (C) 1-chloro-3-nitrobenzene: benzene ring with Cl and NO2 in the meta positions (D) CH2=CH−CH2Cl (allyl chloride)
›Reveal solutionSolution
This tests reactivity toward nucleophilic substitution for benzylic, allylic, and differently-substituted aryl halides. The answer is (C): the meta-nitro chlorobenzene, since -NO2 only activates the ring toward nucleophilic aromatic substitution from the ortho/para position, not meta.
Concept and Intuition
Nucleophilic substitution reactivity depends heavily on the type of C–X bond and any stabilizing/activating groups. Benzylic and allylic halides are highly reactive because the developing positive charge (SN1) or the transition state (SN2) is stabilized by resonance with the adjacent pi system. Plain aryl (vinyl-type) C–Cl bonds are normally very unreactive to nucleophilic substitution because of partial double-bond character (resonance donation from the ring) and the difficulty of forming an aryl cation or backside attack on an sp2 carbon. This inertness can be overcome ('activated') by a strong electron-withdrawing group like −NO2, but ONLY if it is positioned ortho or para to the leaving group, because only then can its resonance structures place negative charge directly on the carbon bearing the leaving group (stabilizing the Meisenheimer-type intermediate in nucleophilic aromatic substitution).
Step-by-Step Solution
- (A) Benzyl chloride: the −CH2Cl carbon is benzylic; ionization gives a resonance-stabilized benzylic carbocation, making it highly reactive via SN1 (also reactive via SN2). Reactive.
- (D) Allyl chloride: the C–Cl carbon is allylic; ionization gives a resonance-stabilized allylic carbocation, also highly reactive. Reactive.
- (B) 1-chloro-4-nitrobenzene: the −NO2 is para to Cl, so its resonance structures put negative charge directly on the ring carbon bearing Cl, strongly stabilizing the anionic intermediate of nucleophilic aromatic substitution — this activates the ring, making it noticeably more reactive than plain chlorobenzene. …
- AP EAPCET 2024Set eng-2024-05-20-AN1 markMCQQ.The product of which of the following reactions undergo hydrolysis by SN1 mechanism? A. C6H5−CH2−CH=CH2 $\xrightarrow[\text{(iii) }SOCl_2]{\text{(i) }(BH_3)_2\text{(ii) }H_2O_2/OH^-}B.\mathrm{C_6H_5-CH_2-CH=CH-CH_3}\xrightarrow{HBr}C.\mathrm{C_6H_5-C(CH_3)=CH_2}\xrightarrow{HBr}D.\mathrm{C_6H_5-CH_2-CH=CH-CH_3}\xrightarrow{HBr,\ (C_6H_5CO)_2O_2}$ (A) C, D only (B) A, B, C only (C) B, C only (D) A, D only
›Reveal solutionSolution
This tests whether you can connect how an alkyl halide is made (Markovnikov vs anti-Markovnikov addition) to how it will react later (SN1 needs a stabilised — tertiary/benzylic/allylic — carbocation).
Concept and Intuition
SN1 hydrolysis proceeds through a carbocation intermediate, so it is fast only when that carbocation is well stabilised — tertiary, allylic, or benzylic (ring directly or nearly conjugated). A plain primary or an "ordinary" secondary halide, with no such stabilisation, hydrolyses by SN2 instead. So for each reaction you must first work out the structure of the halide product, then judge how stable its carbocation would be.
Step-by-Step Solution
- A: (BH3)2 then H2O2/OH− is hydroboration–oxidation — strictly anti-Markovnikov, so the terminal alkene C6H5−CH2−CH=CH2 gives the terminal (primary) alcohol C6H5CH2CH2CH2OH. SOCl2 then swaps −OH for −Cl with retention of position, giving a primary chloride three carbons from the ring. A primary carbocation is never favourable, so this halide hydrolyses by SN2 only — excluded.
- C: plain HBr (no peroxide) on C6H5−C(CH3)=CH2 (α-methylstyrene) adds Markovnikov-wise: H+ goes to the terminal =CH2, generating the carbocation on the ring-attached carbon, C6H5−C+(CH3)−CH3. This cation is tertiary and directly benzylic (fully conjugated with the ring) — about as stable as a carbocation gets. Br− then attacks there, giving C6H5C(CH3)(Br)CH3, a textbook SN1 substrate (structurally like tert-cumyl bromide). Definitely included.
- B vs D both start from the same internal alkene C6H5−CH2−CH=CH−CH3, so the only difference is where Br ends up.
- B (plain HBr, ionic/Markovnikov): the reaction proceeds through whichever secondary carbocation is more stable, which is the one adjacent to the benzylic CH2 (weak but real stabilisation from the nearby aromatic ring through hyperconjugation). Br− ends up there, giving a secondary bromide close to the ring — favourable enough for SN1 in this comparison. …
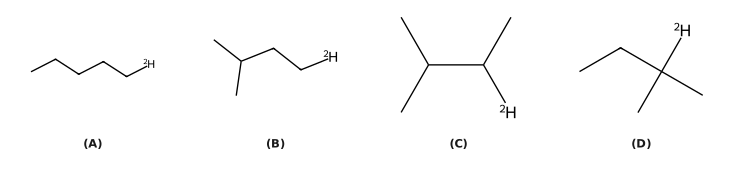
- AP EAPCET 2024Set eng-2024-05-21-AN1 markMCQQ.Hydrolysis of an alkyl bromide X (C5H11Br) follows first order kinetics. Reaction of X with Mg in dry ether followed by treatment of D2O gave Y. What is Y? (A) [FIGURE] (a straight-chain, 5-carbon skeletal structure with D at the terminal carbon, i.e. CH3CH2CH2CH2CH2D) (B) [FIGURE] (a branched 5-carbon skeletal structure: an isopropyl group (two methyls on one carbon) connected via a 2-carbon chain to a terminal D, i.e. (CH3)2CHCH2CH2D) (C) [FIGURE] (a branched 5-carbon skeletal structure with a methyl branch near one end and D attached to an internal carbon rather than at the chain terminus) (D) [FIGURE] (a branched 5-carbon skeletal structure: an ethyl group connected to a carbon bearing two methyl branches and D, i.e. a tertiary carbon, CH3CH2C(CH3)2D)
›Reveal solutionSolution
First-order kinetics for the hydrolysis of a C5H11Br isomer identifies it as the tertiary bromide (S_N1 requires the more stable, more substituted carbocation). Grignard formation + D2O quench installs D exactly where the Br (and hence MgBr) was, with no skeletal rearrangement.
Concept and Intuition
The rate of SN1 solvolysis depends on carbocation stability: tertiary > secondary > primary. Only a tertiary alkyl halide among the C5H11Br isomers would show clean first-order (unimolecular, carbocation-mediated) hydrolysis kinetics at an appreciable rate under mild conditions — this pins down X as the tertiary isomer, 2-bromo-2-methylbutane, CH3CH2C(CH3)2Br.
A Grignard reagent, R–MgBr, is formed by inserting Mg directly into the C–Br bond — the carbon skeleton is completely untouched; only the halogen is replaced by the −MgBr group. Quenching a Grignard reagent with D2O (heavy water) protonates (deuterates) the carbanion-like carbon, replacing −MgBr with −D — again with no skeletal rearrangement. This is a standard method for site-specific deuterium labeling.
Step-by-Step Solution
- C5H11Br hydrolyzing by first-order kinetics ⟹ SN1 mechanism ⟹ the carbon bearing Br must be tertiary (to form the stabilized 3° carbocation). So X=CH3CH2C(CH3)2Br (2-bromo-2-methylbutane).
- X+Mg (dry ether) → Grignard reagent CH3CH2C(CH3)2MgBr, formed at the same tertiary carbon (no rearrangement in Grignard formation).
- Grignard reagent +D2O → the −MgBr is replaced by −D: Y=CH3CH2C(CH3)2D. …
- AP EAPCET 2023Set ap-2023-05-23-FN1 markMCQQ.Observe the two statements Assertion (A): Benzyl chloride undergoes SN1 reaction readily while chlorobenzene does not. Reason (R): C-Cl bond in chlorobenzene has partial double bond due to resonance. The correct answer is (A) Both A and R are correct and R is correct explanation of A. (B) Both A and R are correct but R is not correct explanation of A. (C) A is correct but R is incorrect. (D) A is incorrect but R is correct.
›Reveal solutionSolution
Benzyl chloride's benzylic cation is resonance-stabilized, favouring SN1; chlorobenzene resists substitution because resonance gives its C–Cl bond partial double-bond character — and this resonance argument is exactly why chlorobenzene doesn't react, making R the correct explanation of A.
Concept and Intuition
In benzyl chloride (C6H5−CH2−Cl), the C–Cl bond is on an sp3 carbon attached to the aromatic ring, not directly on the ring itself. When this bond ionizes, it generates a benzylic carbocation, which is strongly stabilized by resonance delocalization of the positive charge into the aromatic ring — making SN1 ionization very favourable.
In chlorobenzene (C6H5−Cl), the chlorine is directly bonded to an sp2 ring carbon. Chlorine's lone pair can delocalize into the ring's π system through resonance, giving the C–Cl bond significant partial double-bond character. This makes the bond shorter and stronger than a typical C–Cl single bond, and also means the ring carbon bearing Cl is not electrophilic in the way needed for nucleophilic attack — so chlorobenzene resists both SN1 (no easy ionization, since breaking a partial-double bond and losing aromatic stabilization is costly) and SN2 (steric and electronic shielding from the ring).
Step-by-Step Solution
- Assess the assertion: benzyl chloride is SN1-reactive (true, benzylic cation is resonance-stabilized), chlorobenzene is not (true, due to bond strengthening). Assertion is correct.
- Assess the reason: chlorobenzene's C–Cl bond has partial double-bond character from resonance with the ring lone pair on Cl. Reason is correct (this is textbook resonance structure reasoning for haloarenes). …
🎓Unlock everything free for 14 days
- ✓Full step-by-step solutions
- ✓Concept-first explanations
- ✓Methods, shortcuts & mistakes
- ✓PYQ mapping + timed mock tests
Full access for 14 days. No credit card required.