Q.Aryl chlorides and bromides can be easily prepared by electrophilic substitution of arenes with chlorine and bromine respectively in the presence of Lewis acid catalysts. But why does preparation of aryl iodides requires presence of an oxidising agent?
🔒You're viewing a preview — the full solution, concept, methods & PYQ mapping are locked.
🔒 Start your 14-day free trial to unlock the full solution →Concept understanding — Nucleophilic Addition
Nucleophilic Addition – From Intuition to Precision
Imagine you have a molecule with a carbon–oxygen double bond — a carbonyl group (C=O). That oxygen is greedy for electrons; it pulls them away from carbon, leaving the carbon slightly positive (δ+) and the oxygen slightly negative (δ−). Now, if you bring a species that is rich in electrons (a nucleophile, meaning "nucleus-loving"), it will naturally be attracted to that electron-deficient carbon. The nucleophile attacks the carbon, the π bond breaks, and the oxygen picks up a proton (or some other electrophile) to become stable. That, in a nutshell, is nucleophilic addition.
The key idea: a nucleophile adds across a polar multiple bond (usually C=O or C≡N), breaking the π bond and forming two new sigma bonds.
The Precise Statement
Nucleophilic addition is a reaction in which a nucleophile (an electron-rich species) forms a sigma bond with an electrophilic carbon atom of a polar multiple bond (typically a carbonyl group, C=O, or a nitrile, C≡N), while the π bond breaks. The resulting intermediate then captures a proton (or another electrophile) to give a neutral product.
In general form:
RX2C=O+NuX−HX+RX2C(OH)Nu
The nucleophile (NuX−) attacks the carbonyl carbon; the oxygen becomes negatively charged; then a proton (HX+) from the medium attaches to the oxygen, yielding an alcohol.
Why It Happens – The Driving Force
The carbonyl carbon is electrophilic because:
- Oxygen is more electronegative than carbon, so the C=O bond is polarised: CXδ+=OXδ−.
- The π bond is weaker than a σ bond, so it can break relatively easily.
A nucleophile (like OHX−, CNX−, or NHX3) has a lone pair or a negative charge. It seeks positive centres. The attack forms a new σ bond, and the π electrons move entirely to oxygen, creating an alkoxide ion (RX2C−OX−). This intermediate is then quenched by a proton.
A common mistake: thinking the nucleophile attacks the oxygen. No — oxygen is already electron-rich; the nucleophile goes to the carbon because it is electron-deficient.
A Concrete Example – Addition of HCN to a Ketone
Take acetone (CHX3COCHX3) and hydrogen cyanide (HCN). In the presence of a base, CNX− (the nucleophile) attacks the carbonyl carbon:
CHX3COCHX3+CNX−CHX3C(OX−)(CN)CHX3
The alkoxide intermediate then picks up a proton from HCN (or from water) to give a cyanohydrin:
CHX3C(OX−)(CN)CHX3+HX+CHX3C(OH)(CN)CHX3
The product is acetone cyanohydrin. Notice: two new sigma bonds formed (C−CN and O−H), and the π bond is gone.
What Makes a Good Nucleophile?
Strong nucleophiles are usually negatively charged or have lone pairs:
- OHX−, CNX−, NHX2X−, CHX3OX−, HX− (from hydride reagents like NaBHX4 or LiAlHX4)
- Neutral but polarisable: NHX3, HX2O (weaker, but can add under acidic conditions) …
Why this formula?
Nucleophilic Addition: Why the Mechanism Works the Way It Does
Let's build this from first principles — understanding why nucleophilic addition happens, not just memorising the steps.
1. The Core Problem: Why Does Addition Happen at All?
A carbonyl group (C=O) has a polarised double bond:
- Oxygen is more electronegative than carbon → it pulls electron density toward itself.
- This creates a partial positive charge on carbon (δ+) and a partial negative charge on oxygen (δ−).
CXδ+=OXδ−
Key insight: The carbon is electron-deficient — it wants electrons. A nucleophile (Nu⁻) is electron-rich — it wants to give electrons. This is a natural match.
2. The Two-Step Mechanism (Why Two Steps?)
Step 1: Nucleophilic Attack (Slow, Rate-Determining)
The nucleophile donates its lone pair to the electrophilic carbonyl carbon.
NuX−+C=O[Nu−C−O]X−
Why this happens:
- The π bond between C and O breaks — the electrons move entirely to oxygen.
- Oxygen now has a full negative charge (alkoxide ion).
- The carbon changes from sp2 (trigonal planar) to sp3 (tetrahedral).
This step is slow because the π bond must break — it requires energy.
Step 2: Protonation (Fast)
The negatively charged oxygen picks up a proton (HX+) from the solvent or acid.
[Nu−C−O]X−+HX+Nu−C−OH
Why this happens:
- The alkoxide ion is a strong base — it wants to neutralise its charge.
- Protonation gives a stable neutral alcohol product.
3. The Key Formula: Rate Law Derivation
For a general nucleophilic addition:
NuX−+RX2C=Okproducts
The rate law comes from the slow step (Step 1):
Rate=k[Nu−][RX2C=O]
Why this form?
- The reaction is bimolecular — two species must collide with correct orientation.
- Doubling either concentration doubles the rate (first order in each).
- This is second order overall.
Exam tip: This is why nucleophilic addition is often called addition-elimination when followed by loss of a leaving group (like in acyl substitution), but here it's just addition.
4. Why the Tetrahedral Intermediate Forms (And Why It's Unstable)
The intermediate is tetrahedral (sp3 hybridised carbon):
- Bond angles: ~109.5°
- Four groups around carbon: Nu, R, R', O⁻
Why it's unstable:
- The negative charge on oxygen is high-energy.
- The tetrahedral geometry is sterically crowded (especially with bulky R groups).
- The intermediate collapses quickly — either back to starting materials or forward to product. …
The key idea is that iodination is reversible because I2 is a very weak electrophile, and the HI byproduct reduces the product back to the arene.
Reasoning:
- In electrophilic aromatic substitution, I2 is far less reactive than Cl2 or Br2 — it cannot polarise sufficiently to act as an electrophile without help.
- Even if a small amount of iodination occurs, the HI produced is a strong reducing agent that rapidly reduces the aryl iodide back to the hydrocarbon. …
Iodination of an arene is reversible: ArH+I2⇌ArI+HI. The HI formed is a good reducing agent — it converts the aryl iodide back to the arene — so on its own the reaction never accumulates product. An oxidising agent (HIO₄, or HNO₃) is added to oxidise the HI away, driving the equilibrium forward. That is why aryl iodides need an oxidising agent while aryl chlorides and bromides do not.
Why the usual method works for Cl₂ and Br₂
Chlorination and bromination of arenes proceed cleanly with just a Lewis acid catalyst (FeCl₃, AlCl₃), which polarises the halogen molecule into a strong electrophile:
Cl2+FeCl3→Clδ+⋯FeCl4δ−
The HCl or HBr released as byproduct does not attack the aryl halide product, so these reactions are effectively irreversible — no extra reagent is needed.
The problem with iodine: the reaction is reversible
Iodination is different in one decisive way. The reaction sits in an equilibrium:
ArH+I2⇌ArI+HI
The HI byproduct is a good reducing agent: it reduces the aryl iodide back to the parent arene (regenerating I₂), pulling the equilibrium backwards. It also doesn't help that I₂ is the weakest electrophile of the halogens, which makes the forward reaction sluggish to begin with — but the equilibrium is the core problem: even the product that does form is destroyed by the HI accumulating in the mixture.
The solution: oxidise away the HI
An oxidising agent — HIO₄ (periodic acid) is the one NCERT names; HNO₃ also works — is added to oxidise the HI as it forms, removing it from the equilibrium:
ArH+I2⇌ArI+HI
The oxidising agent removes HI (for example, 2HI+H2O2→I2+2H2O), so by Le Chatelier's principle the equilibrium shifts to the right and the aryl iodide accumulates. …
Concept: Reversibility of Iodination and Removal of HI
The key idea is that iodination of an arene is a reversible reaction, and the HI byproduct drives it backwards unless it is removed by oxidation.
Method: Equilibrium Analysis of Arene Halogenation
Why the problem arises:
- Chlorination and bromination (with a Lewis acid such as FeCl3/AlCl3) are effectively irreversible — the HCl/HBr byproduct does not attack the product, so no extra reagent is needed.
- Iodination is reversible:
ArH+I2⇌ArI+HI
The HI formed is a good reducing agent — it reduces the aryl iodide back to the arene (regenerating I2). I2 is also the weakest electrophile of the halogens, so the forward reaction is slow to begin with.
The solution:
Add an oxidising agent — HIO₄ (the one NCERT names) or HNO3 — to oxidise the HI byproduct as it forms, removing it from the equilibrium.
Step-by-Step Reasoning
-
Electrophilic attack:
The iodine (polarised/activated in the reaction mixture) attacks the benzene ring, forming a sigma complex.
-
Deprotonation:
Loss of H+ from the sigma complex gives the aryl iodide and HI.
-
The reverse reaction (the problem):
The accumulated HI reduces ArI back to ArH, so the equilibrium yields little product.
-
Oxidation of HI (the fix):
The oxidising agent converts HI back to I2 — for example: …
Here’s a breakdown of the common mistakes students make on this concept, along with how to avoid each.
The Core Concept (Why the Question Exists)
The question tests your understanding of reactivity trends in electrophilic aromatic substitution (EAS) and the redox chemistry of halogens.
- For Cl₂ and Br₂: The halogen molecule is already a strong enough electrophile (when activated by a Lewis acid like FeCl₃ or AlCl₃) to attack the benzene ring.
- For I₂: Iodine is a much weaker electrophile than chlorine or bromine, and the reaction produces HI as a byproduct. HI is a good reducing agent that reduces the aryl iodide back to the arene (regenerating I₂), reversing the reaction.
The fix: An oxidizing agent (like HIO₄ or HNO₃) oxidizes the HI (the byproduct) back into I₂, preventing the reverse reaction and pushing the equilibrium forward.
Common Mistake #1: Confusing "Oxidizing Agent" with "Catalyst"
The Mistake:
Students say: "The oxidizing agent is needed because iodine is a weaker electrophile, so we need a stronger catalyst." They treat the oxidizing agent as if it’s just another Lewis acid catalyst.
Why it’s wrong:
A Lewis acid catalyst (like FeCl₃) activates the halogen by polarizing it (making it more electrophilic). An oxidizing agent does not activate iodine directly. Instead, it removes the byproduct (HI) that would otherwise destroy the aryl iodide product.
How to Avoid:
- Remember the byproduct: For every I₂ that reacts, one HI is produced. HI is a strong reducing agent.
- Trace the electron flow: HI + [O] → I₂ + H₂O. The oxidizing agent regenerates the starting material (I₂), not just activates it.
- Exam tip: If a question asks "Why is an oxidizing agent needed?" your answer must mention preventing the reduction of the aryl iodide back to the arene by HI. Do not just say "to make iodine more reactive."
Common Mistake #2: Thinking the Oxidizing Agent Makes Iodine "More Electrophilic"
The Mistake:
Students write: "The oxidizing agent increases the electrophilicity of iodine."
Why it’s wrong:
Oxidizing agents do not directly increase the positive charge or polarity of I₂. They work indirectly by removing HI. The actual electrophile in the reaction is still I₂ (or I⁺ generated in situ, but that’s a separate mechanism). The oxidizing agent doesn’t touch the I₂ molecule itself.
How to Avoid:
- Use precise language: Say "The oxidizing agent oxidizes the HI byproduct back to I₂, preventing the reverse reaction."
- Draw the equilibrium: Show the reversible arrow: ArH+I2⇌ArI+HI The oxidizing agent shifts the equilibrium to the right by removing HI.
Common Mistake #3: Forgetting the Role of HI as a Reducing Agent
The Mistake:
Students say: "HI is a strong acid, so it protonates the benzene ring and stops the reaction."
Why it’s wrong:
HI is indeed an acid, but the real problem is that it reduces the aryl iodide back to the arene. The reaction is reversible, and HI is the reducing agent that drives it backward. Protonation of the ring is not the main issue here (though it can happen, it’s secondary).
How to Avoid:
- Remember the equilibrium: ArH + I₂ ⇌ ArI + HI. HI provides the electrons that reduce the ArI product back to ArH. …
Showing the 12 most recent of 33 on this concept.
- AP EAPCET 2026Set eng-2026-05-13-FN1 markMCQQ.What are X and Y respectively in the following set of reactions? C6H5CNXA , C6H5CNYB ; A,BH+C6H5−C(CH3)=N−CH2C6H5 (A) H2/cat ; (CH3)2Cd (B) Na/C2H5OH ; CH3MgBr,H2O (C) DIBAL−H,H2O ; CH3MgBr,H2O (D) LiAlH4,H2O ; (CH3)2Cd
›Reveal solutionSolution
The final imine comes from condensing acetophenone with benzylamine. Benzonitrile is reduced by Na/C₂H₅OH to give benzylamine (A), and reacted with CH₃MgBr then hydrolysed to give acetophenone (B); acid catalyses their condensation to the imine.
Concept and Intuition
Nitriles (R−C≡N) are versatile precursors to both amines and ketones depending on the reagent used:
- Full reduction of a nitrile (e.g. dissolving-metal reduction with sodium in ethanol, or catalytic H2/LiAlH₄) adds four hydrogens across the triple bond, converting −C≡N directly into a primary amine, −CH2NH2.
- Grignard addition to a nitrile gives a metallated imine salt, R−C(=NMgBr)−R′, which on aqueous acidic hydrolysis (H2O/H3O+) is hydrolysed straight through to the ketone R−CO−R′ (releasing ammonia).
A ketone and a primary amine then condense under acid catalysis (loss of water) to form an imine (C=N−R) — exactly the target product here.
Step-by-Step Solution
- Target imine: C6H5−C(CH3)=N−CH2C6H5. Break the C=N bond conceptually: the C6H5−C(CH3)= portion comes from a ketone, C6H5−CO−CH3 (acetophenone); the =N−CH2C6H5 portion comes from a primary amine, C6H5CH2−NH2 (benzylamine).
- Route to A (benzylamine) from C6H5CN: full reduction of the nitrile using sodium and ethanol (a dissolving-metal / nascent-hydrogen reduction) directly gives C6H5CH2NH2. …
- AP EAPCET 2026Set eng-2026-05-18-AN1 markMCQQ.Which of the following are the correct statements about D-glucose? I. It forms oxime with hydroxyl amine II. It forms addition product with NaHSO3 III. It forms cyanohydrin with HCN IV. It forms saccharic acid with bromine water The correct answer is (A) II & III only (B) I & III only (C) II & IV only (D) III & IV only
›Reveal solutionSolution
Glucose's structural-elucidation reactions: it does form an oxime and a cyanohydrin (confirming the aldehyde group exists in equilibrium with the ring form), but it does NOT form a bisulphite addition product, and bromine water oxidises it only to gluconic acid, not saccharic acid.
Concept and Intuition
Even though glucose predominantly exists as a cyclic hemiacetal, a small equilibrium amount of the open-chain aldehyde form is always present. This small amount is enough to react with small, reactive nucleophiles like HCN and NH2OH (whose reactions pull the equilibrium forward), giving cyanohydrin and oxime respectively. But bulkier/other characteristic aldehyde tests (like the bisulphite addition with NaHSO3, or Schiff's test) fail for glucose — this mismatch between "should behave like an aldehyde" and "doesn't give all aldehyde tests" was itself key historical evidence for glucose's cyclic hemiacetal structure.
Step-by-Step Solution
- Statement I: Glucose + NH2OH→ glucose oxime (via the open-chain −CHO). This reaction is well-documented and used as evidence for the carbonyl group. True.
- Statement II: Glucose does not give the NaHSO3 addition product — a well-known NCERT fact distinguishing glucose from typical aldehydes despite having a carbonyl group in its open form. False.
- Statement III: Glucose + HCN → glucose cyanohydrin (nucleophilic addition at the carbonyl carbon), again evidence for the −CHO group. True. …
- AP EAPCET 2026Set ap-2026-05-20-AN1 markMCQQ.What are B and C respectively in the given sequence of reactions? Bromocyclohexane Mg ∣ dryether A; A CH3CH2OH B; A (i) HCHO(ii) H3O+ C (A) Cyclohexanol , Cyclohexylmethanol (B) Cyclohexane , Cyclohexylmethanol (C) Cyclohexane , Ethylcyclohexane (D) C2H6 , Ethoxycyclohexane (cyclohexane ring bearing an −OCH2CH3 substituent)
›Reveal solutionSolution
Tests Grignard reagent formation and its two contrasting fates: quenching by an acidic O-H gives the alkane; addition to formaldehyde gives a one-carbon-homologated primary alcohol.
Concept and Intuition
A Grignard reagent (R-MgX) is a strong carbanion-like nucleophile and an extremely strong base. It reacts instantly with any acidic proton (even a weak one like the O-H of an alcohol) faster than it can do anything else — this simply protonates the carbanion and regenerates R-H. Only when there is no acidic proton available (e.g. with a carbonyl compound like HCHO) does the Grignard get the chance to act as a nucleophile and add across the C=O bond, building a new C-C bond and a new alcohol after the acidic workup.
Step-by-Step Solution
- Bromocyclohexane + Mg in dry ether → cyclohexylmagnesium bromide (A), by oxidative insertion of Mg into the C-Br bond.
- A + CH3CH2OH: the O-H proton of ethanol quenches the Grignard immediately (acid-base reaction), giving cyclohexane (B) + CH3CH2OMgBr. No new C-C bond forms. …
- AP EAPCET 2025Set eng-2025-05-21-FN1 markMCQQ.Consider the following sequence of reactions Isopropyl benzene O2XH+H2O phenol +Z The incorrect statement about z is (A) z gives yellow precipitate of CHI3 with NaOH + I2 solution (B) z gives isopropyl alcohol on reduction with H2 in the presence of Pd catalyst (C) z on reaction with CH3MgBr followed by hydrolysis gives 2° alcohol (D) z does not give positive test with Fehling's reagent
›Reveal solutionSolution
Tests the cumene-to-phenol process and acetone's reactions; the answer is (C), since a ketone + Grignard reagent gives a tertiary (not secondary) alcohol.
Concept and Intuition
The industrial cumene process converts isopropylbenzene to phenol in two steps: air oxidation to cumene hydroperoxide (X), followed by acid-catalysed hydrolytic rearrangement to give phenol and acetone (Z) as co-product:
CumeneO2Cumene hydroperoxide (X)H+H2OPhenol+Acetone (Z)
So Z = acetone, (CH3)2C=O, a ketone. To find the incorrect statement, we check each of acetone's known reactions:
- Iodoform test: acetone has a CH3−CO− group, so it gives a positive iodoform test (yellow CHI3 precipitate) with NaOH/I2 — true.
- Catalytic hydrogenation: ketones are reduced to secondary alcohols by H2/Pd; acetone → isopropyl (2°) alcohol — true.
- Grignard addition: a ketone reacting with a Grignard reagent, upon hydrolysis, gives a tertiary alcohol (since the ketone carbon already bears two alkyl/aryl groups, and the Grignard's R group adds a third) — acetone + CH3MgBr → 2-methylpropan-2-ol (tert-butanol), a tertiary alcohol, NOT secondary. This makes the given statement false.
- Fehling's test: ketones (other than special reducing sugars) generally do not give a positive Fehling's test since it requires an oxidisable aldehyde-type group; acetone gives a negative result — true.
Step-by-Step Solution
- Deduce Z = acetone from the cumene process.
- Check (A): acetone + iodoform test → positive (true statement, so not the answer). …
- AP EAPCET 2025Set eng-2025-05-22-FN1 markMCQQ.The correct statement about the product of the following reaction is CH3CHO(i) C2H5MgBr(ii) H2Oproduct (A) It undergoes dehydration with 20% H3PO4 at 358 K (B) It gives ketone on oxidation with CrO3 (C) It does not give positive iodoform test (D) It is a vinylic alcohol
›Reveal solutionSolution
Tests identifying the Grignard-addition product (a secondary alcohol) and its correct chemical behaviour on oxidation, dehydration, and the iodoform test.
Concept and Intuition
An aldehyde plus a Grignard reagent adds the alkyl group to the carbonyl carbon; after aqueous workup this gives a SECONDARY alcohol (aldehyde + RMgX → 2° alcohol; ketone + RMgX → 3° alcohol). Once the product's structure is nailed down, each option can be checked against that structure's real chemistry.
Step-by-Step Solution
- CH3CHO+C2H5MgBr→CH3−CH(OMgBr)−C2H5; hydrolysis with H2O gives CH3−CH(OH)−CH2CH3 = butan-2-ol, a secondary alcohol.
- Secondary alcohols are oxidised by chromium(VI) reagents such as CrO3 to ketones (here, butan-2-one, CH3COCH2CH3) — this statement is straightforwardly true.
- Checking (C): butan-2-ol has the fragment CH3−CH(OH)− (a methyl group directly on the carbinol carbon) — this is exactly the structural requirement for a POSITIVE iodoform test, so the claim that it does "not" give one is false. …
- AP EAPCET 2025Set ap-2025-05-20-FN1 markMCQQ.Glucose on reaction with HCN forms a compound 'A'. Acid hydrolysis of A gives B. The molecular formula of B is (A) C7H14O8 (B) C6H14O7 (C) C7H14O6 (D) C7H12O5
›Reveal solutionSolution
Glucose's cyanohydrin (from HCN addition) hydrolyses its nitrile group to a carboxylic acid, giving B with molecular formula C7H14O8.
Concept and Intuition
This is the classic 'glucose chain-length extension' sequence used historically (Kiliani–Fischer style logic) to probe glucose's structure: HCN adds across the free aldehyde group to form a cyanohydrin (adding one carbon), and subsequent acid hydrolysis converts the newly added nitrile into a carboxylic acid group, adding two oxygens (from two water molecules) while releasing ammonia.
Step-by-Step Solution
- Glucose (open-chain aldose): C6H12O6, with a free −CHO group.
- Reaction with HCN: nucleophilic addition of HCN across the aldehyde carbonyl adds the entire HCN unit to the molecule, forming cyanohydrin A: A=C6H12O6+HCN=C7H13NO6.
- Acid hydrolysis of the nitrile group in A: R−C≡N+2H2O→R−COOH+NH3.
- Applying atom balance: C7H13NO6+2H2O→B+NH3. …
- AP EAPCET 2025Set eng-2025-05-23-AN1 markMCQQ.An alkyl bromide X(C5H11Br) undergoes hydrolysis in a two step mechanism. X is converted to Grignard reagent and then reacted with CO2 in dry ether followed by acidification gave Y. What is Y? (A) (CH3)2CH−CH2−CH2−COOH (skeletal structure: a branched chain with a methyl branch, then two CH2 groups, ending in COOH) (B) CH3−CH2−CH2−CH2−CH2−COOH (skeletal structure: an unbranched chain ending in COOH) (C) CH3−CH2−CH(CH3)−CH2−COOH (skeletal structure: a chain with a methyl branch nearer the COOH end) (D) (CH3)3C−CH2−COOH (skeletal structure: a gem-dimethyl branched carbon adjacent to CH2COOH)
›Reveal solutionSolution
This tests the difference between a rearranging SN1 hydrolysis and a rearrangement-free Grignard carboxylation of the same neopentyl-type halide. Answer: (CH3)3C−CH2−COOH (D).
Concept and Intuition
The phrase "hydrolysis in a two-step mechanism" is the giveaway for identifying X: among the C5H11Br isomers, neopentyl bromide, (CH3)3C−CH2−Br, is the standard example of a primary halide that is nevertheless forced into an SN1 (two-step: ionisation, then nucleophilic capture) pathway on hydrolysis, because the bulky tert-butyl group next to the leaving carbon blocks backside SN2 attack. Once ionised, the resulting primary carbocation is so unstable that it instantly rearranges by a 1,2-methyl shift to a tertiary carbocation, so hydrolysis of X actually gives a rearranged tertiary alcohol.
But the question does not hydrolyse X directly — it first converts X to its Grignard reagent. Formation of a Grignard reagent (insertion of Mg metal into the C–Br bond) is a concerted/radical-pair process at the original carbon and does not proceed through a free carbocation, so no rearrangement occurs; the alkyl skeleton of the Grignard reagent is identical to that of X. Reaction of a Grignard reagent with CO2 (dry ether) forms a magnesium carboxylate at that same carbon, and acidification liberates the carboxylic acid — effectively inserting one new carbon (COOH) onto the unrearranged skeleton.
Step-by-Step Solution
- Identify X: C5H11Br whose hydrolysis is a two-step (carbocation, rearranging) process ⇒ neopentyl bromide, (CH3)3C−CH2−Br.
- Form the Grignard reagent: (CH3)3C−CH2−Br+Mgdry ether(CH3)3C−CH2−MgBr — no rearrangement, skeleton preserved. …
- AP EAPCET 2025Set eng-2025-05-23-AN1 markMCQQ.What is A in the following reaction? CH3−CH=CH−CH2−CH2−CN1) AlH(i−Bu)22) H2O(A) (A) CH3−CH=CH−CH2−CH2−CH2−NH2 (B) CH3−CH2−CH2−CH2−CH2−CH2−NH2 (C) CH3−CH=CH−CH2−CH2−CHO (D) CH3−CH2−CH2−CH2−CH2−CHO
›Reveal solutionSolution
DIBAL-H stops nitrile reduction at the aldehyde stage (via an imine intermediate hydrolysed on work-up) and leaves alkenes untouched. Answer: (C).
Concept and Intuition
DIBAL-H, AlH(i−Bu)2, delivers a single hydride. When it attacks a nitrile (R−C≡N), the hydride adds once to give a stable metalated imine (an aluminium–nitrogen chelate) that does not collapse further at low temperature because the aluminium coordinates and "locks" the intermediate. Aqueous hydrolysis of this imine/enamine–aluminium complex during work-up then releases the corresponding aldehyde, R−CHO. This is the standard method for stopping a nitrile reduction one oxidation level short of the amine that full reduction (e.g. with LiAlH4 or catalytic hydrogenation) would give. DIBAL-H is also a mild, chemoselective hydride source that does not reduce ordinary isolated alkene double bonds, so the CH3−CH=CH− portion of the substrate is unaffected.
Step-by-Step Solution
- Substrate: CH3−CH=CH−CH2−CH2−C≡N.
- Step 1, AlH(i−Bu)2 (1 equivalent, controlled conditions): hydride adds once to the nitrile carbon, forming the imine–aluminium adduct CH3−CH=CH−CH2−CH2−CH=NAl(i−Bu)2 (conceptually); the alkene is untouched throughout. …
- AP EAPCET 2025Set eng-2025-05-24-FN1 markMCQQ.What are X and Y in the following reaction sequence? C6H5N2+Cl−XC6H5CN(i) CH3MgBr (ii) H2OY (A) KCN; C6H5COCH3 (B) KCN; C6H5C(OH)(CH3)2 (C) CuCN|KCN; C6H5CH(OH)CH3 (D) CuCN|KCN; C6H5COCH3
›Reveal solutionSolution
Diazonium → nitrile is a Sandmeyer reaction (CuCN|KCN); a Grignard reagent adding to a nitrile and then hydrolysing gives a ketone, not an alcohol — so Y=C6H5COCH3.
Concept and Intuition
Sandmeyer-type reactions replace the −N2+ group of a diazonium salt with a nucleophile delivered via a copper(I) catalyst; with cyanide (as CuCN, generated from CuCN|KCN) this installs −CN directly onto the aromatic ring, retaining the ring intact — a route to aryl nitriles that plain KCN alone (without Cu(I)) does not reliably achieve on an aryl diazonium.
A nitrile's carbon is only singly electrophilic toward a Grignard: one equivalent of RMgX adds across the C≡N triple bond to give a metalated imine (an imino-magnesium salt). This does not react further with a second equivalent of Grignard (unlike an ester, which can go on to a tertiary alcohol) because the intermediate is stable to the reaction conditions until workup. Aqueous acidic hydrolysis of that imine salt then gives a ketone, not an alcohol.
Step-by-Step Solution
- C6H5N2+Cl−CuCN|KCNC6H5CN — Sandmeyer reaction; X=CuCN|KCN. …
- AP EAPCET 2025Set eng-2025-05-26-FN1 markMCQQ.What are X and Y respectively in the following reaction sequence? (g = వాయువు, dil = విలీన) 4-O2N-C6H4-CHOC2H5OHHCl(g)Xdil. HClY (A) X=4-O2NC6H4CH(OC2H5)2 (diethyl acetal) ; Y=4-O2NC6H4CHO (the aldehyde) (B) X=4-O2NC6H4CH(OH)(OC2H5) (hemiacetal, OH and OC2H5 on the same carbon) ; Y=4-O2NC6H4CH(OC2H5)2 (diethyl acetal) (C) X=4-O2NC6H4CH(OC2H5)2 (diethyl acetal) ; Y=4-O2NC6H4CH(OH)(OC2H5) (hemiacetal) (D) X=4-O2NC6H4CH(OH)(OC2H5) (hemiacetal) ; Y=4-O2NC6H4CH2OH (the benzylic alcohol)
›Reveal solutionSolution
This tests acetal formation (protection) and its acid hydrolysis (deprotection) of an aldehyde. The answer is (A): X is the diethyl acetal, and Y is the regenerated aldehyde.
Concept and Intuition
Aldehydes react with excess alcohol under anhydrous acidic conditions (e.g. dry HCl gas) to form acetals, RCH(OR′)2, via a hemiacetal intermediate that is not isolated under these forcing (excess alcohol, anhydrous acid, often with removal of water) conditions — the reaction proceeds essentially straight through to the acetal. Acetals are a classic protecting group for the carbonyl: they are stable under basic and neutral conditions but are cleaved by aqueous acid, regenerating the original aldehyde plus the alcohol (acetal hydrolysis is simply the reverse of acetal formation, driven by the presence of water).
Step-by-Step Solution
- Step 1 (aldehyde → X): 4-nitrobenzaldehyde reacts with excess ethanol in the presence of dry HCl gas. Under these anhydrous, acid-catalyzed, excess-alcohol conditions, the aldehyde is converted all the way to its diethyl acetal: X=4-O2NC6H4CH(OC2H5)2.
- Step 2 (X → Y): treating the acetal X with dilute (aqueous) HCl reverses acetal formation — water attacks and the acetal hydrolyzes back to the free aldehyde, releasing 2 equivalents of ethanol: Y=4-O2NC6H4CHO, i.e., the original 4-nitrobenzaldehyde is regenerated.
- So overall this sequence is simply 'protect the aldehyde as its acetal, then deprotect it back' — X is the acetal, Y is the aldehyde again. …
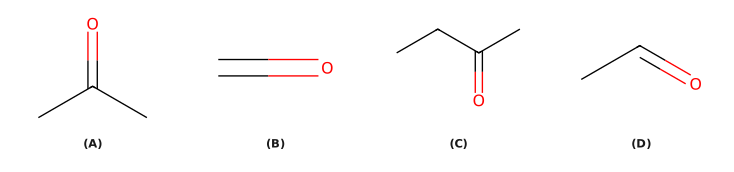
- AP EAPCET 2025Set eng-2025-05-27-FN1 markMCQQ.An alcohol, X (C5H12O) in the presence of Cu/573K gives Y (C5H10). The reactants required for the preparation of X are (A) acetone, CH3COCH3 (drawn as a skeletal structure with the carbonyl O at the bottom), and C2H5MgBr (B) HCHO, (CH3)3CMgBr (C) [FIGURE] (a five-carbon ketone: an ethyl group and a branched, isopropyl-type group attached to a central carbonyl carbon), and C2H5MgBr (D) acetone, CH3COCH3 (drawn as a skeletal structure with the carbonyl O at the top), and (CH3)2CHMgBr
›Reveal solutionSolution
X loses H2O (not H2) over Cu/573K to give C5H10, which is the signature of a tertiary alcohol undergoing dehydration; only acetone + C2H5MgBr builds a tertiary C5H12O alcohol. Answer: (A).
Concept and Intuition
Over copper at 573 K, alcohols react differently depending on their class:
- 1° alcohols dehydrogenate (lose H2) to give aldehydes.
- 2° alcohols dehydrogenate (lose H2) to give ketones.
- 3° alcohols have no H on the carbinol carbon to remove for dehydrogenation, so instead they undergo dehydration (lose H2O) to give alkenes.
Here, X (C5H12O) gives Y (C5H10). Checking the atom balance: C5H12O−H2O=C5H10 — mass balance fits loss of water, confirming X must be a tertiary alcohol.
A tertiary alcohol is made by adding a Grignard reagent to a ketone (not an aldehyde, which gives a secondary alcohol): R2C=O+R′MgX→R2C(OH)R′. We need the resulting alcohol to have exactly 5 carbons.
Step-by-Step Solution
- Determine X's alcohol class from the reaction Cu/573K: since Y=C5H10 (loss of H2O from X, not H2), X must be a tertiary alcohol giving an alkene by dehydration.
- Test option (A): acetone (CH3)2C=O (3 carbons) + C2H5MgBr (2 carbons) → (CH3)2C(OH)C2H5 = 2-methyl-2-butanol, a tertiary alcohol with 5 carbons, formula C5H12O. ✓ Matches on both carbon count and alcohol class.
- Test option (D): acetone (3C) + (CH3)2CHMgBr (isopropyl, 3C) → (CH3)2C(OH)CH(CH3)2 = 2,3-dimethyl-2-butanol, which has 6 carbons (C6H14O) — too many carbons, rejected. …
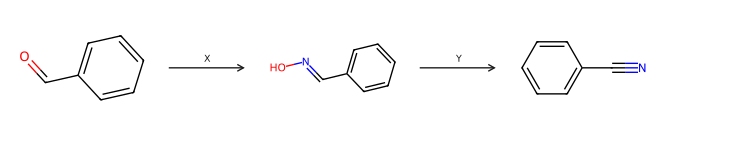
- AP EAPCET 2024Set eng-2024-05-18-FN1 markMCQQ.What are X and Y respectively in the following reaction sequence? [FIGURE] (benzaldehyde X Oxime Y benzonitrile) (A) NH2NH2, C6H5SO2Cl / Pyridine (B) NH2NH2, (CH3CO)2O (C) NH2OH, C6H5SO2Cl / Pyridine (D) NH2OH, (CH3CO)2O
›Reveal solutionSolution
Aldehyde → oxime needs hydroxylamine; oxime → nitrile needs a dehydrating agent, classically acetic anhydride — X, Y = NH2OH, (CH3CO)2O, option (D).
Concept and Intuition
This is the standard two-step laboratory route from an aldehyde to the one-carbon-longer nitrile (useful because nitriles hydrolyse further to carboxylic acids, or reduce to amines):
- Oxime formation: aldehydes and ketones condense with hydroxylamine (NH2OH) to form oximes (C=N−OH), releasing water. This is a standard carbonyl condensation, analogous to hydrazone/semicarbazone formation.
- Dehydration of the oxime to a nitrile: removing a molecule of water from the oxime (−CH=N−OH→−C≡N+H2O) requires a dehydrating agent. The reagent commonly cited in NCERT-level organic chemistry for this step is acetic anhydride, (CH3CO)2O (it also works with reagents like P2O5, SOCl2, or tosyl chloride/pyridine in more advanced contexts, but the textbook-standard answer is acetic anhydride).
Step-by-Step Solution
- Benzaldehyde (C6H5CHO) reacts with NH2OH to give benzaldoxime, C6H5CH=N−OH: this identifies X = NH2OH. …
🎓Unlock everything free for 14 days
- ✓Full step-by-step solutions
- ✓Concept-first explanations
- ✓Methods, shortcuts & mistakes
- ✓PYQ mapping + timed mock tests
Full access for 14 days. No credit card required.